TMT vs Label Free
Tandem mass tag (TMT) fits when relative protein discovery can use one multiplex labeling kit and same-run reporter-ion comparison. Label-free quantification (LFQ) fits when samples stay as independent unlabeled injections, enrollment may continue, or labeling chemistry is impractical. Both usually report relative abundance of the same protein across conditions. A concentration report uses matched standards and a calibration model. TMT mixes labeled channels into one liquid chromatography-tandem mass spectrometry (LC-MS/MS) path and reads reporter ions after fragmentation (Thompson et al. 2003). LFQ aligns and normalizes separately acquired runs, commonly by MS1 peak-area methods such as MaxLFQ or by DIA fragment chromatograms (Cox et al. 2014).
For defined samples and comparison groups, see the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service and Label-Free Quantitative Proteomics Service for project-specific feasibility and analysis planning.
TMT vs Label-Free at a Glance
| TMT | Label-free (DDA or DIA) | |
|---|---|---|
| Sample handling | Amine labeling, then pool channels | No isotopic or isobaric tags |
| Acquisition path | Shared LC-MS/MS run for the plex | Independent injections per sample |
| Quantitative signal | Reporter-ion intensities | Aligned peak areas or fragment chromatograms |
| Enrollment | Closed after mixing | Later compatible injections can be added under the same plan |
| Main design point | Ratio compression from co-isolation | Run-to-run drift, missing values, batch effects |
| What is reported | Relative ratios inside one plex |
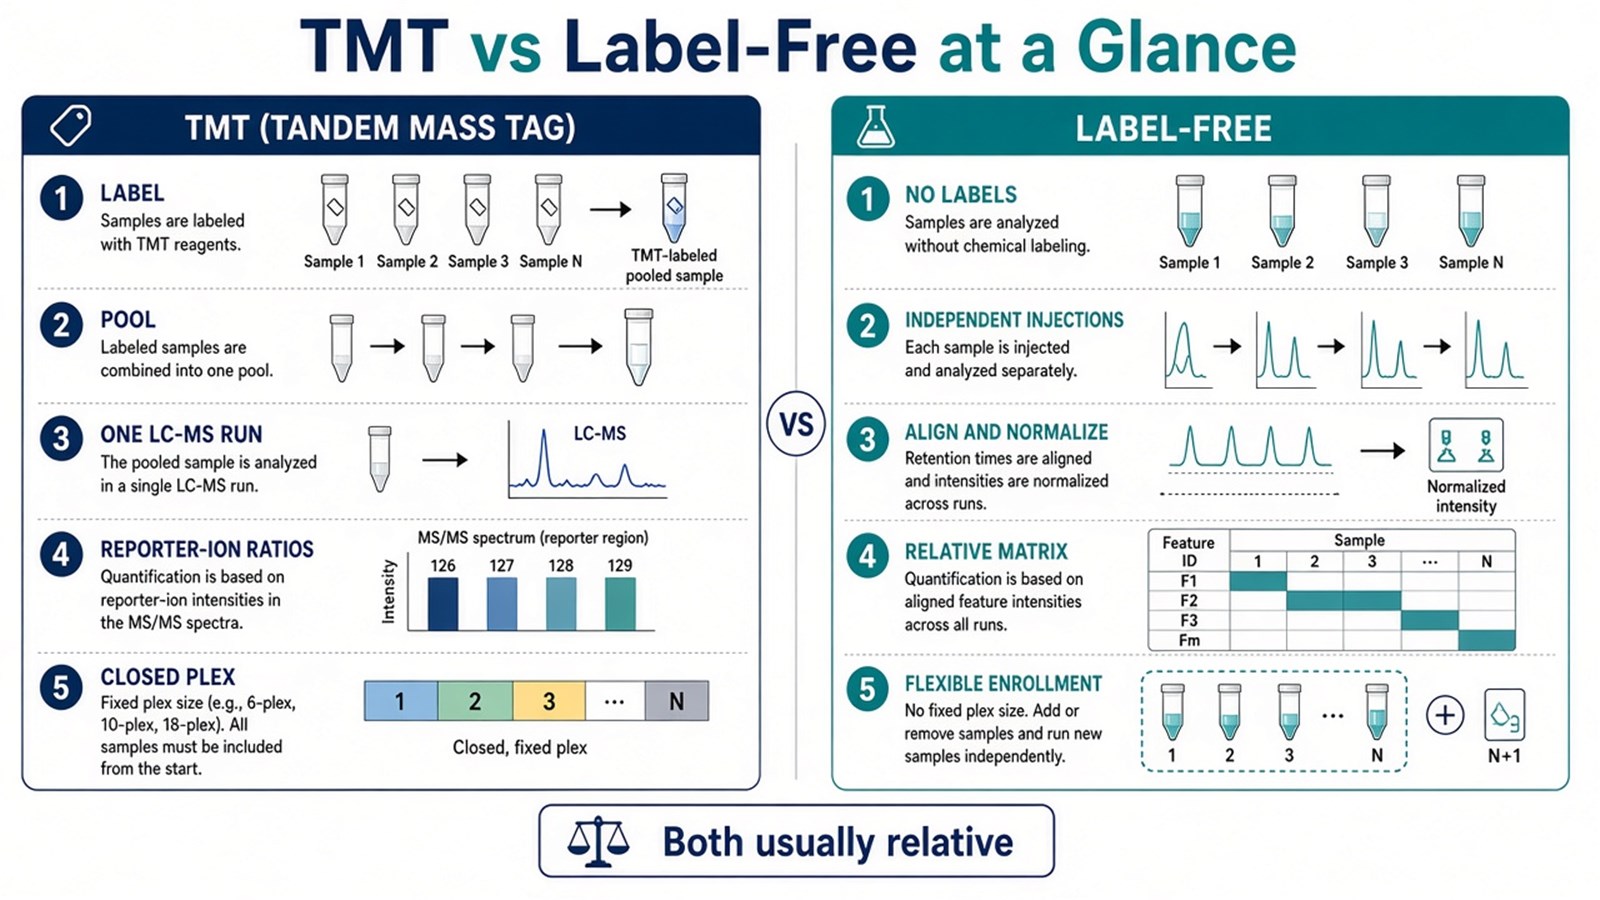
Figure 1. TMT compares channels inside one pooled run. Label-free compares independently acquired runs after alignment and normalization.
Key Trade-Offs That Drive Selection
1. Same-run comparison versus independent enrollment
TMT reduces injection-to-injection drift for channels that share chromatography and precursor selection. The plex size is set by the kit, and mixing closes the channel list. LFQ keeps samples independent, so later compatible injections can be added, and every run carries its own loading and instrument history.
2. Ratio compression versus run-to-run variability
TMT MS2 reporters can underestimate true fold changes when co-isolated peptides dilute channel differences (Ting et al. 2011). Fractionation and SPS-MS3 can reduce compression on compatible instruments. LFQ avoids reporter mixing and uses declared normalization, run-order control, and missing-value review.
3. Chemistry cost versus operational simplicity
TMT adds reagent cost, amine-buffer constraints, and labeling QC. LFQ omits labeling steps and often lowers per-sample chemistry burden, then spends that budget on chromatography stability and analysis discipline.
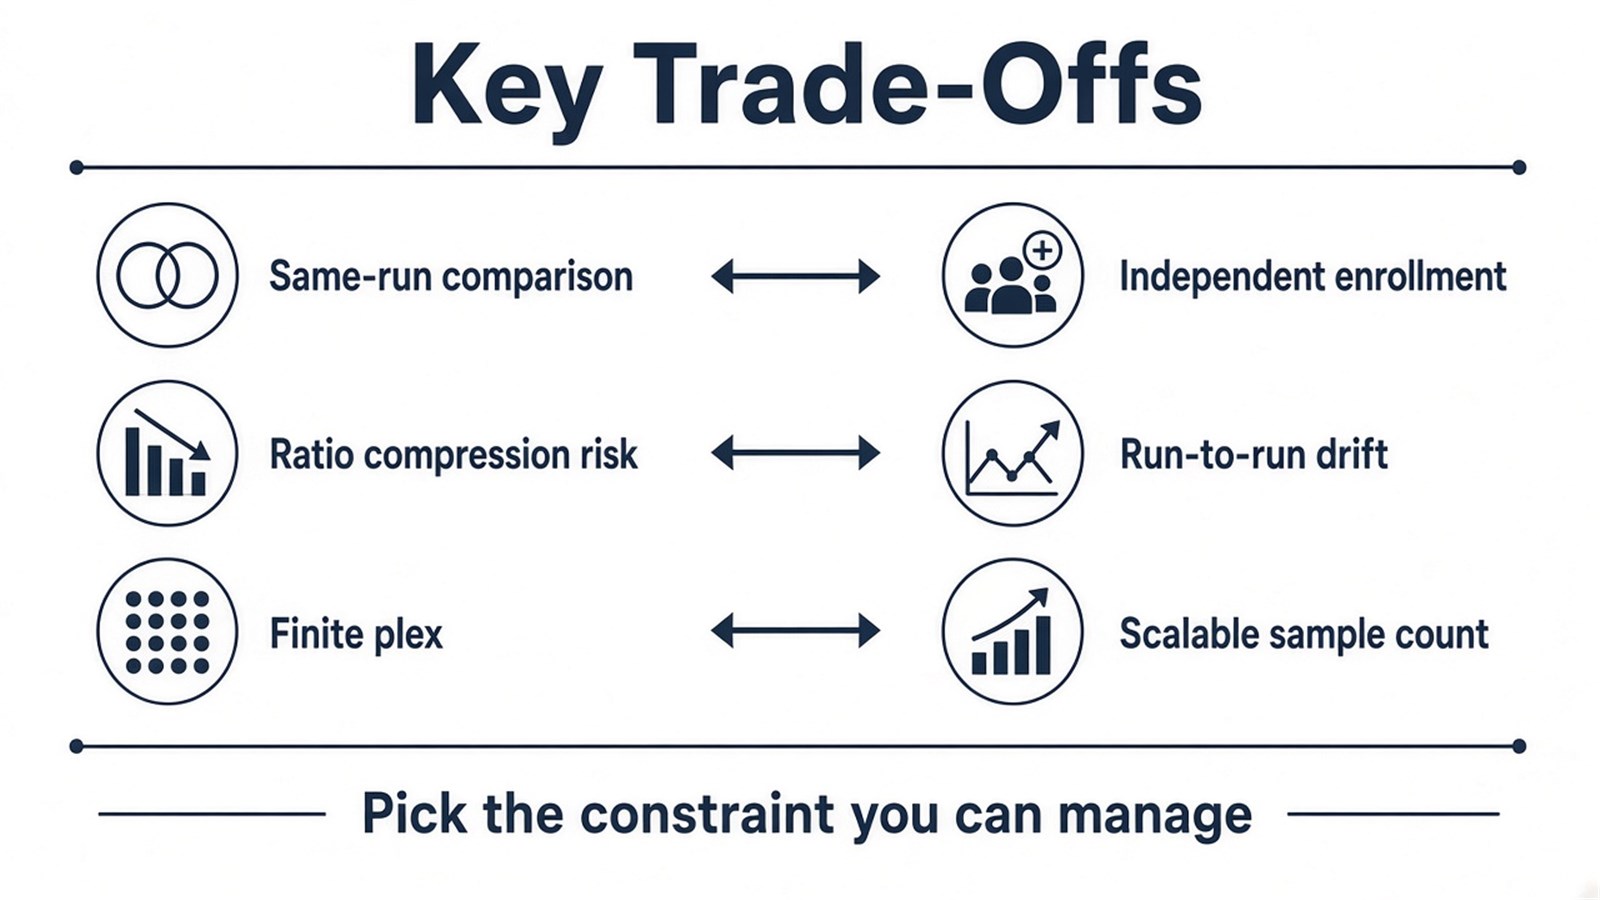
Figure 2. Method choice balances multiplex convenience, the main quantitative interference mode, and how samples enter the study.
For TMT design detail, see Advantages and Disadvantages of TMT-Based Quantitative Proteomics Analysis. For unlabeled design detail, see Advantages and Disadvantages of Label-Free Quantitative Proteomics. For a broader isobaric-versus-LFQ framework, see Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification.
When TMT or Label-Free Fits
| TMT | Label-free | Also consider | |
|---|---|---|---|
| Cohort structure | Groups and replicates fit one kit | Sample number or timing exceeds one plex | A concentration report with matched standards |
| Chemistry | Clean amine labeling is feasible | Labeling would consume scarce material or fail on the matrix | A closed peptide panel already decided |
| Comparison priority | Same-run relative ratios | Independent-run relative discovery | A defined-pair or clinical assay framework |
| Main design point | Manage compression with fractionation or MS3 | Manage drift with QC and normalization | Another relative discovery design |
| Acquisition note | DDA TMT reporter methods | DDA-LFQ or unlabeled DIA |
TMT fits multi-condition panels that can be labeled and mixed together. LFQ fits clinical tissues, biofluids, archival material, or evolving cohorts that stay unlabeled. AQUA or other SIS absolute assays fit when defined units are required. PRM or MRM fits when targets are already closed.
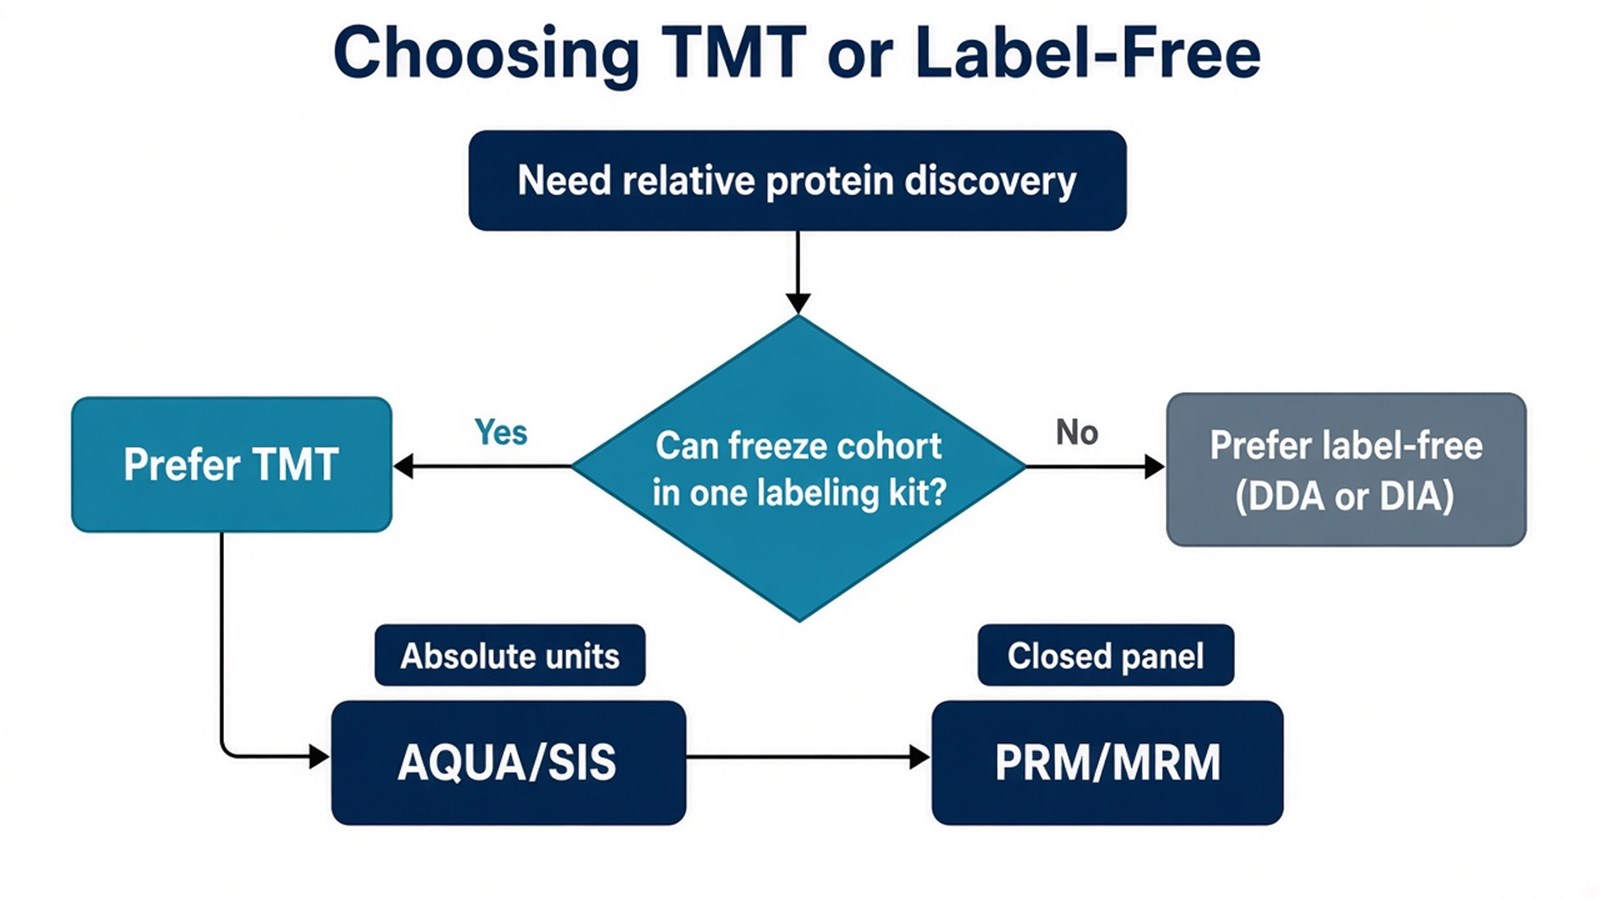
Figure 3. Start from whether the cohort fits one TMT kit. Otherwise keep label-free independent injections, or move to absolute or targeted methods when those readouts are required.
Where Each Route Is Used
TMT reporters carry relative channel intensities; sequence evidence still comes from peptide fragment spectra. LFQ uses replicates, QC pools, and declared missing-value rules with the relative matrix. Both methods support discovery applications such as treatment screens, genotype panels, and biomarker candidate ranking at relative values.
Once plex feasibility, labeling chemistry, enrollment rules, and the relative design are defined, the project can be evaluated as a TMT or label-free study. Researchers can review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service and Label-Free Quantitative Proteomics Service for sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. How should accuracy be compared between TMT and label-free?
TMT can reduce some injection drift inside one plex and can compress ratios. LFQ accuracy depends on chromatography stability, normalization, and missingness. The method that matches the cohort and the readout is the better choice for that study.
2. How do label-free quantification and DIA relate?
Label-free means no isotopic or isobaric tags. DIA is an acquisition mode that can be used in unlabeled workflows; DDA-LFQ is also label-free.
3. How is absolute protein concentration reported from TMT or LFQ?
A concentration report uses matched standards and a calibration model together with the discovery intensities.
4. What if my sample number exceeds one TMT kit?
Plan bridging across plexes, split into multiple kits with care, or switch to label-free if independent enrollment fits better.
5. When should I choose targeted methods instead?
When the peptide list is closed and the next readout is fragment-level confirmation or calibrated units from a scheduled panel.
Reference
- A. Thompson, J. Schafer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- J. Cox, M.Y. Hein, C.A. Luber, I. Paron, N. Nagaraj, M. Mann (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031591
- L. Ting, R. Rad, S.P. Gygi, W. Haas (2011). MS3 eliminates ratio distortion in isobaric multiplexed quantitative proteomics. Nat. Methods, 8, 937-940. https://doi.org/10.1038/nmeth.1714
Support Documents
Quantitative Proteomics: Label-Based and Label-Free Strategies
Compare label-based and label-free quantitative proteomics strategies and explore their applications to help identify a suitable approach for your research.