



The mechanism of pull-down coupled with mass spectrometry (MS) for protein identification is conversion of a ligand eluate into peptide-supported protein names. An immobilized ligand holds a wash-stable neighborhood. Digestion and fragment-spectrum matching name what was present in that neighborhood. A name is not an interactor, a direct contact, or an indirect contact. Empty-handle enrichment is the comparison that licenses interaction language after identification. Without it, tag binders and resin binders are identified as partners of the bait.
The ligand can be a glutathione S-transferase (GST) or His fusion, a biotinylated compound, or a nucleic acid. An antibody is immunoprecipitation, not pull-down. Tag chemistry is handle occupancy. It is not high specificity of the cargo, and it does not keep a near-native conformation. For the six bench steps that implement this mechanism, Workflow of Pull-Down Coupled with MS for Protein Identification provides a complementary workflow. For what those names mean as an identification claim, Identification of Protein Interactions Using Pull-Down and Mass Spectrometry provides a complementary identification discussion.
Researchers who already have a ligand eluate and need names from that mixture can also review the MtoZ Biolabs Pull Down based Protein Analysis Service with Mass Spectrometry for project-specific feasibility and analysis planning.
The neighborhood that will be named depends on the ligand class. A GST or His fusion presents a recombinant cargo. A biotinylated compound presents a small-molecule handle. A nucleic-acid bead presents a sequence. Each class has a matching empty arm: tag-only, linker-only, or scrambled sequence. Immobilization on agarose or magnetic beads does not certify native fold. A misfolded cargo still occupies the resin and still contributes peptides that the search will name as bait.
Capturing and Eluting a Mixture, Not Sorted Partners
Once the ligand is on the solid support, a lysate or a purified prey is offered. Proteins that remain do so because their off-rate is slower than the wash in that buffer. Washing does not leave only high-affinity partners. It is a kinetic filter. Low-affinity true contacts leave. Handle binders can stay.
Elution usually reverses the ligand–resin bond with a competitive ligand, acid, or detergent. Free glutathione or imidazole competes the handle off the bead. SDS or low pH denatures the assembly and can raise peptide yield, including more handle binders. Changing ionic strength can loosen some cargo–prey contacts, but it does not sort the tube into a partner-only fraction. The collected material still contains ligand, cargo, prey, and resin binders. Empty-handle must use the same exit. If the two arms leave by different bonds, later enrichment compares two different mixtures.
For the fusion-specific detection chain, where the question is how a PPI is reported as co-recovery rather than how a name is produced, Mechanism of Pull-Down and MS in Fusion Protein Interaction Analysis provides a complementary fusion mechanism.
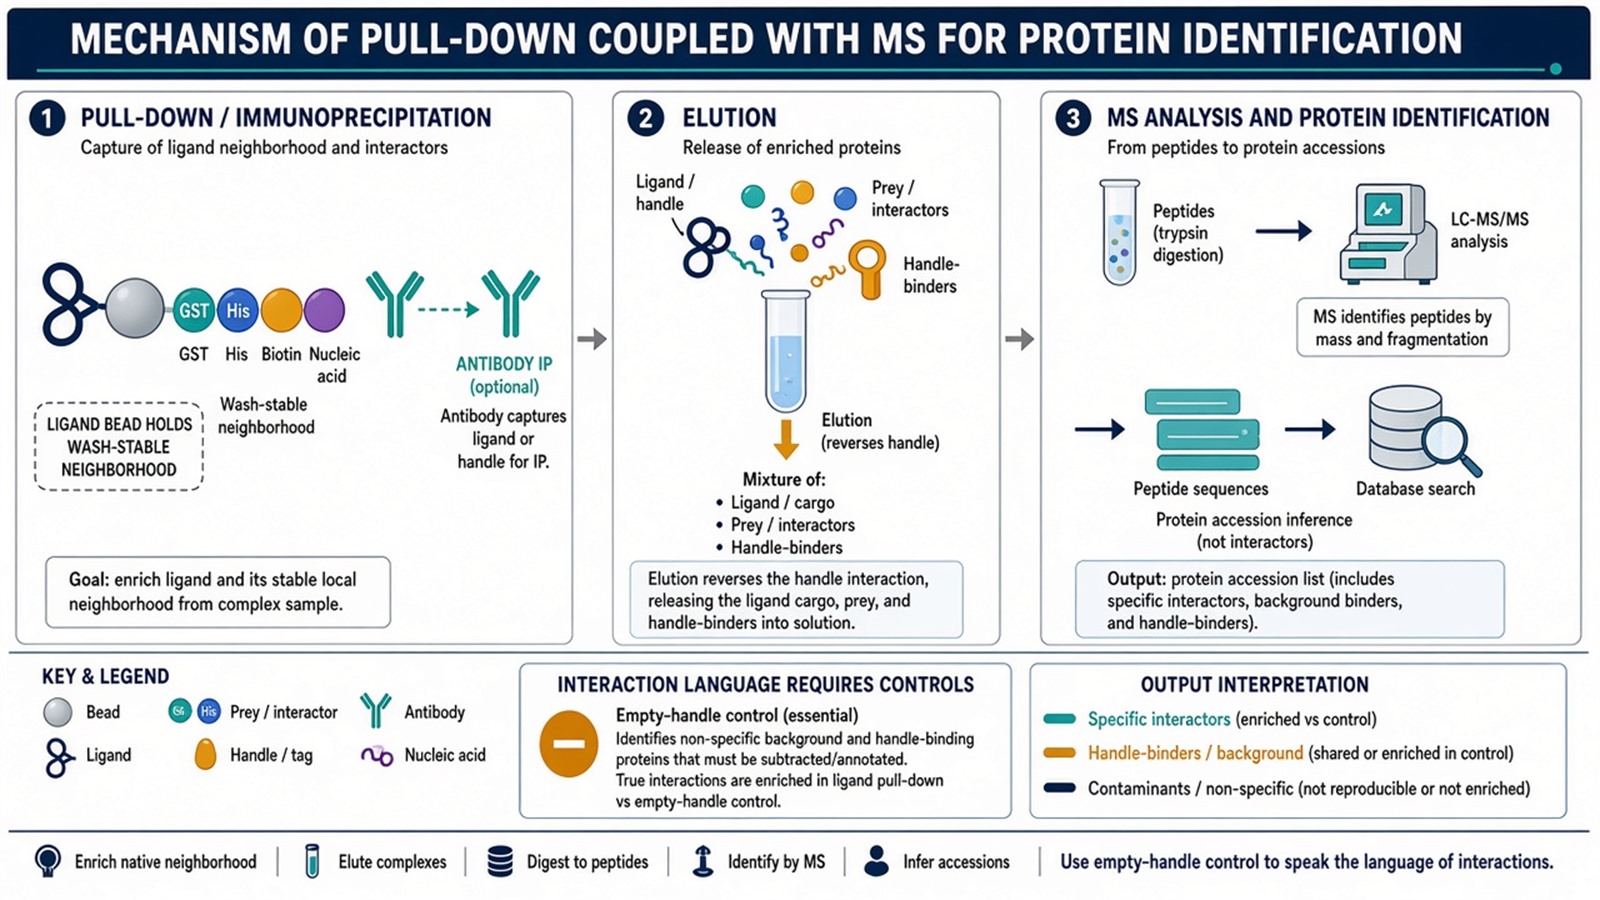
Figure 1. Capture holds a wash-stable neighborhood. Elution reverses the handle more often than it sorts prey from bait. MS then names peptides in the mixture.
Write which bond elution will reverse before calling the eluate a partner-protein fraction. Competitive GSH or imidazole releases the handle, not a purified interactor set.
Mass Spectrometry Analysis and Protein Identification
Identification on that mixture has four operations. They produce accessions. They do not produce interactors.
Proteins are digested, commonly with trypsin. Peptides are separated by liquid chromatography and ionized. The spectrometer records precursor mass-to-charge values and fragment spectra. Sequence assignment comes from the fragments against a protein database. A parent-ion spectrum is not an identification.
A matched accession means that peptides from that protein were present in the digest. It does not mean the protein bound the cargo, bound directly, or bound indirectly. For when this identification method is the right first choice, Advantages and Disadvantages of Pull-Down Coupled with MS for Protein Identification provides a complementary decision.
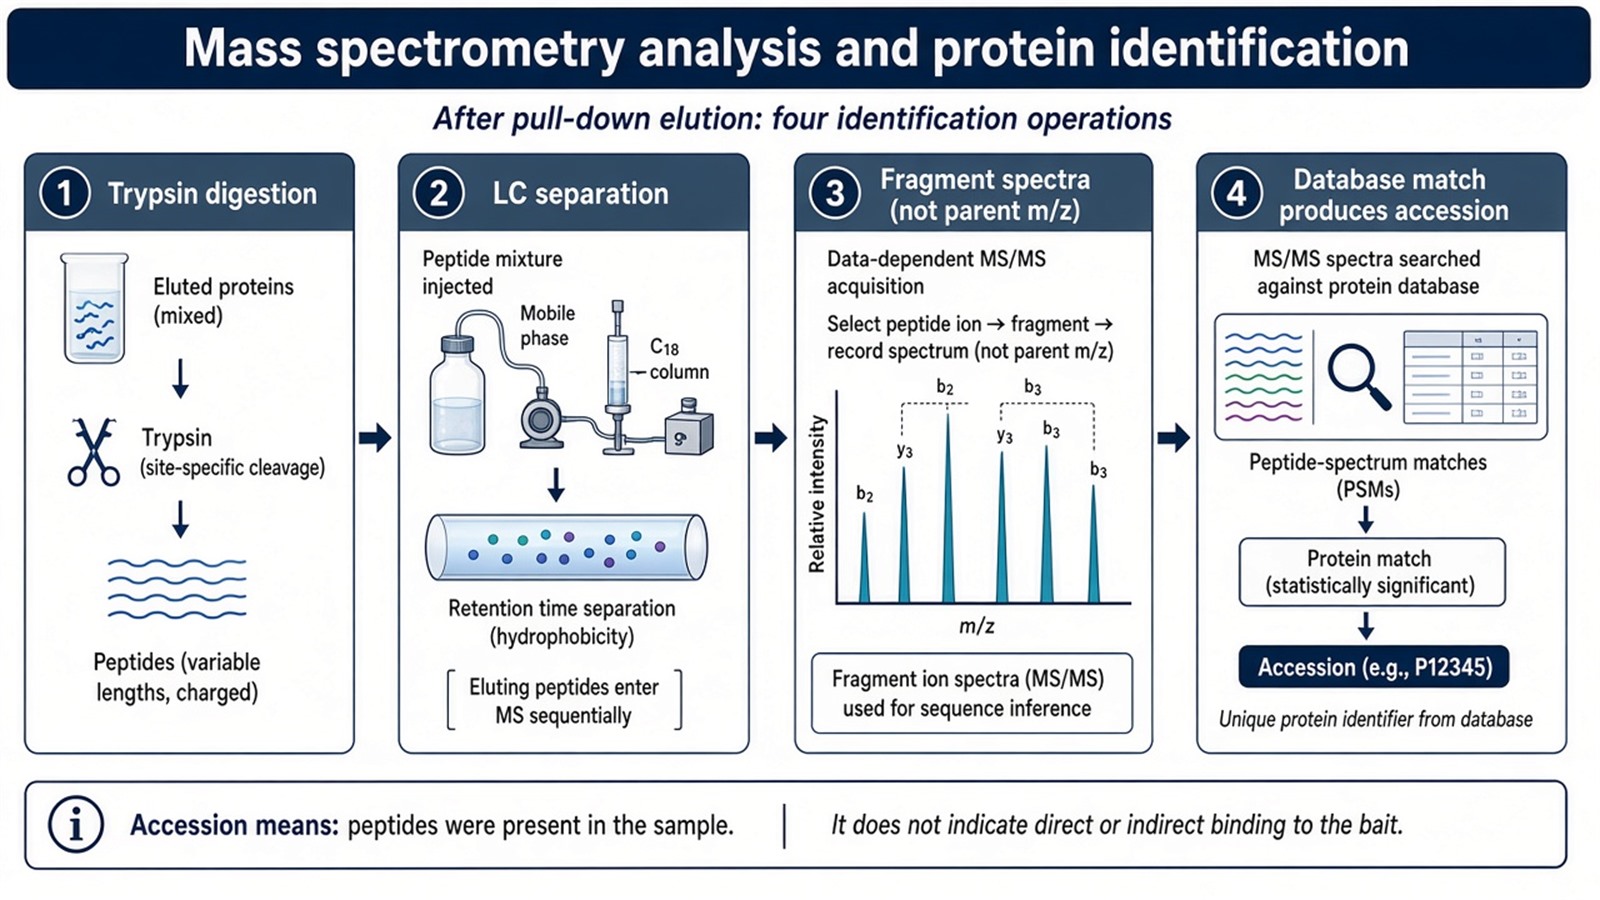
Figure 2. Trypsin, chromatography, tandem MS, and a search engine name proteins in the eluate. Direct versus indirect contact is not a field in that search.
Data Analysis and Interpretation
Peak intensity and relative abundance, when acquired, report occupancy after wash. Statistical comparison against the empty-handle arm can rank enrichment. A p-value on that comparison is still a bead-level rank. It does not determine which interactions are significant in a cell, and it does not replace a missing empty-handle digest.
Annotation and network drawing use literature that sits outside the file. Those overlays can suggest roles to test. They are not part of the identification mechanism and they do not reveal biological significance from one pull-down. For how to read the table after names exist, How to Interpret the Results of a Pull-Down Experiment provides a complementary interpretation order.
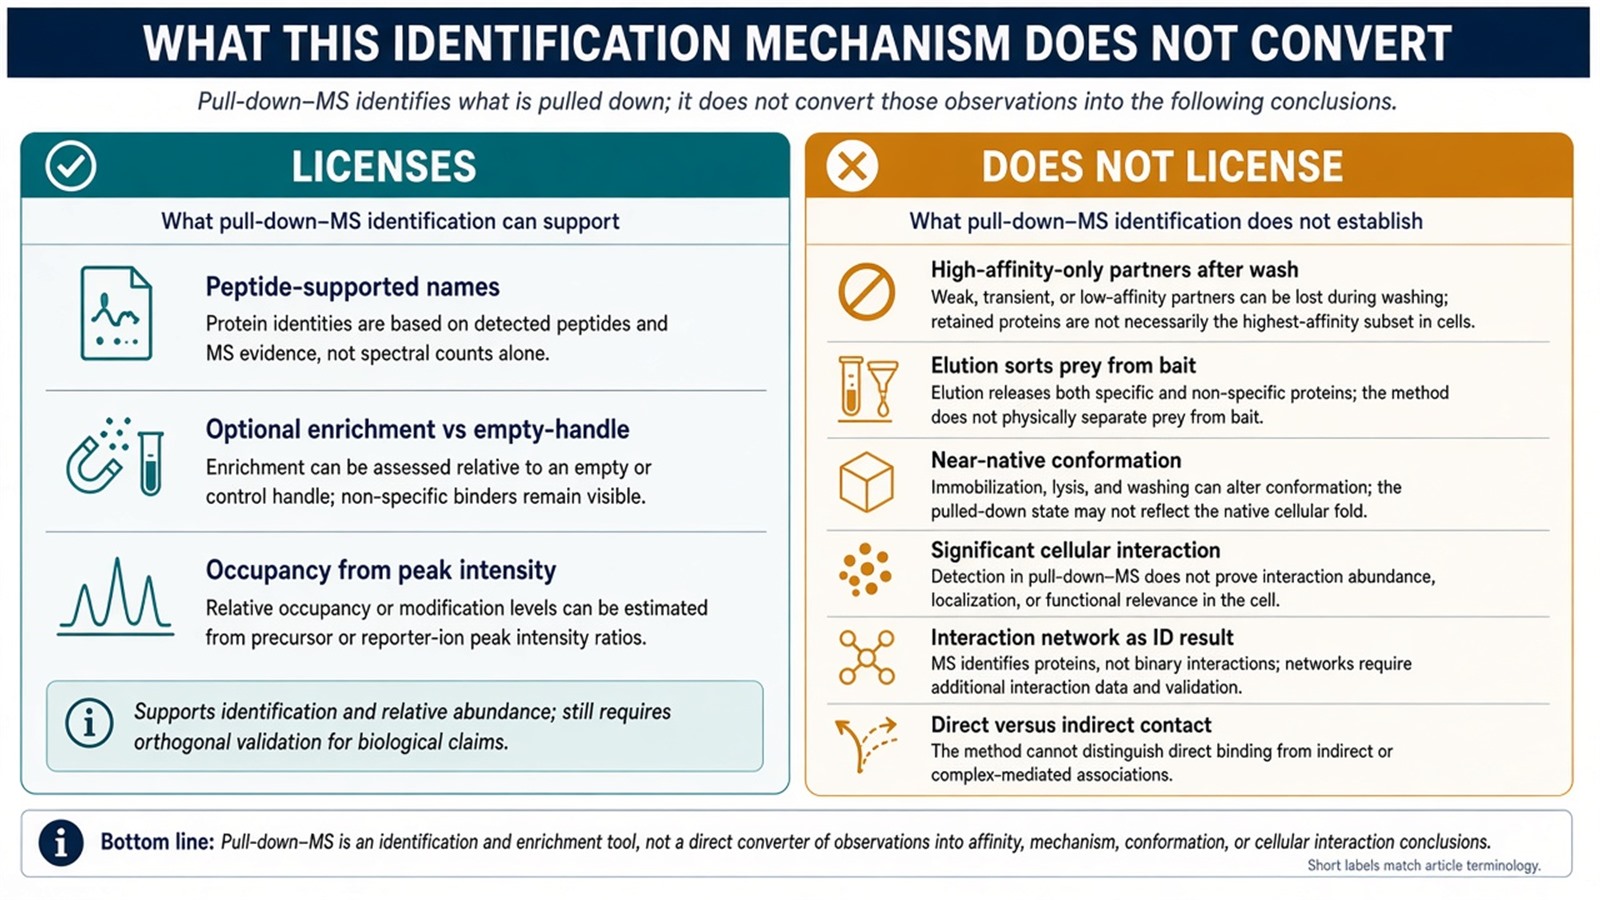
Figure 3. Supported: peptide-supported accessions and, if both arms exist, enrichment. Not converted by this mechanism: high-affinity-only capture, a sorted prey list, or a pathway map.
Experimental Limits
The mechanism reduces a mixture to what the ligand, the wash, and the search engine can name. Diagnostic language sits outside ordinary research reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable identification-mechanism plan names the ligand, the elution bond, the empty-handle arm, and the claim that will be allowed, before the first digest. MtoZ Biolabs scopes pull-down with Western or Orbitrap-class LC-MS/MS. Interpretability still follows this conversion, not the database brand.
Once names are the deliverable around a broader PPI question, the project can be scoped as MS-based interaction analysis. Researchers who need method matching beyond pull-down identification can review the MtoZ Biolabs Protein-Protein Interaction Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What is the identification mechanism of pull-down plus MS?
A ligand holds a wash-stable neighborhood; elution releases a mixture; fragment spectra name proteins present in that mixture.
2. Does washing leave only high-affinity partners?
No. Washing is a kinetic filter. High-affinity language is not a result of the wash.
3. Does elution separate partners from the target?
Usually not. Competitive elution reverses the handle. The tube still contains ligand, cargo, prey, and handle binders.
4. Does an identification mean the protein interacts directly or indirectly?
Neither is licensed by the name. Direct versus indirect is a later, orthogonal question.
5. How does this page differ from the fusion mechanism and the workflow pages?
This page is how a protein name is produced. The fusion mechanism page is how a PPI is detected as co-recovery. The workflow page orders the six bench steps.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Dunham W.H., Mullin M., Gingras A.C. (2012). Affinity-purification coupled to mass spectrometry: basic principles and strategies. Proteomics, 12, 1576-1590. https://doi.org/10.1002/pmic.201100523
- Mellacheruvu D., Wright Z., Couzens A.L., et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
- Perkins D.N., Pappin D.J., Creasy D.M., Cottrell J.S. (1999). Probability-based protein identification by searching sequence databases using mass spectrometry data. Electrophoresis, 20, 3551-3567. https://doi.org/10.1002/(SICI)1522-2683(19991201)20:18<3551::AID-ELPS3551>3.0.CO;2-2
- Phizicky E.M., Fields S. (1995). Protein-protein interactions: methods for detection and analysis. Microbiol. Rev., 59, 94-123. https://doi.org/10.1128/mr.59.1.94-123.1995
