



Pull-down coupled with mass spectrometry (MS) is the right identification method when an immobilized ligand and a matched empty-handle arm can support an open peptide-identified list. It is the wrong first identification method when the sample is an unfractionated proteome, when one nominated band already answers the question, or when the next sentence is a binding constant or a transient live-cell contact. The real advantages are enrichment around one ligand, names that Western blot cannot supply, and relative occupancy if a quantitative method is chosen. The real disadvantages sit in the same chemistry: ligand and resin binders enter the file, recovery limits who can be sequenced, the wash removes weak partners, and condition changes rewrite the list. High specificity, detection of rare transient interactions, and precise quantification of dynamics are not properties of the combination. They are claims that have to be earned by occupancy, the empty-handle arm, and a named contrast.
Tagged fusions, biotinylated ligands, and nucleic-acid baits belong here. An antibody capture is immunoprecipitation (IP) or co-immunoprecipitation (Co-IP) and is controlled with an isotype. Keeping those names separate prevents an IgG lane from being asked to serve as a GST-only control. For how identification itself is defined, Identification of Protein Interactions Using Pull-Down and Mass Spectrometry provides a complementary identification discussion. For when a fusion is the right PPI method rather than the right ID method, Advantages and Disadvantages of Pull-Down and MS in Fusion Protein Interaction Analysis provides a complementary fusion decision.
Researchers who already have a ligand, an empty-handle arm, and an identification question can also review the MtoZ Biolabs Pull Down based Protein Analysis Service with Mass Spectrometry for project-specific feasibility and analysis planning.
Advantages That Hold Only Under Stated Conditions
These points are useful when the condition in the same sentence is true. They are not built-in virtues.
Open names around one ligand, if the empty-handle arm is digested
MS can identify proteins that were not nominated for a blot. Selective capture enriches the ligand neighborhood. It does not reduce background in the file. Sensitivity of the spectrometer recovers peptides that were present. Presence is not a rare or transient interaction. Transient contacts that fail the wash never enter the digest.
Relative occupancy, if a quantitative method is chosen
Label-free quantification on paired runs is the default. SILAC applies when the prey culture can be labeled. iTRAQ or TMT labels after digestion and keeps tube-to-tube capture error inside the ratio. Those methods report occupancy after wash. They do not enhance precision of a dynamic PPI, and they do not report a copy number.
Ligand class can be chosen, if the control matches that class
A tag, a biotinylated compound, or a nucleic acid can be immobilized. That choice is real. It is not unlimited versatility across biological systems. ELISA is not a standard pull-down validator. Western blot can test a nominated pair after the open list is named. For ligand classes and why an antibody is named IP, Affinity Pull-Down Assay provides a complementary ligand-class discussion.
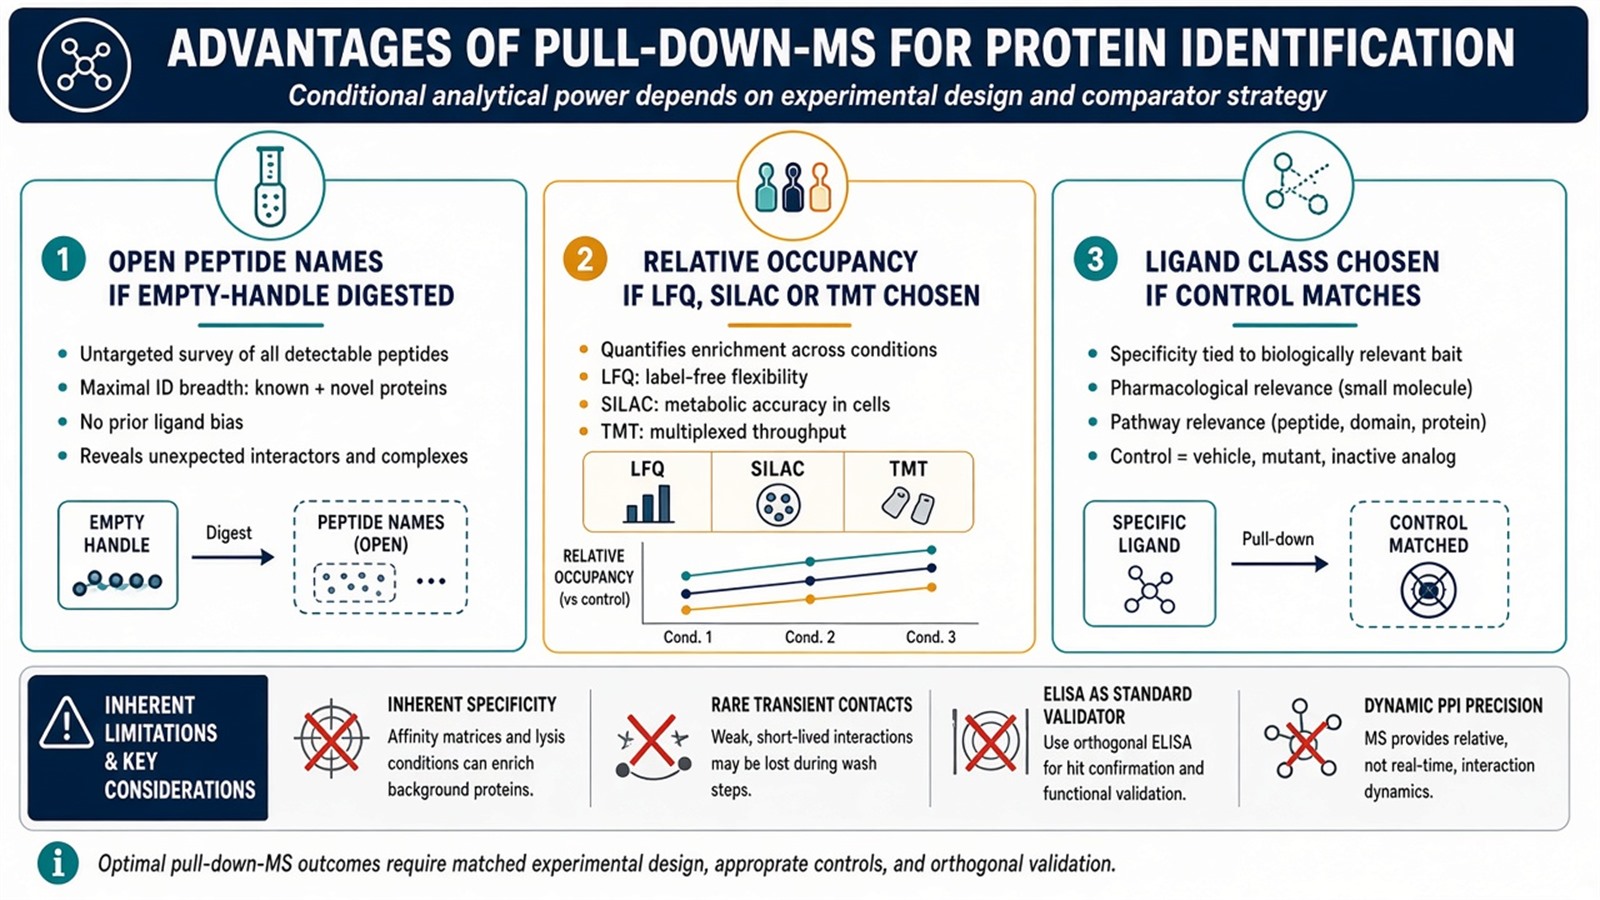
Figure 1. Supported when both arms exist: open peptide names and relative occupancy. Not granted by capture or MS sensitivity: low background, transient contacts, or a dynamic PPI proof.
Write the empty-handle arm before calling the identification specific. More wash cannot invent a control that was not processed.
Disadvantages That Sit in the Identification Chemistry
These limits are structural. They are not beginner errors.
Non-specific identifications remain after wash
The bait does not “bind unrelated proteins” as a special failure mode. Tag binders, linker binders, and resin binders occupy the same eluate. Stringent washing reduces some of those rows and also deletes weak true partners. Control experiments estimate the residue. They do not restore credibility to a single-arm search table.
Recovery limits who can be sequenced
Ligand occupancy, bait amount, and elution efficiency decide the peptide load. Low recovery is an identification limit: proteins that never reached the digest cannot be named. It is not, by itself, an accuracy problem in the search engine.
The wash disrupts complexes that the method is meant to see
Unstable contacts leave during lysis and wash. That is the kinetic filter. It is not a side effect that compromises an otherwise intact interaction network. Post-lysis contacts can also form and appear as false-positive identifications. Empty-handle and, where needed, a no-ligand bead estimate those rows.
The list is condition-dependent
Buffer, salt, detergent, and operator time change who remains. Reproducibility follows a written wash and matched arms, not a more flexible protocol. Laboratory-to-laboratory differences are expected when those variables are unnamed. For how a pull-down blot or table is read after identification, How to Interpret the Results of a Pull-Down Experiment provides a complementary interpretation order.
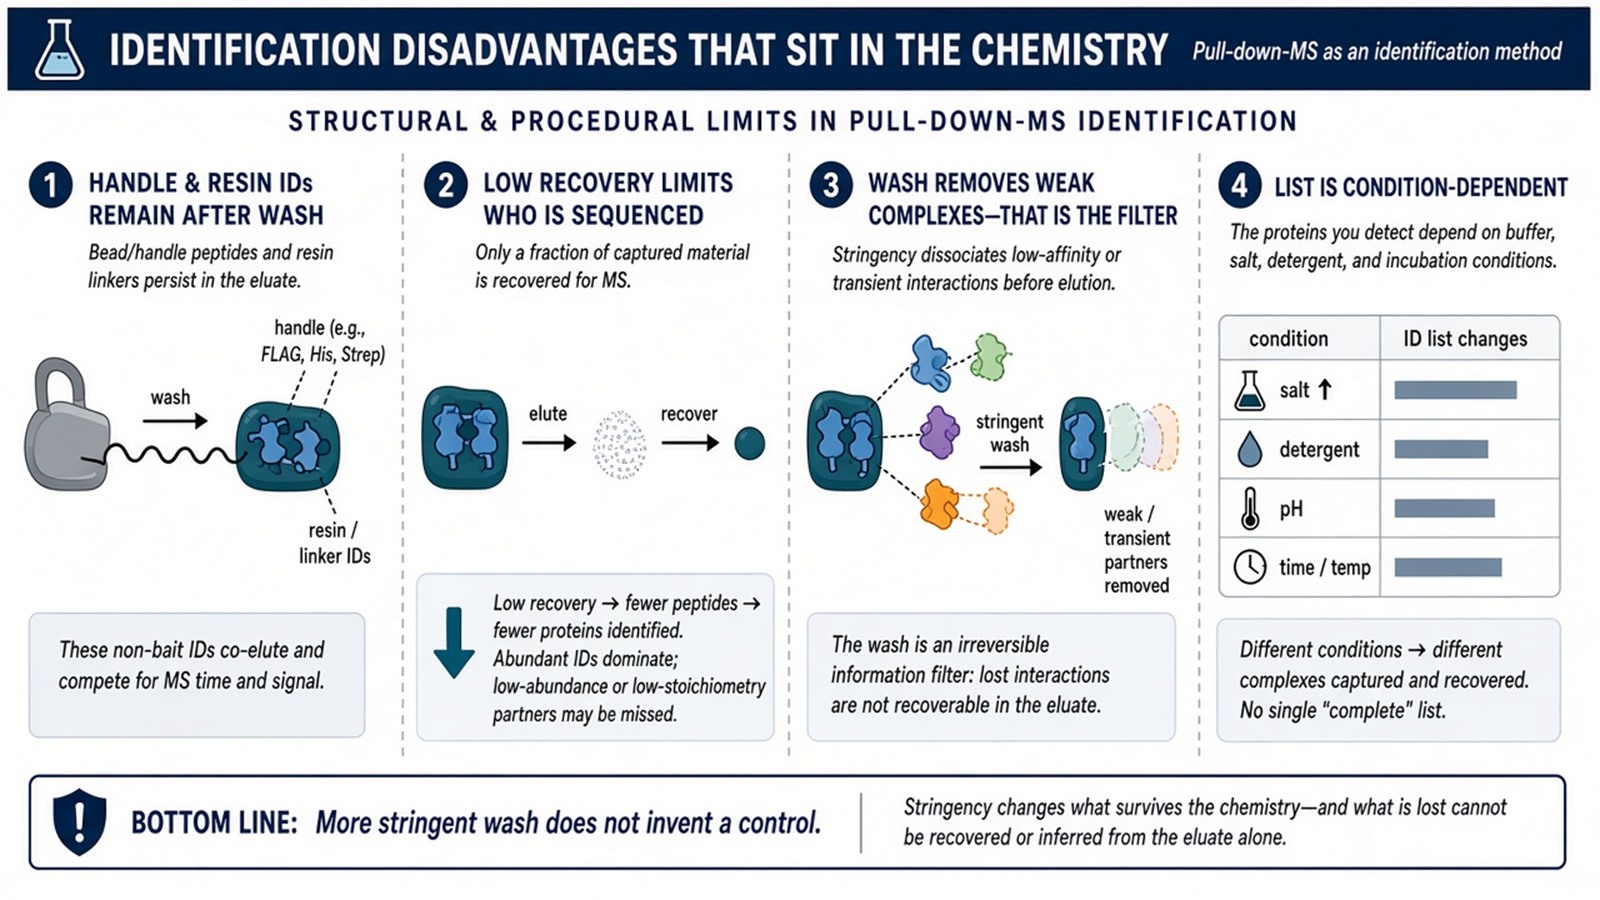
Figure 2. False IDs follow the handle. Missing IDs follow recovery and wash. Reproducibility follows a written condition, not operator independence.
When Pull-Down–MS Is Not the Identification Method
Choose a different first assay when the identification claim is already named.
An unfractionated lysate census is shotgun proteomics, not pull-down. A single nominated pair is a Western after a small-scale capture. Intact-complex mass is native MS or an orthogonal mass method. Endogenous capture of an untagged bait is Co-IP-MS. A GST-specific five-step coupling is a narrower page; GST Pull-Down Assay Combined with Mass Spectrometry for Protein-Protein Interaction Analysis provides that GST–MS workflow.
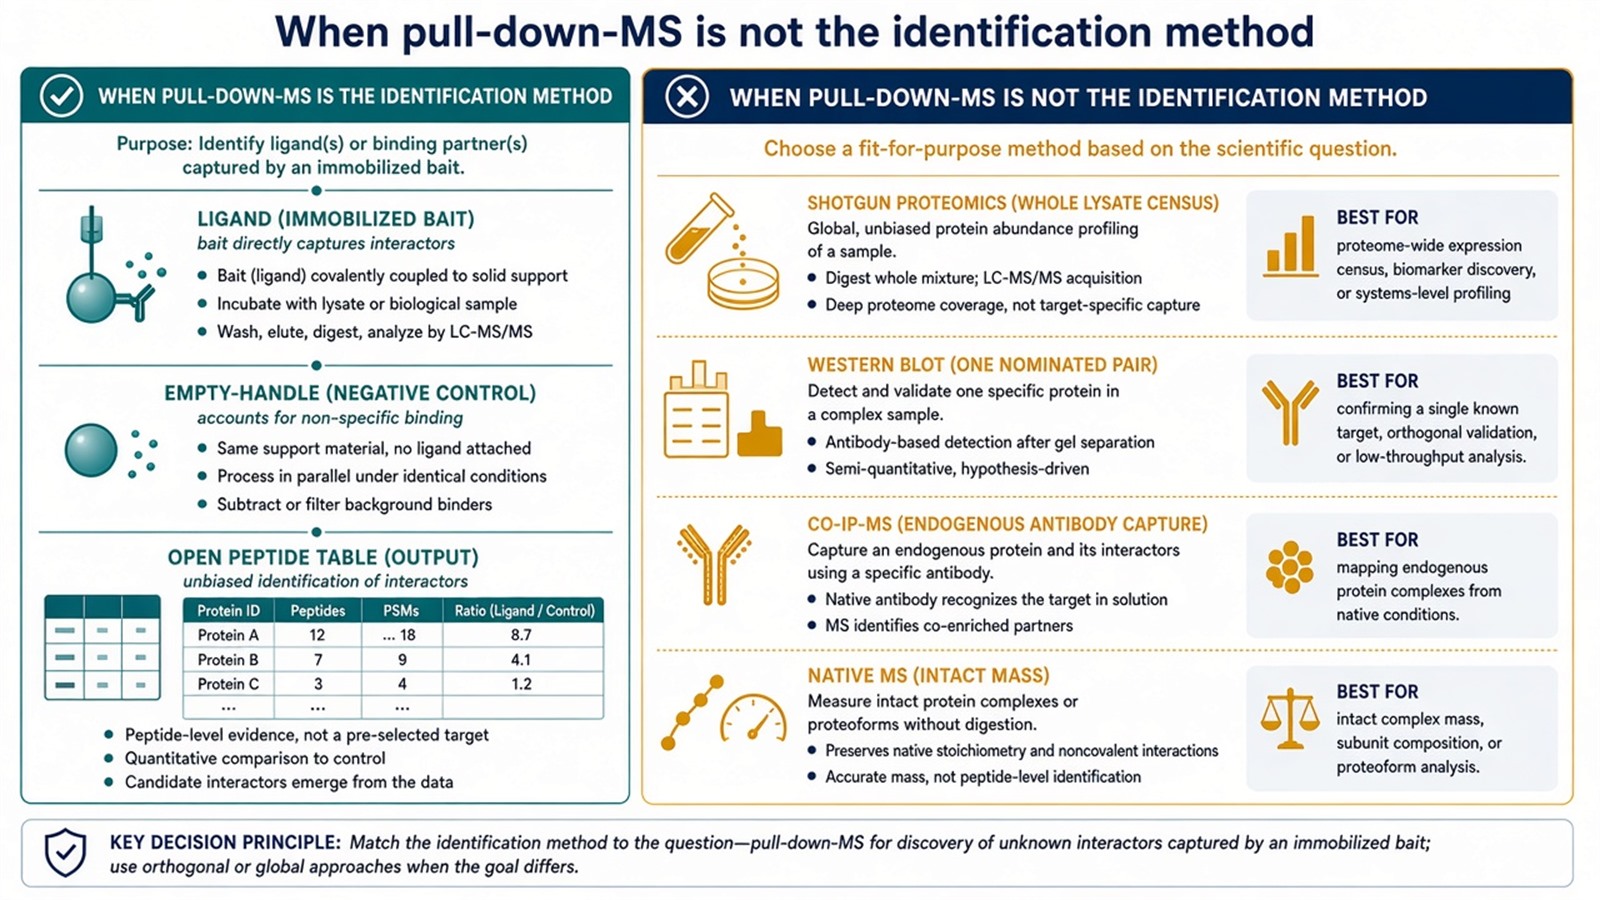
Figure 3. Right first ID method: ligand, empty-handle, open peptide table. Wrong first method: whole-proteome census, one Western pair, intact mass, or endogenous antibody capture.
Experimental Limits
The combination reduces a mixture to what the ligand, the wash, and the search engine can report. Diagnostic language sits outside ordinary research reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable identification decision names the ligand class, the empty-handle arm, and the claim that will be allowed, before the first lysis. MtoZ Biolabs scopes pull-down with Western or Orbitrap-class LC-MS/MS. Interpretability still follows those arms, not the resin brand.
Once the ligand is specified as a GST fusion, the wet capture can be scoped as GST pull-down. Researchers who need a GST-bait plan before identification can review the MtoZ Biolabs GST Pull Down Assay Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What is the main identification advantage of pull-down plus MS?
An open peptide-supported list around one ligand, if the empty-handle arm is run.
2. Does selective capture make identifications specific?
No. Capture enriches the neighborhood. Specificity of a name is the empty-handle comparison.
3. Do SILAC or iTRAQ make the identification quantitative?
They can report relative occupancy after wash. They do not measure a dynamic binding constant.
4. Why do identifications change between laboratories?
Wash, ligand occupancy, and elution are part of the method. Unwritten conditions produce different lists.
5. How does this page differ from the fusion advantages page?
This page scores pull-down–MS as an identification method across ligand classes. The fusion page decides when a tagged bait is the right PPI experiment.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Dunham W.H., Mullin M., Gingras A.C. (2012). Affinity-purification coupled to mass spectrometry: basic principles and strategies. Proteomics, 12, 1576-1590. https://doi.org/10.1002/pmic.201100523
- Mellacheruvu D., Wright Z., Couzens A.L., et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
- Phizicky E.M., Fields S. (1995). Protein-protein interactions: methods for detection and analysis. Microbiol. Rev., 59, 94-123. https://doi.org/10.1128/mr.59.1.94-123.1995
- Vikis H.G., Guan K.L. (2004). Glutathione-S-transferase-fusion based assays for studying protein-protein interactions. Methods Mol. Biol., 261, 175-186. https://doi.org/10.1385/1-59259-762-9:175
