



Identification of protein interactions using pull-down and mass spectrometry (MS) means asking which proteins remain with an immobilized ligand after a written wash, then converting that eluate into a peptide-identified list. The ligand can be a tagged protein, a small molecule, or a nucleic acid. Liquid chromatography–tandem MS (LC-MS/MS) names what was present in the digest. Presence in the digest is not an interaction. A matched ligand-only, tag-only, or linker-only arm is part of the identification. Without it, resin binders and handle binders are scored as partners of the bait.
When the capture reagent is an antibody, the same solid-phase logic is immunoprecipitation (IP) or co-immunoprecipitation (Co-IP) and is controlled with an isotype, not with an empty tag. Keeping those names separate prevents an IgG lane from being asked to serve as a GST-only control. Western blotting after pull-down can only test nominated pairs. Pull-down–MS is used when the next table should be an open inventory around one ligand.
Researchers who already have a tagged bait, a compound handle, or a nucleic-acid ligand and need an identification table can also review the MtoZ Biolabs Pull Down based Protein Analysis Service with Mass Spectrometry for project-specific feasibility and analysis planning.
What Identification Reports
A pull-down holds a known ligand on agarose or magnetic beads and offers a purified prey or a lysate. Proteins that survive detergent, salt, and wash time stay on the resin. MS after digestion reports peptide-spectrum matches and protein accessions. Those accessions are candidates for enrichment against the ligand-only arm. They are not, by themselves, protein–protein interactions, binding sites, or a disease mechanism.
High sensitivity of the spectrometer can recover low-abundance peptides that were present. High throughput does not mean many samples are analyzed in one injection. Specificity is the ligand-only comparison, not the ligand chemistry. For how ligand classes differ and why an antibody is named IP, Affinity Pull-Down Assay provides a complementary ligand-class discussion. For the GST-specific coupling of handle capture to LC-MS/MS, GST Pull-Down Assay Combined with Mass Spectrometry for Protein-Protein Interaction Analysis provides a complementary GST–MS page.
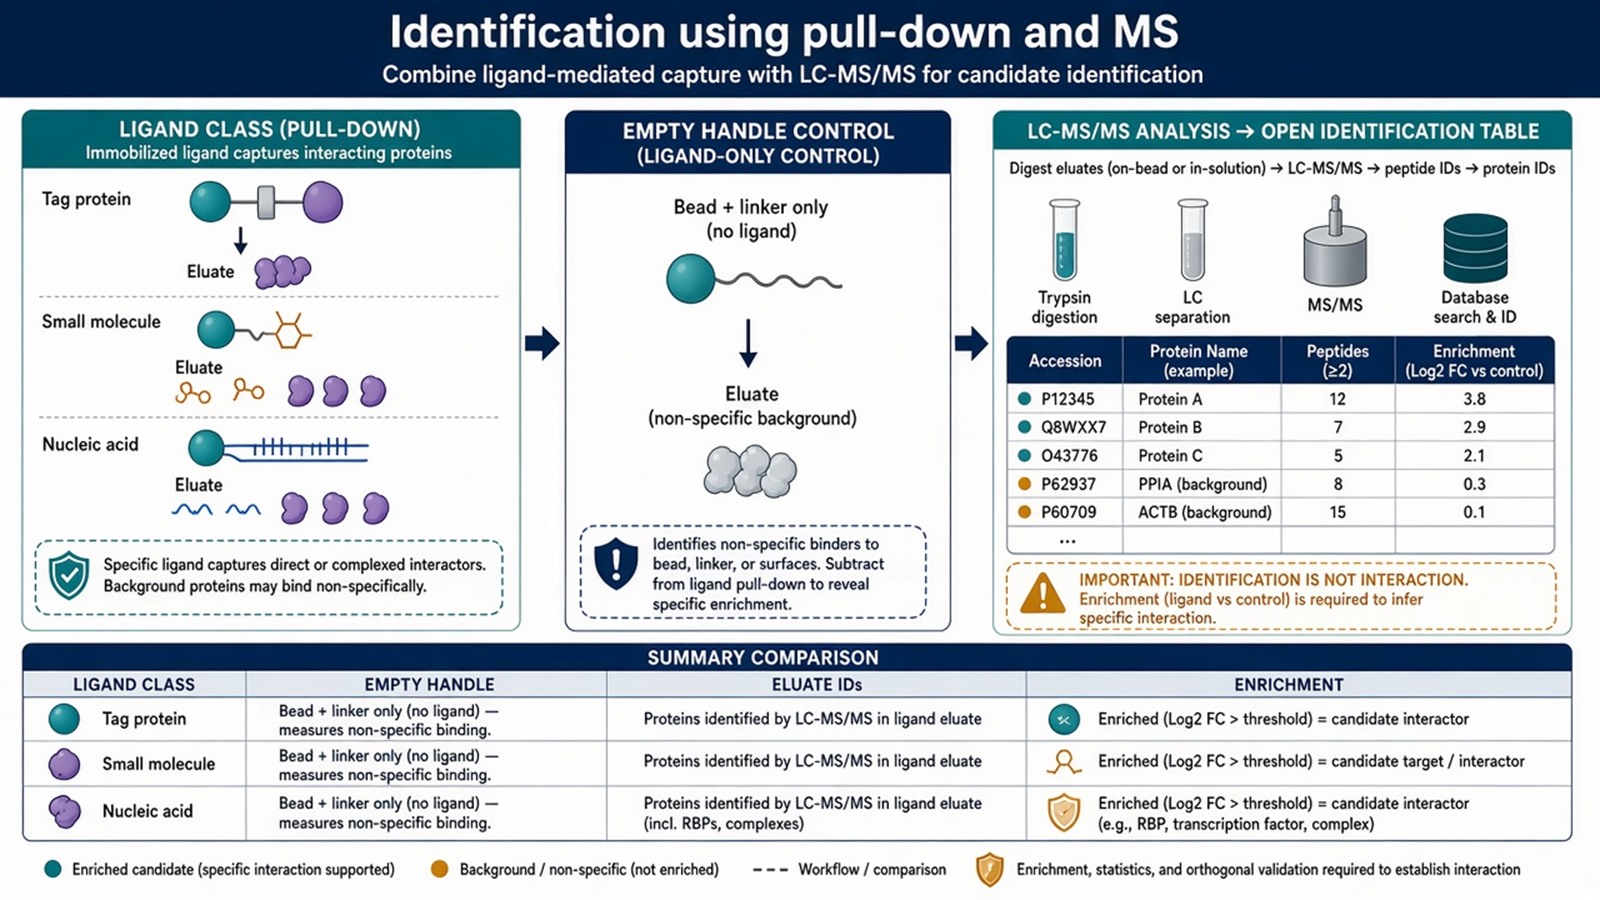
Figure 1. Pull-down retains wash-stable proteins on a ligand bead. MS names the open list. Ligand-only estimates handle and resin binders. Western still tests nominated pairs.
Name the ligand class and the empty-handle arm before the first digest. The search engine cannot invent a control that was not processed.
Principles and Workflow of Target Protein Pull-Down
The principle is occupancy of an immobilized ligand, not antibody–Protein A/G. A GST fusion on glutathione (GSH) beads is the bait. It does not “capture a separate target protein” unless that sentence names a prey. The fusion is presented; prey that remains is asked later.
1. Sample preparation
Cells or tissues are lysed in a buffer that keeps the intended contacts. Total extract is a feed, not a negative control. Input protein is matched across ligand and ligand-only arms.
2. Ligand immobilization
The ligand is bound to the solid phase before, or together with, the prey, depending on the route. Tag pull-downs use GST, His, Strep, or biotin handles. Compound pull-downs use a linker-coupled analog. Nucleic-acid pull-downs use DNA or RNA. Antibody resins belong with IP.
3. Incubation and washing
The same prey is offered to both arms. Multiple washes reduce some resin binders and also strip weak ligands. Wash survival is a kinetic filter. It is not proof that leftover proteins are specific.
4. Elution, then identification
Elution uses a competitive ligand, acid, or detergent. Reducing salt concentration is a wash change, not a standard elution. The eluate is then digested and searched. For how to read the blot or the table after those steps, How to Interpret the Results of a Pull-Down Experiment provides a complementary interpretation order.
Application of Mass Spectrometry in Protein Identification
MS after pull-down is a peptide identification workflow. It is not a second, independent proof of binding.
1. Protein digestion
The eluate, or the washed beads, is reduced, alkylated, and cut with trypsin. Ligand and ligand-only must use the same exit. Cleanup does not rescue an unmatched capture.
2. Mass spectrometry analysis
Peptides are separated by liquid chromatography, ionized, and measured by mass-to-charge ratio and fragment spectra. Sequence assignment comes from those fragments against a protein database. Parent m/z alone does not identify a protein.
3. Data analysis
Database matches produce a protein list. Each row is first asked whether the ligand or bait was recovered and whether the candidate is enriched against the empty-handle arm. Subtracting every ligand-only identification deletes real partners that leak into the handle arm. Annotation and network overlay come last. For antibody capture plus MS, How to Combine Co-IP with Mass Spectrometry for Protein Interaction Analysis provides a complementary Co-IP-MS coupling.
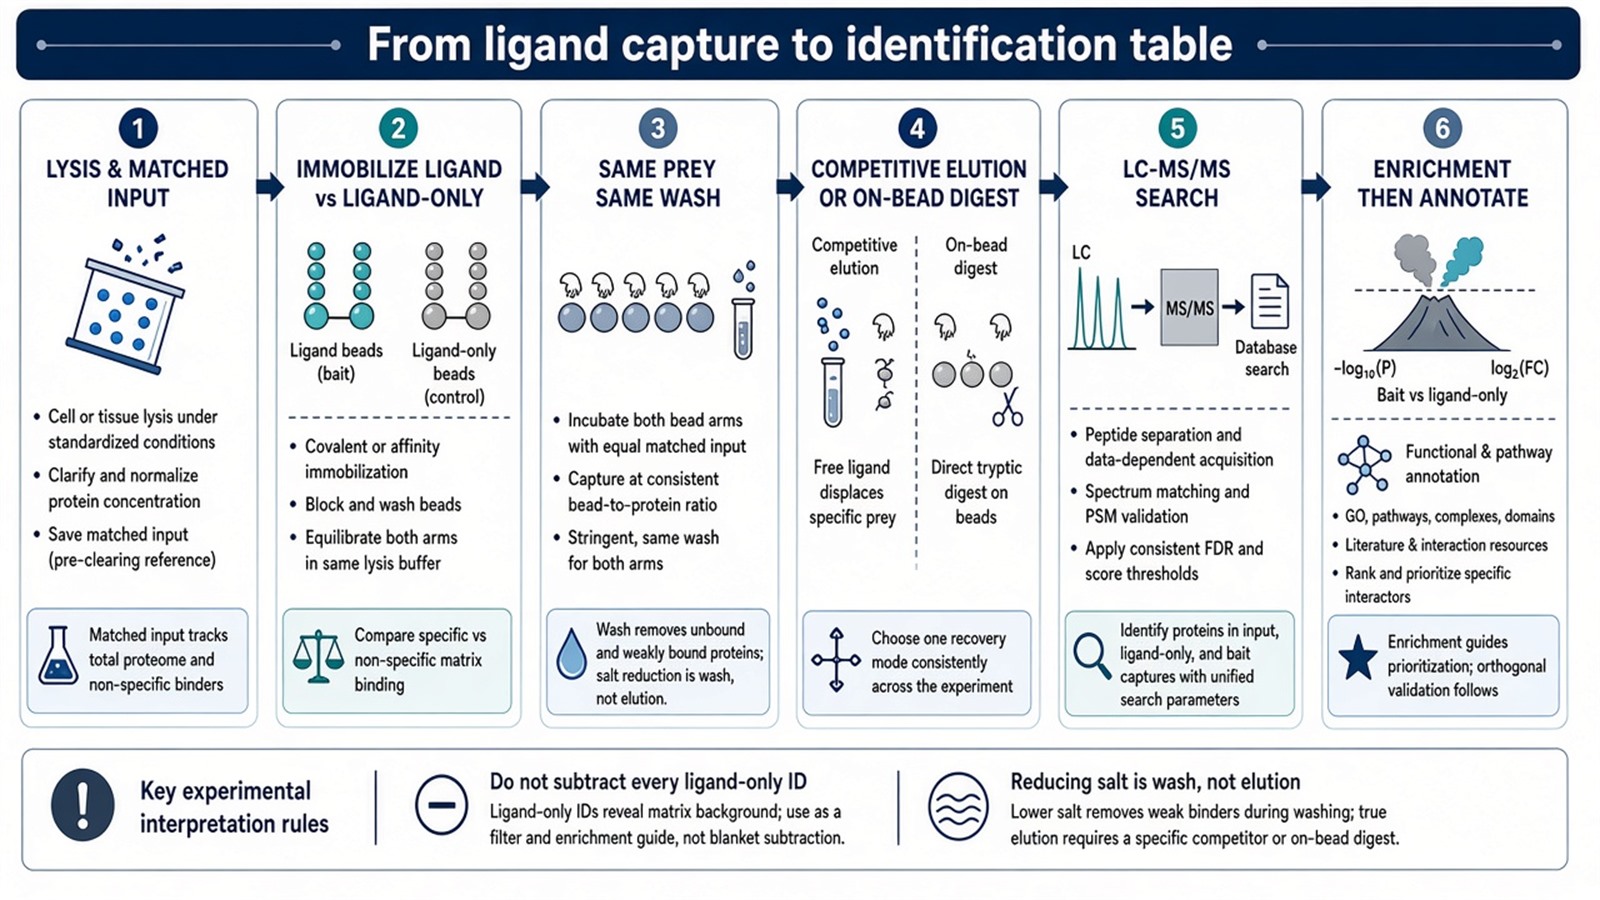
Figure 2. Digestion and LC-MS/MS convert both arms into peptide tables. Identification is the search. Interaction language waits for enrichment.
Advantages of Target Protein Pull-Down Combined with Mass Spectrometry
These points hold only with the condition in the same sentence.
Open identification around one ligand, if the empty-handle arm is run
MS can name proteins that were not nominated for Western blot. The advantage disappears when only the bait arm is digested.
Detection of low-abundance peptides that were present, if bait occupancy is adequate
The spectrometer can report traces in the digest. Traces that also sit in the ligand-only arm remain background. Weak interactions that fail the wash never enter the file.
Relative comparison across matched arms, if a quantitative method is chosen
Label-free quantification on paired runs is the default. SILAC applies when the prey culture can be labeled. Isobaric tags label after digestion and keep tube-to-tube capture error inside the ratio. Pull-down is not inherently non-quantitative. It is uninterpreted until those arms exist.
High specificity is not granted by “specific ligands or antibodies.” High throughput is not granted by running one LC-MS/MS gradient.
Limitations of Target Protein Pull-Down Combined with Mass Spectrometry
Non-specific binding
Background remains after extensive washing. Ligand-only, linker-only, or bead-only estimates that residue. More wash cannot reconstruct a control that was omitted.
Detection limits
Sample purity and complexity affect who is sequenced. Low-abundance or weak partners can be absent because they were washed off, not because the instrument lacked sensitivity.
What the table still cannot say
An identification list does not report a binding constant, a direct binary contact, or a complete complex. Quantitative MS after pull-down reports occupancy after wash. It does not convert the capture into an intracellular network or a drug-target claim.
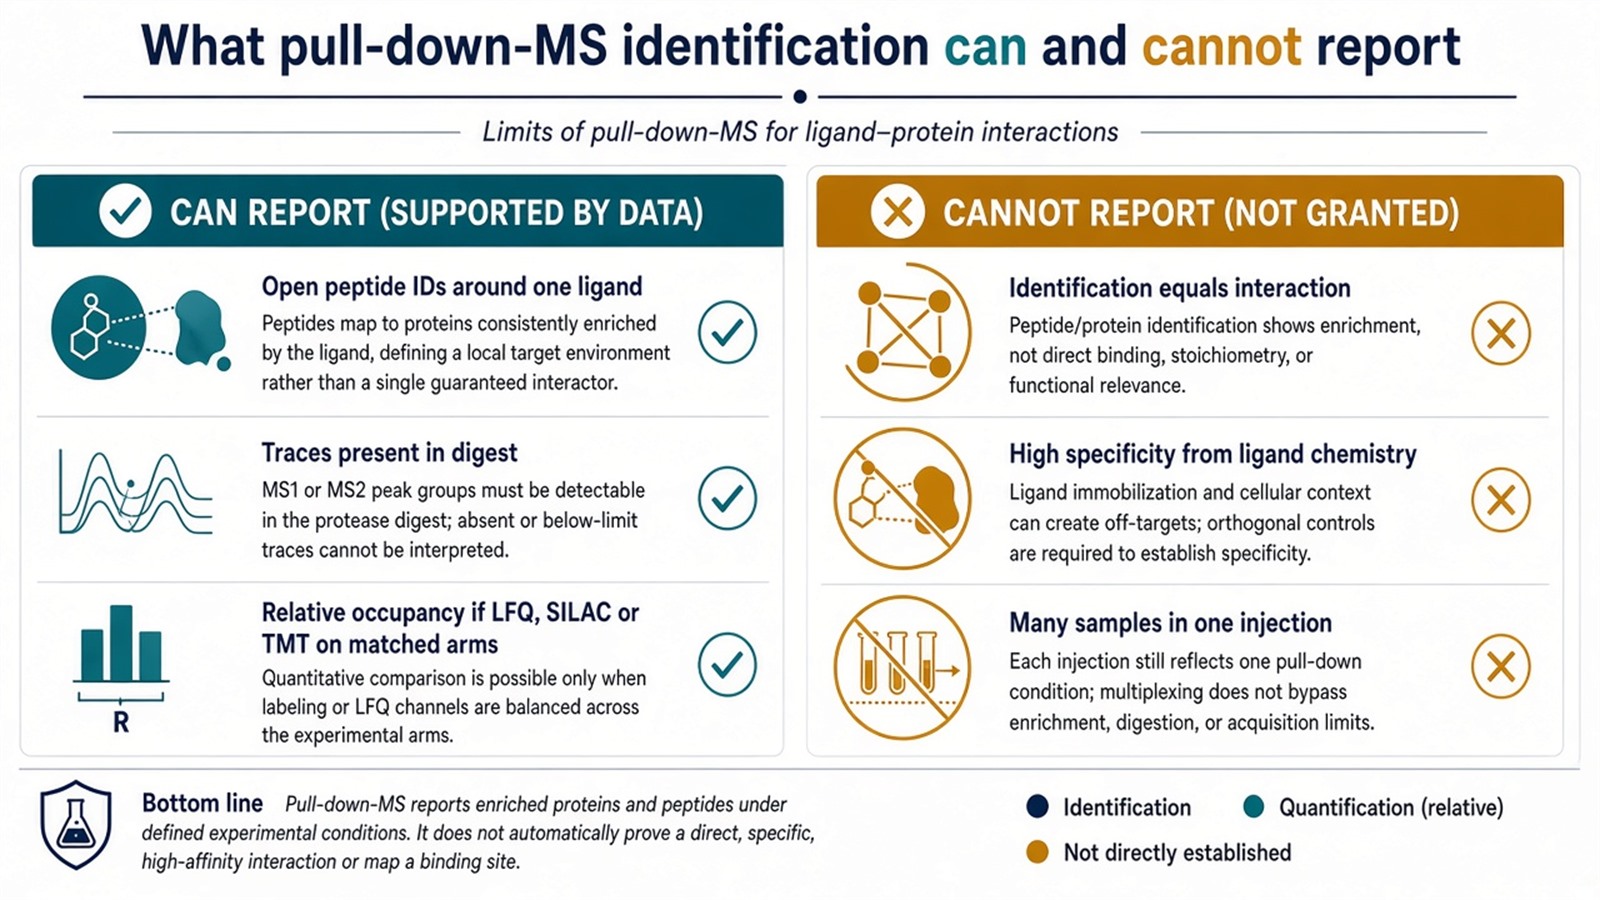
Figure 3. Supported when the empty-handle arm is processed: open IDs and relative enrichment. Not granted by ligand chemistry or MS sensitivity: low background, simultaneous multi-sample throughput, or a PPI proof.
Experimental Limits
The method reduces a mixture to what the ligand, the wash, and the search engine can report. Diagnostic language sits outside ordinary research reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable identification plan names the ligand class, the empty-handle arm, the elution or on-bead digest route, and the claim that will be allowed, before the first lysis. MtoZ Biolabs scopes pull-down with Western or Orbitrap-class LC-MS/MS. Interpretability still follows those arms, not the resin brand.
Once the ligand is specified and a GST fusion is the handle, the wet capture can be scoped as GST pull-down. Researchers who need a GST-bait plan before identification can review the MtoZ Biolabs GST Pull Down Assay Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What does pull-down plus MS identify?
Proteins present in the ligand eluate, as peptide-supported accessions. Interaction language requires enrichment against the empty-handle arm.
2. Is antibody capture a pull-down?
The bead logic is similar. The name and the control are IP or Co-IP, with an isotype, not a tag-only protein.
3. Does a GST fusion capture the target protein?
The GST fusion is the bait being presented. Prey is what may remain with that bait.
4. Can MS analyze many pull-down samples at once?
One injection is one digest, unless isobaric tags multiplex several arms. That is not unlimited throughput.
5. Is pull-down non-quantitative?
The capture reports occupancy after wash. LFQ, SILAC, or TMT can compare matched arms. They do not report a Kd.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Dunham W.H., Mullin M., Gingras A.C. (2012). Affinity-purification coupled to mass spectrometry: basic principles and strategies. Proteomics, 12, 1576-1590. https://doi.org/10.1002/pmic.201100523
- Mellacheruvu D., Wright Z., Couzens A.L., et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
- Vikis H.G., Guan K.L. (2004). Glutathione-S-transferase-fusion based assays for studying protein-protein interactions. Methods Mol. Biol., 261, 175-186. https://doi.org/10.1385/1-59259-762-9:175
- Phizicky E.M., Fields S. (1995). Protein-protein interactions: methods for detection and analysis. Microbiol. Rev., 59, 94-123. https://doi.org/10.1128/mr.59.1.94-123.1995
