



Fusion pull-down plus mass spectrometry (MS) is the right method when a soluble fusion exists, an empty-fusion arm can be processed in parallel, and the next table should be an open identification list around that recombinant bait. It is the wrong first method when the next sentence is an endogenous complex, a binding constant, a contact residue, or a proteome-wide network. The real advantages are capture without an IP-grade antibody, an open peptide list instead of one Western pair, and relative comparison across matched arms. The real disadvantages sit in the same chemistry: the bait is recombinant, tag and resin binders travel with the cargo, the wash removes weak partners, and the MS file is occupancy after wash. High sensitivity, high specificity, and a mapped interactome are not properties of the combination. They are claims that have to be earned by occupancy, the empty-fusion arm, and a named orthogonal assay.
A page that lists those slogans as inherent strengths, then lists data analysis as the only cost, misleads the choice against co-immunoprecipitation–MS (Co-IP-MS), crosslinking MS, or a nominated Western.
Researchers who already have a tagged fusion, an empty-tag control, and an open-list question can also review the MtoZ Biolabs Pull Down based Protein Analysis Service with Mass Spectrometry for project-specific feasibility and analysis planning.
Advantages That Hold Only Under Stated Conditions
These points are useful when the condition in the same sentence is true. They are not built-in virtues.
Handle capture without an antibody, if the fusion is soluble
Glutathione S-transferase (GST), His, Strep, biotin, or Flag can occupy a matching resin and present the cargo. That avoids an IP-grade antibody. The advantage disappears when the fusion is an inclusion body or a truncated stub. For the two linked events behind this capture, Principle of Fusion Protein Interaction Analysis Using Pull-Down and MS provides a complementary principle.
Open identification around one bait, if empty-fusion is digested
MS can name proteins that were not nominated for Western blot. Ligand binding does not, by itself, reduce background in the file. Specificity is the empty-fusion comparison. Without that arm, a large identification table is a larger background table. For how identification is read across ligand classes, Identification of Protein Interactions Using Pull-Down and Mass Spectrometry provides a complementary identification discussion.
Unnominated candidates, if enrichment is scored first
A lysate offered to the fusion can return proteins that no antibody was raised against. That is a candidate list, not analysis of the entire proteome and not a hypothesis-free map of all potential partners. Subtracting every empty-fusion identification deletes real partners that leak into the handle arm.
Relative occupancy across matched arms, if a quantitative method is chosen
Label-free quantification on paired runs is the default. SILAC applies when the prey culture can be labeled. Isobaric tags label after digestion. Pull-down is not inherently non-quantitative. Intensity is still recovery after wash, not a binding constant. For the GST-specific coupling, GST Pull-Down Assay Combined with Mass Spectrometry for Protein-Protein Interaction Analysis provides a complementary GST–MS page.
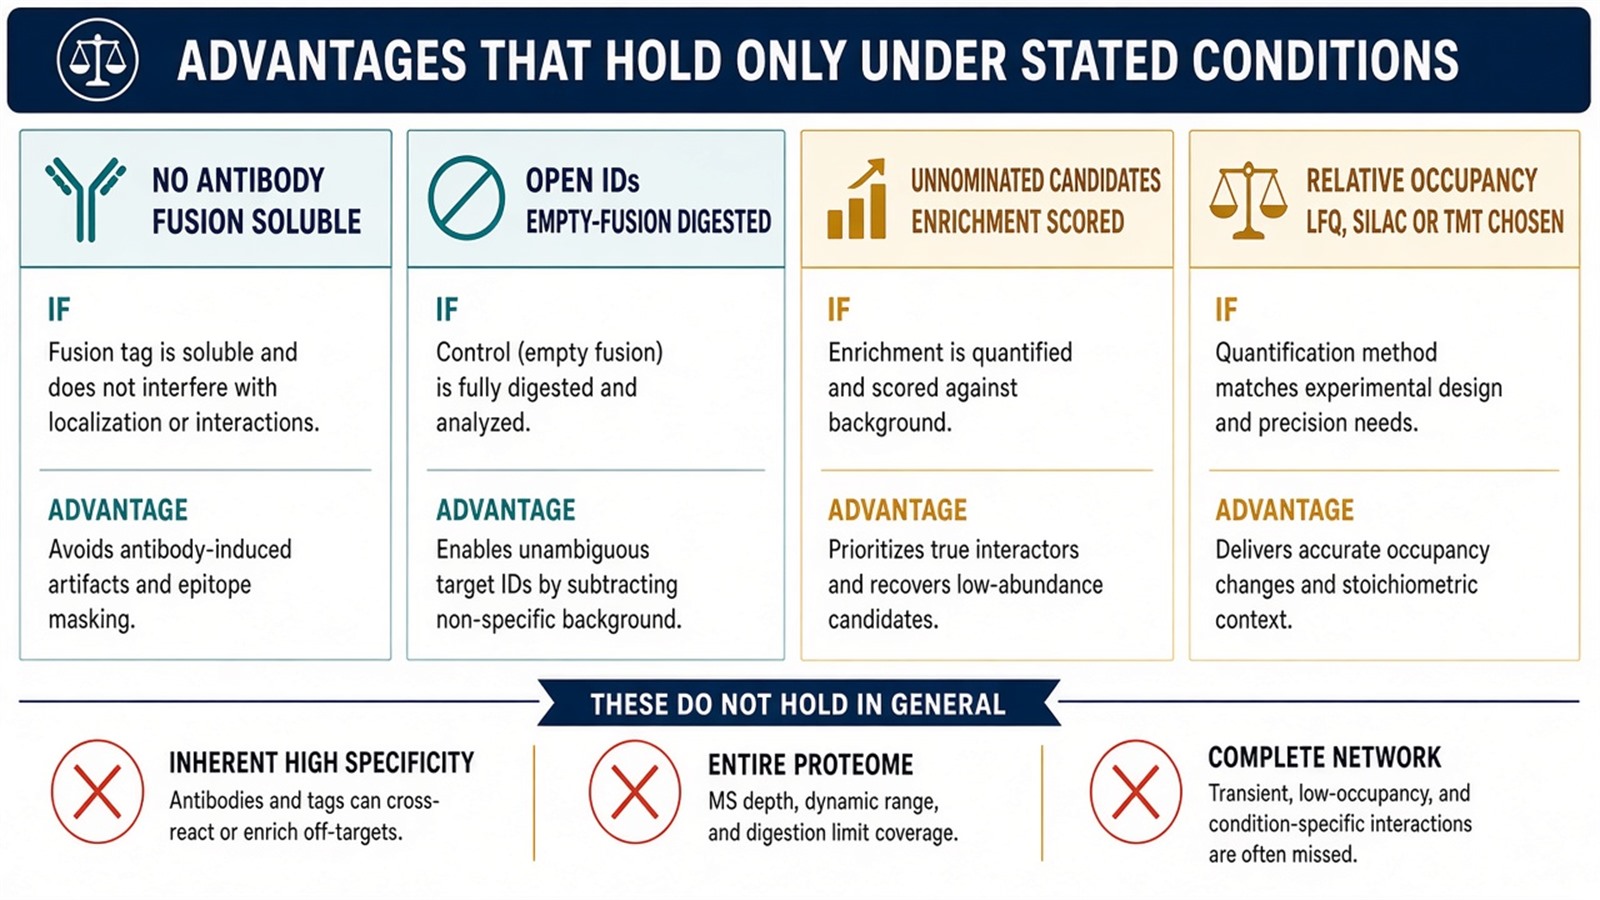
Figure 1. Supported when both arms exist: antibody-free capture, an open ID table, and relative occupancy. Not granted by tag chemistry or MS precision: low background or a complete network.
Write the empty-fusion arm before calling the method specific. Sensitivity of the spectrometer cannot invent a handle control.
Disadvantages That Sit in the Chemistry
These limits are structural. They are not beginner errors, and more bioinformatics time does not remove them.
Recombinant bait, not an endogenous complex
The fusion is expressed and immobilized. A tissue or cell lysate offered to that bead is still a tube reaction. Only an untagged antibody Co-IP can support an endogenous-complex sentence. For when Co-IP is the better first method, Advantages and Disadvantages of Co-Immunoprecipitation in Protein Interaction Analysis provides a complementary Co-IP decision.
Tag and resin binders travel with the cargo
GST dimers, His-binding proteins, streptavidin contaminants, and bead binders occupy the same eluate as cargo partners. False positives are expected. They are estimated by empty-fusion, not eliminated by a more specific ligand slogan.
The wash creates false negatives
Low-abundance or weak partners can be absent because they failed the wash, not because the instrument lacked sensitivity. Raising salt to “remove nonspecific proteins” also strips true weak contacts.
The table is not a network, a structure, or a copy-number assay
Database matching produces accessions. Functional annotation and network drawing are later overlays. Complex, time-consuming analysis is real work. It is not the core disadvantage. The core disadvantage is treating that overlay as a result of the pull-down. For how GST eluates are scored as bait versus GST-only, Analysis of Protein-Protein Interactions Using GST Pull-Down Assay provides a complementary GST analysis path.
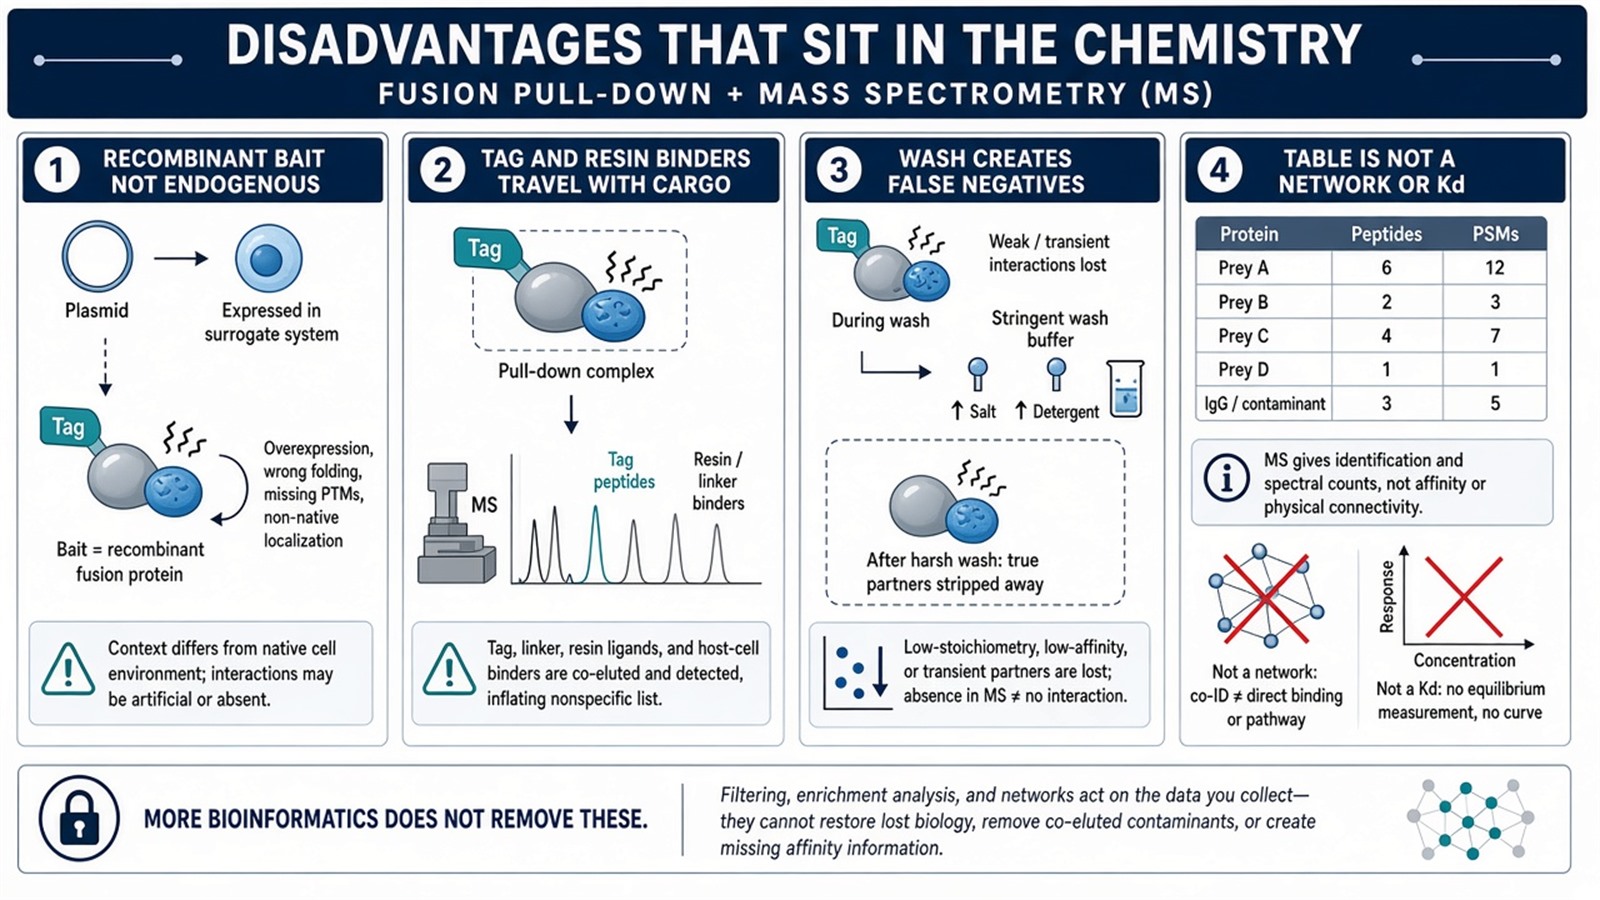
Figure 2. False positives follow the tag. False negatives follow the wash. Dependence on buffer and tag is the method, not an optional complication.
When This Combination Is Not the First Method
Choose a different first assay when the claim is already named.
An endogenous complex needs Co-IP-MS with an IP-grade antibody. A binding site or residue proximity needs crosslinking MS or a structural method. A dissociation constant needs surface plasmon resonance, isothermal titration calorimetry, or an equivalent biophysical assay. Live-cell proximity needs BioID, TurboID, or APEX. A nominated pair that a Western can settle does not need an open MS table.
Buffer, wash, and tag choice change who remains. Protein degradation during digest changes who is sequenced. Those dependencies are why the empty-fusion arm and a written wash belong in the plan, not why the method is too fragile to use.
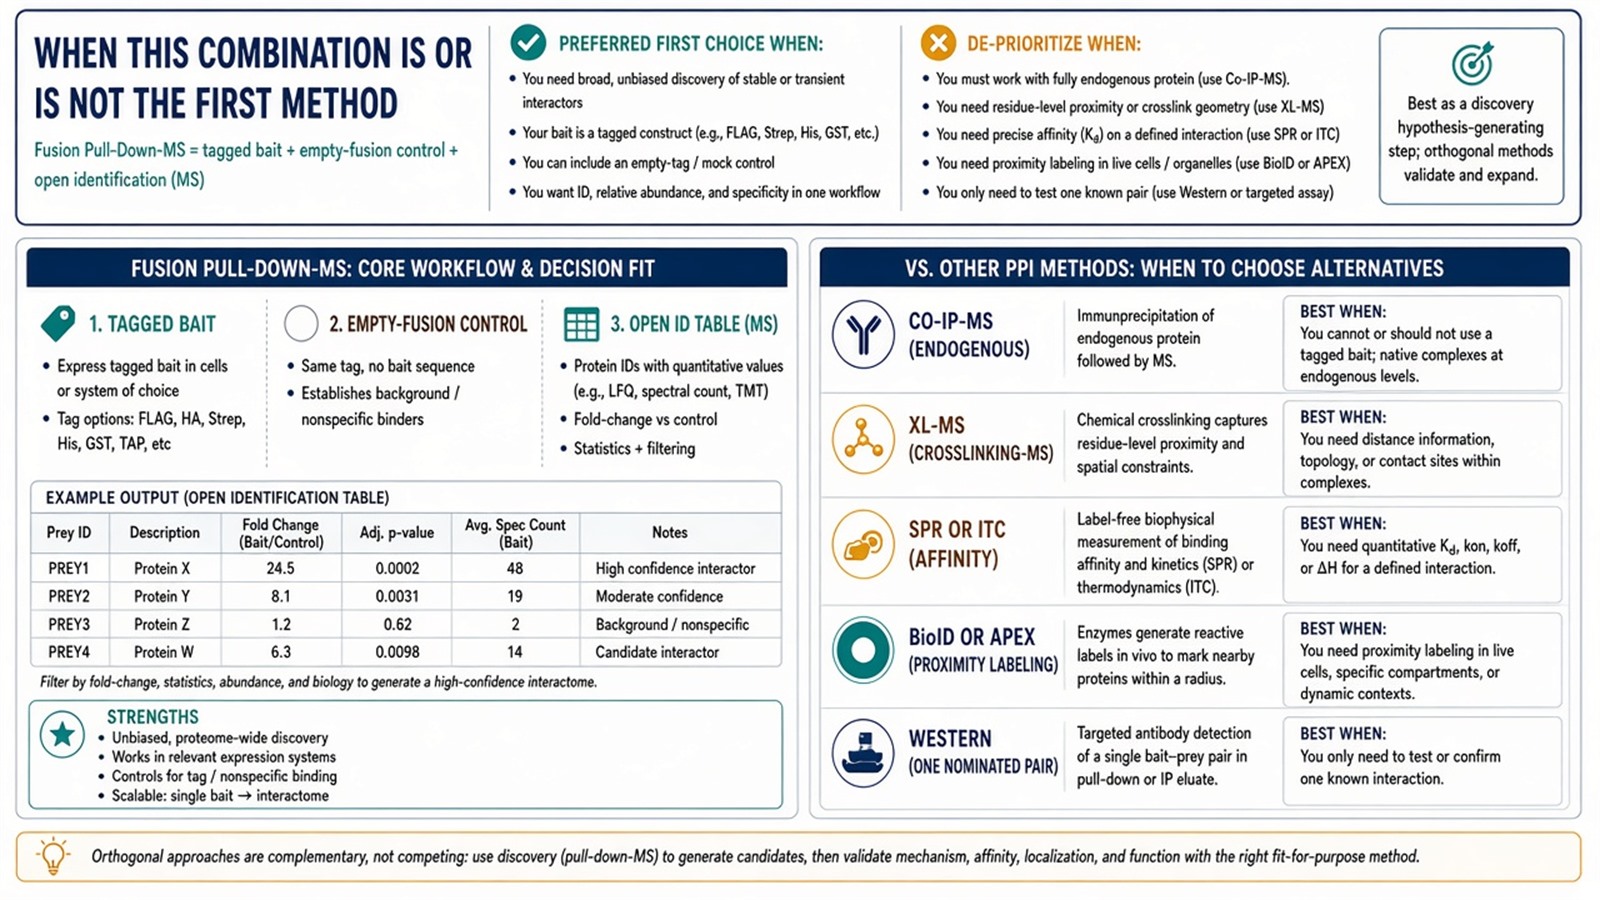
Figure 3. Right first method: tagged bait, empty-fusion, open IDs. Wrong first method: endogenous sentence, Kd, contact residue, or proteome-wide map.
Experimental Limits
The combination reduces a mixture to what the tag, the resin, the wash, and the search engine can report. Diagnostic language sits outside ordinary research reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable decision names the next sentence, the empty-fusion arm, and the claim that will be allowed, before the first induction. MtoZ Biolabs scopes fusion pull-down with Western or Orbitrap-class LC-MS/MS. Interpretability still follows those conditions, not the resin brand.
Once the handle is specified as GST and the wet capture is the bottleneck, the project can be scoped as GST pull-down. Researchers who need a GST-bait plan before identification can review the MtoZ Biolabs GST Pull Down Assay Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What is the main advantage of fusion pull-down plus MS?
An open identification table around a recombinant bait without an IP-grade antibody, if empty-fusion is run.
2. Does ligand specificity make the result specific?
No. The ligand captures the handle. Specificity of a prey is the empty-fusion comparison.
3. Can this method map a protein-interaction network?
One bait returns a wash-stable neighborhood. A network is a later overlay after many baits and orthogonal assays.
4. Why do false positives and false negatives persist?
Tag and resin binders remain after wash. Weak or low-occupancy partners leave during wash. Both are chemistry, not only operator error.
5. When should Co-IP-MS be chosen instead?
When the bait is untagged and the sentence must be endogenous capture from a cell or tissue lysate.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Dunham W.H., Mullin M., Gingras A.C. (2012). Affinity-purification coupled to mass spectrometry: basic principles and strategies. Proteomics, 12, 1576-1590. https://doi.org/10.1002/pmic.201100523
- Mellacheruvu D., Wright Z., Couzens A.L., et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
- Vikis H.G., Guan K.L. (2004). Glutathione-S-transferase-fusion based assays for studying protein-protein interactions. Methods Mol. Biol., 261, 175-186. https://doi.org/10.1385/1-59259-762-9:175
- Phizicky E.M., Fields S. (1995). Protein-protein interactions: methods for detection and analysis. Microbiol. Rev., 59, 94-123. https://doi.org/10.1128/mr.59.1.94-123.1995
