



Co-IP results answer which proteins remain with an endogenous bait after antibody capture, Input and IgG contrast, and either targeted detection or LC-MS/MS identification. A result can be written only after the antibody recovers the native bait and enrichment is scored against matched controls. The readout fits cells or tissues when an IP-competent antibody exists and the claim is co-recovery, not binary direct binding (Hein et al. 2015). Prefer this reading when Input and IgG were processed with the bait IP. Reassess a band, an IP-MS list, or a negative call when the antibody is Western-only, controls are missing, or a pathway map is used to complete the figure.
Researchers who already have Co-IP or IP-MS arms and a written result sentence can also review the MtoZ Biolabs IP-Mass Spectrometry Service and Protein-Protein Interaction Analysis for project-specific result review.
What a Co-IP Result Is Allowed to Say
A Co-IP result is not the blot image, the protein table, or a network drawn from leftover names. The result sentence is narrower: protein B is enriched with bait A versus matched IgG inside this capture recipe, after bait recovery was shown versus Input. That sentence can come from a targeted blot or from IP-MS. It cannot be upgraded to a disease mechanism, a complete interactome, or a direct contact.
Western blot results test a named prey when a detection reagent exists. IP-MS results extend the same claim to an open eluate. Identification depth in a Co-IP-style eluate is typically hundreds to about a thousand proteins, depending on the sample. A longer list is not a stronger result. Readout choice is compared in IP-MS vs Co-IP Western Blot: Which Method Fits Your Study?. How those same gates are used as an analysis workflow is covered in Co-Immunoprecipitation Analysis.
The result fits cultured cells or tissues when the antibody is supplied by the study. It is a poor first readout when tagging is required because no IP-competent antibody exists, or when the claim is reconstituted binding of a purified domain.
A Co-IP result reports candidate co-recovery under stated controls. It does not certify that two proteins contact each other.
Antibody Specificity Before a Result Is Called
Antibody specificity is the first result gate. If the capturing reagent does not recover the intended native bait, later bands or protein names are not Co-IP results. Application-specific validation remains necessary because Western performance does not establish IP performance (Uhlen et al. 2016). How to qualify that reagent is covered in How to Choose the Right Antibody for Co-IP Assays?.
Record clone, lot, host species, and bead pairing next to the result. A lot change can move bait recovery and then look like a biological change. Extra antibody that fills the eluate with chain signal makes both blot and IP-MS results noisier. Empty-tag controls from AP-MS do not replace antibody documentation on a Co-IP result.
If no IP-competent antibody exists, stop calling Co-IP results and change handle. Tagged AP-MS or GST pull-down produce different result classes, compared in What Are the Main Differences Between GST-Pull Down and CoIP.
Input and IgG Decide Whether the Result Stands
Input and IgG are the denominators of a Co-IP result. They are not decorative lanes.
Input is taken before capture. It shows whether the bait, and when relevant the named prey, existed in the extract. A missing IP band or a missing bait protein group is an expression or extraction problem if Input is empty. It is a capture problem if Input is positive.
IgG or bead controls must use the same lysate batch, lysis, wash, elution, and, for IP-MS, the same digestion and search. Preclearing is not a result substitute for IgG. If IgG was washed unlike the bait IP, specificity cannot be read. What those arms must look like as submitted material is covered in Requirements for Samples in Protein Immunoprecipitation-Mass Spectrometry.
Result check |
What a readable result looks like |
What the result cannot say |
|---|---|---|
Antibody specificity |
Intended bait recovered; no dominant off-target |
Partner names from an off-target capture |
Input |
Bait present before capture when expected |
A missing IP read as a missing interaction |
IgG or bead |
Shared background without bait-specific recovery |
Every remaining name is bait-specific |
Enrichment efficiency |
Bait higher in bait IP than in Input and IgG |
A long MS table or a single remaining blot band |
Same-protein comparison |
One protein compared across matched arms |
Unrelated proteins ranked as tighter binders |
A practical reading sequence is:
- Write the co-recovery sentence the figure is asked to support.
- Confirm native bait recovery versus Input.
- Compare the same bait and prey against matched IgG.
- Keep only names that pass that contrast.
- Leave a shortlist for an orthogonal pair assay.
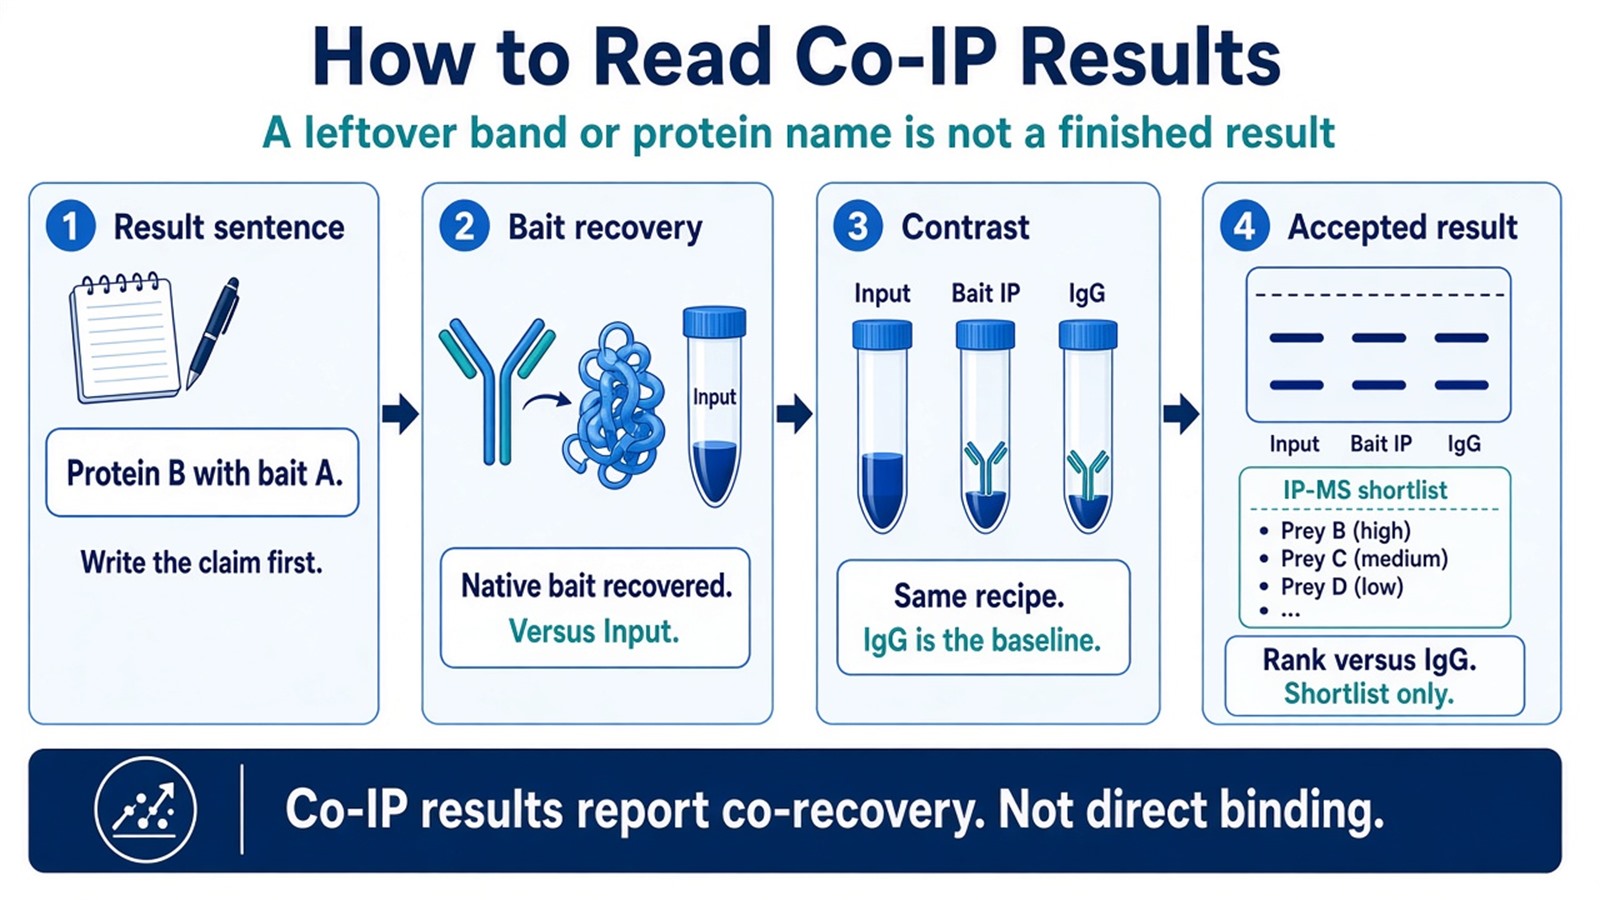
Figure 1. A Co-IP result starts after bait recovery is shown. A leftover band or protein name is not a finished result.
Enrichment Efficiency and Mass Spectrometry Credibility
Enrichment efficiency decides whether capture worked. If the bait is weak versus Input, stop partner interpretation. Relative intensity can compare the same protein across Input, IgG, and bait IP, or across treatments that used one antibody and one wash. Limits of that comparison are covered in Can Co-Immunoprecipitation (Co-IP) Be Used for Quantitative Comparison. Intensity does not rank unrelated proteins as tighter binders.
For blot results, a prey band in the bait IP, reduced in IgG, and consistent with Input can support a relative co-enrichment sentence for that named protein. A missing band can mean the prey was absent, the prey antibody failed, or wash chemistry stripped the pair.
For IP-MS results, identification of a protein is not a confidence call. Ranking requires bait recovery, Input-aware enrichment, and a declared contrast against IgG. Probabilistic scoring such as SAINT uses bait versus control counts to assign a probability that a prey is specifically co-purified (Choi et al. 2011). A contaminant catalog such as the CRAPome is a prior on control frequency (Mellacheruvu et al. 2013). Pathway or network software can annotate a shortlist. It cannot repair a missing IgG arm or convert a table into a validated interactome.
After those gates, a protein-level value can support: protein B is enriched with bait A versus matched IgG in this Co-IP result. Peptide support, antibody clone, and wash chemistry belong next to that statement.
If the bait is missing from the result table, diagnose before ranking, as in Why Was the Target Protein Not Detected by IP-MS?. Protocol-level search matching is covered in Immunoprecipitation Mass Spectrometry Protocol.
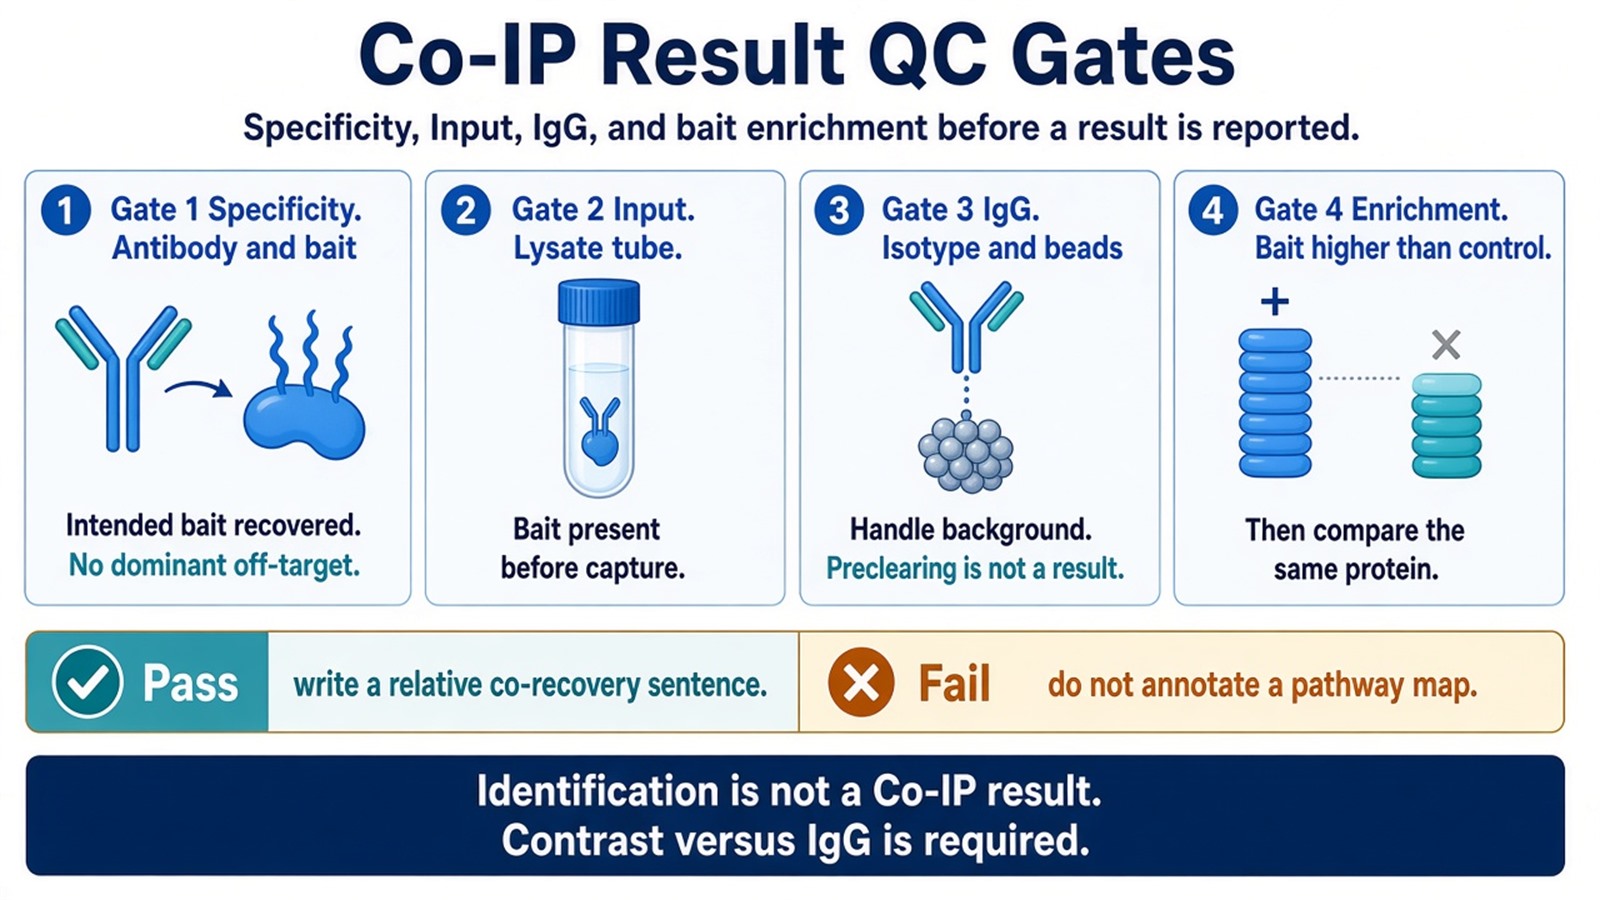
Figure 2. Mass spectrometry credibility is a result outcome. Network enrichment does not replace Input and IgG.
Common Misreadings of Positive and Negative Results
The common result bias is writing Co-IP as highly specific because an antibody was used. Another bias is treating every co-eluting protein, or every darker band, as a validated partner. Over-washing can erase the association the claim needed and then look like a clean negative. Under-washing makes bait IP and IgG similar, so subtraction makes every name look specific.
Do not write a successful bait IP as proof that remaining proteins contact the bait. Do not write a negative Co-IP result as proof that no interaction exists in the cell. Absence can mean the wrong antibody, epitope masking, over-washing, or a bait that is not expressed in the tested state. How the wet-lab protocol is designed before the result is generated is covered in Designing an Efficient Co-immunoprecipitation (Co-IP) Protocol.
Prefer Co-IP or IP-MS results when native bait recovery can be shown. Prefer GST pull-down or tagged AP-MS when no IP-competent antibody exists. Prefer a defined-pair assay when the next sentence is direct contact. Mixed enrichment-MS projects can be reviewed under the MS-Based Protein-Protein Interaction Analysis Service.
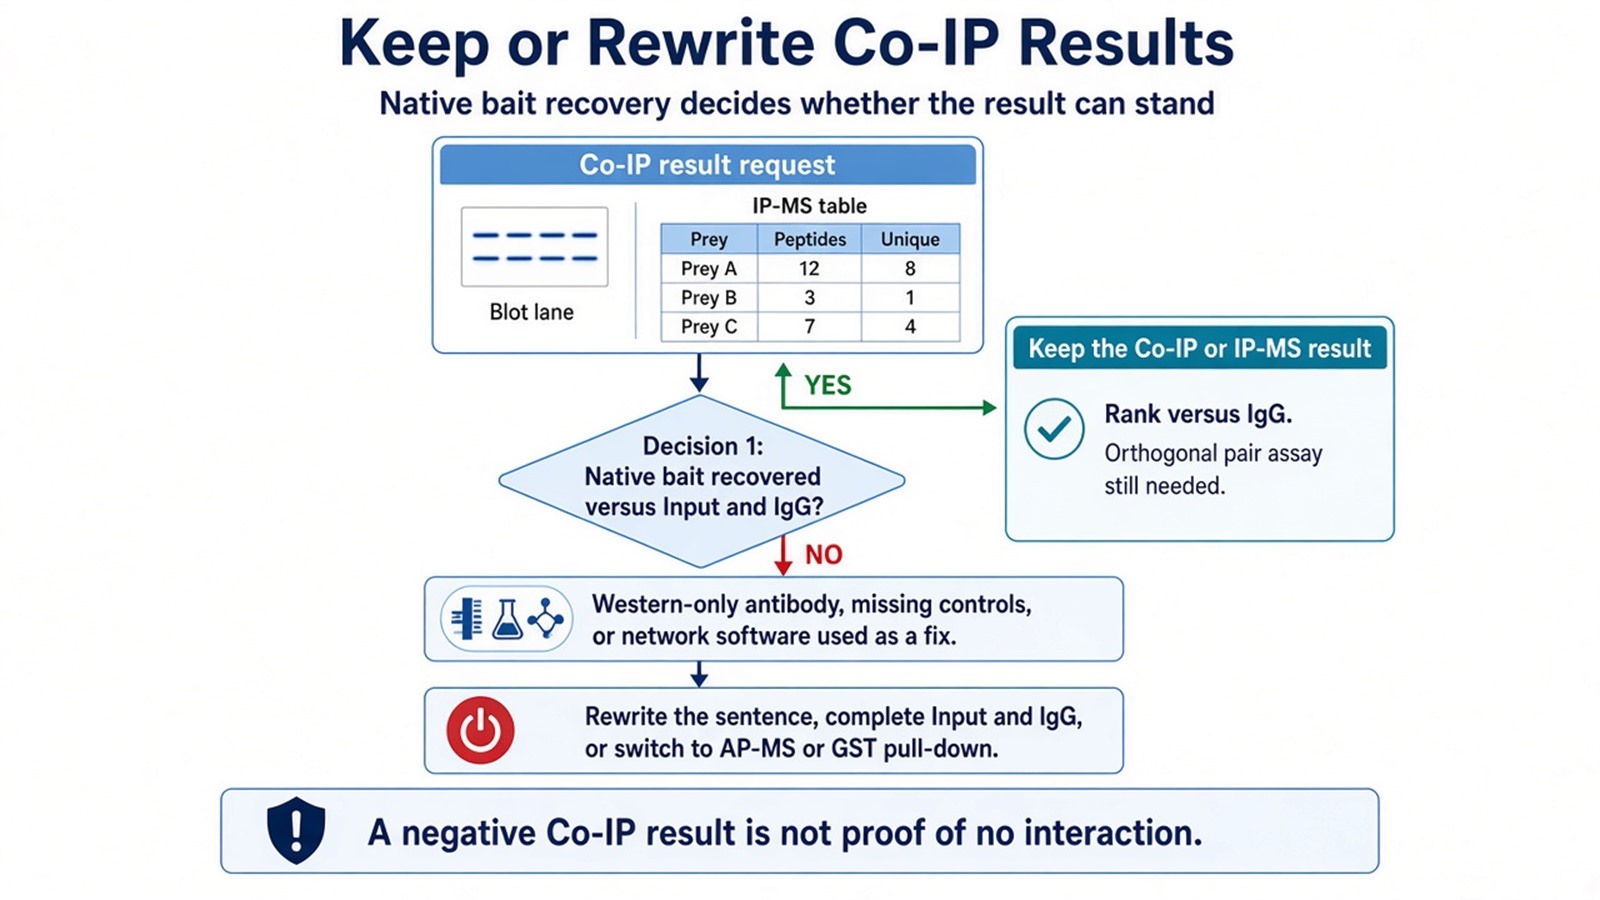
Figure 3. Keep a Co-IP result only while native bait recovery is documented. A pathway map is not a completed result.
A Co-IP result supports candidate co-recovery after antibody, Input, IgG, and bait enrichment are shown. It does not certify a binary interactome.
Once the claim, antibody status, Input and IgG results, and the blot or IP-MS readout are defined, the project can be scoped as Co-IP or IP-MS result review. Researchers can review the MtoZ Biolabs IP-Mass Spectrometry Service for sample evaluation and result planning.
Frequently Asked Questions
1. Does a band in the bait-IP lane count as a Co-IP result?
No. The same band must be read against Input and matched IgG. A single lane is an observation, not a result.
2. Does identification in an IP-MS table count as a Co-IP result?
No. Identification shows a protein was present in the eluate. A result requires bait recovery and enrichment versus IgG.
3. Can a negative Co-IP result prove that two proteins do not interact?
No. Absence can come from a weak antibody, over-washing, epitope masking, or a bait that was not expressed in the tested state.
4. Can pathway software finish Co-IP or IP-MS results?
No. Pathway tools annotate a shortlist. They do not replace bait recovery or IgG contrast.
5. When should a Co-IP result move to a pair assay?
Move when the next written sentence is direct contact or affinity. Keep the Co-IP shortlist as the input to that assay, not as the proof.
Reference
- M. Uhlen, A. Bandrowski, S. Carr, A. Edwards, J. Ellenberg, E. Lundberg, D. L. Rimm, H. Rodriguez, T. Hiltke, M. Snyder, T. Yamamoto (2016). A proposal for validation of antibodies. Nat. Methods, 13, 823-827. https://doi.org/10.1038/nmeth.3995
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- H. Choi, B. Larsen, Z.Y. Lin, A. Breitkreutz, D. Mellacheruvu, D. Fermin, Z.S. Qin, M. Tyers, A.C. Gingras, A.I. Nesvizhskii (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
