



Co-immunoprecipitation analysis, or Co-IP analysis, asks which proteins remain with an endogenous bait after antibody capture, Input and IgG contrast, and either targeted detection or LC-MS/MS identification. Analysis starts only after the antibody recovers the native bait and enrichment is scored against matched controls. The method fits cells or tissues when an IP-competent antibody exists and the claim is co-recovery, not binary direct binding (Hein et al. 2015). Prefer this analysis when Input and IgG were processed with the bait IP. Reassess when the antibody is Western-only, controls are missing, or a network figure is used to repair a weak bait.
Researchers who already have Co-IP or IP-MS arms and a written interaction claim can also review the MtoZ Biolabs IP-Mass Spectrometry Service and Protein-Protein Interaction Analysis for project-specific analysis planning.
What Co-Immunoprecipitation Analysis Can Claim
Co-IP analysis is not a bead workflow plus a blot or a search report. The analysis sentence is narrower: protein B is enriched with bait A versus matched IgG inside this capture recipe, after bait recovery was shown versus Input. That sentence can be read from a targeted blot or from IP-MS. It cannot be upgraded to a disease mechanism, a complete interactome, or a direct contact.
IP-MS extends the same analysis to an open eluate. Identification depth in a Co-IP-style eluate is typically hundreds to about a thousand proteins, depending on the sample. Depth is not a stronger analysis. Readout choice is compared in IP-MS vs Co-IP Western Blot: Which Method Fits Your Study?. How the wet-lab protocol is designed before analysis is covered in Designing an Efficient Co-immunoprecipitation (Co-IP) Protocol.
The analysis fits cultured cells or tissues when the antibody is supplied by the study and a protein reference database exists for an MS ending. It is a poor first analysis when tagging is required because no IP-competent antibody exists, or when the claim is reconstituted binding of a purified domain.
Co-immunoprecipitation analysis reports candidate co-recovery under stated controls. It does not certify that two proteins contact each other.
Antibody Specificity Before Any Partner Is Analyzed
Antibody specificity is the first analysis gate. If the capturing reagent does not recover the intended native bait, later bands or protein names are not Co-IP results. Application-specific validation remains necessary because Western performance does not establish IP performance (Uhlen et al. 2016). How to qualify that reagent is covered in How to Choose the Right Antibody for Co-IP Assays?.
Record clone, lot, host species, and bead pairing next to the analysis. A lot change can move bait recovery and then look like a biological change. Extra antibody that fills the eluate with chain signal makes both blot and IP-MS analysis noisier. Empty-tag controls from AP-MS do not replace antibody documentation in a Co-IP analysis.
If no IP-competent antibody exists, stop the Co-IP analysis and change handle. Tagged AP-MS or GST pull-down are different analyses, compared in What Are the Main Differences Between GST-Pull Down and CoIP.
Input and IgG Are the Analysis Baseline
Input and IgG are the denominators of Co-IP analysis. They are not decorative lanes.
Input is taken before capture. It shows whether the bait, and when relevant the named prey, existed in the extract. A missing IP band or a missing bait protein group is an expression or extraction problem if Input is empty. It is a capture problem if Input is positive.
IgG or bead controls must use the same lysate batch, lysis, wash, elution, and, for IP-MS, the same digestion and search. Preclearing is not an analysis substitute for IgG. If IgG was washed unlike the bait IP, specificity cannot be analyzed. What those arms must look like as submitted material is covered in Requirements for Samples in Protein Immunoprecipitation-Mass Spectrometry.
Analysis check |
What a pass looks like |
What the result cannot say |
|---|---|---|
Antibody specificity |
Intended bait recovered; no dominant off-target |
Partner names from an off-target capture |
Input |
Bait present before capture when expected |
A missing IP read as a missing interaction |
IgG or bead |
Shared background without bait-specific recovery |
Every remaining name is bait-specific |
Enrichment efficiency |
Bait higher in bait IP than in Input and IgG |
A long MS table or a single remaining blot band |
Same-protein comparison |
One protein compared across matched arms |
Unrelated proteins ranked as tighter binders |
A practical analysis sequence is:
- Confirm the written co-recovery claim.
- Confirm native bait recovery versus Input.
- Compare the same bait and prey against matched IgG.
- Rank only names that pass that contrast.
- Keep a shortlist for an orthogonal pair assay.
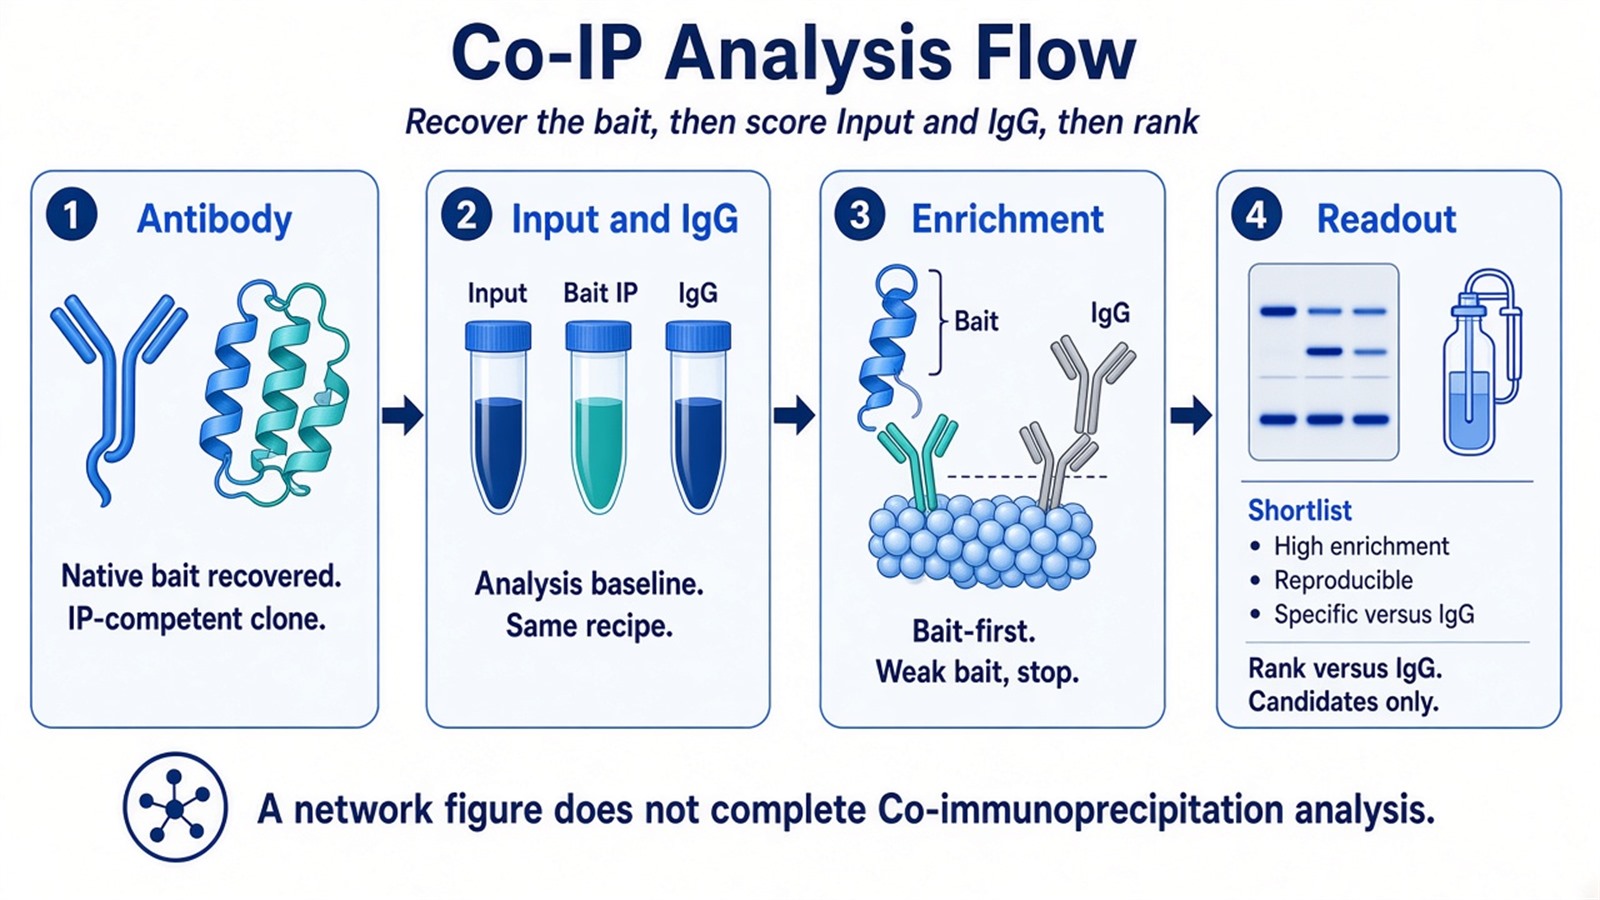
Figure 1. Co-IP analysis starts after bait recovery is shown. A bead cartoon is not an interpreted result.
Enrichment Efficiency and Mass Spectrometry Credibility
Enrichment efficiency is the analysis of whether capture worked. If the bait is weak versus Input, stop partner interpretation. Relative intensity can compare the same protein across Input, IgG, and bait IP, or across treatments that used one antibody and one wash. Limits of that comparison are covered in Can Co-Immunoprecipitation (Co-IP) Be Used for Quantitative Comparison. Intensity does not rank unrelated proteins as tighter binders.
For blot analysis, a prey band in the bait IP, reduced in IgG, and consistent with Input can support a relative co-enrichment sentence for that named protein. A missing band can mean the prey was absent, the prey antibody failed, or wash chemistry stripped the pair.
For IP-MS analysis, identification of a protein is not a confidence call. Ranking requires bait recovery, Input-aware enrichment, and a declared contrast against IgG. Probabilistic scoring such as SAINT uses bait versus control counts to assign a probability that a prey is specifically co-purified (Choi et al. 2011). A contaminant catalog such as the CRAPome is a prior on control frequency (Mellacheruvu et al. 2013). Pathway or network software can annotate a shortlist. It cannot repair a missing IgG arm.
After those gates, a protein-level value can support: protein B is enriched with bait A versus matched IgG inside this Co-IP analysis. Peptide support, antibody clone, and wash chemistry belong next to that statement.
If the bait is missing from the analysis table, diagnose before ranking, as in Why Was the Target Protein Not Detected by IP-MS?. Protocol-level search matching is covered in Immunoprecipitation Mass Spectrometry Protocol.
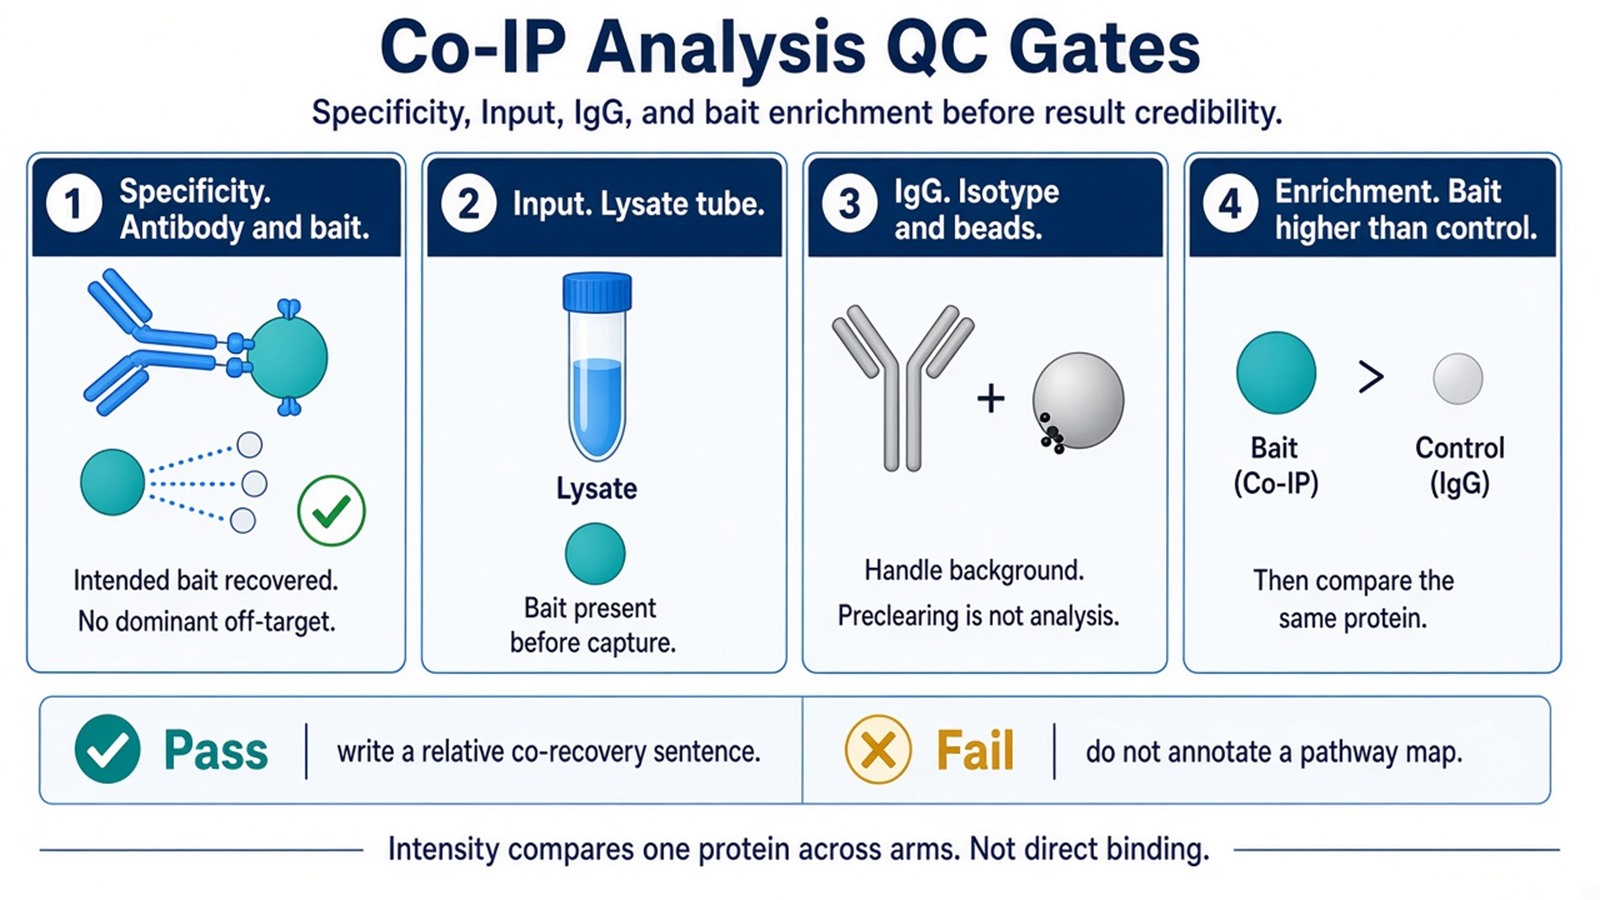
Figure 2. Mass spectrometry credibility is an analysis outcome. Network enrichment does not replace Input and IgG.
Common Biases and Result Boundaries
The common analysis bias is writing Co-IP as highly specific because an antibody was used. Another bias is treating every co-eluting protein, or every darker band, as a validated partner. Over-washing can erase the association the claim needed and then look like a clean negative. Under-washing makes bait IP and IgG similar, so subtraction makes every name look specific.
Do not write a successful bait IP as proof that remaining proteins contact the bait. Do not write a negative Co-IP analysis as proof that no interaction exists in the cell. Absence can mean the wrong antibody, epitope masking, over-washing, or a bait that is not expressed in the tested state.
Prefer Co-IP or IP-MS analysis when native bait recovery can be shown. Prefer GST pull-down or tagged AP-MS when no IP-competent antibody exists. Prefer a defined-pair assay when the next sentence is direct contact. Mixed enrichment-MS projects can be reviewed under the MS-Based Protein-Protein Interaction Analysis Service.
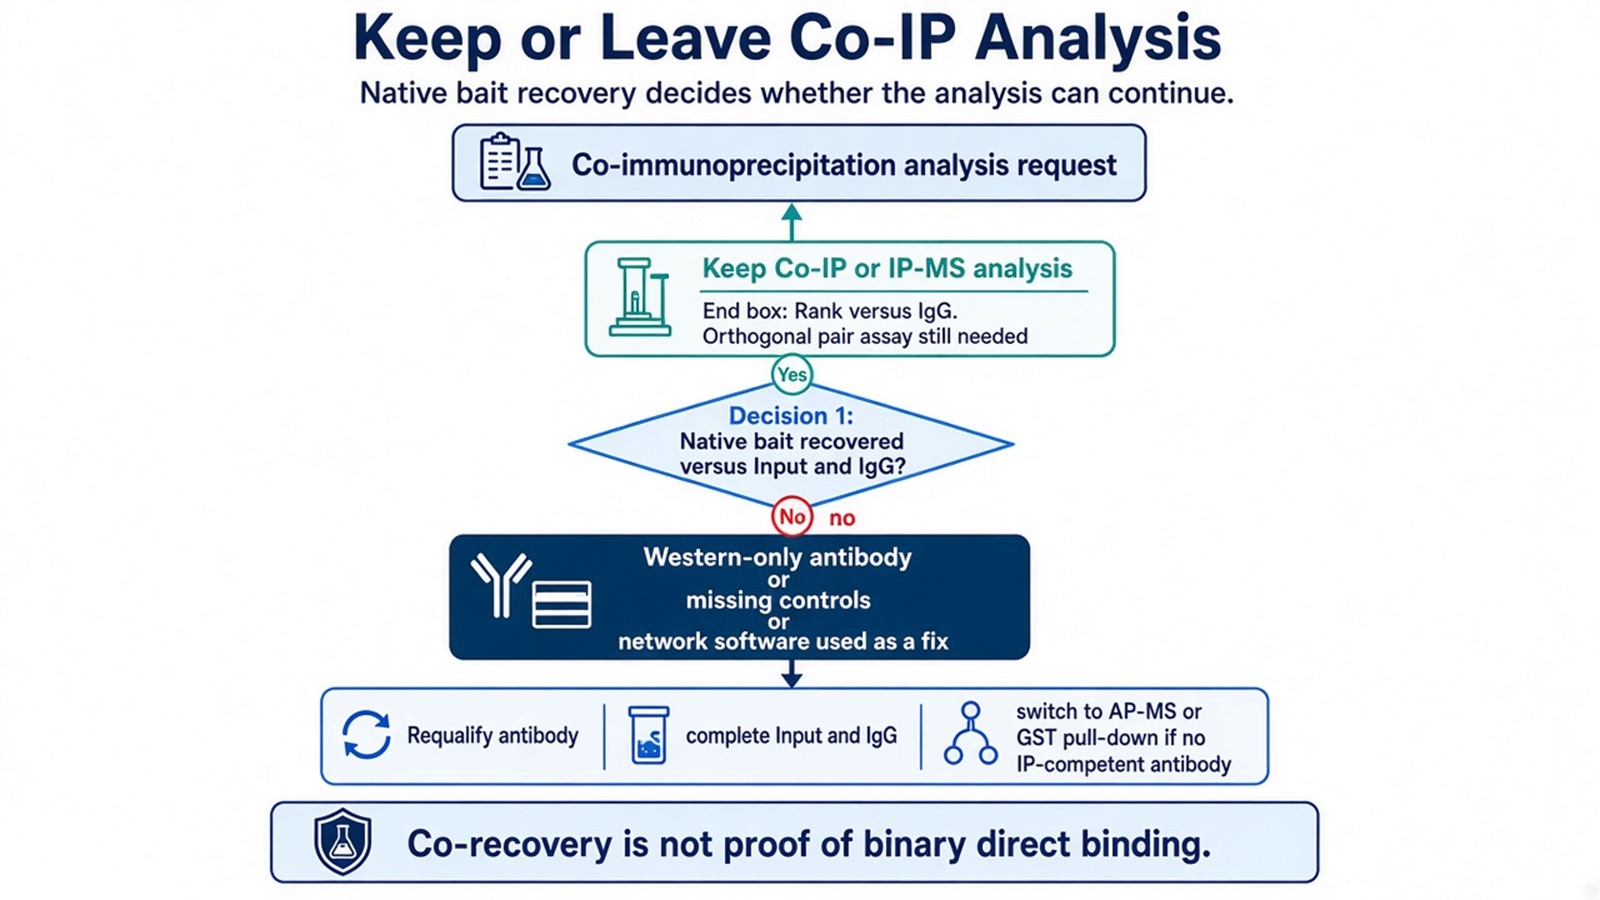
Figure 3. Keep Co-immunoprecipitation analysis only while native bait recovery is documented. A pathway map is not a completed analysis.
Co-immunoprecipitation analysis supports candidate co-recovery after antibody, Input, IgG, and bait enrichment are shown. It does not certify a binary interactome.
Once the claim, antibody status, Input and IgG results, and the blot or IP-MS readout are defined, the project can be scoped as Co-IP or IP-MS analysis. Researchers can review the MtoZ Biolabs IP-Mass Spectrometry Service for sample evaluation and analysis planning.
Frequently Asked Questions
1. Does a band in the bait-IP lane complete Co-IP analysis?
No. The same band must be interpreted against Input and matched IgG. A single lane is an observation, not an analysis.
2. Can pathway software finish an IP-MS Co-IP analysis?
No. Pathway tools annotate a shortlist. They do not replace bait recovery or IgG contrast.
3. Is Co-IP analysis more specific than GST pull-down because it uses an antibody?
No. Specificity is shown by bait recovery versus Input and IgG. An antibody handle can still pull off-targets.
4. Can I analyze two treatments if bait recovery differs?
Not as a partner comparison. Analyze bait recovery first. Prey intensity will follow capture if the bait collapsed in one arm.
5. When should Co-IP analysis move to a pair assay?
Move when the next written sentence is direct contact or affinity. Keep the Co-IP shortlist as the input to that assay, not as the proof.
Reference
- M. Uhlen, A. Bandrowski, S. Carr, A. Edwards, J. Ellenberg, E. Lundberg, D. L. Rimm, H. Rodriguez, T. Hiltke, M. Snyder, T. Yamamoto (2016). A proposal for validation of antibodies. Nat. Methods, 13, 823-827. https://doi.org/10.1038/nmeth.3995
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- H. Choi, B. Larsen, Z.Y. Lin, A. Breitkreutz, D. Mellacheruvu, D. Fermin, Z.S. Qin, M. Tyers, A.C. Gingras, A.I. Nesvizhskii (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
