



Analysis of protein complexes using AP-MS reports bait-centered co-enrichment after affinity capture and liquid chromatography–tandem mass spectrometry (LC-MS/MS). The complex sentence it can support is that a set of names was recovered with this bait under these wash and control conditions. That sentence is narrower than listing the subunits of a defined complex, assigning stoichiometry, or explaining how the assembly works in a pathway.
Signal transduction, metabolic regulation, and gene expression remain contexts in which a bait may be tested. They are not outputs of the spectrometer. One eluate does not become “the complex” because the heading says protein-complex research. For the technique definition behind the capture, What Is the AP-MS Technique? provides a complementary definition page. For the interaction sentence that stops at recovery with a bait, AP-MS Protein Interaction provides a complementary claim page.
Researchers who already have a tagged bait and need a ranked co-enrichment table can also review the MtoZ Biolabs Affinity Purification-Mass Spectrometry Service for project-specific feasibility and analysis planning.
What One Bait Eluate Is Not
A protein complex is a multi-subunit assembly that can be asked about experimentally. AP-MS asks a different question first: which proteins travel with this bait through the handle and the wash. Shared recovery across related baits can later mark a module. That overlap is still a co-enrichment pattern. It is not a complete subunit list, and it does not sort core members from hitchhikers.
Washing does not leave only the target protein complexes. The wash is a kinetic filter. Harsh salt or detergent can drop real neighbors. Gentle washing keeps bead and tag binders. Empty-tag, bead-only, or a matched non-bait arm ranks enrichment. Identification of a name in one purification is not membership. For that evidence split, Understanding AP-MS: A Comprehensive Guide provides a complementary evidence page.
Tag capture is the common AP-MS format on this site. Antibody capture of an endogenous subunit is usually planned as IP-MS. Those two formats do not share a control. An overexpressed tagged subunit can assemble a different object than the endogenous complex.
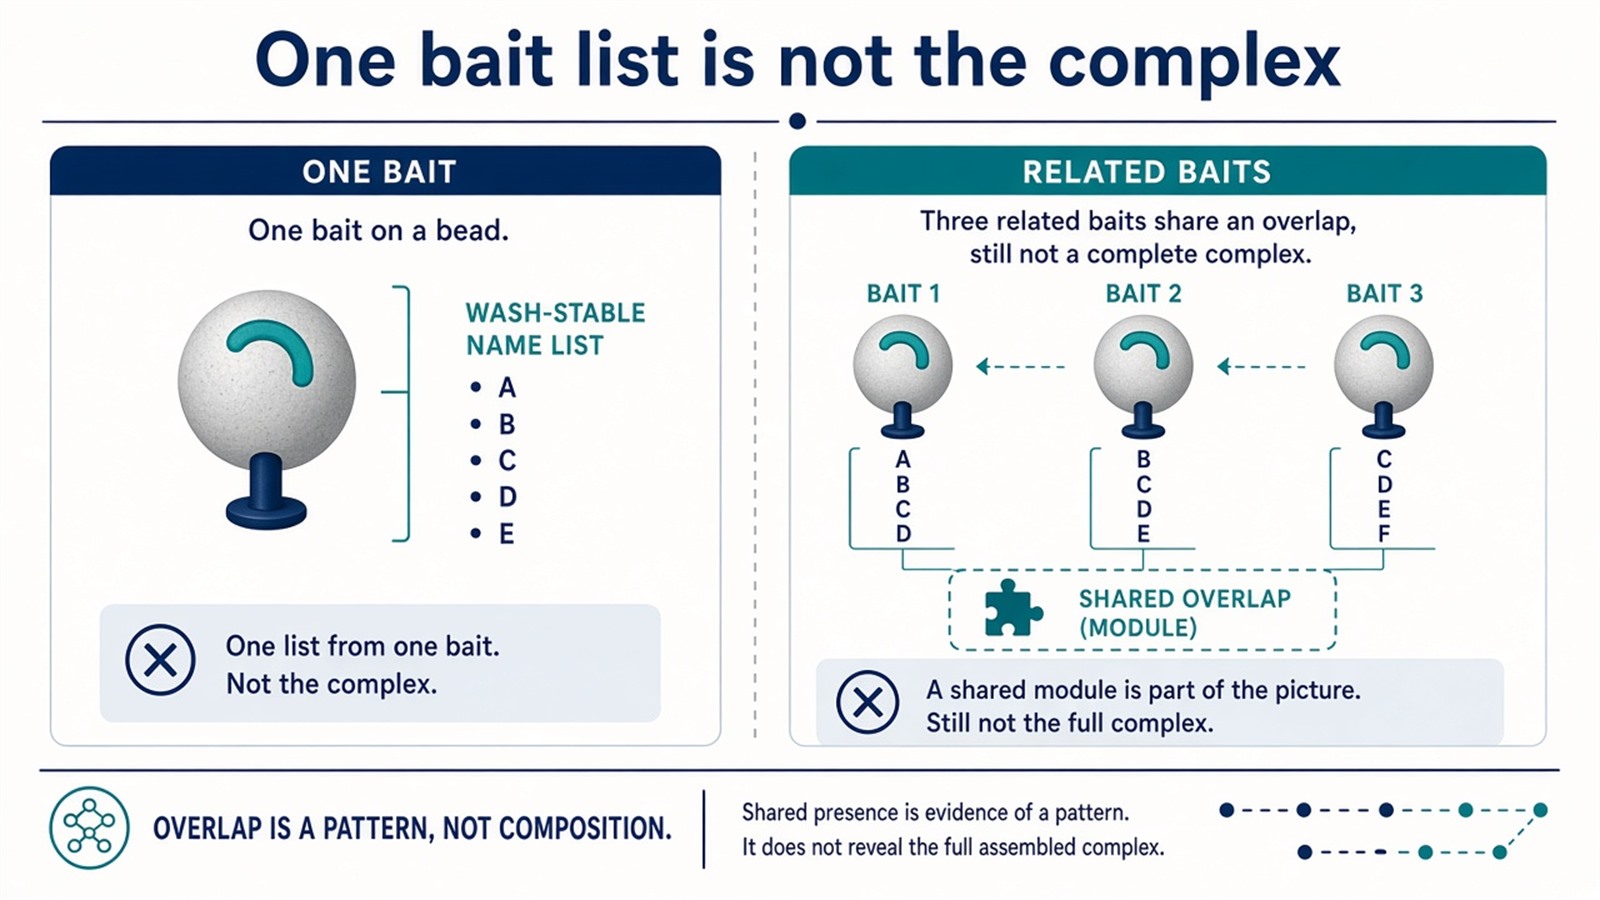
Figure 1. A single bait eluate is a wash-stable neighborhood around that bait. Overlap across related baits can mark a module. Neither drawing is a complete complex.
Write the complex sentence as recovery with this bait, or as overlap across named baits. Do not write it as composition, stoichiometry, or function.
Capture, Wash, and Peptide Naming
The original three-step description remains if each step is read as an evidence event. A handle occupies one subunit. The wash decides which associations remain. Digestion and LC-MS/MS name peptides in the eluate. The spectrometer does not separate intact complex proteins by mass and charge and then call that identification. Bottom-up spectra support peptide-to-protein assignment. They do not quantify a complex as an object.
Quantification, when designed, is a contrast: bait versus control, or condition A versus condition B under matched capture. It is not proof that complex composition changed in the cell. For the capture–wash–elute chain, How AP-MS Works provides a complementary mechanism page.
Two-handle TAP was built to reduce some first-resin binders around a tagged subunit. The second resin still does not return a native-state inventory. For that cassette, Tandem Affinity Purification Mass Spectrometry provides a complementary TAP page.
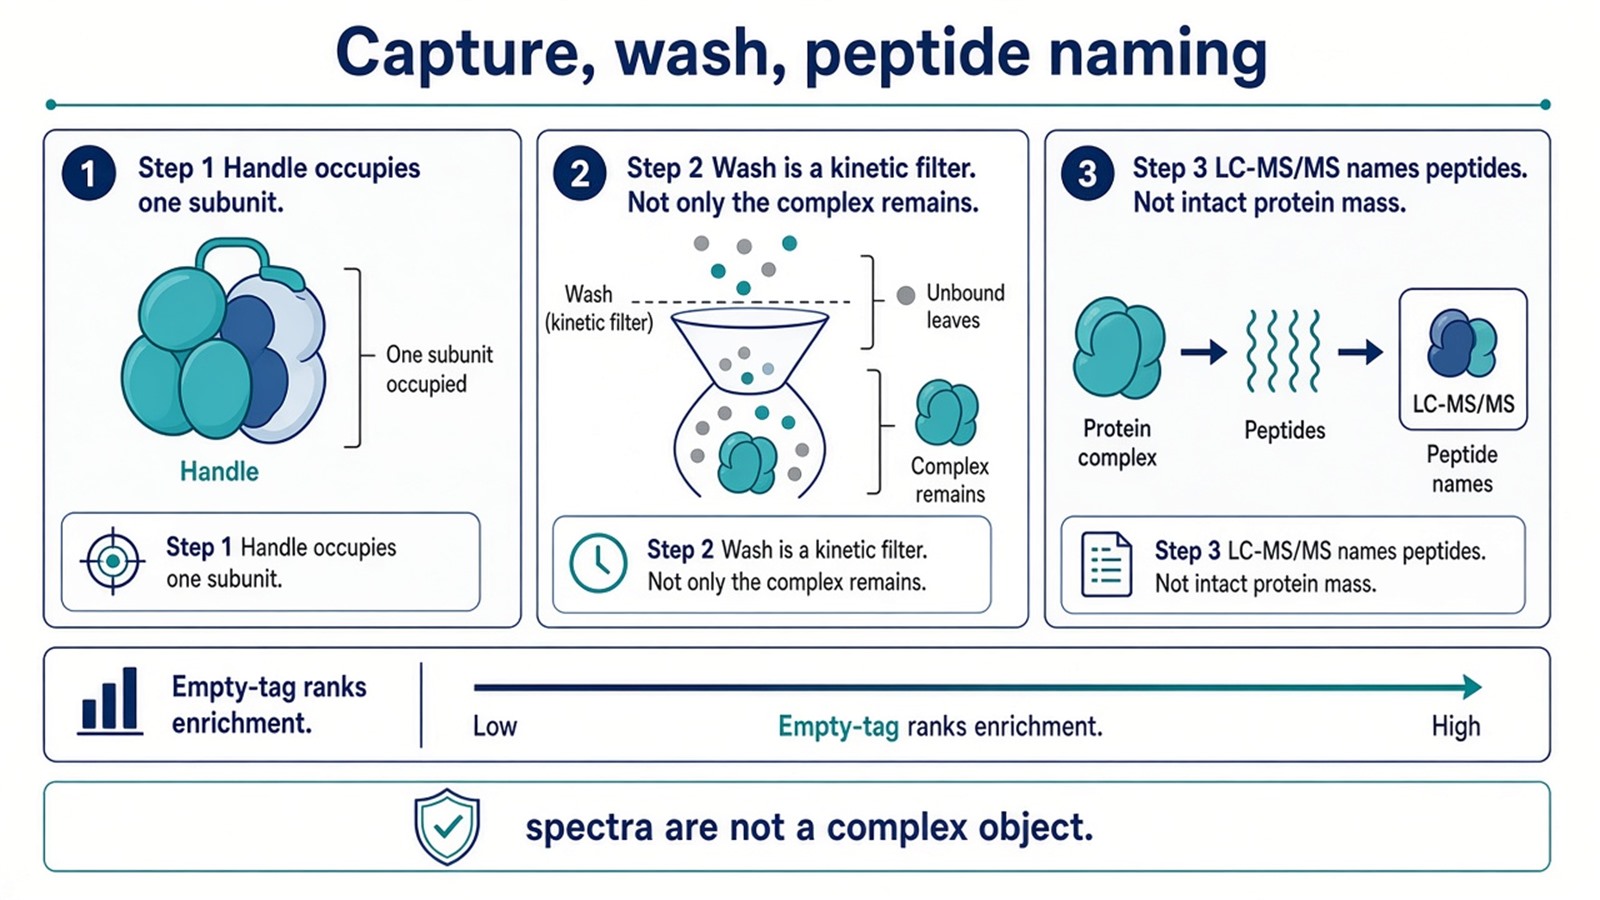
Figure 2. Capture occupies one subunit. Wash is a kinetic filter, not a step that leaves only the complex. LC-MS/MS names peptides. Empty-tag ranks enrichment.
Advantages and Limits, Recast
The original advantage list does not hold as written. AP-MS can detect peptides from proteins that are scarce in the eluate. That is not detection of low-abundance protein complexes as a class, and it is not high specificity. Versatility does not extend to every complex regardless of cellular abundance. Tagged lines are the usual sample. Tissues and clinical specimens are possible only when a handle and a matched control can be written; they are not a granted applicability.
The original limitation list is true and incomplete. Nonspecific binders produce candidates that look like subunits until empty-tag is compared. Sample complexity is a processing note. The scientific limits that belong on the same list are missing subunits after a harsh wash, indirect co-recovery of a hitchhiker, and the absence of stoichiometry or architecture in a ranked table.
Signaling-pathway and disease settings remain allowable as study backgrounds. A project can ask which names follow this subunit after a written stimulus, or which names change rank when capture chemistry is matched across two lysates. AP-MS does not uncover the molecular mechanism of signal transduction, identify disease-related complexes as a class, or name therapeutic targets. Those sentences need a separately validated assay. Efficient and accurate complex analysis is not granted by combining affinity purification with a spectrometer.
Experimental Limits
AP-MS complex analysis reduces a project to one subunit as bait, one wash, and one control class. Function, architecture, biomarker, and therapeutic-target language sits outside ordinary research reporting unless a separately validated assay is already named. A long identification list does not become a complex by being labeled protein-complex research.
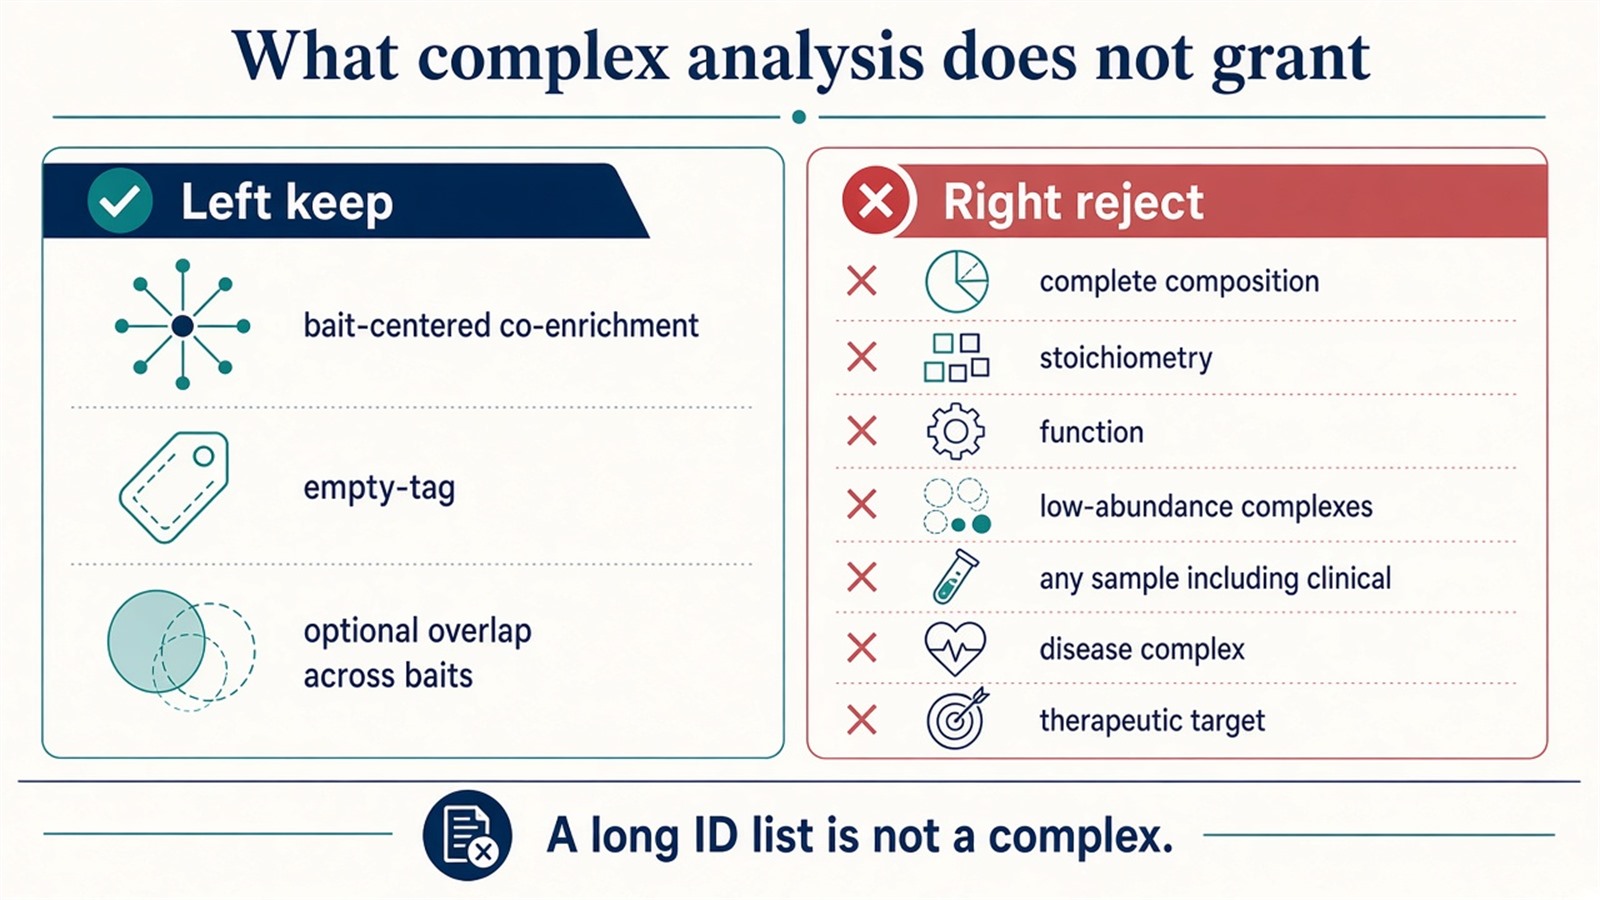
Figure 3. Keep: bait-centered co-enrichment and optional overlap across baits. Not granted: complete composition, stoichiometry, low-abundance complexes, or therapeutic targets.
Project Planning and Service Scope
A usable request names the subunit used as bait, whether other subunits will be baited for overlap, the matched control, and the orthogonal assay that would be required for a composition or function sentence. If only one predefined subunit pair is in scope, a focused Co-IP is the shorter route and should not be written as complex analysis by AP-MS. MtoZ Biolabs scopes tagged-bait AP-MS on Orbitrap-class LC-MS/MS. Interpretability still follows those arms, not a protein-complex heading.
Once the next sentence is a named subunit that only needs a yes-or-no recovery check, this page should hand off. Researchers who need that blot or targeted pellet can review the MtoZ Biolabs Co-IP MS Protein Interaction Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. Does AP-MS determine the composition of a protein complex?
Not from one bait eluate. It ranks names that survived the wash with that bait. Overlap across related baits is a later pattern, still not a complete list.
2. Does washing leave only the target complex?
No. The wash is a kinetic filter. Real neighbors can fall off. Bead binders can remain.
3. Can AP-MS detect low-abundance protein complexes?
It can detect peptides that reached the spectrometer. Low cellular abundance is not a granted complex class.
4. Does AP-MS explain complex function?
No. Function is a later sentence. Annotation of recovered names is not a mechanism.
5. Are clinical specimens a standard AP-MS complex sample?
Only when a handle and a matched control can be written. They are not a default sample type.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Gavin A.C., Bösche M., Krause R., et al. (2002). Functional organization of the yeast proteome by systematic analysis of protein complexes. Nature, 415, 141-147. https://doi.org/10.1038/415141a
- Krogan N.J., Cagney G., Yu H., et al. (2006). Global landscape of protein complexes in the yeast Saccharomyces cerevisiae. Nature, 440, 637-643. https://doi.org/10.1038/nature04670
- Mellacheruvu D., Wright Z., Couzens A.L., et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
- Dunham W.H., Mullin M., Gingras A.C. (2012). Affinity-purification coupled to mass spectrometry: basic principles and strategies. Proteomics, 12, 1576-1590. https://doi.org/10.1002/pmic.201100523
