



Top-down proteomics quantifies intact proteins and proteoforms by liquid chromatography-tandem mass spectrometry (LC-MS/MS) without obligatory proteolytic digestion, preserving combinatorial post-translational modifications and sequence variants that bottom-up peptides often split (Catherman et al. 2014; Toby et al. 2016). Shared advantages are proteoform-resolved relative comparison, direct intact-mass evidence, and clearer isoform discrimination when ionization and chromatography support the target. Shared limitations are harder denaturation and desalting than peptide digests, narrower dynamic range, lower throughput for complex lysates, and relative-only intensities unless matched intact standards are added. Prefer top-down when the claim needs intact proteoform ratios and sample chemistry can stay MS-compatible; keep bottom-up LFQ, DIA, or TMT for deep discovery coverage across large cohorts.
Researchers who already have defined intact-protein targets or a specific project objective can also review the MtoZ Biolabs Top-Down Proteomics Service and LC-MS Based Intact Protein Analysis Service for project-specific feasibility and sample planning.
Advantages and Limitations at a Glance
Topic |
Advantage when controls hold |
Matching limitation |
|---|---|---|
Analyte form |
Intact proteoform kept together |
Complex lysates resist intact separation |
PTM and variants |
Combinatorial modifications stay on one chain |
Site assignment still needs informative fragments |
Quantification logic |
No peptide-to-protein roll-up for the named form |
Dynamic range and ionization bias remain |
Isoform discrimination |
Intact mass separates forms peptides can conflate |
Throughput usually below deep bottom-up discovery |
Claim type |
Relative proteoform ratios under matched prep |
Not absolute concentration without intact standards |
Sample chemistry |
Strong when desalting and MS buffers succeed |
Detergents, salts, and aggregates block signal |
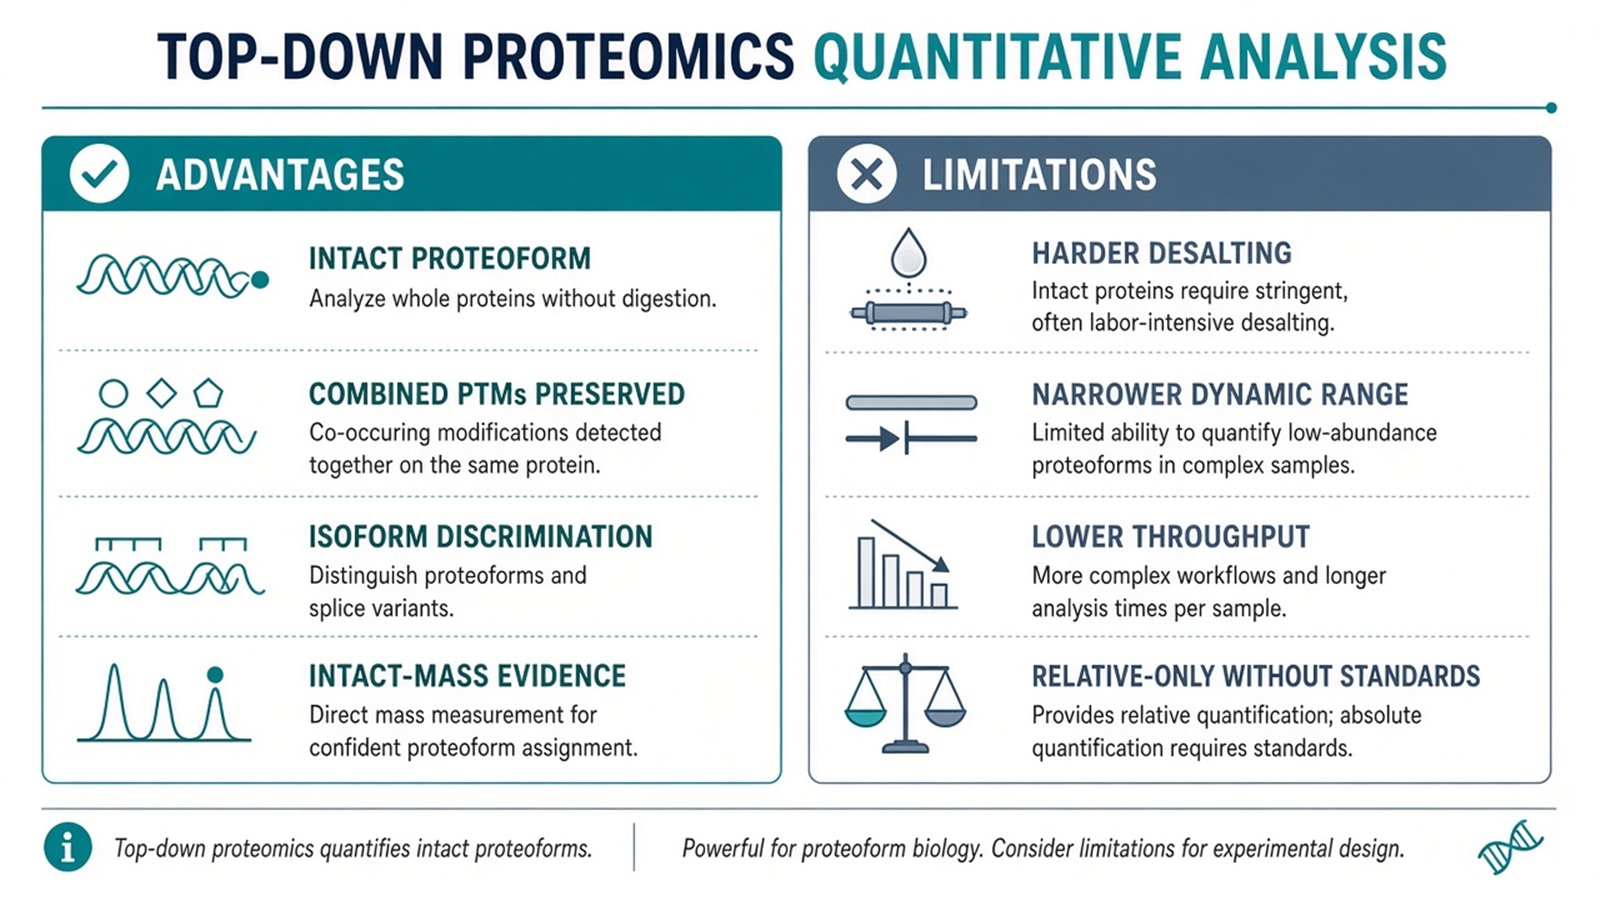
Figure 1. Top-down quantitative advantages and limitations arise together: intact proteoform resolution requires MS-compatible intact prep that complex lysates may not support.
Calling top-down universally more accurate than bottom-up without stating proteoform claim, sample complexity, and the relative readout overstates method class alone.
Sample and Prep Gates Before Top-Down Quantification
Top-down advantages only hold when intact proteins reach the instrument in a usable state. Sample type and prep decide feasibility before acquisition is named.
Sample or prep situation |
Top-down relative claim often open |
Reassess or prefer bottom-up |
|---|---|---|
Purified protein, subunit, or simple mixture |
Intact mass and proteoform ratios often workable |
Residual detergent or polymer adducts dominate |
Enriched proteoform family (for example histones) |
Intact comparison after matched extraction |
Incomplete enrichment mixes unrelated forms |
Whole-cell or tissue lysate without fractionation |
Usually blocked for deep intact coverage |
Prefer bottom-up LFQ, DIA, or TMT discovery |
High-salt or strong-detergent extracts |
Blocked until exchange into MS-compatible buffers |
Peptide digests tolerate broader cleanup chemistry |
Scarce clinical aliquots needing many contrasts |
Often better as bottom-up discovery |
Intact methods may consume input for limited depth |
Absolute amount required |
Needs matched intact standards and calibration |
Ordinary intact intensities stay relative |
Shared early prep still matters: declare extraction chemistry, reduce complexity when needed, exchange into volatile or MS-compatible buffers, and keep denaturation or native conditions aligned with the claim. Intact desalting and concentration replace the peptide cleanup path used in bottom-up work. Unmatched loads, incomplete buffer exchange, and adduct formation become quantitative artifacts that later software cannot repair.
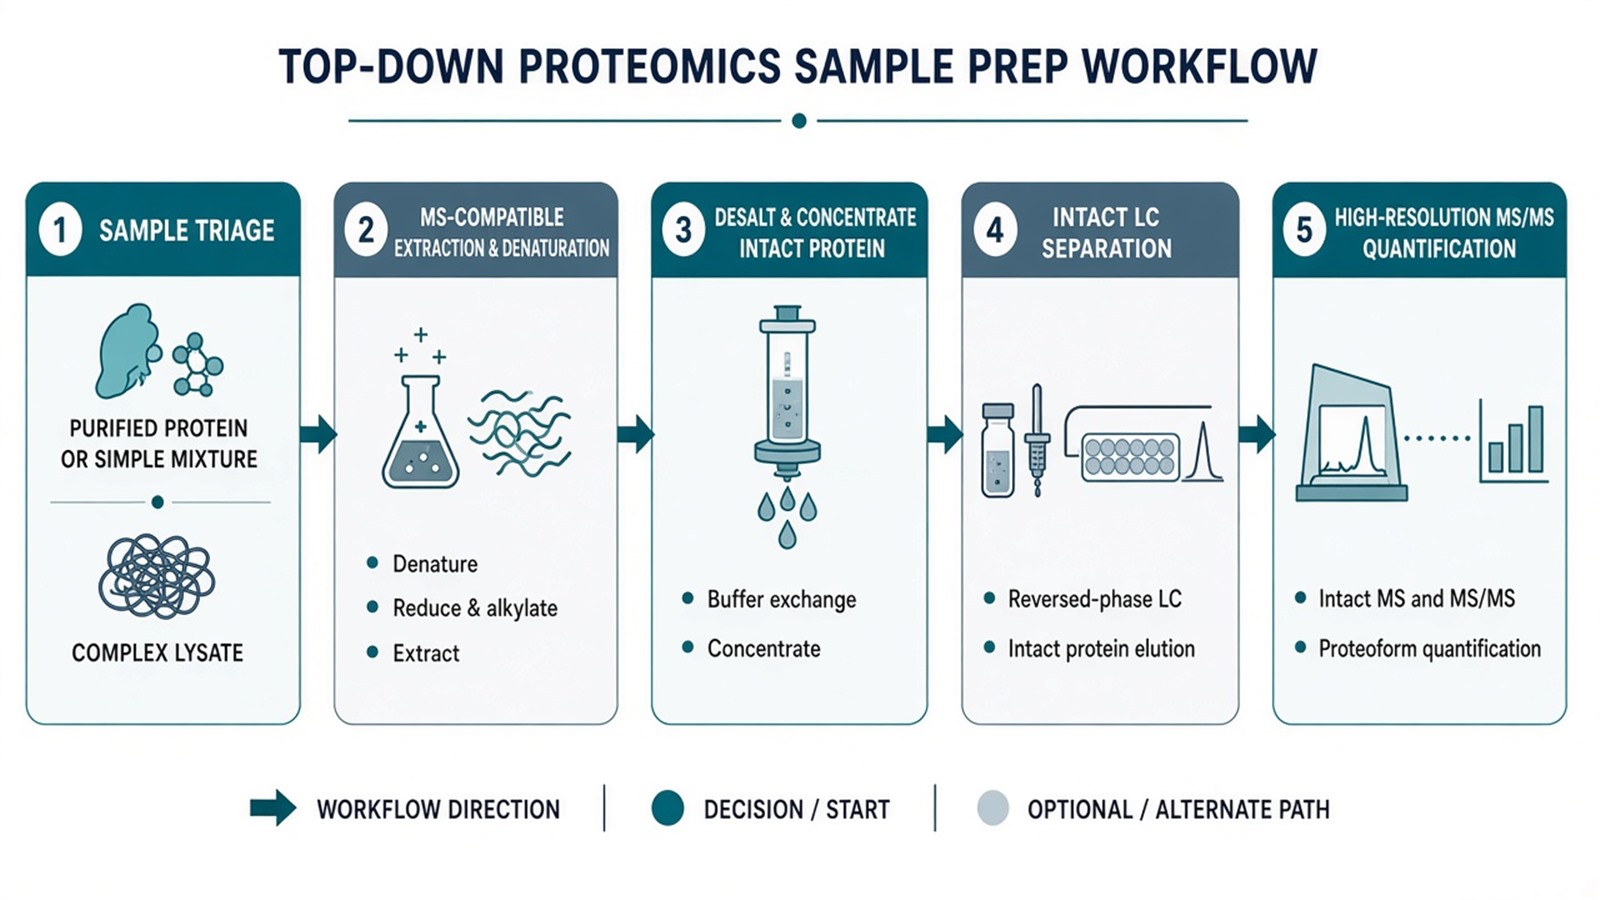
Figure 2. Top-down quantification depends on intact-protein prep: MS-compatible extraction and desalting precede intact LC-MS/MS comparison.
For intact-protein service context, see Top-down Proteomics Analysis Service. For bottom-up discovery when intact prep fails, see What Is Label-Free Analysis? LFQ Principles, Workflow, Advantages, and Limitations and Absolute and Relative Quantitative Proteomics Analysis. For MS protein quantification framing, see Protein Quantification Measured by Mass Spectrometry.
Judgment Criteria: When Top-Down Prep Fits
Study need |
Top-down quantification often fits |
Prefer bottom-up or redesign |
|---|---|---|
Claim |
Relative ratios of named intact proteoforms |
Deep peptide coverage across thousands of proteins |
Sample |
Purified, enriched, or low-complexity intact targets |
Unfractionated complex lysates as the only material |
Chemistry |
Desalting and MS-compatible buffers are enforceable |
Strong detergents or salts cannot be exchanged |
Throughput |
Focused proteoform panels |
Large multi-arm discovery cohorts |
Standards |
Relative intact intensities acceptable |
Absolute concentration required without intact SIS |
Prefer top-down when the scientific question is proteoform-resolved and prep can keep intact proteins soluble and desalted. Prefer bottom-up LFQ, DIA, or TMT when the question is deep relative discovery across complex matrices. Prefer middle-down or subunit strategies when partial cleavage better localizes a biotherapeutic variant. Prefer PRM, MRM, or AQUA when a closed peptide shortlist needs amount units after discovery.
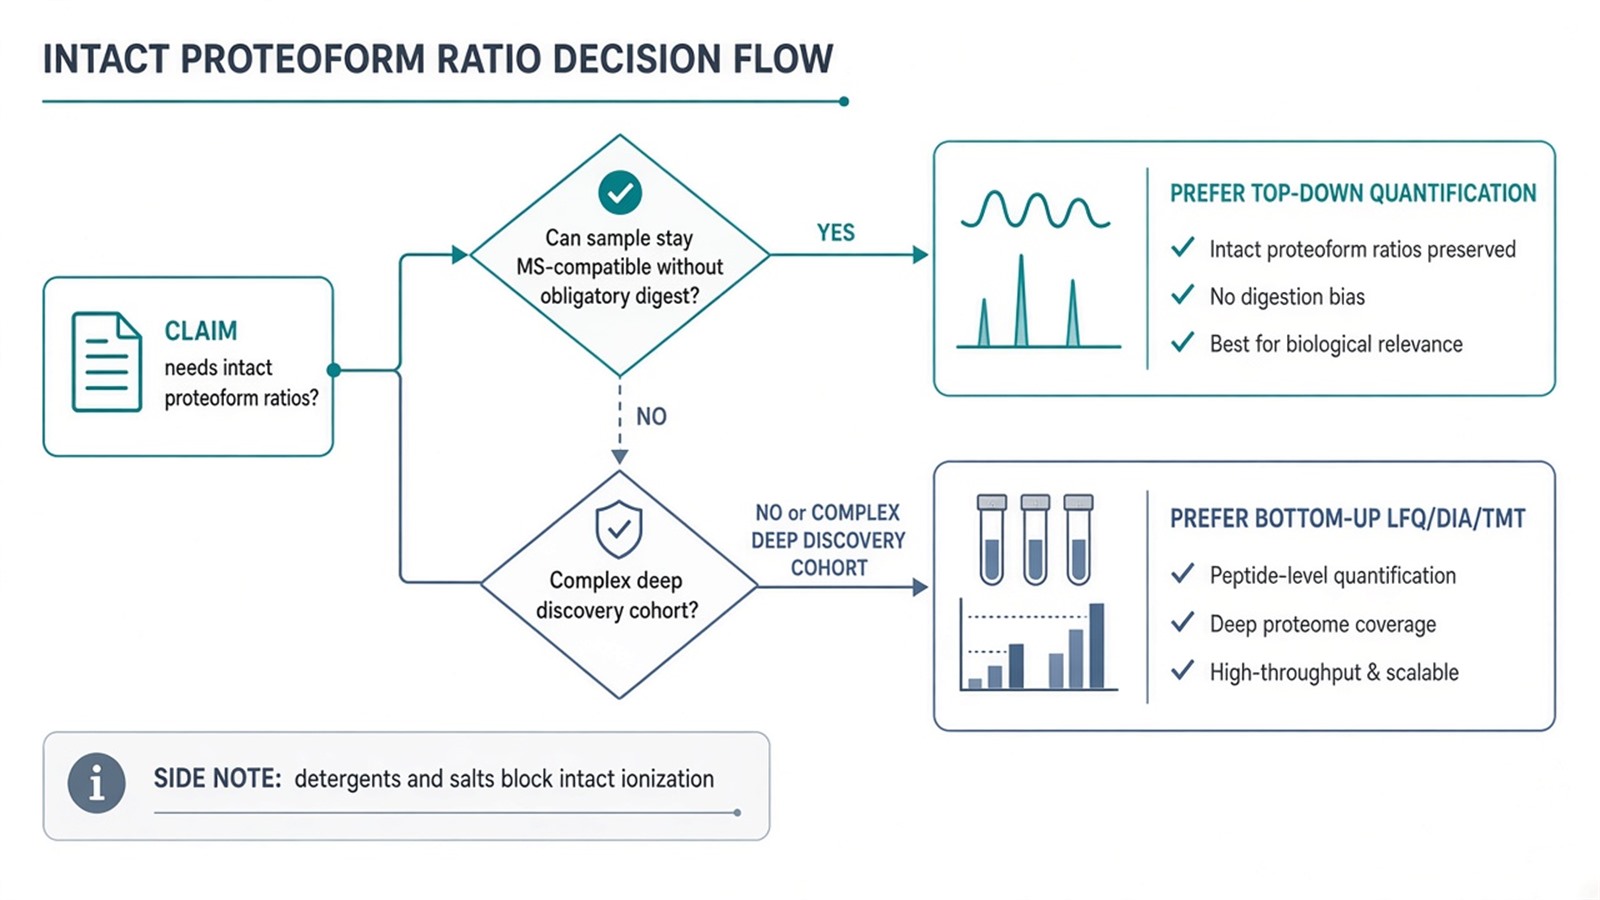
Figure 3. Select top-down from the intact proteoform claim and MS-compatible prep; change route when lysate complexity or buffer chemistry blocks intact analysis.
Limits and Application Boundaries
Top-down does not erase ionization bias among proteoforms. Intact relative intensities are not molar concentrations unless matched intact standards and calibration are added. Pathway enrichment from a differential list still reflects overrepresentation relative to the quantified background, not proven pathway activity. Native MS assembly claims are a related but separate experiment from denatured top-down proteoform quantification.
Typical applications tied to sample and prep:
- Purified recombinant proteins or antibody subunits needing intact mass and proteoform ratios.
- Enriched histone or other PTM-dense families after matched extraction.
- Biotherapeutic variant monitoring when intact or subunit chemistry is controlled.
- Focused mechanism studies where a small intact panel answers the claim better than a peptide inventory.
Diagnostic cutoffs and companion-diagnostic claims sit outside ordinary research top-down reporting unless a separately validated assay framework is in place.
Once sample complexity, buffer exchange, and the intact relative claim are defined, the project can be scoped as a top-down quantitative study. Researchers can review the MtoZ Biolabs Top-Down Proteomics Service for sample evaluation and workflow planning. For intact LC-MS options, see LC-MS Based Intact Protein Analysis Service.
Frequently Asked Questions
1. Is top-down always more accurate than bottom-up for quantification?
No. Top-down is stronger for intact proteoform identity. Accuracy still depends on matched prep, ionization, and QC. Bottom-up often remains better for deep discovery coverage.
2. What sample-prep failure most often blocks top-down quantification?
Residual detergents, high salt, and incomplete desalting are common. Intact proteins then ionize poorly or fail chromatography.
3. Can top-down replace TMT or LFQ for a whole tissue cohort?
Usually not as the sole discovery route. Complex lysates typically need bottom-up depth; top-down fits focused intact targets or enriched fractions.
4. Does skipping digestion automatically give absolute protein concentration?
No. Intact intensities remain relative unless matched intact standards and calibration are added.
5. When should I choose middle-down instead of full top-down?
When partial cleavage better localizes a subunit or biotherapeutic variant while still avoiding a full bottom-up peptide inventory.
Reference
- A.D. Catherman, O.S. Skinner, N.L. Kelleher (2014). Top Down proteomics: facts and perspectives. Biochem. Biophys. Res. Commun., 445, 683-693. https://doi.org/10.1016/j.bbrc.2014.02.041
- D.T. Toby, L. Fornelli, N.L. Kelleher (2016). Progress in Top-Down Proteomics and the Analysis of Proteoforms. Annu. Rev. Anal. Chem., 9, 499-519. https://doi.org/10.1146/annurev-anchem-071015-041550
