



Targeted proteomics measures a predefined protein or peptide list by scheduling those analytes in liquid chromatography-tandem mass spectrometry (LC-MS/MS), rather than reporting whatever an open search returns (Picotti and Aebersold 2012). Methods and applications of targeted proteomics name the same closed-list design. Application of targeted proteomics is not a second assay. Selected reaction monitoring (SRM), multiple reaction monitoring (MRM), and parallel reaction monitoring (PRM) implement the schedule. Later extraction of a few proteins from a data-independent acquisition (DIA) file does not make that file a targeted method. The valid result is relative change of the scheduled analytes, or an amount only with a matched standard and a calibration model. The design does not issue a diagnostic cutoff or a personalized treatment plan.
Researchers who already have defined protein or peptide targets, comparison groups, or a specific project objective can also review the MtoZ Biolabs Targeted Proteomics Service for project-specific feasibility and analysis planning.
What a Targeted Proteomics Experiment Can Claim
The first decision is whether the protein list is closed before acquisition. A targeted run spends instrument time on scheduled precursors, and often on scheduled product ions. The table can support a statement about those analytes inside one chromatographic and scheduling plan. It cannot invent proteins that were never isolated.
A peptide remains a surrogate of a protein or of a modification site. Shared sequence, missed cleavage, and incomplete occupancy can break that surrogate relationship. An application title such as biomarker work, drug-response follow-up, or pathway panel does not repair a peptide that is not unique.
Discovery intensities from label-free quantification, tandem mass tags (TMT), or stable isotope labeling by amino acids in cell culture (SILAC) often supply the candidate list. Targeted acquisition then tests that short list. Candidate confirmation is not clinical validation. Enzyme-linked immunosorbent assay (ELISA) and Western blot remain useful when an antibody and a linear range already exist. Targeted MS does not require an antibody. It still requires LC-MS-compatible peptides that can be scheduled.
Targeted proteomics reports scheduled analytes from a closed list, not a renamed discovery matrix.
Methods That Implement the Closed List
SRM and MRM isolate one precursor-to-product transition at a time on a triple quadrupole (Lange et al. 2008). The first quadrupole passes the precursor, the collision cell fragments it, and the third quadrupole passes a prechosen fragment. Method building includes transition selection and collision-energy work for each pair. Throughput is high once the method is locked. Interference that sits on the chosen fragment is not visible as a full product-ion spectrum.
PRM keeps quadrupole isolation of a scheduled precursor and records a high-resolution product-ion spectrum of that precursor in one scan (Peterson et al. 2012). Fragments can be chosen after acquisition from the same record. Naming the analyzer family does not define the scientific claim.
Researchers can refer to PRM-MS Mass Spectrometry for how PRM records product ions relative to MRM.
DIA with later targeted extraction is a different experiment. Windowed fragmentation records many precursors first (Gillet et al. 2012). A library or peptide query can then extract chromatograms for selected proteins. That reanalysis can be useful. It does not convert the original acquisition into a scheduled SRM, MRM, or PRM method, and it does not inherit the same interference model.
In MtoZ targeted-proteomics offerings, MRM or PRM is typically built on a closed peptide list supplied by the study. Service configuration is not an academic rule that other laboratories must copy, and it does not include a diagnostic or GMP release intended use.
Design |
What is scheduled |
What the table can support |
What it is not by default |
|---|---|---|---|
SRM or MRM |
Precursor-product pairs |
Relative change of those transitions |
A proteome-wide discovery matrix |
PRM |
Precursors; fragments chosen from the product-ion record |
Relative change of those peptides |
Unlimited sensitivity in every matrix |
DIA plus later extraction |
Windows, not a locked transition list |
A query of a recorded DIA file |
A scheduled targeted assay |
Targeted MS plus a matched standard |
The same ions plus a calibrant |
An amount at the standard's level |
Tissue concentration if spike-in was late |
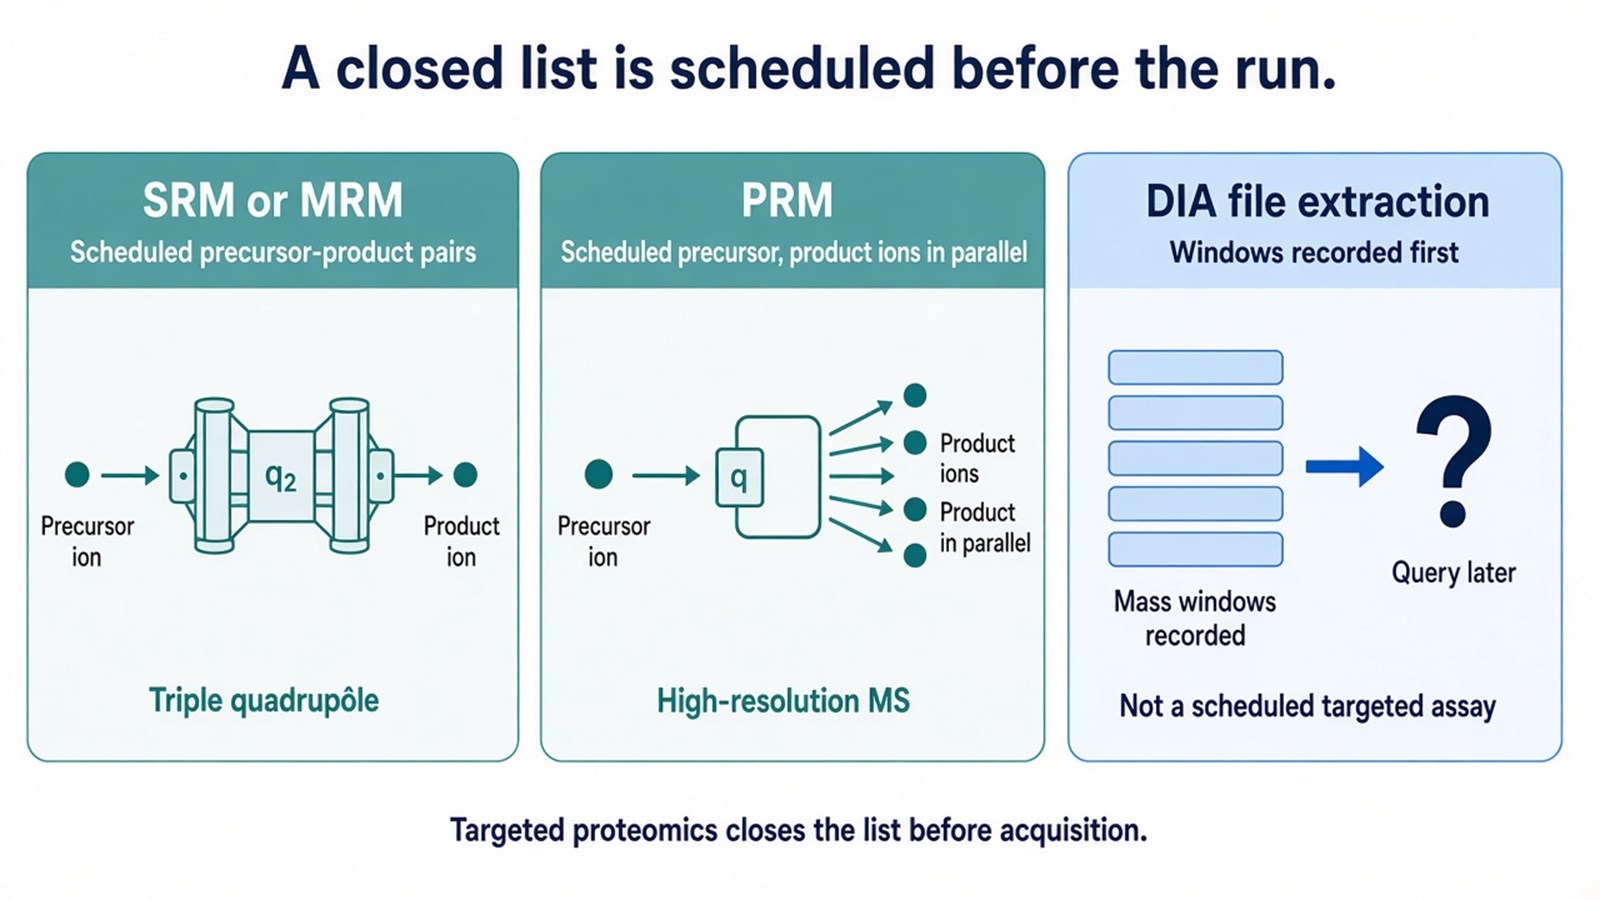
Figure 1. Targeted proteomics schedules a closed list before the run; extracting selected proteins later from a DIA file is a different experiment.
When an Application Fits the Assay
An application fits when the protein or peptide list is already closed, the peptides can be scheduled, and the paper needs repeated measurement of those analytes rather than a new open search. Confirmation of discovery candidates, a defined enzyme or modification panel, and research follow-up of a drug, genotype, or time contrast are common cases.
The assay does not fit an open proteome, a request for early diagnosis, or a personalized treatment certificate. Disease names such as cancer, neurodegeneration, or metabolic syndrome describe the biology that motivated the list. They do not, by themselves, make a targeted table a diagnostic cutoff. Environmental protein surveys and developmental time courses can use the same MS methods only when the list, matrix, and claim are defined the same way. Listing those fields as proof of the method adds catalog text, not a stronger claim.
Unlabeled or TMT discovery remains the matching route when the list is still open. An immunoassay remains the matching route when a validated antibody method already answers the same protein in that matrix. Absolute copy-number or concentration language needs a matched isotope standard and a calibration model.
Researchers can refer to Absolute Protein Quantification Mass Spectrometry for how spike-in level changes that amount claim.
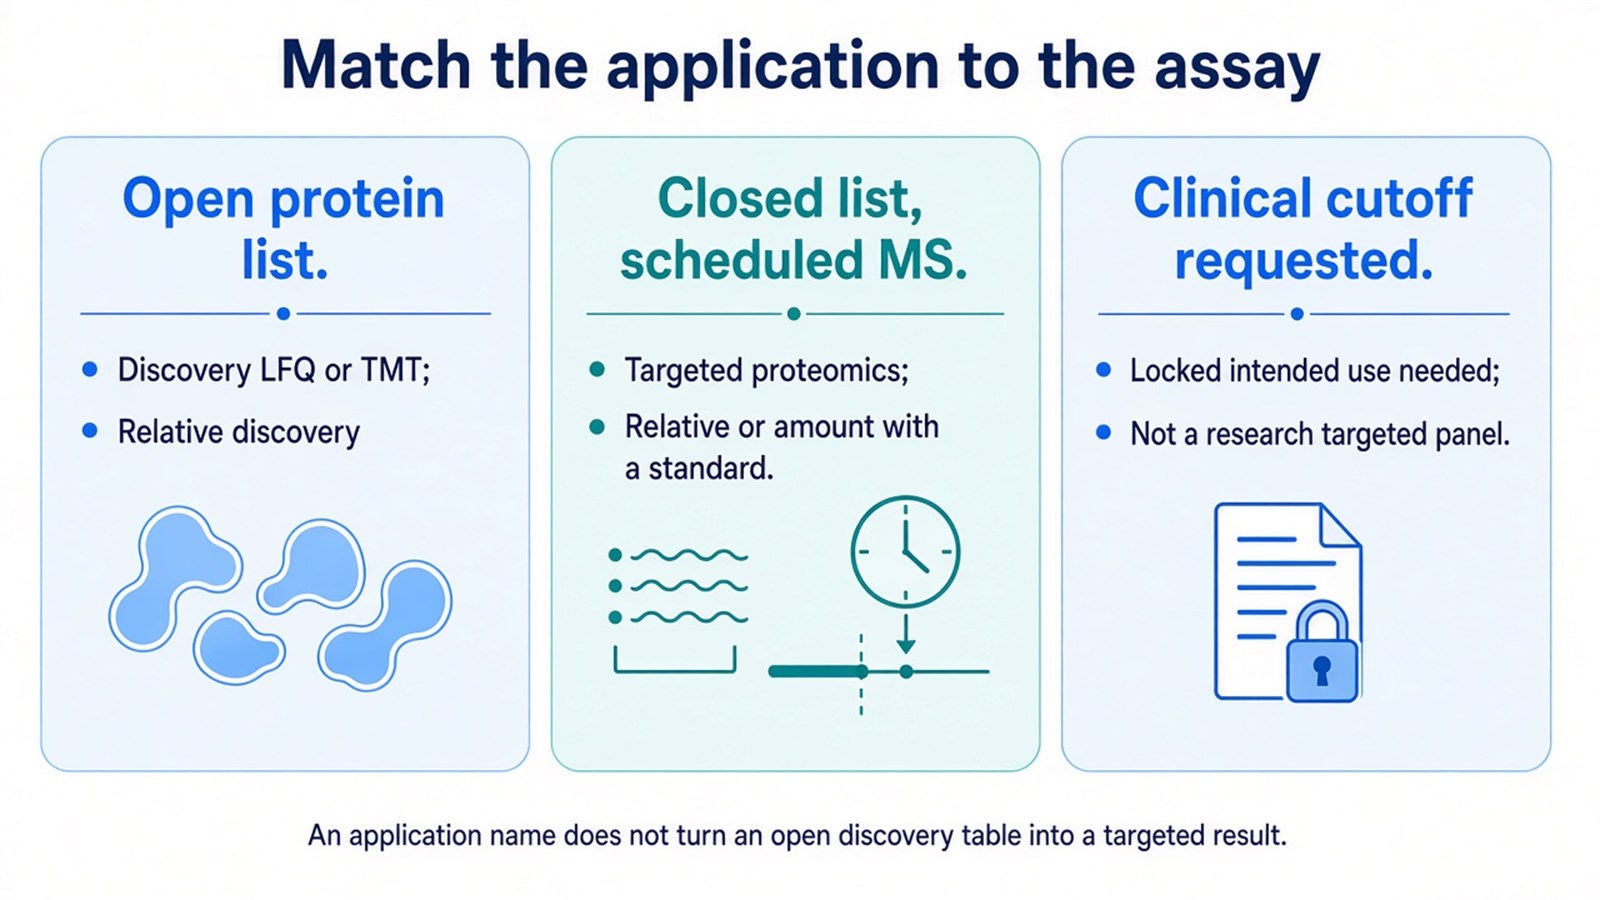
Figure 2. Targeted proteomics fits a closed, scheduled list; an open discovery list stays on LFQ or TMT, and a diagnostic cutoff is a different intended use.
An application name does not turn an open discovery table into a targeted result.
How Targeted Results Should Be Read
A peak group supports the peptide that was isolated and the fragments that were integrated. Coeluting ions with similar mass can still occupy an SRM transition. PRM can often reject one interfered fragment by inspecting neighbors at high resolving power. Neither scan type proves that digestion and extraction were complete.
Scheduling capacity is a method limit. Isolation width, dwell or fill time, and the number of concurrent precursors set how many targets fit in one run. Adding proteins until the cycle time exceeds the chromatographic peak width degrades the peak shape. Multiplex size is not evidence that the assay monitored a proteome.
A single-point heavy-peptide spike can be a planning check. A calibration curve in the study matrix is a stronger amount claim. A heavy peptide added after digestion cannot correct digestion loss that already occurred. Multi-laboratory transfer of an SRM method is a separate reproducibility problem; a local research calibration does not become a locked clinical assay.
Pathway maps, interaction cartoons, and multi-omics overlays are annotations added after the targeted table. They do not convert peptide ratios into a mechanism certificate or a treatment recommendation.
Frequently Asked Questions
1. Is extracting proteins from a DIA file the same as targeted proteomics?
No. DIA records windows first and queries peptides later. Targeted proteomics schedules the closed list during acquisition as SRM, MRM, or PRM.
2. Does a targeted proteomics application in a disease study create a diagnostic test?
No. A research panel in a defined matrix is not a diagnostic cutoff. Diagnostic use needs a locked intended use, a specified specimen, and evidence outside a research calibration.
3. When is a targeted result relative, and when is it an amount?
Relative change needs only the scheduled analytes and a defined contrast. An amount needs a matched standard, a spike-in point that matches the claim, and a calibration model.
4. Should every discovery candidate move to targeted follow-up?
Only when the list is short enough to schedule and the paper needs repeated measurement of those peptides. A still-open proteome stays on discovery acquisition.
Once the analyte list, acquisition method, contrast, and expected relative or calibrated claim are defined, the project can be evaluated as a targeted proteomics study. Researchers can review the MtoZ Biolabs Targeted Proteomics Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
