



Protein interaction screening is a first-pass search for candidate partners, not a finished interaction map. The screen asks which names survive a stated bait, library or proteome, and control set. Validation asks whether a shortlisted pair still co-recovers, is retained on an immobilized bait, or meets a biosensor test. Those jobs are sequential. Scale decides whether the first pass is a yeast binary library, a tagged-cell AP-MS list, or a live-cell proximity list. A hit rule must be written before names leave the screen. Bait format and the reserved validation assay belong in the same plan. A later Co-IP, pull-down, or SPR run is the second stage, not a synonym for the first.
Researchers who already have a bait, a library or tagged-cell plan, and a written screen-then-validate claim can review MtoZ Biolabs Protein-Protein Interaction Analysis for feasibility. Evidence-type choice among discovery and pair assays is covered in Measuring Protein-Protein Interactions.
Screening and Validation Are Two Stages
A screen produces a ranked list. Y2H reports preys that reconstitute a reporter with the bait in yeast (Fields and Song 1989). AP-MS reports proteins that co-purify with a tagged bait versus empty-tag and bead controls. Proximity labeling reports biotinylated neighbors versus empty-enzyme and no-substrate arms. None of those lists is a validated pair.
Validation is a new experiment on a shortlist. Reciprocal Co-IP tests whether the pair can be recovered from a physiological lysate. Pull-down tests in-vitro retention. SPR tests a prepared pair on a sensor. Drawing every screen name as an edge skips that stage. Co-enrichment and proximity can still be indirect (Hein et al. 2015).
Do not treat a methods catalog as a screen plan. Protein chips, FRET, and unnamed high-throughput platforms are not interchangeable first passes. Choose one primary screen from bait format and scale, then name the validation assay before the hit table is exported.
Screening Scale Sets the First Method
Scale is the number of prey hypotheses the first pass is allowed to test. A binary Y2H library tests many coded preys against one bait in yeast. AP-MS tests the proteins that survive capture from one tagged-cell or tagged-organism condition. Proximity labeling tests who was near one fusion during a labeling window. The three scales are not the same throughput claim.
Y2H fits soluble nuclear-competent baits and a cDNA or fragment library. Membrane proteins and pairs that need a modification yeast lacks are a poor first screen. Planning material for library construction from client tissue is typically at least 1x10^7 cells, 500 mg of tissue, or 200 ug of total RNA. The Yeast 2 Hybrid Analysis Service is the library-scale binary route.
AP-MS fits when the bait can be tagged in the cell line or organism of interest and the tag does not destroy localization. Primary cells and clinical tissue that cannot be tagged should not be forced into AP-MS. Empty-tag samples travel with the bait. Identification depth is typically hundreds to thousands of proteins and depends on the sample. The usable set is the bait-enriched subset after controls.
Proximity labeling fits weak or compartment-restricted associations that washing may remove. It needs a fusion that still reaches the intended compartment and at least 1x10^7 cells per labeled sample plus matched negative controls. Enzyme-class names stay off the service label. Intake uses the Proximity Labeling Mass Spectrometry path.
Y2H, AP-MS, and Proximity Labeling as First Screens
The three screens answer different first questions. Y2H asks whether two fusion proteins can meet in the yeast nucleus and turn on a reporter. AP-MS asks who remains with a tagged bait after lysis and wash. Proximity labeling asks who was close enough to be biotinylated before lysis.
Use Y2H when the need is a large binary search and the bait is expected to fold in yeast. Use AP-MS when the need is a physiological-lysate partner list around a tag. Use proximity labeling when the need is a live-cell neighbor map and AP-MS washing is expected to lose the pair. Mixed AP-MS and other enrichment-MS projects can be reviewed under the MS-Based Protein-Protein Interaction Analysis Service.
Do not run all three as one blended screen. A Y2H prey name and an AP-MS accession are not automatically the same biological event. If two screens are required, plan two hit tables and two validation queues.
The comparison below keeps library-scale binary search and cell-context proteomic screens on separate tracks.
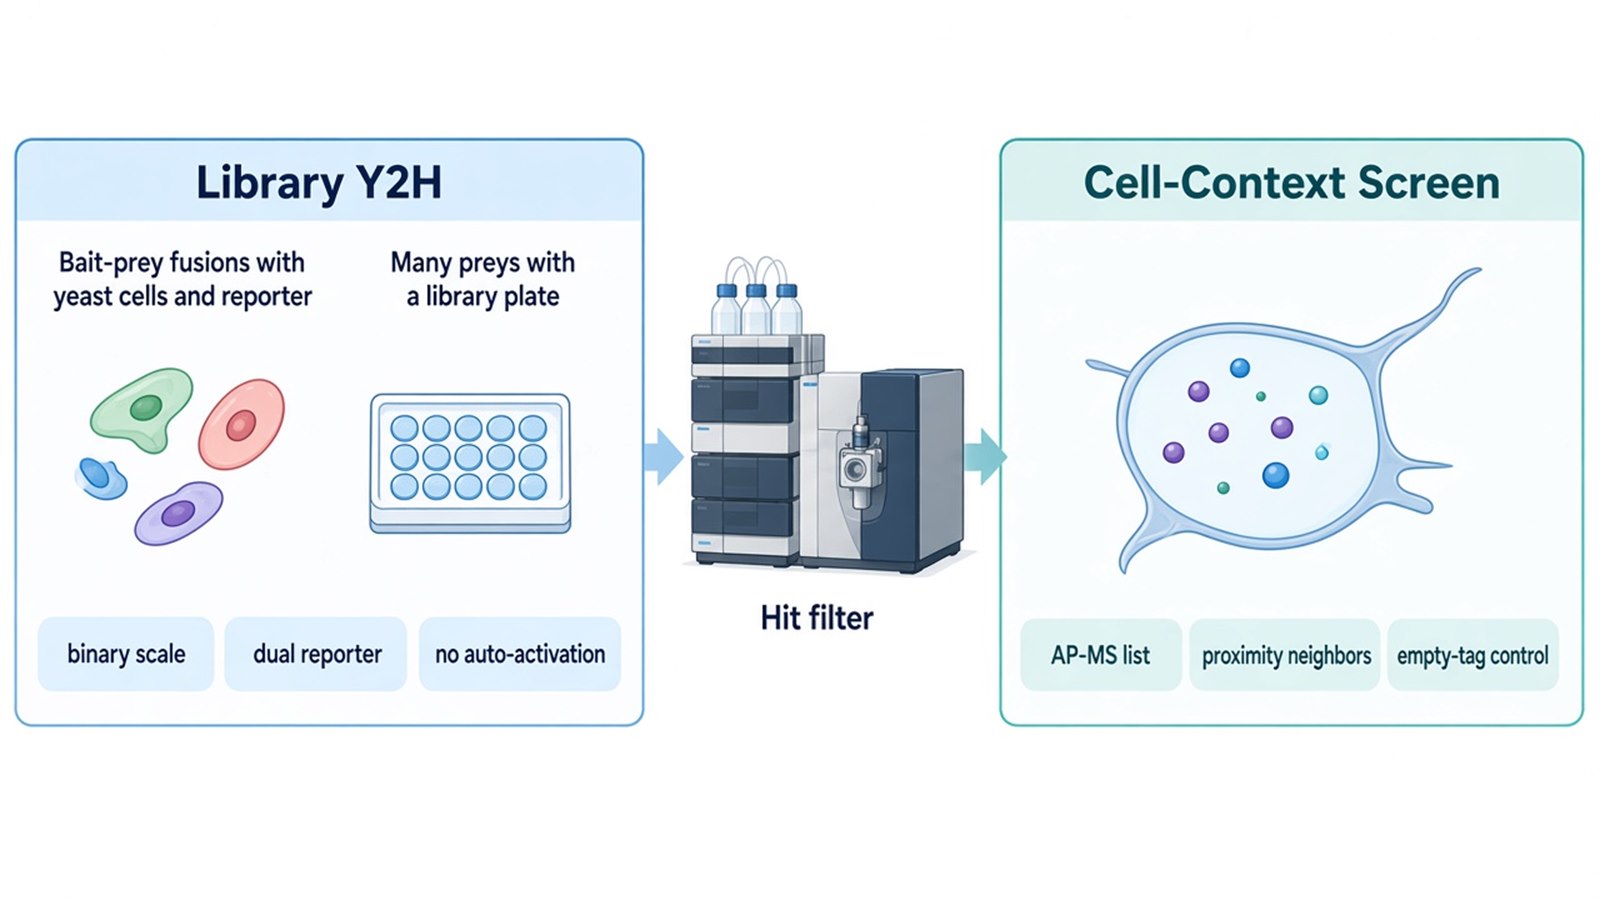
Figure 1. Library-Scale Y2H vs Cell-Context AP-MS or Proximity Screens.
Hit Thresholds Before a Candidate Is Named
A protein interaction screening hit is a name that passes the screen-specific rule, not any protein that appeared once. For Y2H, require reporter activity on the planned selective media and a second readout such as LacZ when that reporter is in the strain. Discard baits that auto-activate without prey. A single colony from a leaky bait is not a hit.
For AP-MS, compare bait tubes with empty-tag and bead-only tubes. Proteins equal in both stay annotated. Recurrent-contaminant catalogs help interpret leftovers after experimental subtraction (Mellacheruvu et al. 2013). They do not replace the empty-tag arm. Replicate enrichment is part of the threshold when more than one bait culture exists.
For proximity labeling, compare the bait fusion with the empty-enzyme and no-substrate arms. Proteins equal in those arms are not proximal hits. Localization of the fusion must be checked before the list is ranked.
Write the threshold in the notebook before the search is opened. Changing the cutoff after seeing favorite names is not a second experiment. Names that fail the rule stay off the validation queue.
Secondary Validation After the Screen
Secondary validation after protein interaction screening uses a different geometry from the first pass. A Y2H hit should be tested as protein, not as another yeast reporter. Reciprocal Co-IP or IP-MS asks whether each partner can recover the other from lysate. Intake for that step is the IP-Mass Spectrometry Service. A defined-pair pull-down asks whether the pair is retained on an immobilized bait. That path is the Fusion Protein Interaction Analysis Service | Pull-Down and MS
An AP-MS or proximity hit that is already a named pair can skip library construction and go straight to reciprocal capture or pull-down. If the next sentence is affinity, use SPR on the prepared pair through Surface Plasmon Resonance Technology. Do not convert a screen intensity into KD.
|
Stage |
Question |
Typical first method |
Hit rule |
Second-stage test |
|---|---|---|---|---|
|
Library binary screen |
Which preys reconstitute a reporter with this bait? |
Y2H |
Dual reporter, no auto-activation |
Co-IP or pull-down of the named pair |
|
Tagged-cell partner list |
Who co-purifies with this tag? |
AP-MS |
Enriched vs empty-tag and beads |
Reciprocal IP-MS or pull-down |
|
Live-cell neighbor list |
Who was near this fusion? |
Proximity labeling |
Enriched vs empty-enzyme and no substrate |
Co-IP of a named neighbor |
|
Affinity of a hit |
How tight is the prepared pair? |
Not a screen |
Not a screen threshold |
SPR or BLI |
|
Cellular complex of a hit |
Does the pair recover from lysate? |
Not a screen |
Not a screen threshold |
Co-IP / IP-MS |
The table is a stage rule, not a ranking of platforms.
Ship the bait sequence, library or tag plan, control arms, and the validation assay reserved for hits. Do not ship a request that asks for a screen and a finished network in one tube.
The diagram below links scale, first method, hit rule, and the second-stage assay to a screening request.
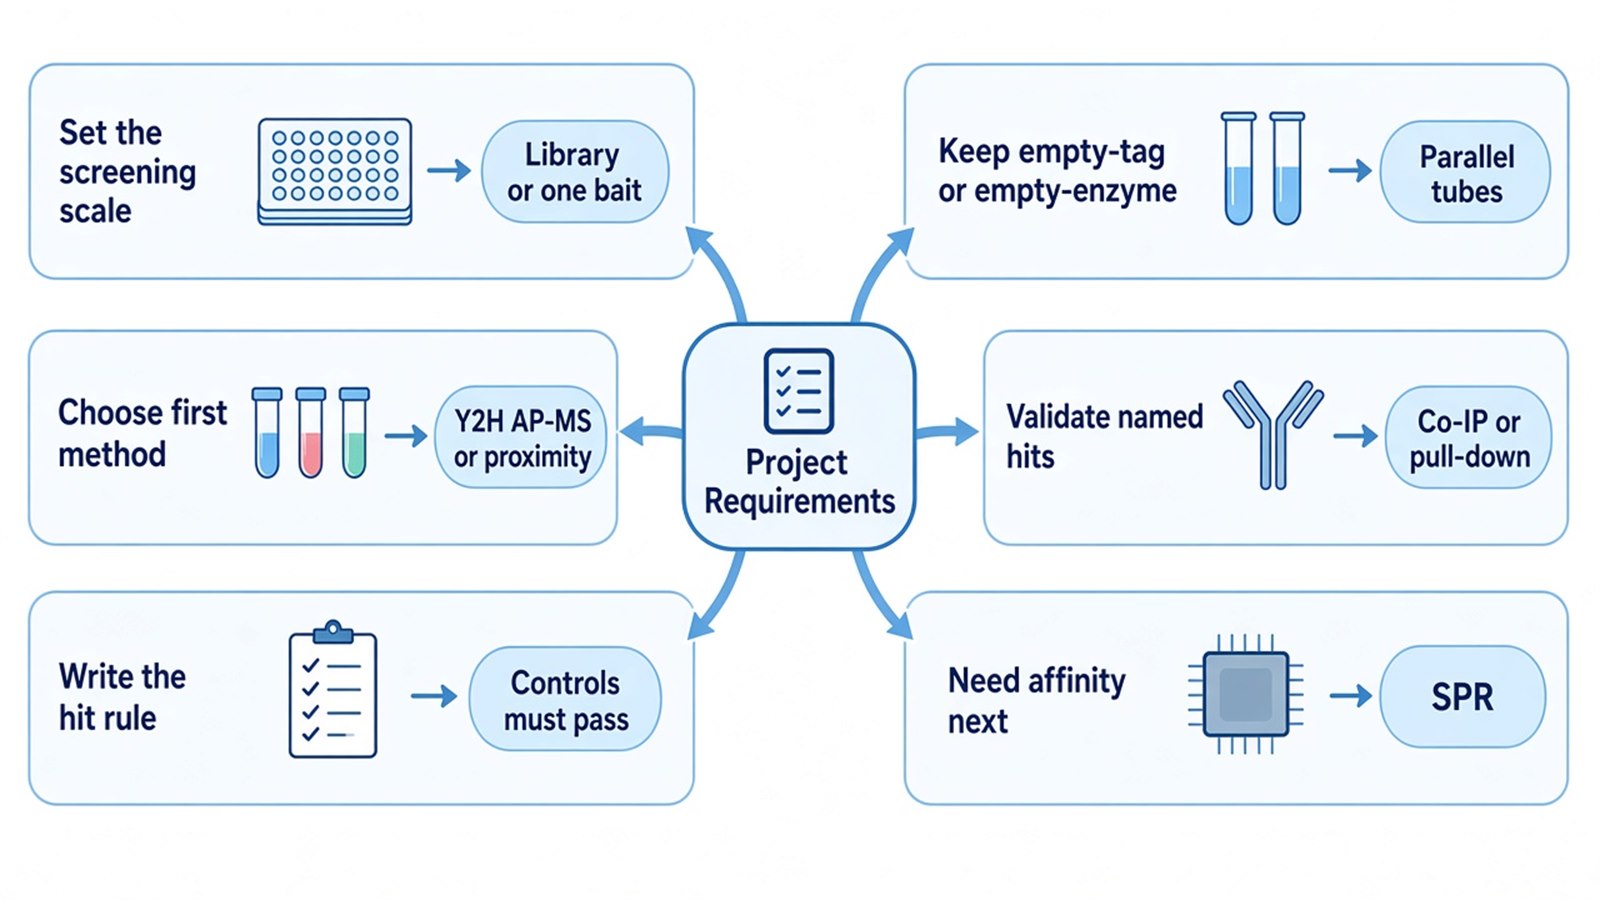
Figure 2. Matching Project Requirements With Screen-Then-Validate Stages.
Deliverables for the first stage typically include the raw list, the control comparison, and the names that passed the stated threshold. Deliverables for the second stage are scoped separately and may include reciprocal capture tables, pull-down retention, or fitted biosensor constants. Deliverables do not include a claim that every screen name is a direct cellular binder. To start intake, send the bait, intended scale, hit rule, and reserved validation assay to info@MtoZ-Biolabs.com or use the site inquiry form so the two stages can be split before work starts.
Frequently Asked Questions
1. Can one protein interaction screening method replace later Co-IP?
No. The screen ranks candidates under its own geometry. Co-IP tests whether a named pair can be recovered from lysate. The two stages answer different sentences.
2. How should Y2H, AP-MS, and proximity labeling be chosen for the first pass?
Choose Y2H for a large binary library and a yeast-compatible bait. Choose AP-MS for a tagged bait in the cell system of interest. Choose proximity labeling when neighbors must be marked before lysis.
3. What is a usable hit threshold for AP-MS?
Require enrichment versus empty-tag and bead-only tubes. Do not promote names that are equal in the controls, and do not replace those tubes with a published contaminant list.
4. Does a Y2H colony count as validation if two reporters are positive?
No. Dual reporters reduce leaky bait artifacts inside the yeast screen. Protein-level Co-IP or pull-down is still the second stage.
5. When should SPR be added after protein interaction screening?
Add SPR after a pair is named and the next sentence is affinity or kinetics of that prepared pair. Do not fit KD to a colony count or an AP-MS intensity.
Researchers can keep protein interaction screening and secondary validation as two stages, then confirm bait, scale, and the reserved follow-up through MtoZ Biolabs Protein-Protein Interaction Analysis.
Reference
- S. Fields, O. Song (1989). A novel genetic system to detect protein-protein interactions. Nature, 340, 245-246. https://doi.org/10.1038/340245a0
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, S.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
