



Applied quantitative proteomics analysis gains leverage when sample pretreatment keeps peptide yields matched across groups. It loses leverage when matrix difficulty, unequal cleanup, or labeling chemistry that fights the sample map creates biases before liquid chromatography-tandem mass spectrometry (LC-MS/MS) starts (Bantscheff et al. 2012; Wisniewski et al. 2009). The guide below states prep-linked advantages and disadvantages for application work. It is not a disease success-story catalog, not a method-definition encyclopedia, and not a platform throughput promise.
Application routes still depend on labeling chemistry, acquisition mode, and claim type. Unlabeled digests can be acquired as data-dependent acquisition (DDA) or data-independent acquisition (DIA). Isobaric tandem mass tag (TMT) work needs amine-compatible peptides before mixing. Named proteins are typically followed by parallel reaction monitoring (PRM) or multiple reaction monitoring (MRM). In MtoZ Biolabs offerings, unlabeled discovery is commonly run as DDA label-free or DIA, multiplexed isobaric work is typically evaluated as TMT, and closed lists are typically followed by PRM, with MRM when a scheduled triple-quadrupole panel already exists.
Where Matched Pretreatment Supports Applied Quantification
Three prep-linked advantages appear repeatedly in applied projects.
Comparable digests across conditions. Shared lysis, reduction, alkylation, and digestion rules let unlabeled label-free quantification (LFQ) or DIA compare the same proteins between treatment arms, genotypes, or time points (Cox et al. 2014; Gillet et al. 2012). The advantage is cross-group comparability, not a longer protein name list.
Cleanup that stabilizes LC and ionization. Removing detergents, salts, and lipids before injection protects chromatography and peak shape. Stable retention helps cross-run alignment when separate injections carry the contrast (Rappsilber et al. 2007).
Route-specific prep that matches the design. TMT labeling after digest can reduce run-order drift inside a plex when the full set can be frozen at tagging (Thompson et al. 2003). Targeted assays benefit when the same cleanup yields scheduleable peptides from every cohort file (Lange et al. 2008).
The main prep advantage in applications is matched peptide material that supports a declared contrast, not generic sensitivity language.
Researchers who need the full lysis-to-digest spine can refer to Quantitative Protein Analysis.
Pretreatment and Matrix Limits That Undermine Applications
The same application themes fail when prep principles break.
Unequal input or batch-specific lysis creates fold changes that track preparation, not biology. Normalization after a confounded prep cannot restore a defensible contrast.
Matrix-specific dropout. Plasma and serum often need equal depletion or fractionation before deep discovery. Depletion applied to only one arm creates artificial application signals. Low-abundance proteins in any lysate often appear as missing values, not as zero biology (Lazar et al. 2016).
Labeling chemistry conflicts. TMT needs amine-free peptide buffers. Metabolic labeling such as stable isotope labeling by amino acids in cell culture (SILAC) needs growable cells and is a poor default for tissue or biofluid application sets. Stable isotope-labeled internal standard (SIS) amount claims fail when spike timing sits on an incomplete or group-biased digest (Gerber et al. 2003).
Open enrollment after mixing. A TMT plex cannot absorb late unlabeled harvests as extra channels. Application plans that keep adding samples fit unlabeled separate injections better.
Prep disadvantages in applications are design failures that look like biological advantages after search.
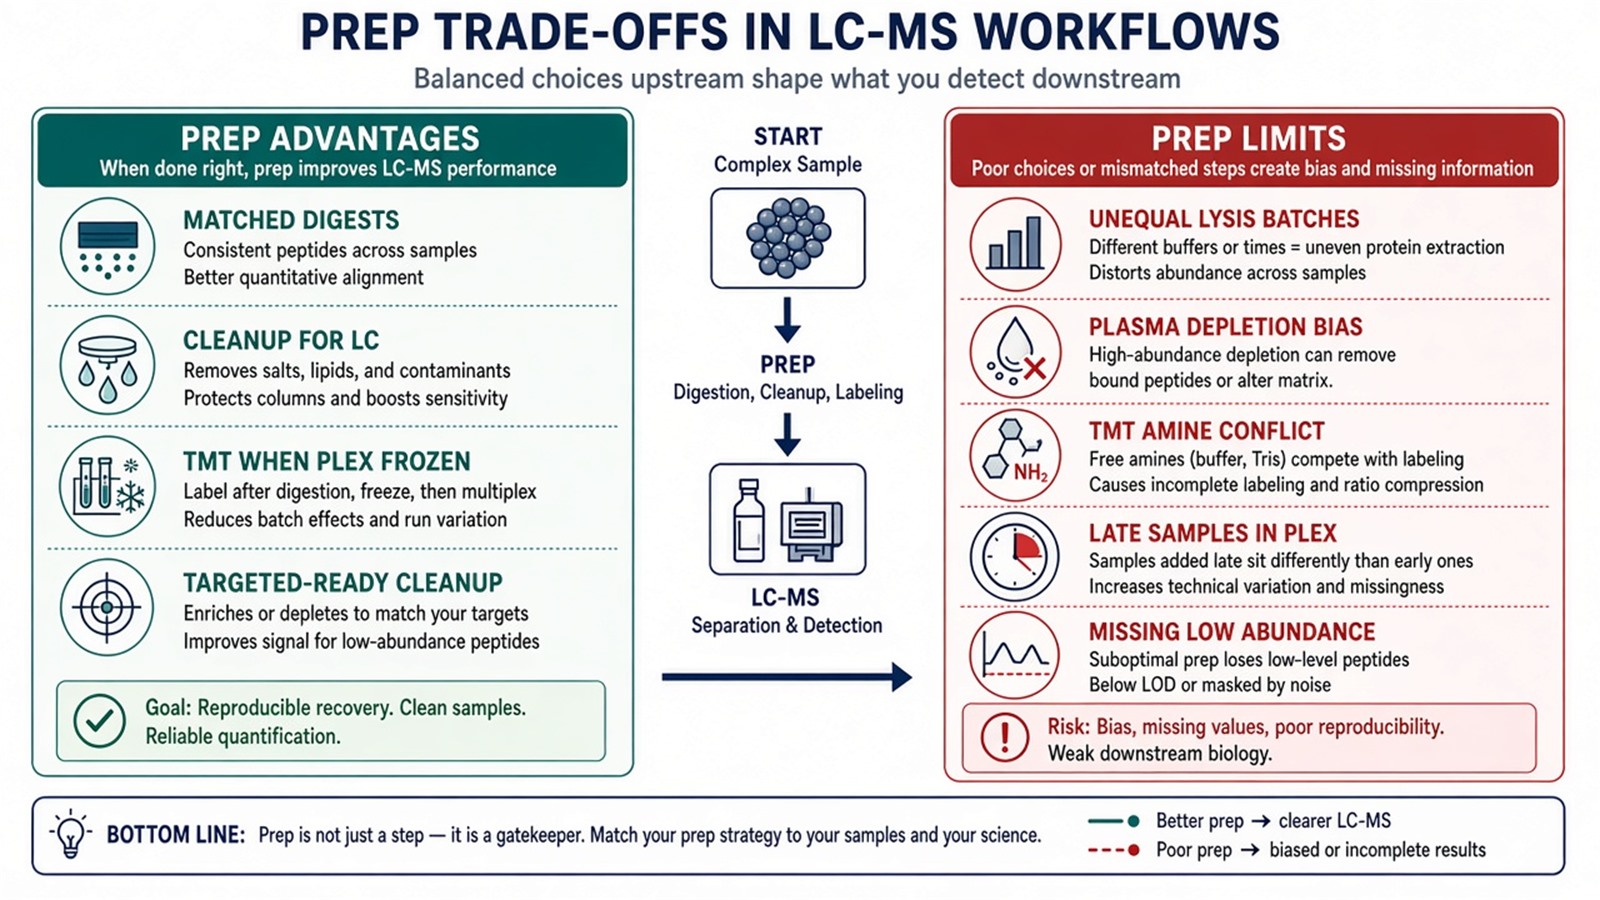
Figure 1. Applied quantification gains from matched prep and loses when matrix, labeling, or enrollment rules conflict with the sample map.
Exploratory enrollment and freeze-thaw checks before large batches are covered in Exploring Quantitative Analysis in Proteomics. This page stays on advantage and disadvantage trade-offs at the prep layer.
Shared Workflow Where Prep Strengths and Weaknesses Surface
Applied quantitative work exposes prep quality at predictable steps.
- Confirm matrix, contrast, and relative versus SIS amount claim.
- Aliquot matched input after a load assay.
- Lyse, digest, and clean under one declared chemistry.
- Desalt and set LC-ready loads; run a pilot when the recipe is new.
- Add TMT or SIS only when the application design requires them.
- Acquire LC-MS/MS and build the matrix under stated rules.
- Test the predefined contrast before application language is attached.
Advantages compound when steps 2 through 4 stay matched across groups. Disadvantages compound when step 4 differs by treatment, when step 5 runs on amine-carryover, or when step 6 starts before a pilot passes.
Pathway or biomarker wording only fits after step 7 returns a defensible relative matrix. Enrichment cannot repair step 3 batch confounding.
Parameters That Turn Prep into an Advantage or a Liability
Parameter |
Prep advantage when matched |
Liability when ignored |
|---|---|---|
Input (cells, mass, total protein) |
Stable peptide yield across arms |
Load-driven false changes |
Lysis and digestion batch |
Shared stoichiometry |
Batch equals treatment |
Cleanup recovery |
Equal peptide recovery |
Group-specific losses |
Detergent and salt removal |
Clean LC peaks |
Suppression and drift |
Amine-free buffer before TMT |
Complete reporters |
Soft channel ratios |
SIS spike timing |
Valid amount on targets |
Relative peaks as concentration |
Pilot before cohort LC |
Catches bad loads early |
Full batch repeat |
Limits follow the same table. Application advantages such as treatment response, mechanism contrast, or bioprocess comparison require peptide-level LC-MS/MS with matched prep. They do not follow from bulk dye assays, mRNA proxies, or identification depth alone. Targeted confirmation advantages appear only when discovery prep left scheduleable peptides.
Researchers who need route selection after prep trade-offs are clear can refer to Protein Quantification Method. For LC injection handoff details, see Quantitative Analysis of Proteins by Liquid Chromatography.
How to Choose Prep for Applied Quantitative Projects
Use matrix, enrollment flexibility, and claim unit together.
Application need |
Prep plan that often fits |
Prep disadvantage to avoid |
|---|---|---|
Open cohort, cells or tissue |
Matched unlabeled prep, separate injections |
Treatment-confounded lysis batches |
Freezable multi-arm set |
Matched digest, TMT, amine cleanup |
Late samples forced into the plex |
Plasma or serum discovery |
Equal depletion plus matched digest |
Depletion on one group only |
Pre versus post treatment |
Interleaved prep and LC order |
All treated samples in one prep day |
Named target follow-up |
Clean digest for PRM or MRM |
Oversized panel after poor cleanup |
Amount on selected proteins |
Matched digest plus assigned SIS |
Spike before digestion QC |
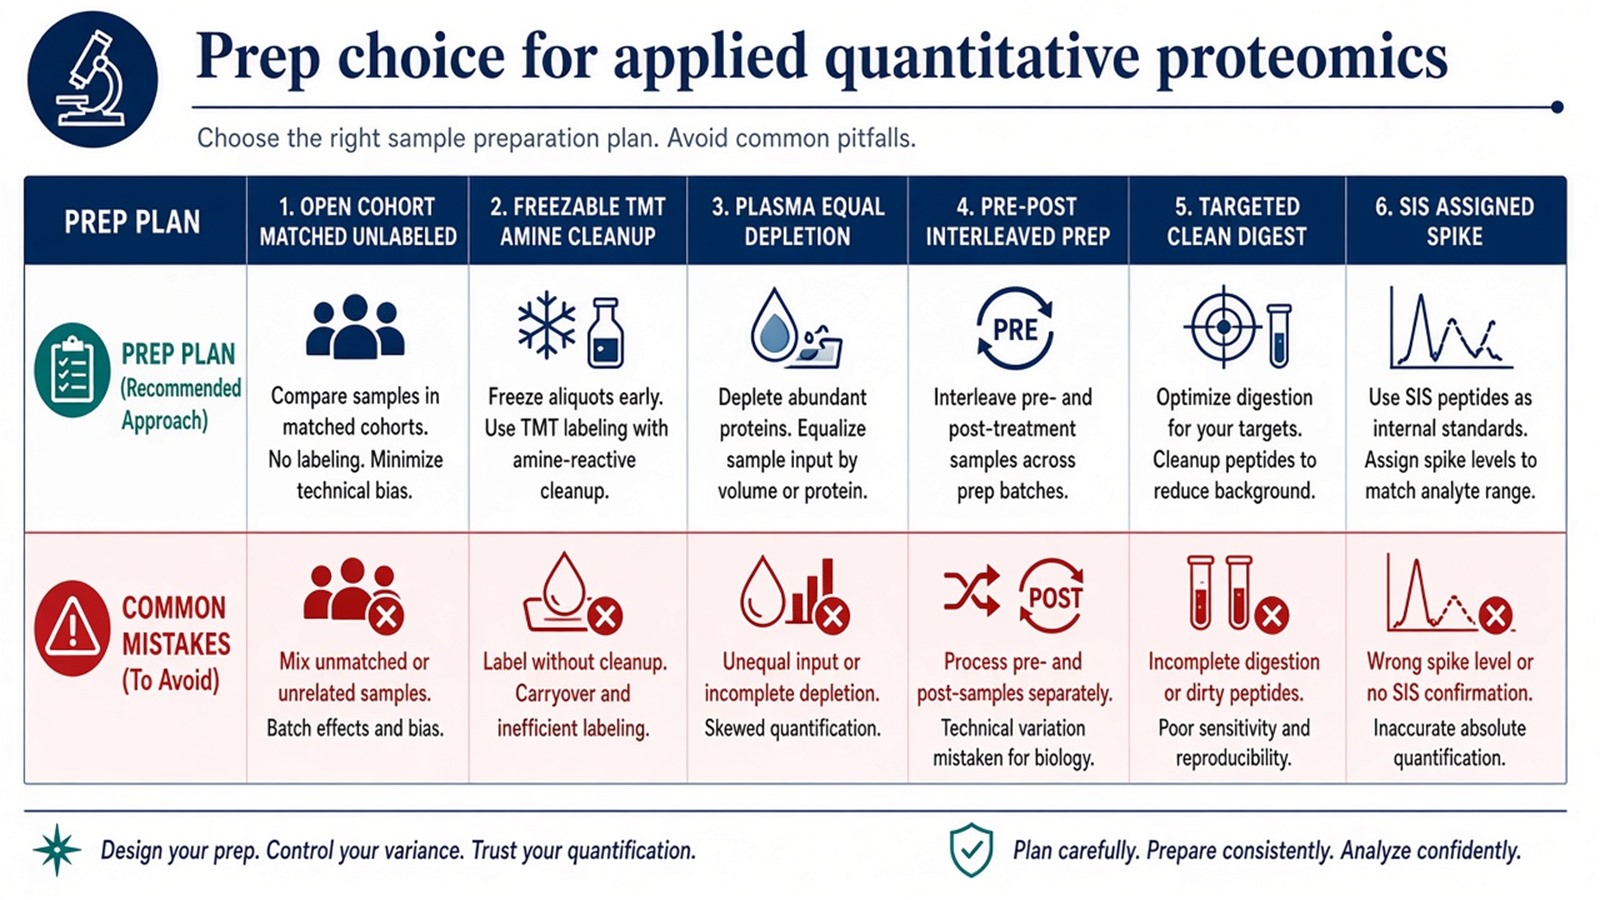
Figure 2. Choose prep to preserve application advantages; unequal cleanup, amine conflict, and amount-without-SIS are common disadvantages.
Unlabeled prep more often fits open application cohorts. TMT prep more often fits freezable multi-arm panels. Targeted prep more often fits after names freeze. Exceptions exist when a short closed cohort must share one LC method to limit drift, or when a pilot shows cleanup failure before the main application batch.
Teams that already have matrix, contrast, and a requested claim can also review the MtoZ Biolabs Quantitative Proteomics Service for project-specific prep planning in applied studies.
Frequently Asked Questions
1. Does quantitative proteomics always outperform Western blot in applications?
No. Western blot fits small configured targets when antibodies exist. Open or multi-protein application contrasts more often need matched LC-MS prep and quantification.
2. Is TMT prep always an advantage for multi-group applications?
No. TMT reduces within-plex run-order issues when the set can be frozen at labeling. Open enrollment applications more often stay unlabeled.
3. Can unequal cleanup still support biomarker application language?
No. Unequal depletion or recovery creates list differences that are prep artifacts. Application claims need matched prep across groups.
4. When does SIS prep become an advantage over relative intensities?
When the application decision needs a mass or molar unit on named peptides and a calibration plan exists. Relative matrices remain enough for many mechanism or treatment contrasts.
5. Does a successful pilot prep remove the need for cohort QC?
No. Pilots catch load and cleanup problems early. The full cohort still needs the same matched rules and QC flags before contrast testing.
MtoZ Biolabs can support applied quantitative proteomics prep across unlabeled DDA or DIA discovery, TMT relative plexes, and targeted PRM or MRM follow-up when sample type, study design, and the intended claim are available.
If further evaluation of matrix-matched prep, depletion strategy, TMT compatibility, or SIS spike planning for an applied contrast is needed, project details can be submitted for a feasibility review.
Reference
- M. Bantscheff, S. Lemeer, M.M. Savitski, B. Kuster (2012). Quantitative mass spectrometry in proteomics: critical review update from 2007 to the present. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-6203-4
- J. Cox, M.Y. Hein, C.A. Luber, I. Paron, M. Mann (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031031
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- A. Thompson, J. Schafer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, R. Johnstone, A.K. Mohammed, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- J. Rappsilber, M. Mann, Y. Ishihama (2007). Protocol for micro-purification, enrichment, pre-fractionation and storage of peptides for proteomics using StageTips. Nat. Protoc., 2, 1896-1906. https://doi.org/10.1038/nprot.2007.261
- J.R. Wisniewski, A. Zougman, N. Nagaraj, M. Mann (2009). Universal sample preparation method for proteome analysis. Nat. Methods, 6, 359-362. https://doi.org/10.1038/nmeth.1322
