



Protein targeted quantification measures named proteins by scheduling proteotypic peptides for liquid chromatography-tandem mass spectrometry (LC-MS/MS) and extracting fragment chromatograms. The project starts with a closed list, not with an unknown proteome. Parallel reaction monitoring (PRM) and multiple reaction monitoring (MRM) / selected reaction monitoring (SRM) are the usual acquisition routes (Lange et al. 2008; Peterson et al. 2012). The default result is a same-peptide relative value. An amount in mass or molar units requires a matched stable isotope-labeled internal standard (SIS) and a declared calibration.
In MtoZ Biolabs offerings, targeted quantification is typically acquired as high-resolution PRM. An MRM panel remains a valid route when a scheduled triple-quadrupole assay is already the production design. PRM is not automatically cheaper, simpler, or more accurate than MRM for every panel.
What Protein Targeted Quantification Can Report
A targeted assay can remeasure listed peptides across a cohort with a locked schedule. That supports confirmation of discovery candidates, monitoring of a pre-specified pathway set, and, when a unique peptide exists, distinction of some homologs or modified forms. Sequence-defined fragments do not require a commercial antibody. They also do not automatically match an immunoassay that reports a different epitope.
The assay cannot inventory proteins that were never scheduled. It cannot rank different proteins as copy numbers from peak height. It cannot treat a relative PRM or MRM ratio as ng per mL. Hundreds of samples can be acquired after a method exists; that throughput is not a property of an unbuilt panel.
If the remaining question is whether a closed list is even the right class of experiment, researchers can refer to Advantages and Disadvantages of Targeted Proteomics. If the remaining question is the six-step closed-list sequence, researchers can refer to Targeted Proteomics Workflow: How Does Targeted Proteomics Work?.
How Named Proteins Become a Scheduled Assay
A practical order is stable even when the instrument class changes. Protein names are frozen. Proteotypic peptides are chosen for uniqueness, digestion yield, and detectable fragments. Matched extracts are digested and cleaned up. SIS peptides are added only when an amount claim is planned, with the spike point recorded. Precursors are scheduled on a PRM or MRM method. Peaks that pass shape, qualifier, and interference checks are integrated. Only then should a group contrast or a calibration be reported.
PRM isolates each scheduled precursor and records a high-resolution fragment spectrum, so fragment choice can continue after acquisition (Peterson et al. 2012). MRM monitors predefined precursor-to-product transitions on a triple quadrupole, which can support larger locked production panels (Lange et al. 2008). Both routes still need unique peptides, chromatography that samples each peak, and a claim that matches whether SIS is present.
Protein targeted quantification reports only peptides that were scheduled and accepted; identification of a protein name in an earlier discovery file is not a targeted result.
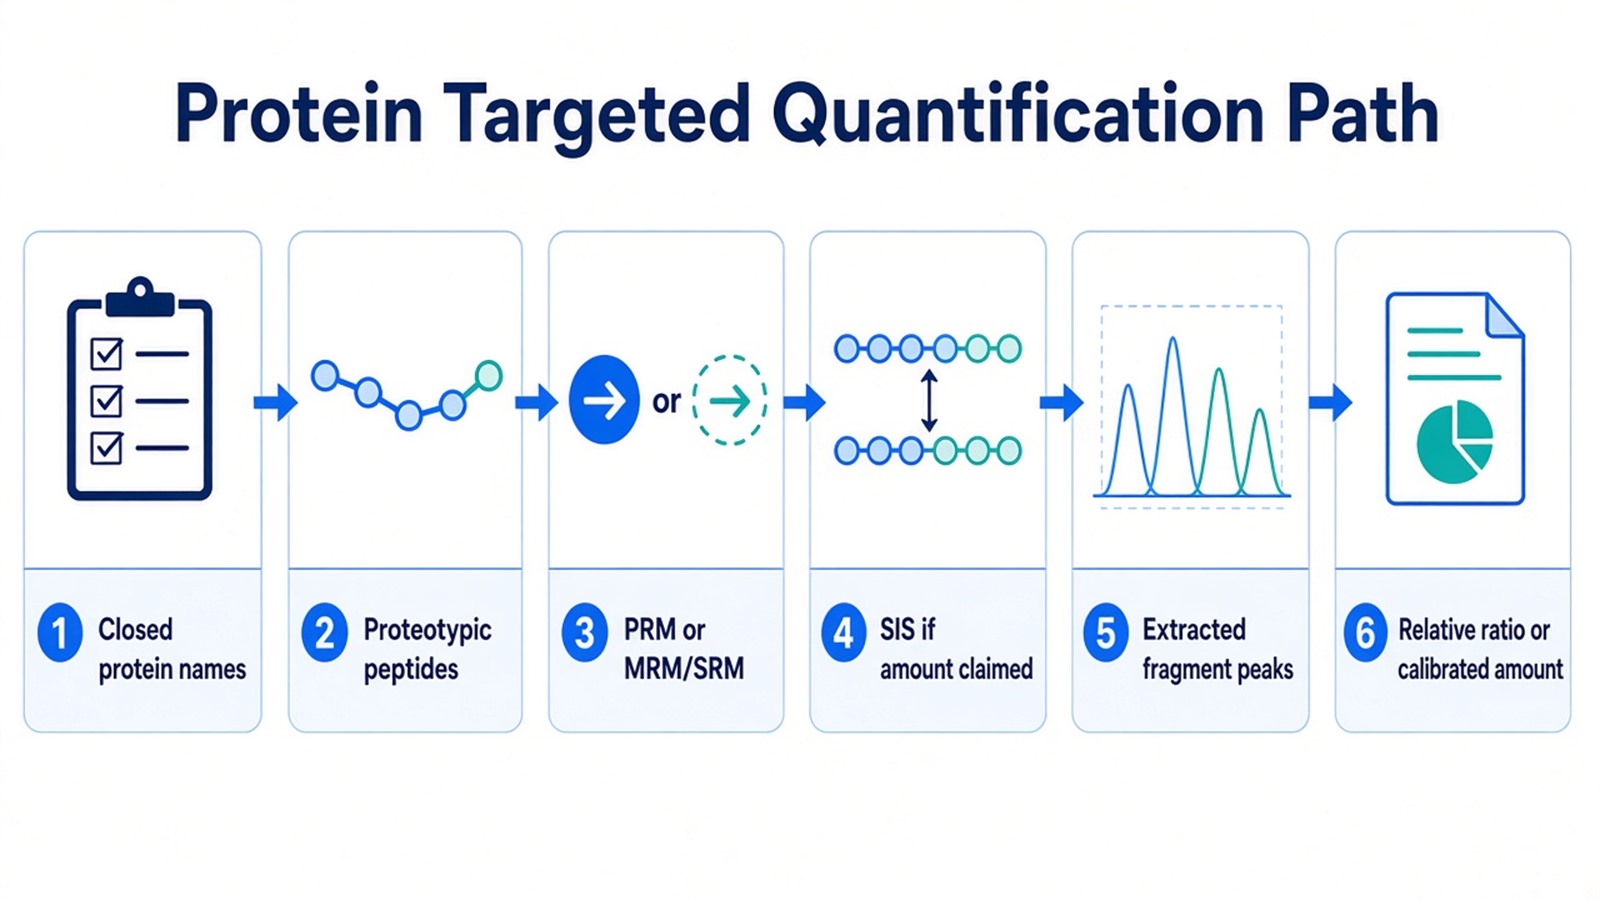
Figure 1. Protein targeted quantification turns a closed name list into extracted fragment peaks before any relative ratio or SIS-supported amount is reported.
Discovery matrices such as unlabeled DDA, DIA, or TMT can supply candidate names. They do not replace peptide detectability checks in the new digest. A prior DDA or DIA dataset is often useful. Sequence-clear targets in a simple matrix can still be evaluated without that file.
Parameters That Control Detectability, Interference, and Units
Peptide uniqueness controls which gene products are reported. Shared peptides belong to a protein group. Site-specific modified peptides report that modified sequence, not occupancy, unless the unmodified partner is measured on the same scale.
Scheduling controls whether the listed peptide is actually sampled. Retention-time windows, isolation width, collision energy, and the number of fragments or transitions per peptide must fit the chromatographic peak width. A panel that is too long for one duty cycle should be split or shortened. There is no universal protein-count capacity that holds for every gradient.
Interference control includes qualifier fragments, peak-shape rules, and SIS co-elution when a heavy peptide is present. Fit-for-purpose documentation should lock those rules before the study set is quantified (Carr et al. 2014). Carryover checks belong after high standards or high-abundance samples.
Unit control is a separate parameter. Relative confirmation uses light peak areas, optionally with a heavy peptide as a loading reference. Amount reporting uses an SIS of assigned concentration. Researchers can refer to Absolute Protein Quantification when the remaining task is calibration, not closing the name list.
Limits of Reporting a Protein From One or a Few Peptides
One accepted peptide is still a peptide result. Proteolytic processing, splice forms, and post-translational modifications can split one protein name across several sequences. A targeted number for an unmodified peptide does not report the modified fraction.
Method development time sits in front of cohort acquisition. Peptide failure, poor chromatography, and matrix suppression are common reasons a named protein never becomes a reportable assay. Enrichment can help some low-abundance or modified targets. It is an extra experimental block, not a software setting.
Targeted quantification does not replace discovery while names are still being added. It also does not replace a well-characterized immunoassay when the clinical or kit epitope is the object of the study. Cross-laboratory transfer requires the same peptides, the same schedule, and the same acceptance criteria.
When to Start a Targeted Quantification Project
The useful test is whether names can freeze, whether unique peptides are known or can be tested, and whether the unit is relative or an amount.
Study need |
Check first |
Route that often fits |
|---|---|---|
Discovery candidates to confirm |
Unique, detectable peptides |
Targeted PRM or MRM, relative |
Pre-specified pathway panel |
List length versus cycle time |
Targeted method, split if needed |
Amount or concentration |
SIS availability and spike point |
Targeted plus matched SIS |
Fragments still under review |
Interference in the matrix |
PRM more often, until transitions lock |
Locked production panel, many samples |
Transition maturity |
MRM more often on a triple quadrupole |
Open protein inventory |
Whether names can freeze |
Discovery first, then a short targeted list |
Shared peptide only |
Homologs and isoforms |
Do not report a single-gene amount from that peptide |
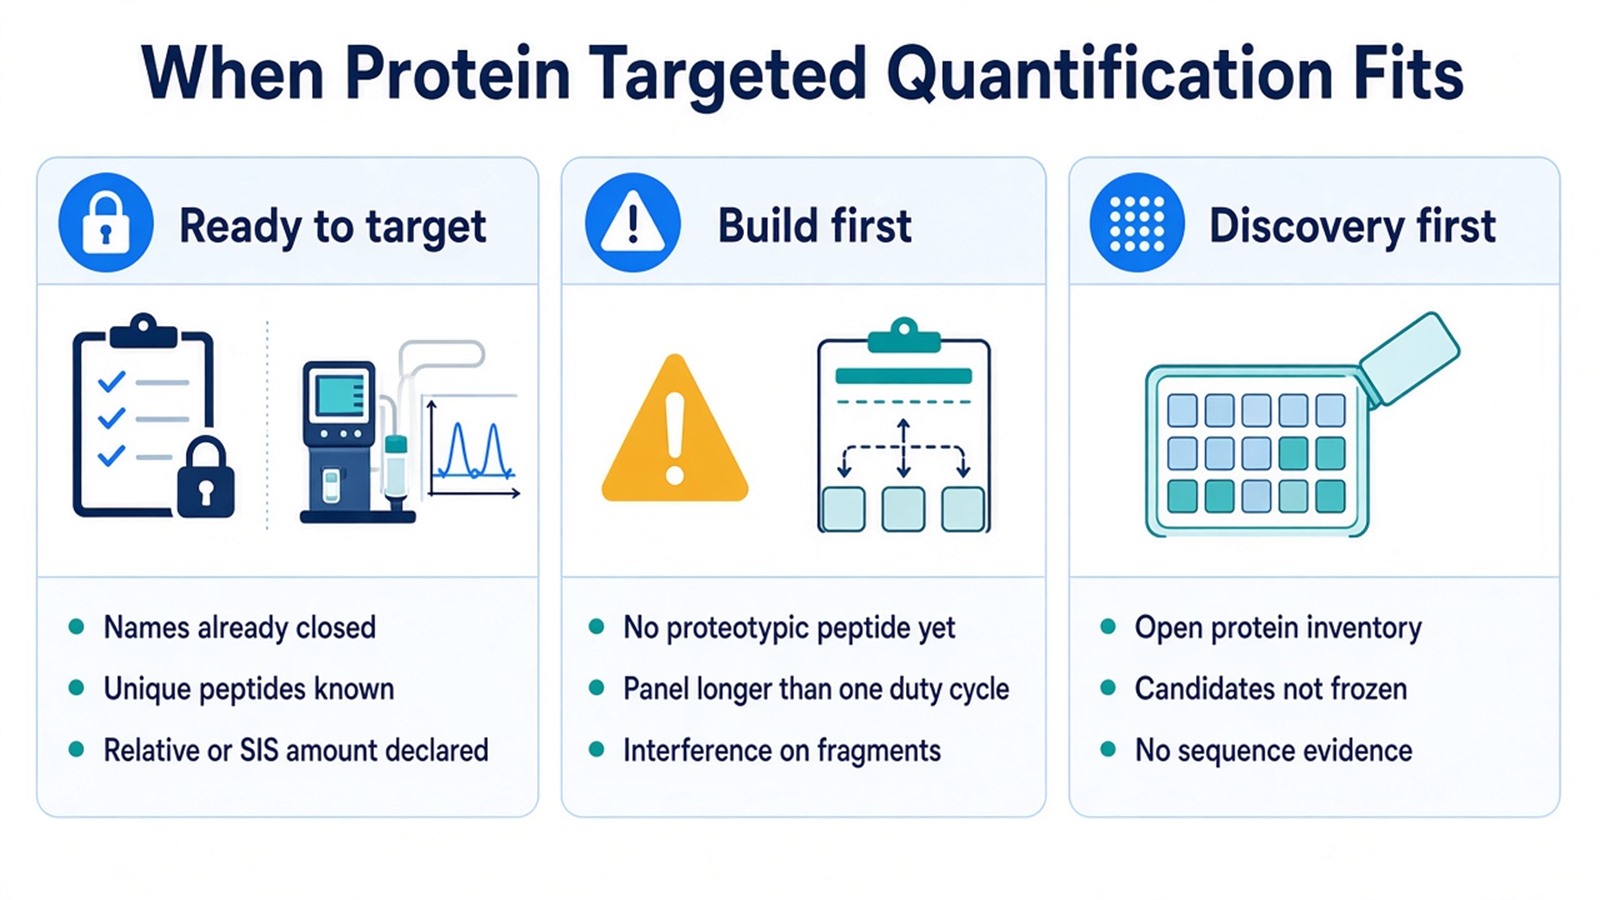
Figure 2. Protein targeted quantification fits a closed, detectable peptide list with a declared relative or SIS-supported claim.
PRM more often fits a new panel that still needs fragment review. MRM more often fits a locked transition set built for repeated cohorts. An exception is a short, clean PRM panel that is already production-ready, or an MRM method that must be rebuilt because interference appeared in a new matrix.
Researchers who already have named proteins, sample type information, and a requested relative or SIS-supported claim can also review the MtoZ Biolabs Targeted Proteomics Service for project-specific peptide evaluation and assay planning.
Frequently Asked Questions
1. Must a discovery DDA or DIA dataset exist before targeted quantification?
A matched discovery file often supplies detectable peptides and retention-time clues. Sequence-clear targets in a simpler matrix can still be evaluated without that file. Undetected peptides in the actual digest cannot be rescued by a literature sequence alone.
2. How many proteins can one targeted method quantify?
The limit is set by peak width, cycle time, and how many fragments or transitions each peptide needs. When the schedule is too dense, split the method or shorten the list. A single published protein-count capacity does not transfer across gradients.
3. Can a phosphorylation or other PTM site be targeted?
Yes, when a site-specific peptide can be scheduled and, for many low-stoichiometry sites, when enrichment is evaluated. A modified SIS reports that modified peptide. Occupancy still needs the unmodified partner on the same scale.
4. Are targeted fold changes interchangeable with an earlier discovery matrix?
They confirm the same peptide in a new assay. They can diverge from discovery intensities because isolation, interference, and digestion batches differ. Treat them as a confirmation layer, not as a silent replacement of the discovery table.
5. Does targeted quantification replace Western blotting for every protein name?
It replaces the need for a commercial antibody when a unique peptide works. It does not automatically reproduce an epitope-based result. Report the peptide sequence that was measured.
MtoZ Biolabs can provide protein targeted quantification using PRM or MRM/SRM for closed protein or peptide lists, including proteotypic peptide evaluation, optional SIS-supported calibration, and documented peak QC when project files and study design are available.
If further evaluation of panel size, PRM versus MRM routing, or whether discovery should run first is needed, project details can be submitted for a feasibility review.
Reference
- S.A. Carr, S.E. Abbatiello, B.L. Ackermann, C. Borchers, B. Domon, E.W. Deutsch (2014). Targeted peptide measurements in biology and medicine: best practices for mass spectrometry-based assay development using a fit-for-purpose approach. Mol. Cell. Proteomics, 13, 907-917. https://doi.org/10.1074/mcp.M113.036095
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O111.013698
