



A laboratory that already named proteins often still has to decide whether a closed-list mass spectrometry assay is worth building. Targeted proteomics schedules predefined peptides for parallel reaction monitoring (PRM) or multiple reaction monitoring (MRM) / selected reaction monitoring (SRM), and quantifies those sequences from extracted fragment signals (Lange et al. 2008; Peterson et al. 2012). The method is not an antibody panel and not an open discovery matrix. Gains come from measuring only what was scheduled. Costs come from the same closure.
Relative same-peptide comparison is the default claim. An amount in mass or molar units requires a matched stable isotope-labeled internal standard (SIS) and a declared calibration. In MtoZ Biolabs offerings, targeted panels are typically acquired as high-resolution PRM. An MRM panel remains a valid route when a scheduled triple-quadrupole assay is already the production design.
What Targeted Proteomics Is Designed to Do
Targeted proteomics is built for confirmation and monitoring of a locked peptide list. Once proteotypic peptides are chosen, the instrument spends its cycle on those precursors. That design can remeasure the same sequences across many samples with fewer stochastic missing values than data-dependent acquisition (DDA) discovery for those peptides (Picotti and Aebersold 2012). Sequence-defined fragments give a chemical identity that an epitope assay does not provide.
The method is not designed to inventory an unknown proteome. Proteins that never entered the inclusion list stay unreported, even if they are abundant in the digest. Speed per named peptide can be high after the method exists. Speed per new protein name is low, because each addition needs peptide selection and scheduling. Cost is not automatically lower than discovery: method development, SIS synthesis when used, and chromatography QC are real project costs.
Researchers who need the operational closed-list sequence can refer to Targeted Proteomics Workflow: How Does Targeted Proteomics Work?.
Where a Closed Peptide List Helps
A usable advantage appears only when names are already closed and the peptides can be detected.
Repeatability for listed targets is the main gain. The same transitions or fragments can be extracted in every sample, which supports longitudinal or multi-site relative comparison when the method is locked (Carr et al. 2014). Specificity is peptide-sequence specificity. Homologs can still share a peptide; uniqueness has to be declared, not assumed.
Duty-cycle use is another gain. Discovery methods spend time on unidentified or one-hit precursors. A targeted method spends time on the panel. Low-abundance listed peptides can become measurable when isolation is scheduled, provided the digest and chromatography support a clean peak.
SIS-supported amount is an optional gain, not a built-in property. AQUA-style heavy peptides convert a light-to-heavy ratio into peptide amount. Researchers can refer to Absolute Protein Quantification when the remaining claim is concentration, not a targeted relative ratio.
PRM and MRM realize those gains by different isolation rules. PRM records a high-resolution fragment spectrum for each scheduled precursor and allows fragment choice after acquisition (Peterson et al. 2012). MRM monitors predefined precursor-fragment transitions on a triple quadrupole and can carry larger production panels when transitions are already validated (Lange et al. 2008). Acquisition class is not a separate biology.
Targeted proteomics improves remeasurement of listed peptides; it does not expand the protein inventory.
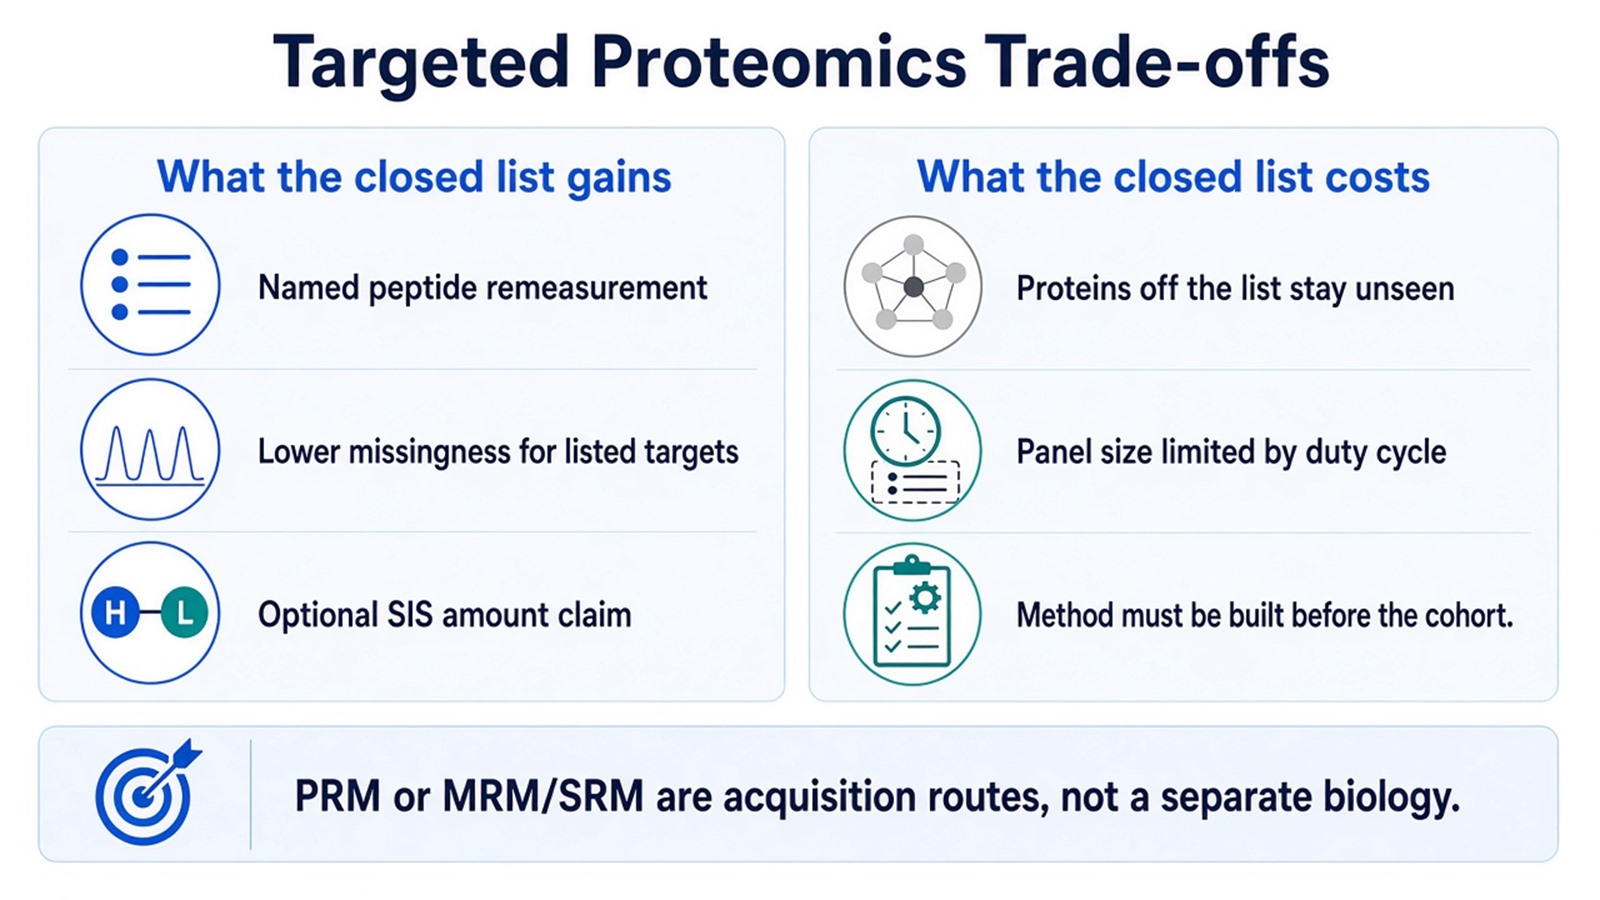
Figure 1. Targeted proteomics trades an open protein list for repeatable measurement of scheduled peptides.
Where the Same Closure Constrains the Project
The closed list is also the main limitation. New biology that was not named before digestion will not appear. Discovery follow-up has to freeze candidates first. A panel that is still growing is a poor targeted project.
Panel size is limited by chromatography and cycle time. Too many precursors in one method thin the points across each peak. The practical response is to split methods, shorten the list, or accept lower sampling of each peptide. There is no universal protein-count capacity that holds for every gradient.
Interference remains. Co-eluting ions can still occupy a PRM isolation window or an MRM transition. Qualifier fragments, peak-shape rules, and, when used, SIS co-elution are part of the method, not optional graphics. Cross-laboratory agreement is not automatic. It depends on a locked schedule, the same peptides, and documented acceptance criteria (Carr et al. 2014).
Relative targeted peak areas are not concentrations. Treating a PRM or MRM ratio as ng per mL without SIS overclaims the assay. Immunoassay disagreement is expected when the epitope and the peptide report different proteoforms.
Parameters That Decide Whether the Trade-off Holds
Peptide uniqueness, detectability, and modification state decide whether the listed protein is actually what is measured. Shared or ragged peptides weaken the protein claim. Spike timing of SIS, when used, decides which losses are captured.
Scheduling parameters decide whether the listed advantage survives acquisition. Retention-time windows, isolation width, collision energy, and the number of fragments or transitions per peptide should be locked before the study set is acquired. Qualifier agreement and carryover checks belong in the QC record.
Claim type should be declared with the panel: relative confirmation, SIS-supported amount, or site-specific modified peptide. Mixing those claims in one table without labeling the unit confuses later statistics.
When PRM fragment choice is the remaining decision, researchers can refer to PRM LC-MS Targeted Proteomics. When a large, scheduled quadrupole panel is the remaining decision, the MRM trade-off is the closer next read.
When Targeted Proteomics Is the Better Next Step
The useful test is whether names can freeze, whether the peptides are unique and detectable, and whether the claim matches the presence or absence of SIS.
Study need |
Targeted proteomics often fits |
Recheck or choose discovery first |
|---|---|---|
Named proteins, remeasure across samples |
Closed PRM or MRM panel |
Protein list still open |
Discovery candidates to confirm |
Short follow-up list from a prior matrix |
Hundreds of new names still required |
Relative fold change for listed peptides |
Targeted without SIS |
Amount unit required with no SIS plan |
Amount or concentration for listed peptides |
Targeted with matched SIS |
Ordinary peak areas treated as ng per mL |
Homolog or isoform question |
Unique peptides for each product |
Shared peptide used as a gene-level result |
Unknown proteome inventory |
Not the first method |
DDA or DIA discovery matrix |
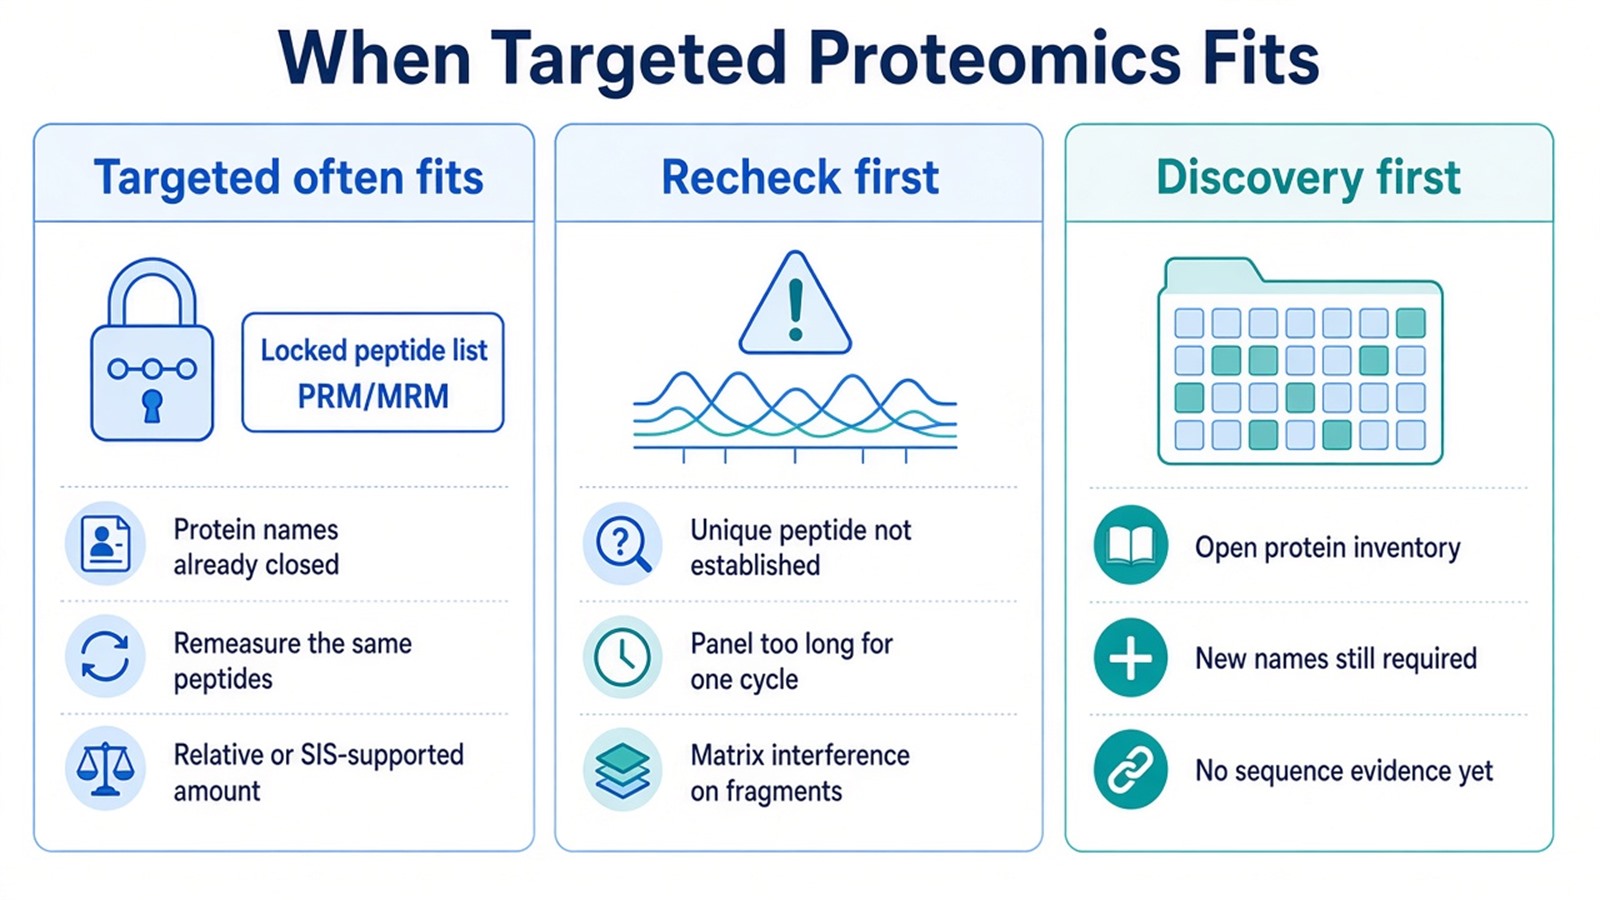
Figure 2. Targeted proteomics fits a closed, detectable peptide list; an open inventory still belongs on a discovery matrix.
Targeted acquisition more often fits after a discovery matrix has produced a short candidate list. An exception is a pre-specified pathway panel that never needed discovery. Discovery more often fits when new names are still required. An exception is a tiny, already-validated set that would waste a full proteome run.
Researchers who already have named proteins, sample type information, and a requested relative or SIS-supported claim can also review the MtoZ Biolabs Targeted Proteomics Service for project-specific peptide evaluation and assay planning.
Frequently Asked Questions
1. Is targeted proteomics the same as an immunoassay for the same protein name?
No. Mass spectrometry reports a defined peptide sequence. An immunoassay reports an epitope. The two numbers can diverge when proteoforms, matrix, or antibody cross-reactivity differ.
2. Does a targeted MS assay always report absolute concentration?
No. PRM or MRM without an SIS is a targeted relative assay. Absolute reporting starts when a matched SIS of assigned amount and a calibration rule are present.
3. Can a targeted panel discover proteins that were never scheduled?
No. Acquisition will not measure a peptide that was never placed on the method. Keep an open inventory on DDA or DIA discovery, then freeze a short list for targeted follow-up.
4. Is targeted proteomics always cheaper or faster than DIA or TMT?
Not at the project level. After a method exists, listed peptides can be remeasured efficiently. Building uniqueness, scheduling, and QC for a new panel can exceed the cost of one discovery cohort. Compare against the number of names and whether they are already closed.
5. Should every targeted project use PRM instead of MRM?
No. PRM more often fits when fragment identity still needs review after acquisition. MRM more often fits a locked, larger transition panel on a triple quadrupole. Panel size, existing methods, and interference, not a single default, should decide.
MtoZ Biolabs can provide targeted proteomics using PRM or MRM/SRM for closed protein or peptide lists, including peptide evaluation, optional SIS-supported calibration, and documented peak QC when project files and study design are available.
If further evaluation of panel size, unique-peptide choice, or whether discovery should run first is needed, project details can be submitted for a feasibility review.
Reference
- S.A. Carr, S.E. Abbatiello, B.L. Ackermann, C. Borchers, B. Domon, E.W. Deutsch (2014). Targeted peptide measurements in biology and medicine: best practices for mass spectrometry-based assay development using a fit-for-purpose approach. Mol. Cell. Proteomics, 13, 907-917. https://doi.org/10.1074/mcp.M113.036095
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O111.013698
- P. Picotti, R. Aebersold (2012). Selected reaction monitoring-based proteomics: workflows, potential, pitfalls and future directions. Nat. Methods, 9, 555-566. https://doi.org/10.1038/nmeth.2015
