



Protein extraction for TMT/iTRAQ begins before any isobaric tag is opened. Tandem mass tags (TMT) and isobaric tags for relative and absolute quantitation (iTRAQ) attach amine-reactive labels to digests so peptides from different samples share one precursor mass on liquid chromatography-tandem mass spectrometry (LC-MS/MS), then release reporter ions for within-plex ratios (Thompson et al. 2003; Ross et al. 2004). In project language, extraction, buffer exchange, matched protein load, digest, and amine cleanup must be parity-locked across channels before labeling. Reporter software cannot repair a channel that lost membrane proteins in lysis or still contains free Tris competing for N-hydroxysuccinimide (NHS) esters. The guide below maps extraction and pretreatment decisions for TMT/iTRAQ workflows, locked parameters, limits, and project fit. It is not a biomedical proteomics primer, not a claim that ultrasonic lysis alone improves accuracy, and not a catalog of future extraction inventions.
Unmatched extraction recipes across arms produce peptide-pool shifts that look like biology after reporter normalization (Bantscheff et al. 2012). Relative within-plex ratios remain the default claim unit unless a matched stable isotope-labeled internal standard (SIS) design is added for amount.
In MtoZ Biolabs quantitative offerings, TMT/iTRAQ extraction work is scoped after matrix, lysis recipe, cleanup path to labeling-compatible buffers, channel map, and quality control (QC) checks are written in the brief. Service intake appears in TMT Quantitative Proteomics Analysis Service and iTRAQ.
What Extraction Scope Must Cover for TMT/iTRAQ
Four sample layers belong in the brief and stay separated.
Matrix-and-harvest layer names tissue, cells, biofluid, or other matrices, plus collection and storage history. Harvest timing must match the declared contrast.
Lysis-and-extraction layer names physical disruption if used, chaotropes or detergents, protease inhibitors when justified, and whether membrane, nuclear, or secreted fractions are intentionally targeted. “Release all proteins” without a recipe version is not a locked step.
Cleanup-to-digest layer names how detergents and strong chaotropes are reduced to digest-compatible levels (precipitation, filter-aided methods, bead-based workflows, or declared equivalents) and how protein amount is measured for matched load.
Labeling-prep layer names reduction, alkylation, digest enzyme, and removal of primary-amine buffers before NHS tagging. Residual Tris or excess ammonium salts compete with peptide amines for TMT/iTRAQ reagents.
Extraction success for TMT/iTRAQ means matched protein recovery into a labeling-compatible peptide pool. It does not mean every lysis buffer is MS-ready.
Plex design after prep appears in Protein Quantification Technology-TMT Labeling Quantitation. Combined iTRAQ/TMT service framing appears in iTRAQ-TMT. This page stays on sample and pretreatment gates.
How Extraction Chemistry Gates Digest and Isobaric Labeling
Lysis chemistry and labeling chemistry are one planning object.
Detergent-rich extracts can solubilize more proteins and still fail downstream if SDS or similar surfactants remain at levels that suppress digestion or chromatography. Chaotrope concentration must be brought into the declared digest window before trypsin or alternative enzymes run.
Amine-reactive TMT/iTRAQ reagents require peptide N-termini and lysine amines. Extraction or neutralization buffers that leave high primary-amine background steal reagent and leave incomplete labeling. Matched protein micrograms per channel must be fixed after cleanup, not only after crude lysis, or channel offsets appear as reporter bias.
iTRAQ and TMT share this pretreatment logic even when kit plex sizes differ. Changing extraction mid-cohort splits the isobaric record the same way for either tag family.
End-to-End Workflow From Extraction to Labeling-Ready Digests
A practical extraction-first TMT/iTRAQ spine usually runs as follows.
- Freeze contrast, matrix, and relative claim for the plex.
- Lock one lysis and disruption recipe for every channel; version the buffer composition.
- Extract proteins; clarify debris under matched conditions.
- Remove or dilute detergents and chaotropes to the declared digest-compatible state.
- Quantify protein and normalize load across channels before reduction and alkylation.
- Digest under locked enzyme and missed-cleavage rules; desalt into a low-amine labeling buffer.
- Confirm labeling-buffer compatibility; reserve pilot aliquots for labeling-completeness checks.
- Label, quench, and mix only after prep QC passes; acquire the declared MS2 or MS3 method.
- Stop and re-extract if one arm shows systematic recovery loss, incomplete digest, or amine interference.
Steps 1 through 7 protect the sample record. Steps 8 through 9 produce the isobaric quantitative record.
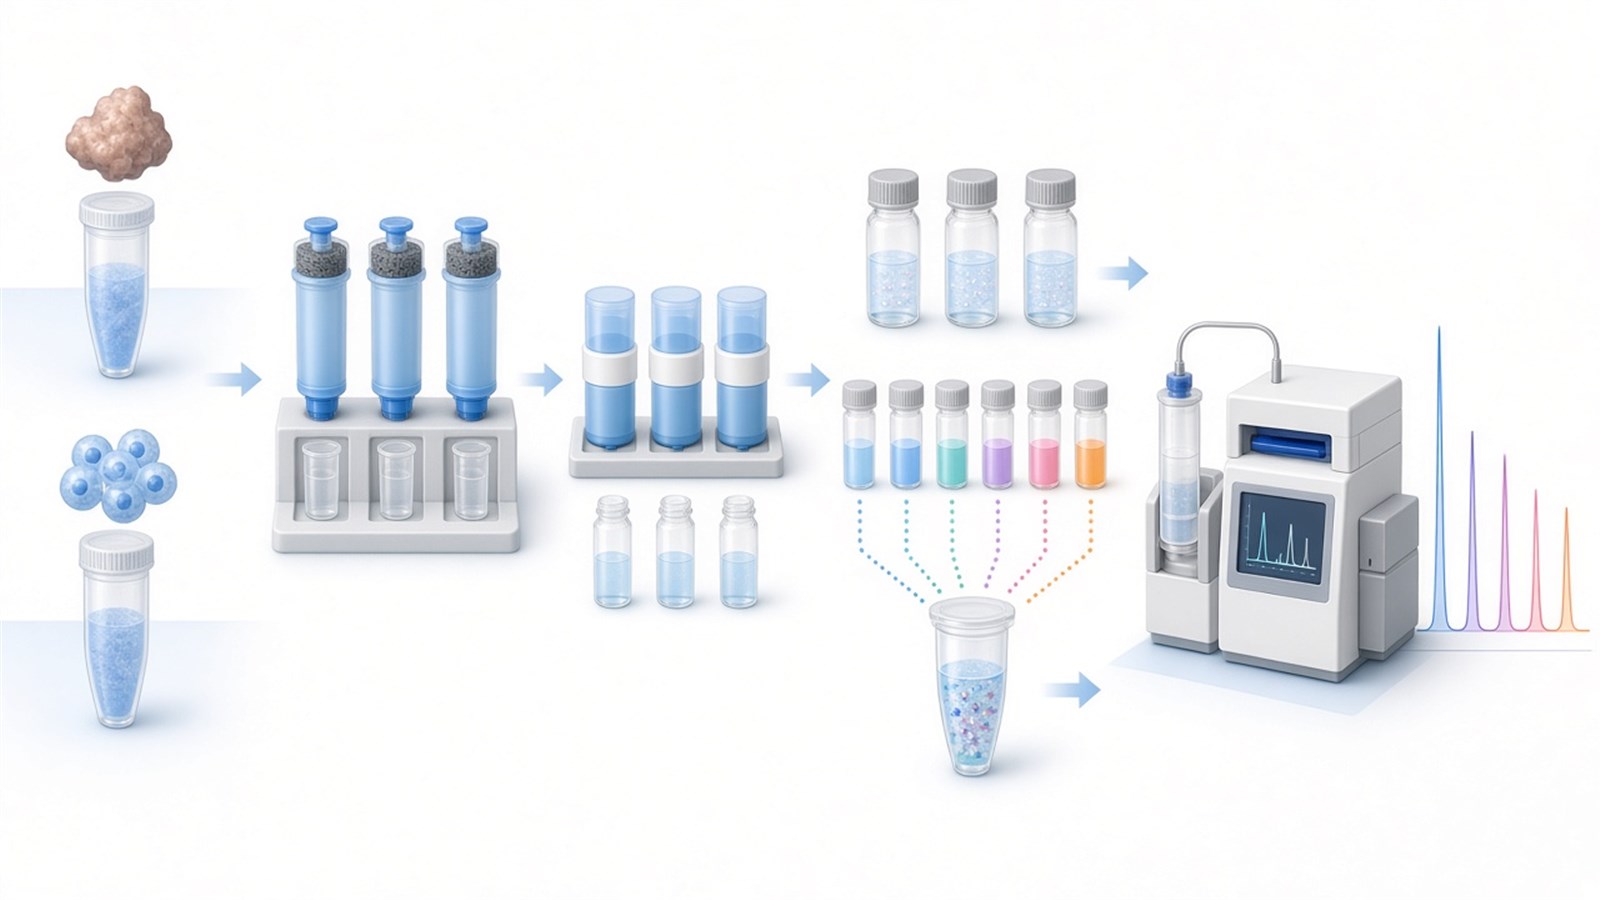
Figure 1. TMT/iTRAQ quantification depends on matched extraction and labeling-compatible cleanup before reporter ratios are tested.
Extraction and Pretreatment Parameters That Must Be Locked
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
Matrix |
Sample type, harvest, storage |
Hidden composition shift |
Lysis |
Buffer version, detergent/chaotrope, disruption |
Unequal proteome slices |
Inhibitors |
Protease inhibitor policy |
Selective protein loss |
Cleanup |
Detergent/chaotrope removal method |
Digest or LC failure |
Load |
Matched protein amount post-cleanup |
Channel offset as biology |
Digest |
Enzyme, time, missed-cleavage policy |
Unequal peptides for tagging |
Amine cleanup |
Buffer exchange before NHS labeling |
Incomplete TMT/iTRAQ labeling |
Channel map |
Which extract occupies which tag |
Prep bias locked into plex |
QC |
Extraction yield, digest, labeling completeness |
Undetected arm failure |
Deliverable |
Extraction SOP and buffer versions |
Report rebuilt after acquisition |
Limits follow the same table. Stronger lysis is not always better if it introduces surfactants that never leave. Absolute quantitation language needs SIS designs beyond standard relative TMT/iTRAQ. Clinical or biomarker promises are outside the extraction table.
How to Choose Extraction Paths for TMT/iTRAQ Projects
Use matrix, detergent burden, amine compatibility, and plex status together.
Project need |
Extraction path that often fits |
Recheck before committing |
|---|---|---|
Matched cells or tissue, standard relative plex |
One locked lysis + cleanup to labeling buffer |
Different lysis per treatment arm |
High detergent needed for membrane proteins |
Declared surfactant removal before digest |
Inject SDS-rich extract into labeling |
Residual Tris or amine buffers after cleanup |
Extra buffer exchange before tagging |
Label through competing amines |
Load still unequal after crude lysis |
Normalize after cleanup quantification |
Label on OD estimates from dirty lysate |
Recipe still changing mid cohort |
Stop channel lock |
Mix plex under drifting extraction |
Ultra-low single-cell input |
Single-cell TMT nano-prep design |
Bulk extraction SOP on one-cell claims |
Goal is expandable unlabeled cohort |
Consider LFQ/DIA instead of forcing tags |
Extract for TMT when multiplex is unnecessary |
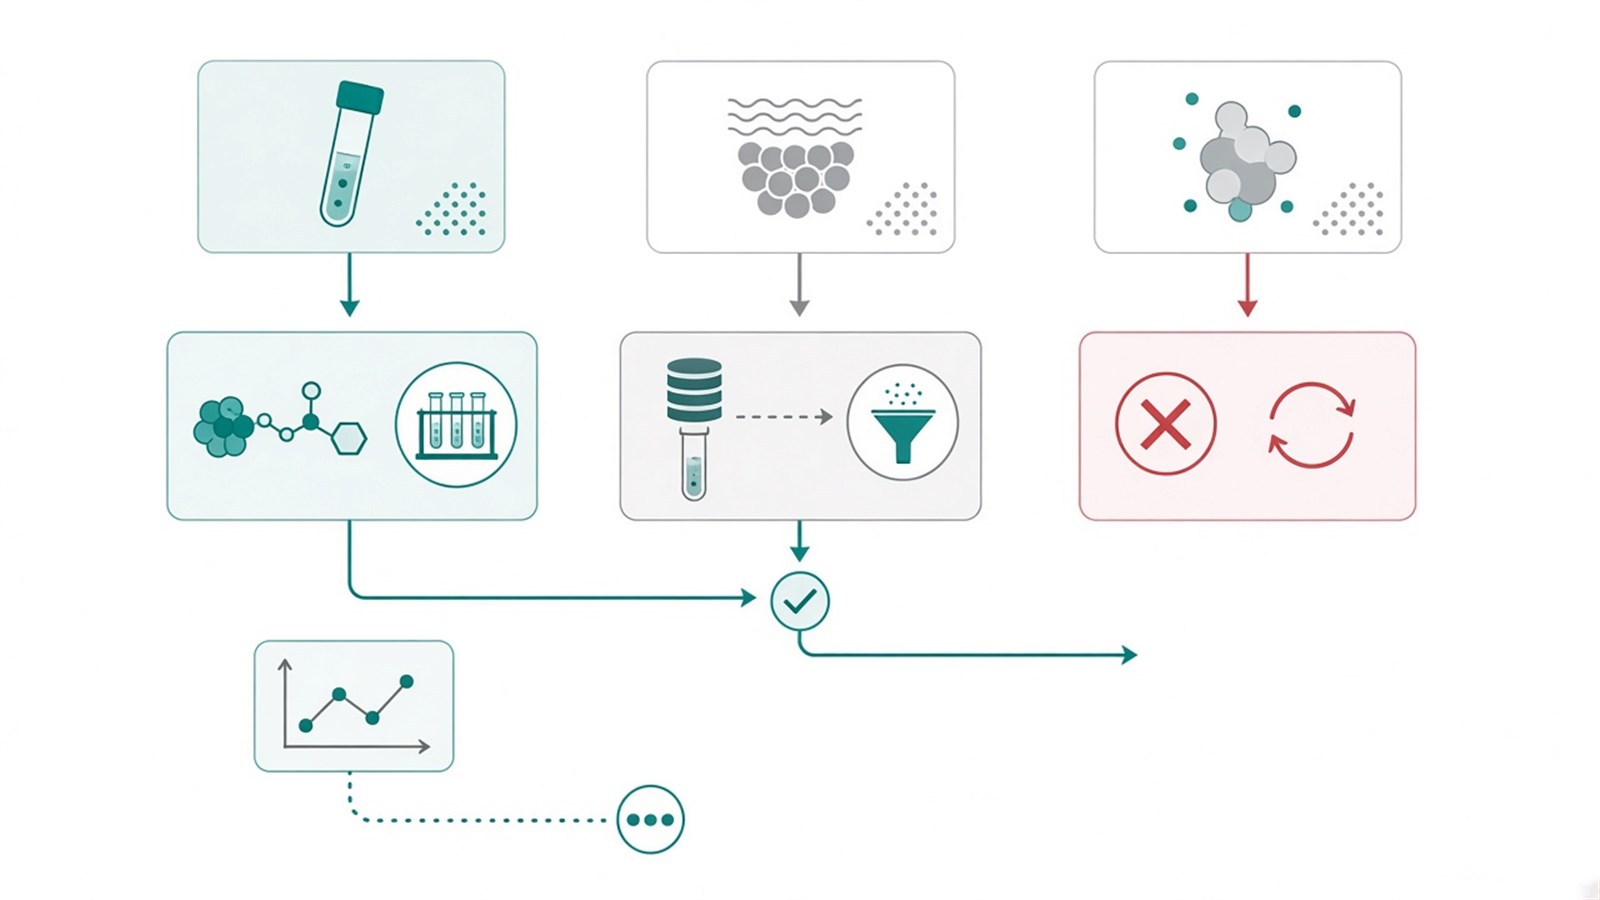
Figure 2. Lock extraction and cleanup before TMT/iTRAQ labeling; change route when detergent, amines, or load parity cannot be held.
Teams with matrix documentation, extraction SOPs, and matched-load plans can review MtoZ Biolabs TMT and iTRAQ service entries when those fields are declared in the project record.
Frequently Asked Questions
1. Why does protein extraction decide TMT/iTRAQ success before tags are applied?
Tags label peptides that survive the declared lysis, cleanup, and digest. Unmatched extraction changes the peptide pool and appears as reporter biology after mixing.
2. Can any lysis buffer be used if samples are later desalted?
Only if detergents, chaotropes, and primary amines are brought into the declared digest and labeling windows. Desalting alone does not fix every incompatible additive.
3. Should protein amount be matched on crude lysate or after cleanup?
Match after the cleanup step that defines the material entering digest and labeling. Crude lysate readings often miss selective losses.
4. Do TMT and iTRAQ need different extraction chemistries?
Core extraction and amine-cleanup logic are shared. Kit plex size and vendor protocols differ; pretreatment parity rules do not.
5. When should projects avoid forcing TMT/iTRAQ after extraction review?
When the cohort must stay unlabeled and expandable, or when extraction cannot deliver matched, labeling-compatible digests across channels.
MtoZ Biolabs can discuss protein extraction scoping for TMT/iTRAQ when matrix, lysis recipe, and channel design are documented.
If further evaluation of detergent removal, amine cleanup, or load parity pilots is needed, project details can be submitted for a feasibility review.
Reference
- A. Thompson, J. Schäfer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, R. Johnstone, A.K. Mohammed, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- P.L. Ross, Y.N. Huang, J.N. Marchese, B. Williamson, K. Parker, S. Hattan, N. Khainovski, S. Pillai, S. Dey, S. Daniels, S. Purkayastha, P. Juhasz, S. Martin, M. Bartlet-Jones, F. He, A. Jacobson, D.J. Pappin (2004). Multiplexed protein quantitation in Saccharomyces cerevisiae using amine-reactive isobaric tagging reagents. Mol. Cell. Proteomics, 3, 1154-1169. https://doi.org/10.1074/mcp.M400129-MCP200
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
