



TMT-based quantitative proteomics analysis starts after peptides from different samples have been tagged and mixed. The analysis principle is dual. The mixed precursor is identified from peptide fragments, and relative channel abundance is read from reporter-ion intensities in the same scan or in a later scan (Thompson et al. 2003). Those two operations can fail independently. A confident peptide-spectrum match with unusable reporters is not a protein ratio. Strong reporters without a unique peptide assignment are not a protein claim.
The chemical meaning of isobaric tagging is a separate question. Researchers can refer to What Is TMT Isobaric Labeling? for how tags keep one precursor mass until fragmentation. Labeling chemistry and acquisition mode remain separate decisions. In MtoZ Biolabs offerings, TMT analysis typically reads reporters from data-dependent acquisition (DDA) spectra at MS2 or MS3.
What TMT Quantitative Analysis Reports
TMT analysis reports a same-peptide relative ratio across channels that were mixed into one plex. After identification, reporter intensities are corrected, peptides are summarized to proteins, and a predefined group contrast can be tested. The quantitative object is the reporter ratio, not the MS1 chromatographic peak area of the intact precursor.
TMT analysis cannot recover a protein that was never identified. It cannot rank different proteins as copy numbers from reporter height. It cannot treat an unlabeled file as an extra channel. It does not report ng or fmol unless a matched stable isotope-labeled internal standard (SIS) was included for that peptide.
Pathway enrichment is optional after the ratio matrix passes quality checks. Researchers can refer to How to Analyze Quantitative Proteomics Data for generic protein-matrix statistics once TMT channel ratios already exist.
How Identification and Reporter Readout Are Coupled
A practical analysis order is stable even when the search environment changes. Spectra are searched against an appropriate protein database with peptide and protein false discovery rate (FDR) thresholds declared. For each accepted peptide, reporter intensities are extracted from the scan that was designated as the quantitative readout. Impurity correction is applied using kit-specific factors. Peptides are rolled up to protein groups. Only then should a group contrast be tested.
Identification uses peptide fragment ions. Quantification uses reporter ions. In MS2 TMT, both can sit in one spectrum. Co-isolated precursors can still mix reporters and pull ratios toward one (Savitski et al. 2013). An additional MS3 reporter scan, including synchronous precursor selection MS3 (SPS-MS3) on compatible instruments, can reduce that mixing. It does not change the analysis principle: the reported number remains a relative channel ratio. Researchers who need acquisition choices against compression can refer to TMT MS3 and SPS-MS3: How to Mitigate Ratio Compression and Enhance Quantitative Accuracy.
TMT analysis joins a peptide identification to a reporter-ion ratio; neither half is a quantitative result on its own.
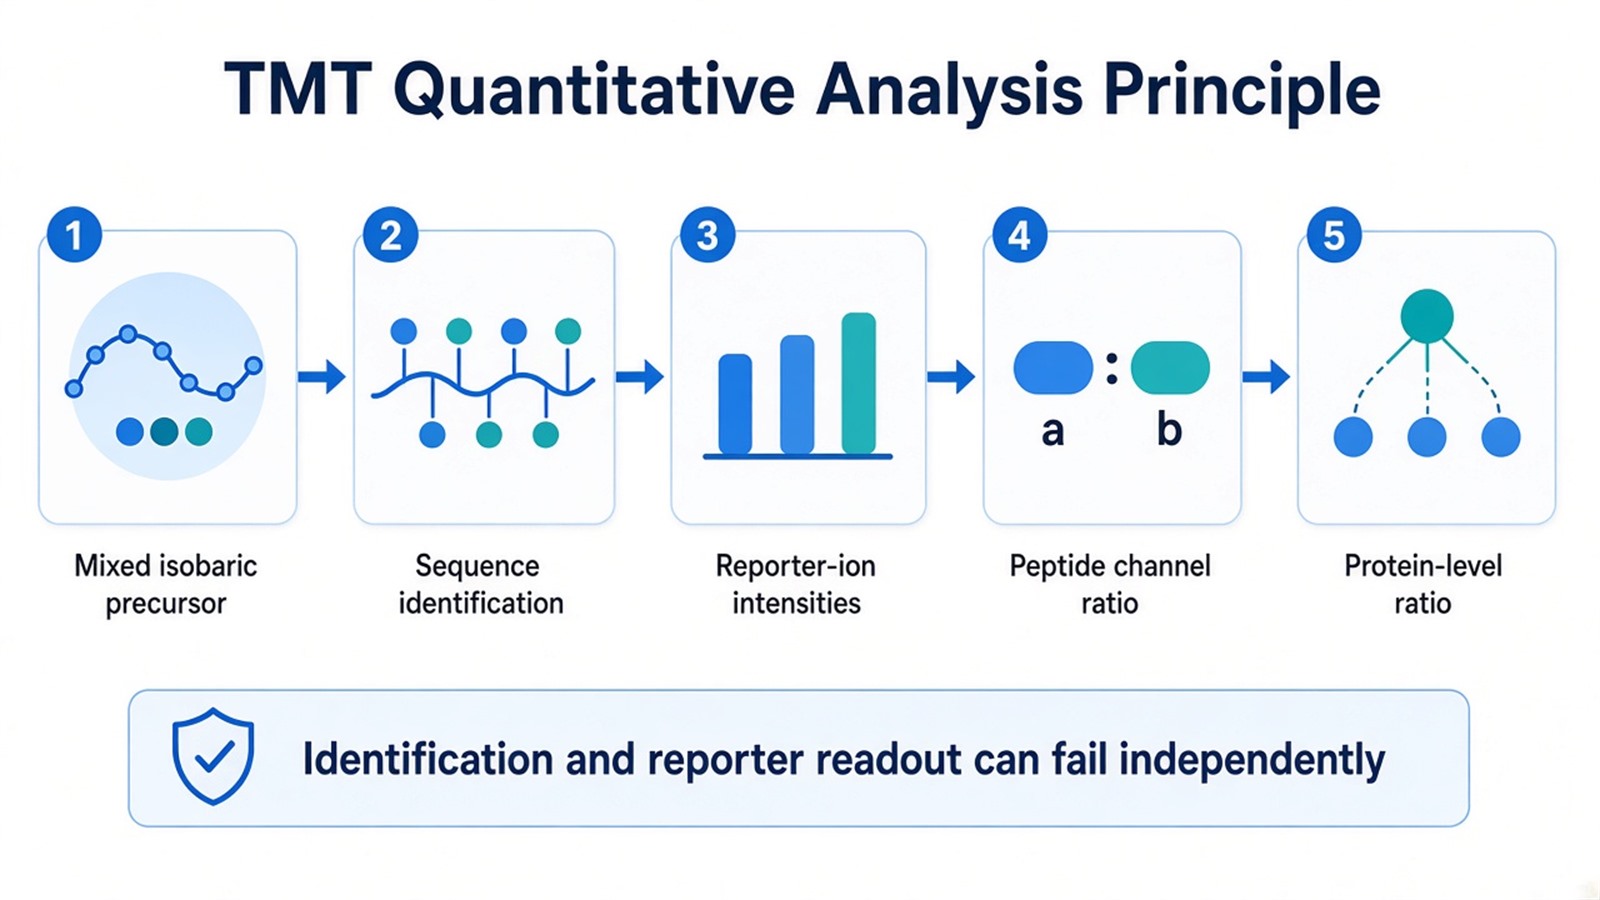
Figure 1. TMT quantitative analysis couples peptide identification with reporter-ion intensities before any protein-level ratio is reported.
Software that extracts reporters does not define the science. Search settings, the reporter scan type, impurity correction, and the protein-inference rule do.
From Channel Intensities to a Protein Ratio
Reporter heights are not yet a protein result. Shared peptides can pull homologs into the same group or split one gene product across several rows (Nesvizhskii and Aebersold 2005). A protein-level TMT value depends on which peptides are accepted as unique, how missing reporters are handled, and whether summation, signal-to-noise, or another summary is used (O'Connell et al. 2018). Those choices should be reported with the matrix.
Within-plex normalization adjusts channel scale after mixing. It assumes that a stated fraction of the proteome is stable across tags. It does not replace matched protein input before labeling. Across-plex comparison needs a bridge or reference channel. Two independent plexes without a shared reference are two experiments, not one ratio table.
Minimum reporter quality rules belong in the methods record. Channels with reporters at noise, or peptides that lack a reporter in most samples, are weak inputs to a protein ratio. Filling those cells by imputation can enable a test and can invent a difference that was never measured.
Parameters That Change the Analyzed Ratio
Several analysis settings change the number that will be written for a protein, even when the same raw files are used.
Database, enzyme specificity, and variable modifications change which spectra become identifications. Unique-peptide rules change which gene products carry the ratio. Impurity correction changes neighboring-channel leakage. The reporter scan type, MS2 or MS3, changes how much co-isolation enters the ratio (Savitski et al. 2013).
Normalization method and its assumption should be locked before the ranked list is inspected. So should the exact contrast, the rule for proteins with too few quantitative peptides, and the FDR thresholds. Quality checks on identification depth, reporter completeness, and sample clustering belong before biological language is added.
When channel offsets still follow load or digestion, the failure is usually upstream of search settings. Re-extracting reporters with a new software version does not repair unmatched input.
When This Analysis Principle Fits the Files You Have
The useful test is whether reporters exist, whether the channel map is known, and whether the claim is a relative ratio inside that plex.
Files and claim |
Check first |
Analysis that often fits |
|---|---|---|
Mixed TMT plex, open protein list |
Channel map and reporter scan type |
Reporter extraction, impurity correction, protein rollup |
Protein-level ratio required |
Unique-peptide and summary rule |
Declare inference and rollup before the contrast |
Fold changes look smaller than expected |
Isolation purity and MS2 mixing |
Review MS3 options; do not impute the compression away |
Two plexes, one contrast |
Shared bridge or reference |
Connect plexes only with a declared bridge |
Unlabeled DDA or DIA files |
Acquisition and labeling |
Not TMT reporter analysis |
Named peptides, confirmation required |
List length |
Targeted PRM or MRM in a new digest |
Amount or concentration unit |
SIS availability |
Ordinary TMT ratios are not sufficient |
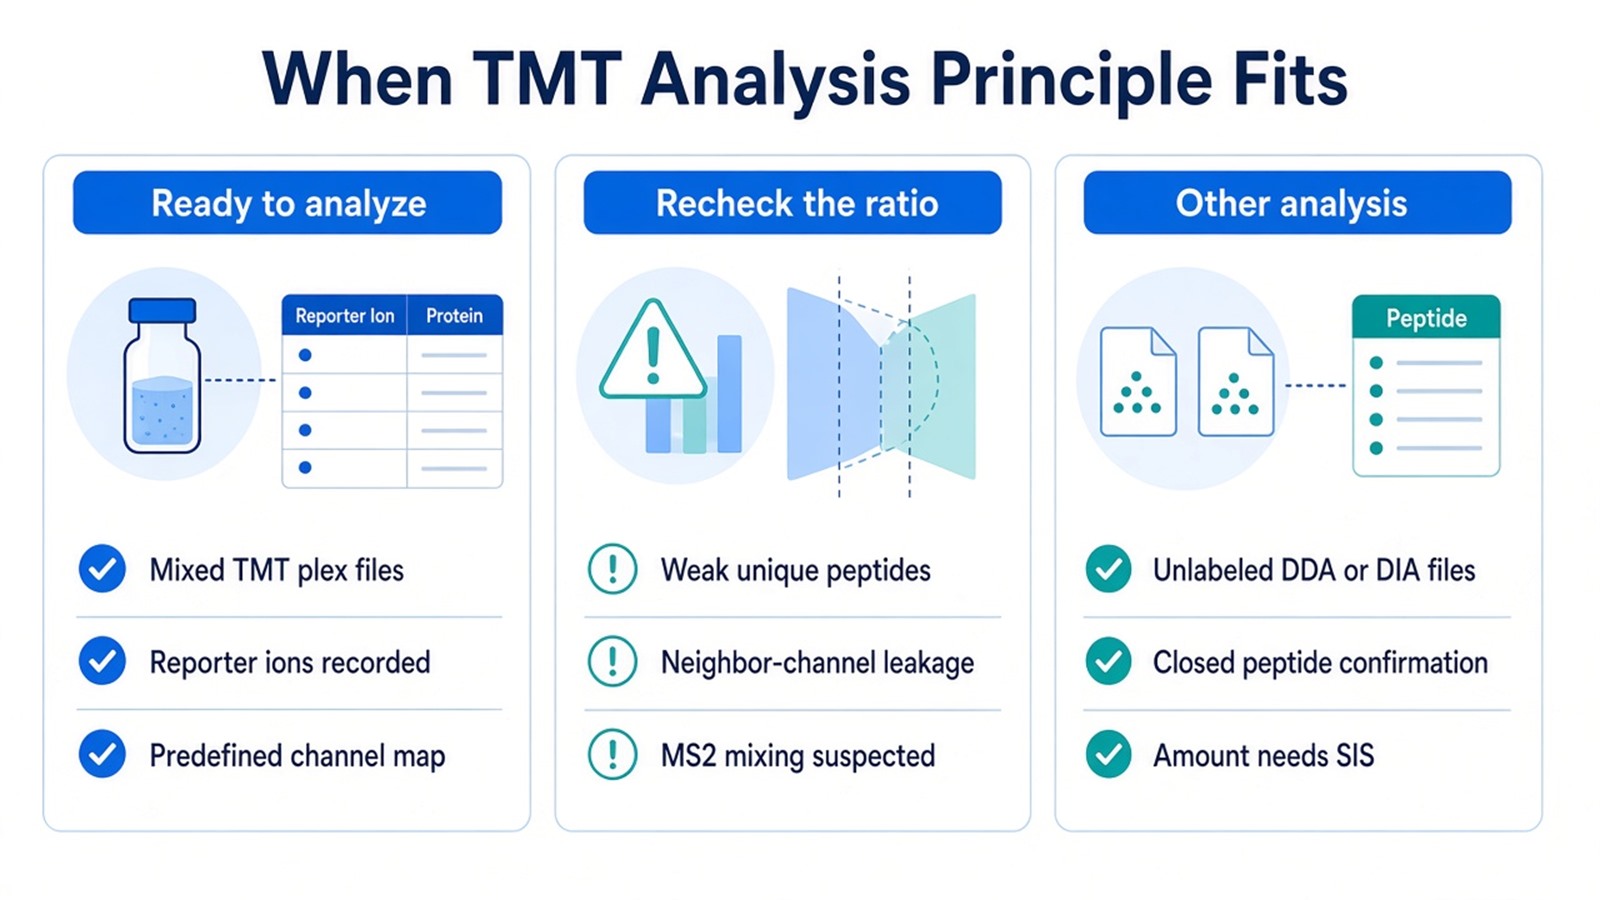
Figure 2. TMT quantitative analysis fits mixed plex files with recorded reporters; unlabeled files, closed confirmation lists, and amount claims need a different plan.
MS2 reporter analysis more often fits a discovery plex when speed and identification depth are the priority. MS3 reporter analysis more often fits when co-isolation is expected to hide a real fold change. Neither scan type is required in every study, and neither converts TMT into an absolute assay.
Researchers who already have mixed TMT files, a channel map, and a requested relative contrast can also review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific search and reporter-extraction planning.
Frequently Asked Questions
1. Is the MS1 peak area the TMT quantitative signal?
No. The mixed precursor is isobaric, so MS1 area does not separate channels. Relative abundance comes from reporter ions after fragmentation.
2. Does a search program define the TMT analysis principle?
No. A program can extract reporters and assemble protein groups. The scientific choices remain the reporter scan, impurity correction, unique-peptide rules, and the contrast.
3. How should TMT channels be normalized compared with separate unlabeled runs?
Within a plex, channels already share one chromatographic path, so normalization is a tag-scale adjustment under a stated stability assumption. Unlabeled runs need cross-run alignment as well. Do not copy an unlabeled pipeline onto reporter tables without checking that assumption.
4. Can two TMT plexes be analyzed as one experiment without a bridge sample?
Not as a single ratio matrix. A shared reference or bridge channel is the usual way to place plexes on one scale. Without it, each plex should keep its own contrast.
5. If unique peptides are scarce, should reporters still be rolled up to a gene name?
A shared peptide reports a protein group, not necessarily one gene product (Nesvizhskii and Aebersold 2005). Declare the inference rule. Do not present a homolog family as a single protein ratio without that note.
MtoZ Biolabs can provide TMT quantitative proteomics analysis for mixed plex projects, including reporter extraction, impurity correction, protein-level rollup, and predefined group contrasts when project files and study design are available.
If further evaluation of reporter scan type, unique-peptide rules, or whether unlabeled analysis is the better match is needed, project details can be submitted for a feasibility review.
Reference
- A.I. Nesvizhskii, R. Aebersold (2005). Interpretation of shotgun proteomic data: the protein inference problem. Mol. Cell. Proteomics, 4, 1419-1440. https://doi.org/10.1074/mcp.R500012-MCP200
- J.D. O'Connell, J.A. Paulo, J.J. O'Brien, S.P. Gygi (2018). Proteome-wide evaluation of two common protein quantification methods. J. Proteome Res., 17, 1934-1942. https://doi.org/10.1021/acs.jproteome.8b00016
- M.M. Savitski, G. Sweetman, M. Askenazi, J.A. Marto, M. Lang, N. Zinn (2013). Measuring and managing ratio distortion for accurate quantitative proteomics using isobaric tags. Mol. Cell. Proteomics, 12, 3409-3419. https://doi.org/10.1074/mcp.M113.030437
- A. Thompson, J. Schafer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, R. Johnstone, A.K. Mohammed, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
