



Targeted proteomics is an acquisition principle. The mass spectrometer is given an inclusion list of peptide precursors, isolates those ions on a chromatographic schedule, and quantifies them from fragment traces (Lange et al. 2008; Peterson et al. 2012). Discovery acquisition decides what to fragment from the live run: the most intense precursors in data-dependent acquisition (DDA), or every ion inside sequential m/z windows in data-independent acquisition (DIA). Targeting is not a labeling chemistry, not an antibody method, and not a requirement that the protein already be highly abundant.
In MtoZ Biolabs offerings, that principle is typically implemented as high-resolution parallel reaction monitoring (PRM). Multiple reaction monitoring (MRM) / selected reaction monitoring (SRM) remains a valid implementation when a scheduled triple-quadrupole transition list is already the production design.
What Targeted Means in Acquisition
Targeted means the precursor list is closed before the injection. Only those peptides are isolated for quantitative fragments. Proteins outside the list stay unmeasured, even if they dominate the digest. The quantitative object is an extracted fragment chromatogram of a proteotypic peptide, not a protein-ion full scan and not a dye-binding total-protein value.
Discovery and targeted principles answer different jobs (Domon and Aebersold 2010). Discovery is for an open protein inventory. Targeted is for remeasurement of named sequences. DIA schedules windows, not a protein name list. Windowed DIA is still discovery acquisition. It should not be described as targeted proteomics because isolation windows were used.
Relative same-peptide comparison is the default claim under this principle. An amount unit requires a matched stable isotope-labeled internal standard (SIS) and a declared calibration. Adding SIS does not change the acquisition principle. It changes the unit of the fragment ratio.
Researchers who need how a closed list is turned into a project plan can refer to Protein Targeted Quantification.
How a Scheduled Precursor Becomes a Quantitative Fragment Trace
The working order of the principle is short. A unique peptide is placed on the inclusion list with an expected retention-time window. During elution, the instrument isolates that precursor. Fragments are recorded either as predefined precursor-to-product transitions (SRM/MRM) or as a full high-resolution fragment spectrum (PRM). The peak area of selected fragments is the quantitative signal (Lange et al. 2008; Peterson et al. 2012).
MRM/SRM and PRM are two bookkeeping methods for the same isolation principle. MRM filters one product ion at a time on a triple quadrupole, which favors a locked transition list. PRM stores many fragments for each scheduled precursor, which favors fragment review after the run. Neither route is a different biology, and neither is required for every panel.
Targeted proteomics isolates listed precursors on purpose; quantification starts only when accepted fragment traces are integrated.
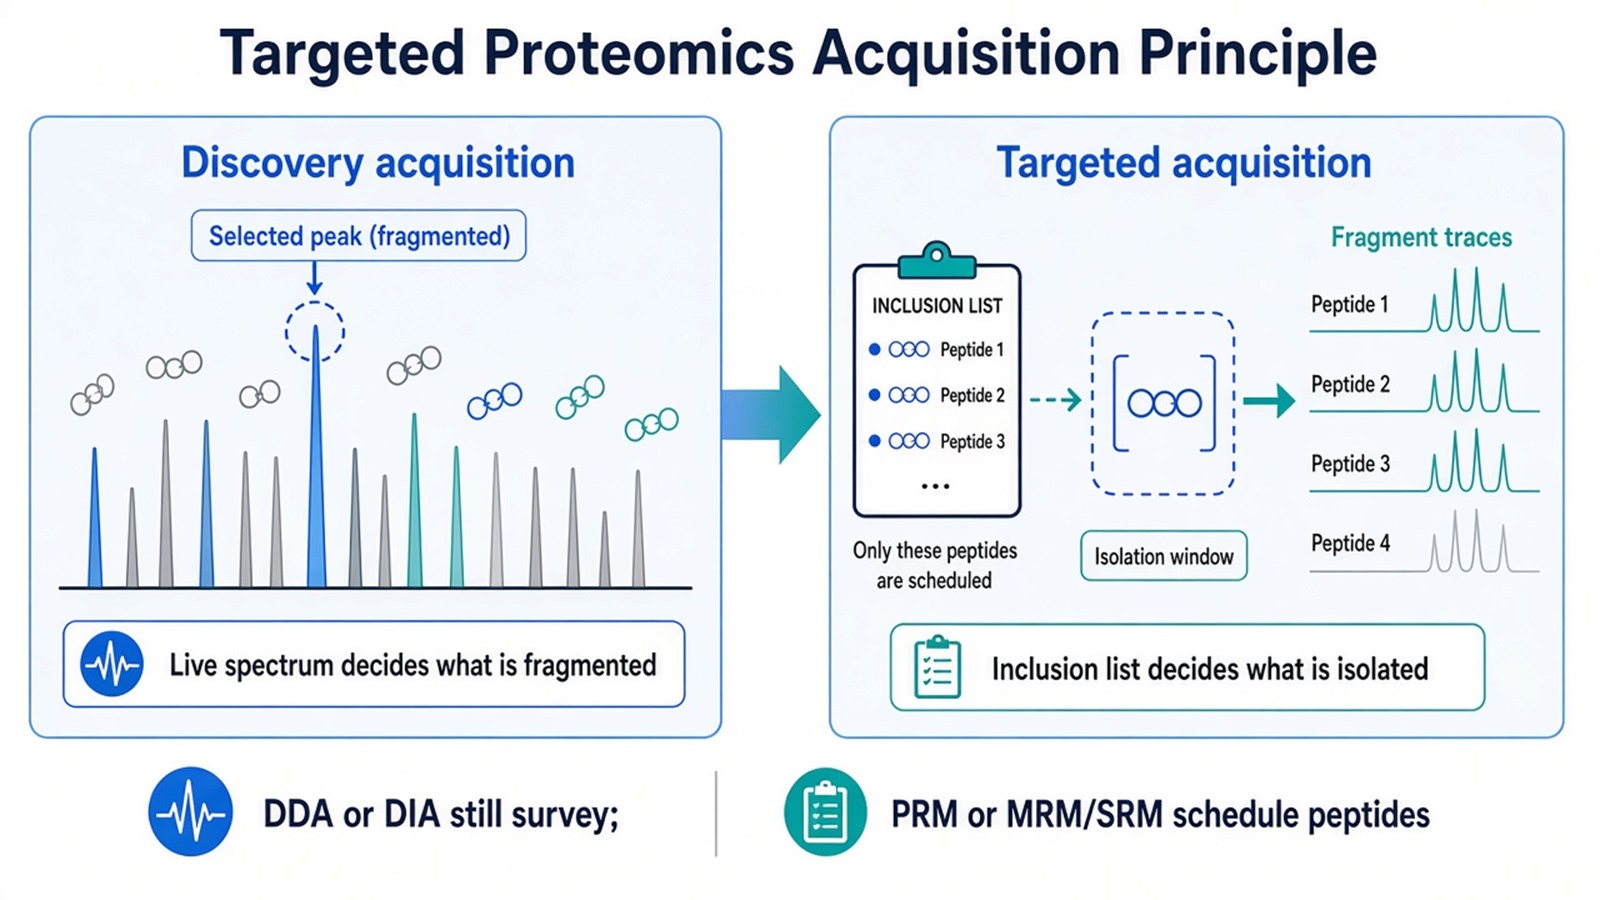
Figure 1. Targeted proteomics is defined by a closed inclusion list that decides which precursors are isolated, not by labeling chemistry.
Sample lysis, precipitation, and digestion are shared with other LC-MS/MS methods. They enable the peptide to exist. They are not the targeting principle. A peptide that never ionizes, or that was never scheduled, will not appear as a targeted result.
Parameters That Must Hold for the Principle to Apply
The inclusion list must contain sequences that map to the intended protein. Shared peptides report a group. Modified peptides report that modified sequence. "Biological relevance" does not replace uniqueness or detectability. High abundance is not a prerequisite. Scheduled isolation can help a listed low-abundance peptide if a clean peak exists. It cannot create a peptide that is absent from the digest.
Isolation width and cycle time decide whether the scheduled ion is what enters the fragment scan, and whether enough points fall across the chromatographic peak. Too many precursors in one method starve each peak. Split the method or shorten the list.
Fragment identity decides whether the trace belongs to the listed peptide. MRM needs qualifier transitions. PRM needs fragments that co-elute and match the spectrum. Interference inside the isolation window remains possible. The principle of targeting reduces stochastic sampling. It does not grant infinite selectivity.
If the remaining task is PRM schedule construction, researchers can refer to PRM LC-MS Targeted Proteomics. If the remaining task is whether a closed list is worth building, researchers can refer to Advantages and Disadvantages of Targeted Proteomics.
Limits Built into a Closed Inclusion List
The same closure that defines targeting also caps the experiment. New proteins cannot be discovered from a targeted file. A prior discovery matrix can propose names. Those names still need peptide-level evidence in the targeted digest.
Isolation is imperfect. Co-eluting ions can share a window or a transition. Qualifier rules and, when used, SIS co-elution are part of applying the principle, not optional extras. A protein name on a slide is not a quantitative result until an accepted peptide trace exists.
The principle does not set a universal protein-count capacity, a fixed sensitivity rank of PRM over MRM, or a concentration unit. Those are assay-design outcomes. Fit-for-purpose documentation still has to declare peptides, schedule, and acceptance criteria.
When This Principle Fits, and When Discovery Acquisition Fits
The useful test is whether peptide names can freeze, and whether the instrument should spend its cycle on that list.
Study need |
Acquisition principle that often fits |
What targeting will not do |
|---|---|---|
Open protein inventory |
DDA or DIA discovery |
Report unscheduled proteins |
Named peptides, remeasure |
PRM or MRM/SRM targeting |
Expand the inventory |
Windowed survey of many peptides |
DIA discovery |
Substitute for a protein inclusion list |
Fragments still under review |
PRM targeting |
Remove all isolation interference |
Locked transition panel |
MRM/SRM targeting |
Skip uniqueness and QC |
Amount for listed peptides |
Targeting plus matched SIS |
Convert light peak area into ng per mL by itself |
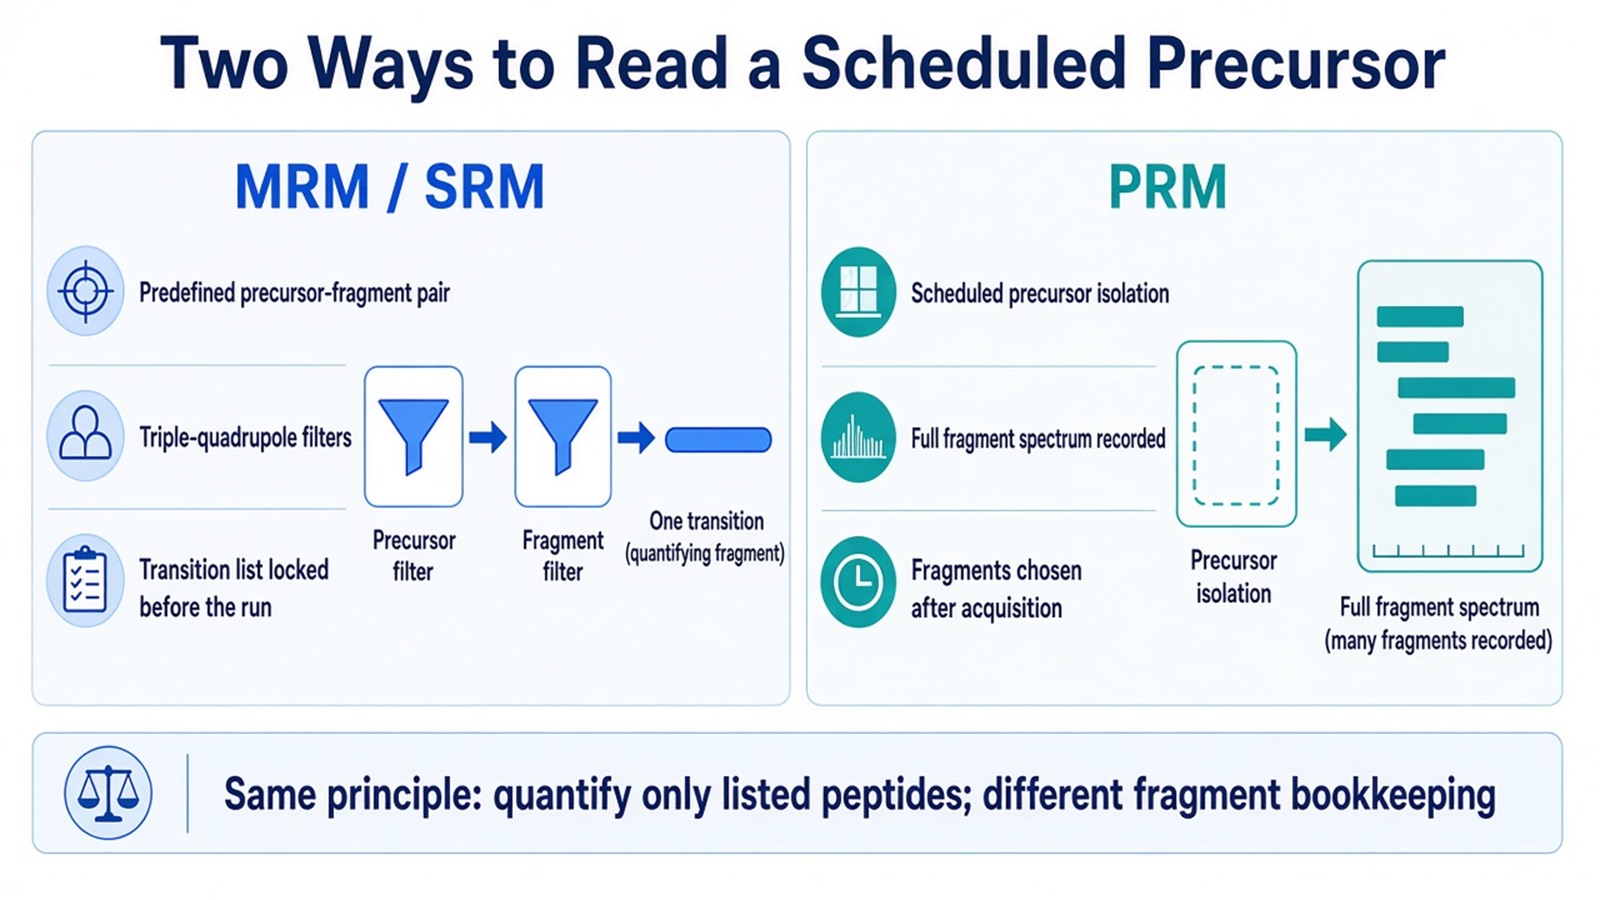
Figure 2. MRM/SRM and PRM implement the same targeting principle with different fragment bookkeeping.
Targeting more often fits after names freeze. Discovery more often fits while new names are still required. An exception is a pre-specified pathway panel that never needed a discovery experiment, or a tiny closed set that would waste a full proteome survey.
Researchers who already have named peptides and a requested relative or SIS-supported claim can also review the MtoZ Biolabs Targeted Proteomics Service for project-specific feasibility of a scheduled assay.
Frequently Asked Questions
1. Is targeted proteomics a labeling method?
No. Targeting is how precursors are selected for fragmentation. Unlabeled or labeled digests can both be acquired with a targeted method. TMT reporter chemistry is a different axis.
2. Must the target protein already be highly abundant?
No. Scheduled isolation is often used because a listed peptide is hard to sample stochastically. The peptide still has to exist, ionize, and pass interference checks in that matrix.
3. Is DIA a targeted method because windows are scheduled?
No. DIA schedules m/z windows to survey many peptides. Targeted proteomics schedules named peptide precursors. Windowed discovery and peptide inclusion lists are different principles.
4. Does isolating a precursor prove protein amount?
No. Isolation creates a fragment trace for that peptide. Protein inference still depends on uniqueness. Amount still depends on a matched SIS and calibration.
5. Do MRM and PRM use different targeting principles?
They share the closed-list isolation principle. They differ in whether one product ion is filtered live (MRM/SRM) or a full fragment spectrum is recorded (PRM).
MtoZ Biolabs can provide targeted proteomics based on scheduled peptide isolation by PRM or MRM/SRM when the protein or peptide list is closed and the intended claim is a relative ratio or an SIS-supported amount.
If further evaluation of inclusion-list design, PRM versus MRM implementation, or whether discovery acquisition is the better first step is needed, project details can be submitted for a feasibility review.
Reference
- B. Domon, R. Aebersold (2010). Options and considerations when selecting a quantitative proteomics strategy. Nat. Biotechnol., 28, 710-721. https://doi.org/10.1038/nbt.1667
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O111.013698
