



The mechanism of MS-based protein-protein interaction analysis is a conversion chain. A written capture or covalent mark concentrates a bait and whatever travels with it. Enzymatic digestion, usually with trypsin, turns that eluate into peptides. Liquid chromatography–tandem mass spectrometry (LC-MS/MS) records mass-to-charge ratios (m/z) and fragment spectra. A search returns accessions. A matched control ranks which names are bait-enriched. The chain answers which proteins were recovered under those conditions. It does not accurately identify or quantify protein–protein interactions, and it does not uncover cellular mechanisms, drug targets, or disease pathogenesis.
High sensitivity, throughput, and specificity are instrument properties. They are not interaction-specificity claims. Signal transduction and metabolic pathways remain contexts in which a bait may be tested. They are not outputs of the spectrometer. For the three physical bases that can feed this chain, How to Identify Protein-Protein Interactions Using Mass Spectrometry provides a complementary method-choice page.
Researchers who already have a bait or an immunoprecipitate and need a ranked recovery table can also review the MtoZ Biolabs Protein-Protein Interaction Analysis Service for project-specific feasibility and analysis planning.
Sample Preparation Is a Filter, Not Prey Isolation
The original first step remains if it is read as enrichment, not as isolation of interacting proteins. Immunoprecipitation or affinity purification captures a bait. On this site, antibody capture is usually planned as IP-MS. A tagged bait is usually planned as AP-MS. Prey, in the original wording, is whatever survived the wash. It is not a confirmed partner set. Crosslinking and proximity labeling use a different first event, a covalent mark, and still enter the same digest.
Trypsin produces peptides. Those peptides are the reporter. The spectrometer does not later measure intact protein mass, and it does not see a complex as an object. For the handle–wash–elute chain inside the tagged format, How AP-MS Works provides a complementary AP-MS mechanism page.
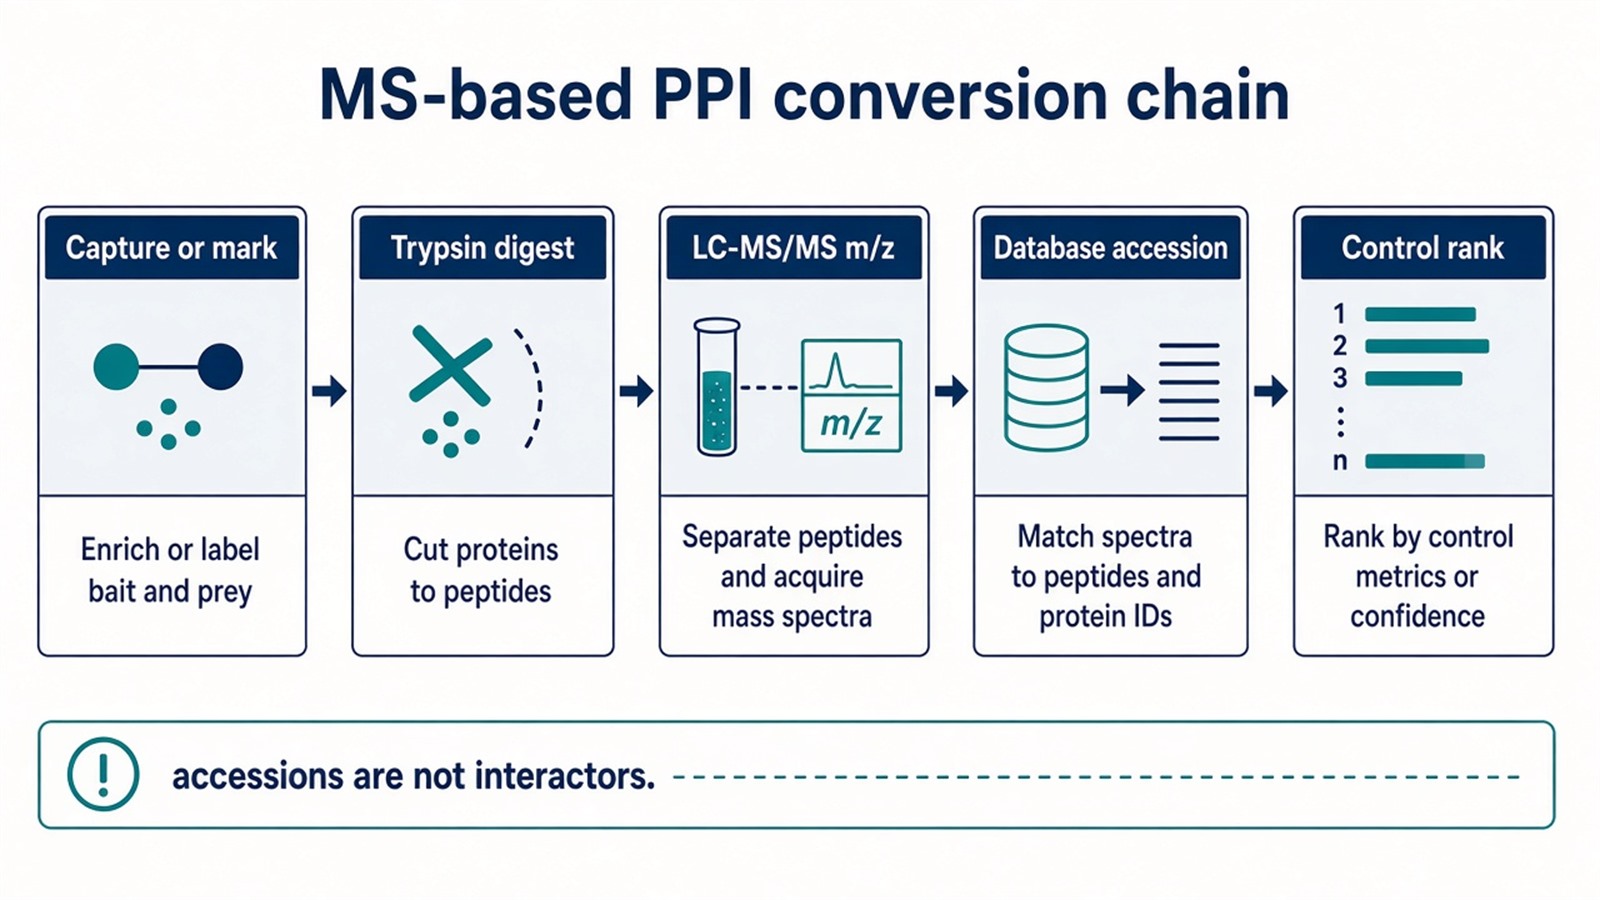
Figure 1. A handle or a covalent pulse writes the input. Trypsin writes peptides. LC-MS/MS writes m/z and fragments. A control ranks accessions. Accessions are not interactors.
Name the first event, the digest, and the control before the first injection. Isolation of prey is not a step in this chain.
What the Spectrometer Is Doing
Peptides are ionized and separated. The instrument records precursor m/z and fragment spectra. Those spectra can be matched to peptide sequences in a database. Parent m/z is not an identification, and a match is not a deduced sequence in the chemical sense of de novo proof. Mascot, Sequest, or another engine returns peptide-spectrum matches and protein accessions. Those accessions are not prey proteins that interact with the bait.
Labeled and label-free designs sit on top of that search. They do not change the physical basis of the capture. Stable isotope labeling by amino acids in cell culture (SILAC) places light and heavy amino acids into two cultures so that the same peptide appears at two masses. The heavy-to-light ratio ranks a written contrast. It is not relative quantification of protein interactions, and it does not report a binding constant. For the mix-before versus mix-after decision, Principle of SILAC-Based Co-IP-MS in Protein Interaction Analysis provides a complementary SILAC page.
Label-free comparison of peptide intensities across samples is also a contrast. It does not infer interaction dynamics. A change in intensity can be bait occupancy, wash-off, digestion, or injection depth. Dynamics is a later overlay.
Data Analysis Does Not Validate or Draw a Network
The original analysis list overreaches at the last two items. Matching spectra, identifying peptides, and inferring proteins are search steps. Constructing interaction networks is not. Statistical filters and bioinformatics tools can rank enrichment. They do not validate interactions.
Yeast two-hybrid, surface plasmon resonance, and bimolecular fluorescence complementation remain allowable orthogonal assays for a shortlisted name. They do not provide additional confirmation and quantification of the PPIs discovered by MS as a class. Each orthogonal method has its own physical basis and its own false-positive profile. A Y2H hit on a name from an IP-MS table is two evidence types, not a validation stamp.
False positives and missed low-abundance names remain real limits. They are why the control arm is part of the mechanism, not a later quality note. An empty-tag, isotype, or ligase-only file has to travel the same digest and the same search. Refining sample preparation and data analysis will not, as a future program, enhance precision and reliability of interaction claims. A longer identification list is still a longer list.
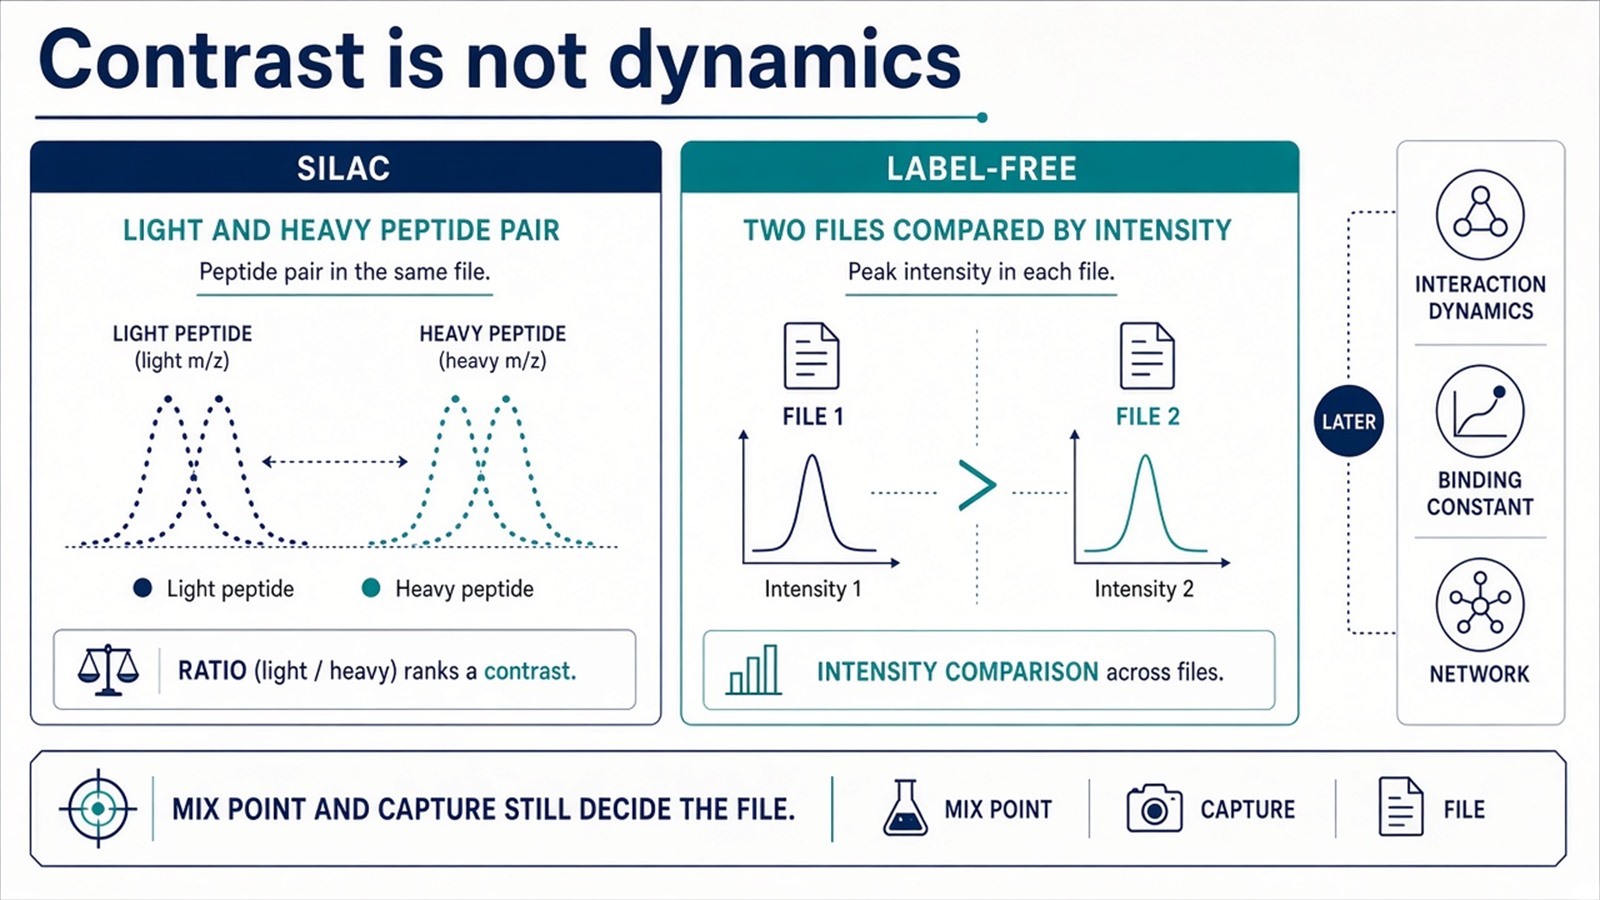
Figure 2. Mascot or Sequest matches peptides. A labeled or label-free ratio ranks a contrast. Network layout and orthogonal assays are later overlays, not validation inside the search.
Experimental Limits
This mechanism reduces a project to one first event, one digest, and one ranked file. Accurate PPI quantification, dynamic cellular processes, disease mechanisms, and drug discovery sit outside ordinary research reporting unless a separately validated assay is already named. A powerful means to study complex biological systems is a slogan, not a step.
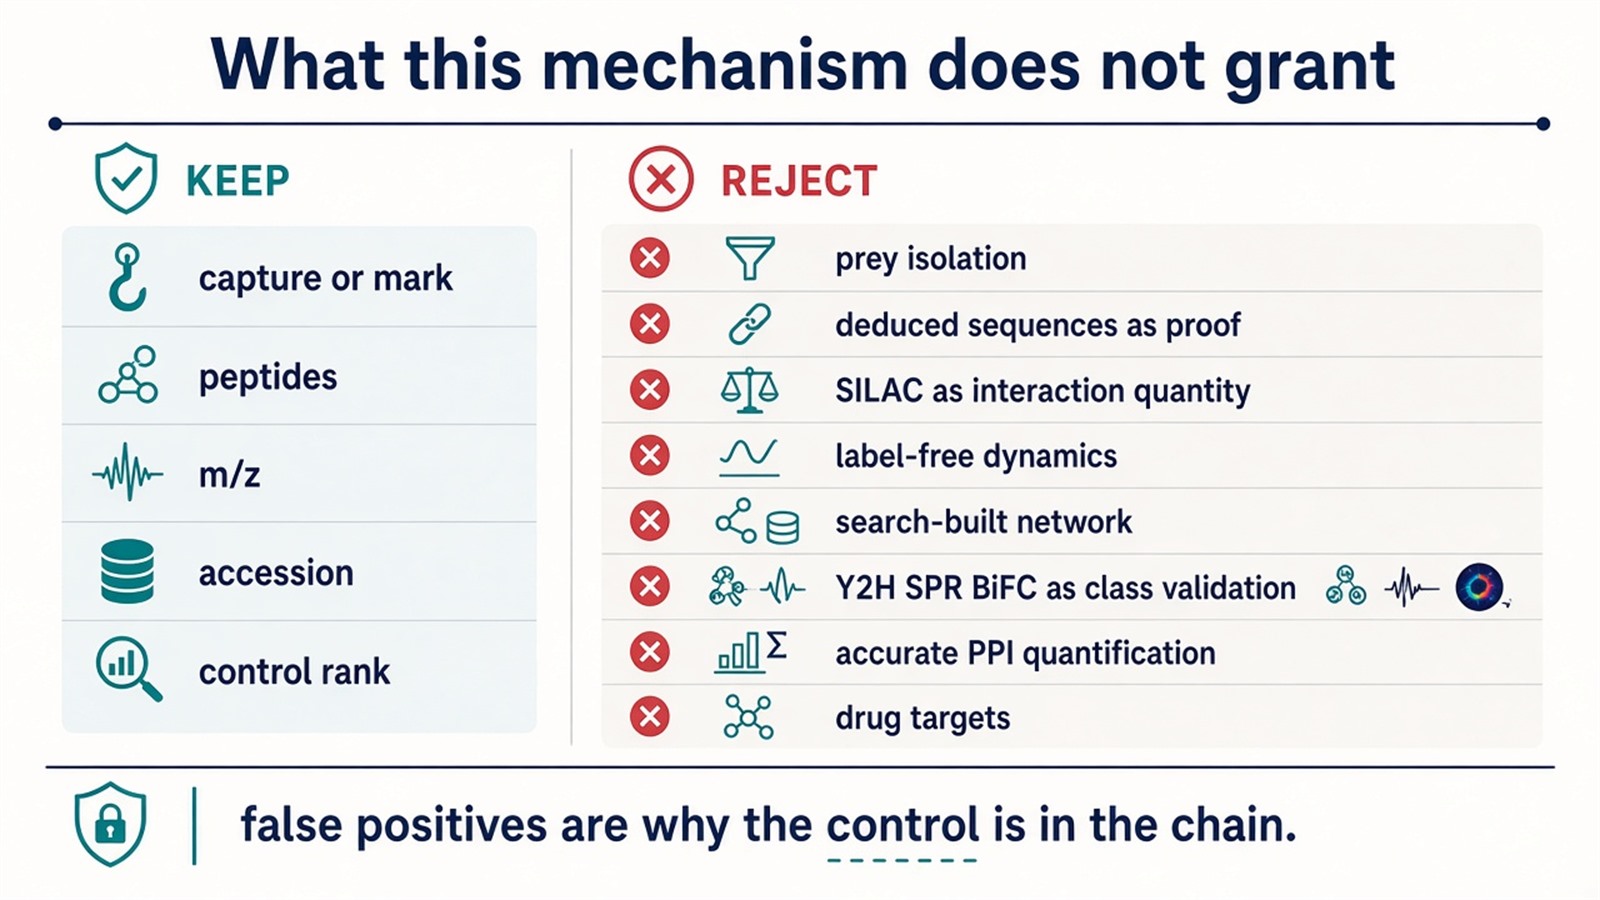
Figure 3. Keep: capture or mark, peptides, m/z, accession, control rank. Not granted: prey isolation, deduced binding, interaction dynamics, search-built networks, or orthogonal assays as automatic confirmation.
Project Planning and Service Scope
A usable request names the first event, whether a tag or an antibody will be used, the labeled or label-free contrast, and the orthogonal assay that would be required for a binding sentence. If the first event is a cross-link pulse or a ligase fusion, this IP-or-AP mechanism page should not force the sample through affinity capture. MtoZ Biolabs scopes IP-MS and AP-MS on Orbitrap-class LC-MS/MS. Interpretability still follows those arms, not a high-specificity heading.
Once the next sentence is a tagged-bait co-enrichment table, this mechanism page should hand off. Researchers who need that table can review the MtoZ Biolabs Affinity Purification-Mass Spectrometry Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. Does MS-based PPI analysis isolate interacting prey with the bait?
It enriches a bait and whatever survived the wash or the covalent mark. Prey is a later claim.
2. Do spectra deduce peptide sequences and identify interacting proteins?
Spectra are matched to a database. Accessions are not interactors.
3. Does SILAC quantify protein interactions?
It reports a heavy-to-light ratio under a written contrast. That is not a binding quantity.
4. Does label-free MS infer interaction dynamics?
It compares peptide abundance across files. Dynamics is a later overlay.
5. Do Y2H, SPR, or BiFC validate MS-discovered PPIs?
They can test a shortlisted name. They do not confirm the MS table as a class.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Dunham W.H., Mullin M., Gingras A.C. (2012). Affinity-purification coupled to mass spectrometry: basic principles and strategies. Proteomics, 12, 1576-1590. https://doi.org/10.1002/pmic.201100523
- Ong S.E., Blagoev B., Kratchmarova I., et al. (2002). Stable isotope labeling by amino acids in cell culture, SILAC, as a simple and accurate approach to expression proteomics. Mol. Cell. Proteomics, 1, 376-386. https://doi.org/10.1074/mcp.M200025-MCP200
- Mellacheruvu D., Wright Z., Couzens A.L., et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
- Choi H., Larsen B., Lin Z.Y., et al. (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
