



Is DDA considered as untargeted quantitative proteomics? Usually yes when acquisition is stochastic top-N shotgun without a closed analyte list—but DDA names an acquisition mode, not a complete project class. Data-dependent acquisition (DDA) on liquid chromatography-tandem mass spectrometry (LC-MS/MS) selects intense precursors for fragmentation as they appear; label-free or labeled discovery workflows often use that mode for open proteome profiling (Michalski et al. 2011). Targeted proteomics instead freezes a closed peptide or protein panel and schedules selected reaction monitoring (SRM), multiple reaction monitoring (MRM), or parallel reaction monitoring (PRM) measurements (Lange et al. 2008; Peterson et al. 2012). In project language, “untargeted” means the analyte list is not closed before acquisition; “targeted” means it is. The guide below answers the DDA question inside a comprehensive targeted-versus-discovery frame with workflow, locked parameters, limits, and fit. It is not a claim that DDA equals all label-free work, not a claim that DDA never uses inclusion lists, and not a sensitivity brochure.
Directed DDA with inclusion lists sits between open discovery and fully scheduled MRM/PRM. Data-independent acquisition (DIA) is another discovery-scale route that fragments windows rather than top-N precursors (Gillet et al. 2012). Unmatched prep still breaks any of these quantitative claims (Bantscheff et al. 2012).
In MtoZ Biolabs quantitative offerings, targeted panels and discovery DDA/DIA routes are scoped after list status, matrix, claim unit, and quality control (QC) design are written in the brief. Targeted intake appears in Targeted Proteomics Service.
What Targeted Proteomics Must Cover (Versus Untargeted DDA)
Four layers belong in the brief and keep targeted work separated from open DDA.
List-status layer freezes named proteins or proteotypic peptides. If the list is open, the project is discovery—even if the instrument is set to DDA.
Claim layer names relative fold change versus amount with stable isotope-labeled internal standard (SIS) peptides. Discovery DDA and targeted MRM/PRM can both be relative; only the declared unit and calibrants decide amount claims.
Acquisition layer names DDA top-N, directed inclusion-list DDA, DIA windows, or scheduled MRM/PRM transitions. Mode choice follows list status; it does not redefine it.
Prep-and-QC layer names matched matrix handling, load, blanks, and QC pools. Stochastic missingness in DDA is an acquisition limit, not a license to skip prep parity.
DDA is typically untargeted quantitative proteomics when precursors are chosen by intensity without a closed panel. Targeted proteomics begins when that panel is frozen and scheduled.
How DDA, DIA, and MRM/PRM Split the Same Question
Acquisition mode and targeting status must be read together.
Classic DDA discovery ranks precursors by intensity each cycle. Low-abundance peptides may be sampled inconsistently across runs, which weakens reproducibility for verification-style claims. That pattern is why many teams call shotgun DDA untargeted or discovery quantification when no inclusion list is used.
DIA fragments predefined m/z windows and supports discovery-scale extraction without top-N competition in the same way. It is still not MRM/PRM unless a closed panel and extraction rules are declared afterward.
MRM/PRM targeted proteomics schedules known peptides regardless of whether they would have won a DDA top-N contest. Route contrast appears in Comparison of Mass Spectrometric MRM with Other Quantitative Methods in Proteomics. PRM versus MRM fit appears in Parallel Reaction Monitoring vs MRM: Which Fits?. Label-free discovery intake appears in Label-Free Quantitative Proteomics Service.
End-to-End Workflow: Decide Targeting Status Before Locking DDA
A practical targeted-versus-DDA spine usually runs as follows.
- Freeze the scientific contrast and whether analytes are already named.
- If the list is open, scope discovery DDA or DIA with matched prep and QC.
- If the list is closed, scope MRM or PRM (or directed DDA only as a bridge) with matrix-matched pilots.
- Declare relative versus SIS amount claims before acquisition method lock.
- Match harvest, digest, and load across arms for whichever route is chosen.
- Build DDA top-N settings or scheduled transition/inclusion methods from pilots—not from slogans.
- Acquire interleaved study and QC injections; stop if missingness or transition drift exceeds rules.
- Quantify only under the declared claim unit; do not treat discovery IDs as a targeted panel after the fact.
- Hand discovery hits into targeted verification only when a new closed list is written.
Steps 1 through 4 set targeting status. Steps 5 through 9 produce the quantitative record for that status.
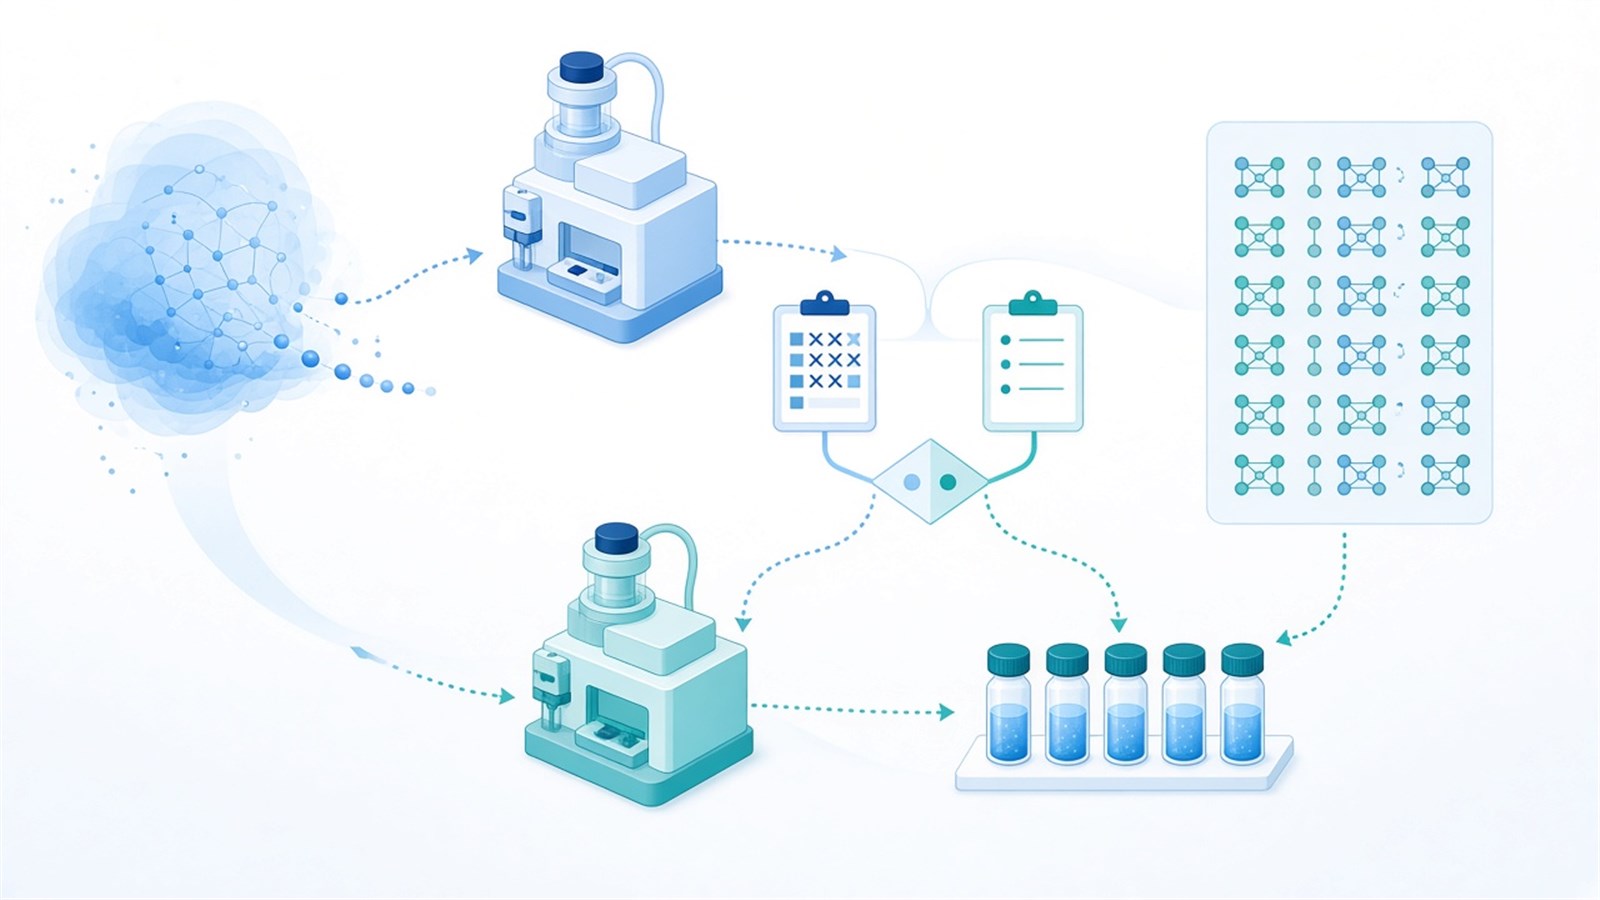
Figure 1. Call DDA untargeted when the analyte list is open; move to MRM/PRM when the panel is closed and scheduled.
Parameters That Lock Targeted Versus Untargeted Claims
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
List status |
Open discovery vs closed panel |
DDA sold as targeted verify |
Acquisition |
DDA top-N, directed DDA, DIA, MRM/PRM |
Mode confused with claim class |
Claim unit |
Relative vs SIS amount |
Peak height as concentration |
Prep parity |
Matched digest and load |
Prep drift as biology |
DDA settings |
Top-N, exclusion, cycle time |
Irreproducible sampling |
Targeted schedule |
Transitions or precursors + windows |
Undersampled peaks |
Missingness |
Rules for discovery matrices |
Stochastic gaps as regulation |
QC |
Pools, blanks, brackets |
Undetected batch drift |
Hand-off |
Discovery-to-targeted list version |
Moving targets mid study |
Deliverable |
Method and list files |
Report rebuilt after acquisition |
Limits follow the same table. DDA is not automatically the best “easy” quantitative method for every matrix. Low-abundance sensitivity and run-to-run consistency are known discovery constraints. Label-free status does not equal untargeted status if an inclusion list is closed.
How to Choose After Answering the DDA Question
Use list status, reproducibility need, and claim unit together.
Project need |
Route that often fits |
Recheck before committing |
|---|---|---|
Open proteome discovery, unlabeled |
DDA or DIA LFQ |
Call it targeted without a panel |
Closed peptide panel, scheduled quantify |
MRM or PRM |
Keep stochastic DDA as verify |
Named list but still surveying fragments |
Directed DDA or PRM bridge |
Assume top-N DDA will hit every target |
Amount on named peptides |
Targeted SIS design |
Discovery intensities as concentration |
Expandable unlabeled cohort |
Discovery LFQ/DIA |
Force a closed MRM list too early |
Discovery hits ready to verify |
Hand off to MRM/PRM |
Re-run open DDA as confirmation |
Prep recipe still changing |
Fix prep first |
Optimize only acquisition mode |
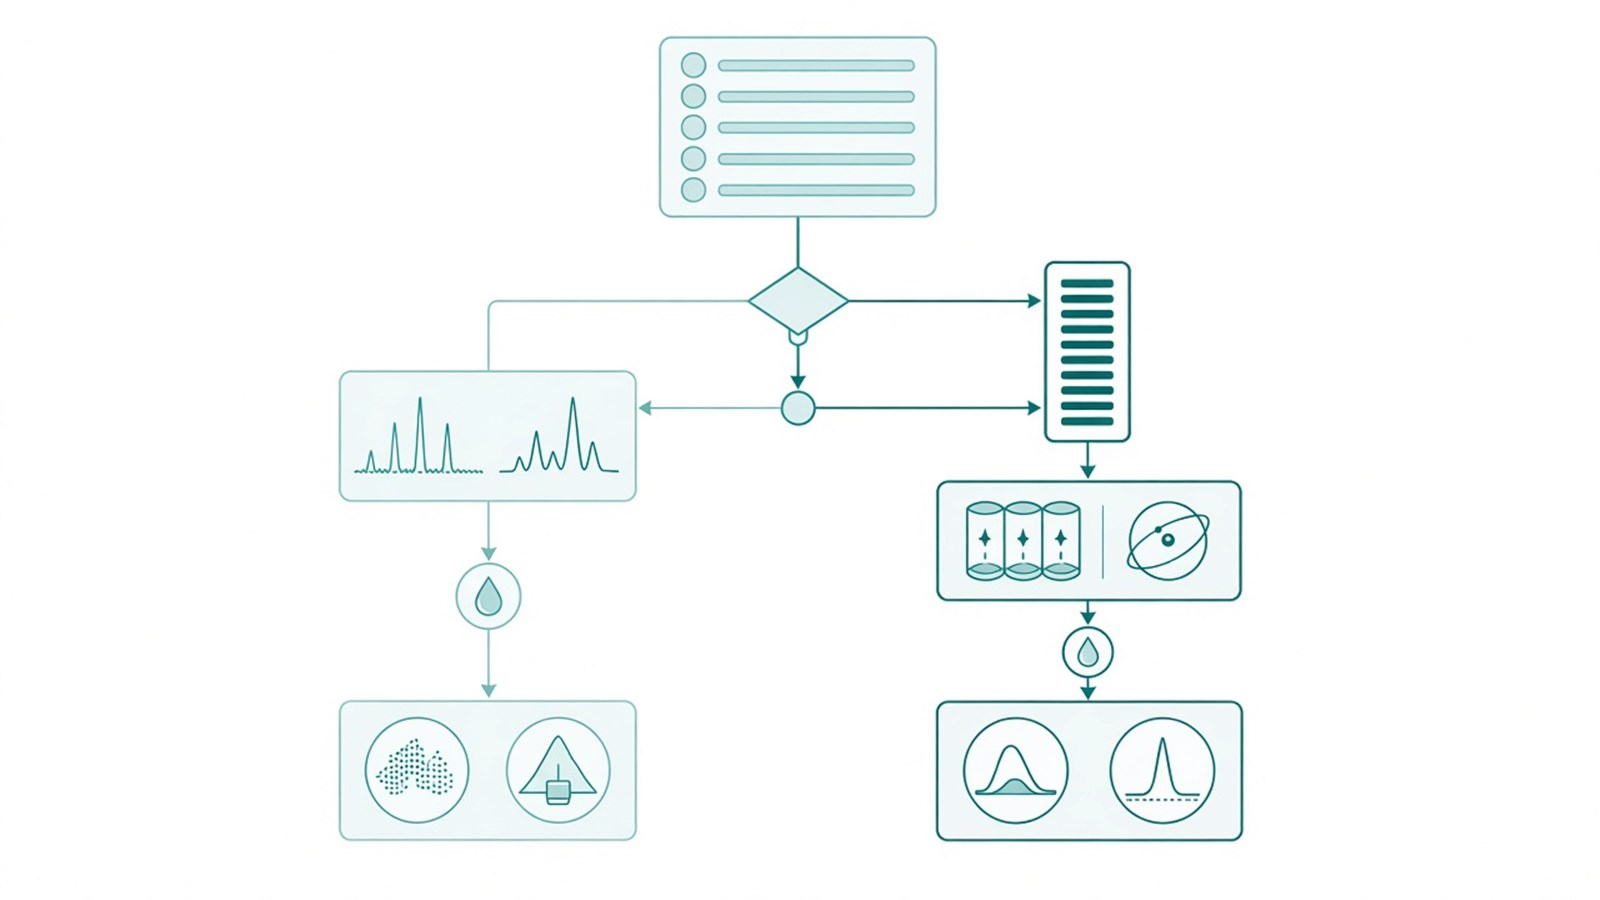
Figure 2. Keep DDA in the untargeted/discovery lane when the list is open; choose targeted proteomics when the panel and schedule are locked.
Teams with list status, matrix documentation, and QC-inclusive designs can review MtoZ Biolabs targeted or label-free service entries when those fields are declared in the project record. PRM service intake appears in Parallel Reaction Monitoring (PRM) Service.
Frequently Asked Questions
1. Is DDA considered untargeted quantitative proteomics?
Yes in the common shotgun case: intensity-driven precursor selection without a closed analyte panel. The label “untargeted” follows list status, not the DDA acronym alone.
2. Can DDA ever be used in a targeted-style project?
Directed DDA with inclusion lists can prioritize named precursors. Fully scheduled MRM/PRM remains the clearer targeted class when verification reproducibility is the claim.
3. Does label-free mean the same thing as untargeted?
No. Label-free describes chemistry. Untargeted describes whether analytes were pre-specified. Labeled discovery (for example TMT) can also be untargeted at the protein-list level.
4. Why is DDA often weaker than MRM/PRM for verification?
Top-N sampling is competitive and can miss the same peptide across runs. Targeted methods schedule that peptide regardless of ranking among co-eluting ions.
5. Should every DDA discovery study continue into targeted proteomics?
Only when a closed peptide list and verification claim are written after discovery. Open DDA results are not a targeted assay by relabeling.
MtoZ Biolabs can discuss whether a project is untargeted DDA or targeted proteomics when list status, matrix, and claim unit are documented.
If further evaluation of DDA versus DIA discovery or MRM/PRM verification is needed, project details can be submitted for a feasibility review.
Reference
- A. Michalski, J. Cox, M. Mann (2011). More than 100,000 detectable peptide species elute in single shotgun proteomics runs but the majority is inaccessible to data-dependent LC-MS/MS. J. Proteome Res., 10, 1785-1793. https://doi.org/10.1021/pr101060v
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, A.A. Klammer, P.F. Doubleday, H. Liu, J.N. Anderson, M.J. MacCoss (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020271
- L.C. Gillet, P. Navarro, S. Tate, H. Röst, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
