



Protein-peptide interactions should be tested as a defined ligand-protein pair, not as a synonym for IP-MS of a native complex. A short peptide bait is a motif, epitope, or designed ligand. An antibody IP recovers a protein and whatever the wash still allows. Tryptic peptides in an antibody-MS table are digestion products used to identify proteins. They are not the peptide ligands under study. Treating peptide capture and whole-complex immunoprecipitation as one method mixes two questions. IP-MS can test whether a motif-dependent protein partner is co-enriched, or whether a competing peptide weakens that recovery. It cannot, by itself, certify that a peptide binds a named protein. Immobilized-peptide pull-down, competition, and a pair-level affinity assay are the usual validation path.
Researchers who already have a defined peptide, a control peptide, and a written binding claim can review the MtoZ Biolabs Peptide-Peptide/Protein Interaction Analysis Service for feasibility. Broader pair-method choice is covered in Measuring Protein-Protein Interactions.
Protein-Peptide Binding Is Not Whole-Complex Immunoprecipitation
A protein-peptide interaction asks whether a short sequence contacts a protein domain or pocket. The peptide may be a linear motif, a phosphorylated epitope, a designed inhibitor, or a viral fragment. The protein may be purified or present in a lysate. The claim is pair-level retention or affinity under a stated buffer.
Whole-complex antibody capture plus MS asks a different question. An antibody recovers an endogenous or tagged protein. LC-MS/MS then identifies proteins that survived capture and washing. The digest is cut so that proteins can be named. Those tryptic fragments are identification reagents. They are not evidence that a peptide ligand bound the bait (Hein et al. 2015).
Do not equate the two workflows. An IP that lists many peptides has identified proteins. It has not mapped protein-peptide interactions. A biotinylated peptide on streptavidin beads is a pull-down bait. It is not an immunoprecipitation. Mixing the words hides the control set that each experiment actually needs.
If the project starts from a protein antibody and no peptide sequence, stay on protein-complex discovery through the IP-Mass Spectrometry Service. If the project starts from a peptide sequence, change the bait geometry before the first capture.
When IP-MS Can Answer a Peptide Question
Antibody-MS is useful for protein-peptide interactions only in a narrow window. The first fit is motif dependence of a protein-protein pair. A competing wild-type peptide is added to a Co-IP. Loss of a named partner, while a mutant peptide leaves that partner in place, supports a motif-sensitive association. The readout is still protein recovery, not a peptide identification table.
The second fit is native-context follow-up after a peptide pull-down hit. A candidate protein recovered on an immobilized peptide can be tested by Co-IP of the endogenous protein, with and without the competing peptide. That run asks whether the protein sits in a recoverable complex. It does not repeat the peptide-bait discovery.
Antibody capture is a poor first assay when the only bait is a short peptide, when no IP-grade antibody exists, or when the claim is affinity of a purified pair. A whole-complex IP will not isolate a free peptide ligand the way a bead-immobilized peptide isolates proteins. Tryptic peptides that appear after digestion should not be rewritten as ligands.
Vector construction, overexpression, and knockout are outside Co-IP intake. A commercial IP-grade antibody supplied by the client remains the usual reagent when antibody-MS is the chosen follow-up.
Peptide Immobilization and Competition Designs
Immobilization turns the peptide into the bait for protein-peptide interactions. A biotin handle on a spacer, a cysteine for thiol coupling, or a tagged recombinant peptide can be parked on streptavidin, maleimide, or affinity resin. The handle should sit away from the motif. A buried biotin or a short spacer can hide the sequence or raise bead background.
Competition tests whether capture depends on that sequence. Excess free wild-type peptide should reduce recovery of sequence-specific binders. A scrambled or point-mutant peptide should not compete to the same degree. Running the mutant peptide as a second immobilized bait is stronger than relying on bead-only resin. Histone-peptide pull-downs that later supported reader assignments used matched modification and sequence controls, not a single decorated peptide (Vermeulen et al. 2010). Early peptide-bait screens likewise compared biotinylated sequences rather than treating one peptide as a universal probe (Schulze and Mann 2004).
Write the competition rule before the capture. State peptide concentration, incubation buffer, and whether the free peptide was pre-incubated with the lysate or the protein. A competition that is added only after washing cannot rescue a sticky bead.
The comparison below keeps whole-complex antibody-MS, immobilized-peptide capture, and pair affinity on separate tracks.
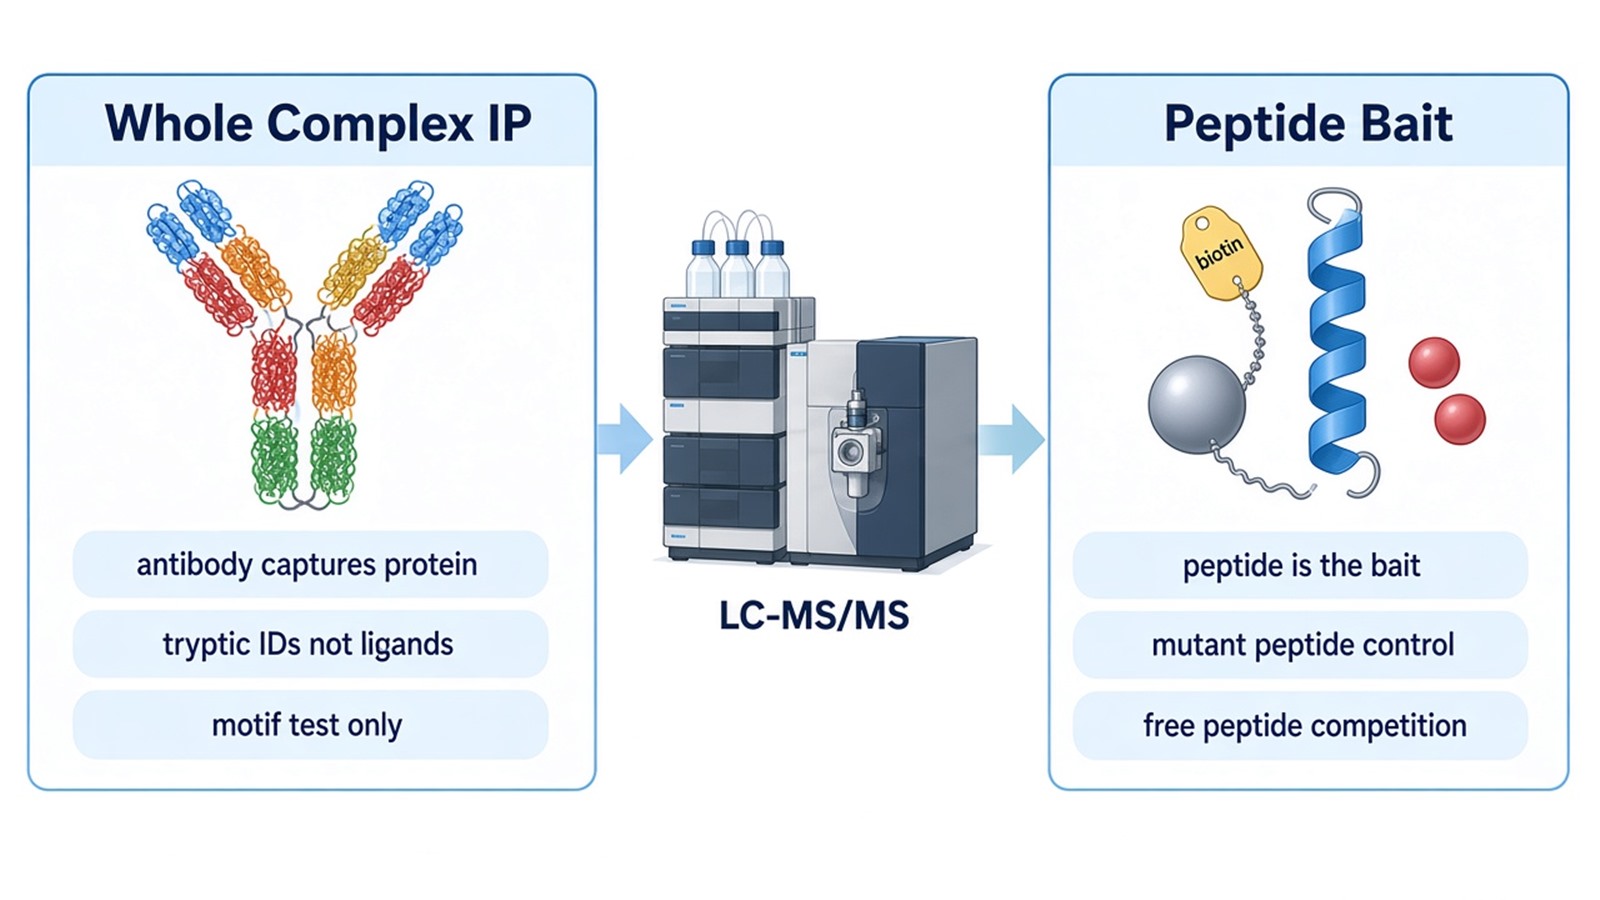
Figure 1. Whole-Complex IP-MS vs Immobilized Peptide Bait vs Pair Affinity.
Pull-Down-MS When the Peptide Is the Bait
Pull-down-MS is the discovery route when the peptide is immobilized and the unknown side is protein identity. The peptide-loaded resin is incubated with a lysate or a purified protein, washed, and digested. LC-MS/MS reports proteins present on the peptide bait versus the control peptide and the empty resin. Relative intensity can be compared across those tubes.
This is an in-vitro retention assay. It can be run on a cell extract, so some captured proteins may still bind through a bridge. It does not report live-cell occupancy. It does not report a cellular KD. Sequence-specific enrichment versus the mutant peptide is the usable discovery call.
Bead binders and abundant lysate proteins still appear. Recurrent-contaminant catalogs such as the CRAPome help interpret leftovers after experimental subtraction (Mellacheruvu et al. 2013). They do not replace the control peptide.
Already-run pull-down products can follow the handoff in the Fusion Protein Interaction Analysis Service | Pull-Down and MS. GST-tagged protein baits, with the peptide as prey, can be scoped through the GST Pull Down Assay Service.
Orbitrap-class high-resolution LC-MS/MS is the usual platform. Search should use UniProt or a client FASTA in MaxQuant or an equivalent disclosed workflow. Identification depth is typically hundreds to thousands of proteins and depends on the sample. After controls, the usable set is the peptide-enriched subset.
Controls and How Binding Evidence Should Be Read
Controls decide whether a name is a peptide-binding candidate. The minimum set for pull-down-MS is the wild-type peptide bait, a sequence-matched mutant or scrambled peptide, and a resin-only tube processed through the same wash, digest, and LC-MS/MS method. Competition with free wild-type peptide is the specificity check on that set.
Antibody-MS follow-up needs its own controls. A bait IP, a matched IgG or empty-bead IP, and the peptide-competition arm should be run together. A peptide that collapses one partner in the Co-IP supports motif sensitivity of that protein pair. It does not convert the rest of the IP list into peptide ligands.
Affinity assays answer a third sentence. SPR can report association, dissociation, and an equilibrium constant for a prepared peptide-protein pair. That constant is a biosensor value. It is not the intracellular affinity. Binding on the chip does not equal functional regulation. Pair-level SPR intake is described under Surface Plasmon Resonance Technology.
|
Evidence |
What it can support |
What it cannot support |
|---|---|---|
|
Tryptic peptides in an IP-MS table |
Protein identity in that digest |
A peptide ligand inventory |
|
Peptide pull-down vs mutant peptide |
Sequence-biased protein retention |
Live-cell occupancy |
|
Free-peptide competition |
Motif sensitivity of capture |
A unique binding site map |
|
Co-IP plus competing peptide |
Motif-sensitive protein recovery |
Direct contact of every co-enriched name |
|
SPR on a prepared pair |
In-vitro affinity and kinetics |
Cellular KD or a functional outcome |
The table is a reading rule, not a ranking of platforms. Mixed enrichment-MS projects that already include peptide-bait and protein-bait arms can also be reviewed under the MS-Based Protein-Protein Interaction Analysis Service.
Samples, Deliverables, and Feasibility Intake
State the peptide sequence, modification, handle, and the exact sentence the result is allowed to support. Send a matched mutant or scrambled peptide with the same handle. For pull-down-MS, cells or tissue that express the suspected protein, or a purified protein, should be declared with lysis buffer. For already-run captures, send visible gel bands, washed beads, or liquid products. Planning amounts for liquid pull-down or IP products are at least 20 ug total protein at more than 0.5 ug/uL, with SDS kept below 0.1 percent. High salt, undepleted detergent, and severely degraded material should not be shipped.
For SPR after a named pair exists, planning amounts are typically 50 to 100 ug of protein at 90 percent purity or higher, plus 1 to 2 mg of peptide. Avoid high salt, glycerol, and amine buffers that block coupling. Peptides may arrive lyophilized or in a compatible solution.
Intake is faster when the bait geometry is mapped before shipment. The diagram below links the peptide question to antibody-MS, pull-down-MS, competition, and affinity follow-up.
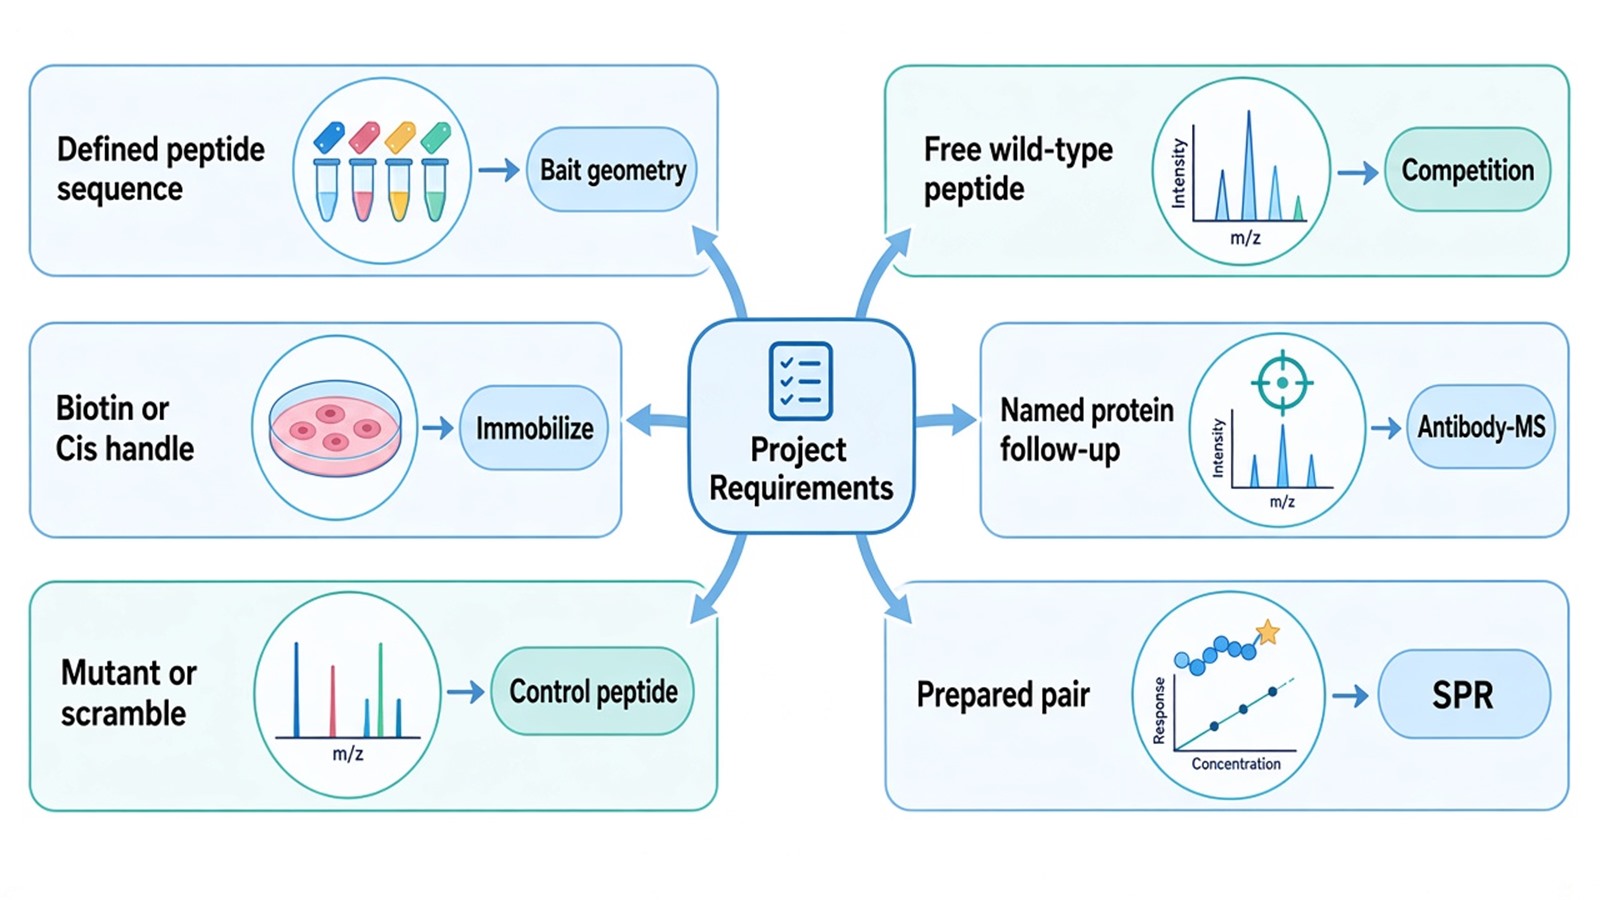
Figure 2. Matching Peptide Checks With IP-MS and Pull-Down Validation.
Deliverables depend on the scoped arm. Pull-down-MS typically returns identification tables, control-peptide comparisons, and a competition note. Antibody-MS follow-up returns bait and control tables plus the peptide-competition contrast. SPR returns sensorgrams and fitted constants for the prepared pair. Deliverables do not include a claim that tryptic peptides were ligands, a cellular KD, or a certificate of direct binding in cells. To start intake, send the peptide sequences, handle, control plan, and intended claim to info@MtoZ-Biolabs.com or use the site inquiry form so bait geometry can be confirmed before shipment.
Frequently Asked Questions
1. Can tryptic peptides in an IP-MS table be treated as peptide ligands?
No. Those peptides are digestion products that identify proteins in the IP. A ligand peptide has to enter the experiment as a designed bait, a competitor, or a purified analyte.
2. Is a biotinylated peptide pull-down enough if no mutant peptide was run?
It can produce an identification table. It cannot separate sequence-specific binders from streptavidin or resin background. Add a scrambled or point-mutant peptide before ranking names.
3. When should SPR be added after a peptide pull-down-MS hit?
Add SPR when one protein has been named and the next sentence is affinity or kinetics of that pair. Keep pull-down-MS as the discovery filter. Do not treat a biosensor KD as the intracellular value.
4. Can a competing peptide in a Co-IP replace immobilized-peptide discovery?
It can test motif dependence of a known protein partner. It cannot screen unknown proteins that recognize the peptide. Discovery still needs the peptide on the bead.
5. What should a feasibility email attach for a protein-peptide project?
Attach wild-type and control peptide sequences, the immobilization handle, whether the protein is purified or in a lysate, and the claim sentence. State if antibody-MS is requested only as a native-complex follow-up.
Researchers can separate protein-peptide interactions from whole-complex IP-MS, then confirm peptide geometry, controls, and the next assay through the MtoZ Biolabs Peptide-Peptide/Protein Interaction Analysis Service.
Reference
- W.X. Schulze, M. Mann (2004). A novel proteomic screen for peptide-protein interactions. J. Biol. Chem., 279, 10756-10764. https://doi.org/10.1074/jbc.M309909200
- M. Vermeulen, H.C. Eberl, F. Matarese, H. Marks, S. Denissov, F. Butter, K.K. Lee, J.V. Olsen, A.A. Hyman, H.G. Stunnenberg, M. Mann (2010). Quantitative interaction proteomics and genome-wide profiling of epigenetic histone marks and their readers. Cell, 142, 967-980. https://doi.org/10.1016/j.cell.2010.08.020
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
