



SILAC-based protein–protein interaction analysis using mass spectrometry is a four-step labeled Co-IP-MS path, not a more precise substitute for every other interactome method. Experimental and control cells are grown with light or heavy amino acids. A bait is captured by co-immunoprecipitation (Co-IP) or a related affinity step. Liquid chromatography–tandem mass spectrometry (LC-MS/MS) reads the same peptides as isotope pairs. Data analysis then ranks proteins by a heavy-to-light (H/L) ratio. The path reports wash-stable recovery under a written contrast. It does not, by itself, measure interaction strength, remodel a network, or prove a direct contact.
Protein–protein interactions (PPIs) sit under most cellular processes, and mass spectrometry (MS) is now a common readout for those complexes. Combining MS with stable isotope labeling by amino acids in cell culture (SILAC) is useful because the label is introduced in living cells and the two states can be compared after one acquisition. The useful first decision is whether the four steps are being used to filter bait-dependent partners or to compare occupancy after a treatment. Calling the combination precise and efficient does not replace that decision.
How an H/L value should be read as specificity or occupancy, and when TMT or label-free quantification is the better label, is covered on Quantitative Analysis of Protein-Protein Interactions Using SILAC and MS. This page stays with the analysis workflow and with the conditions under which its usual advantages hold.
Researchers who already have a proliferating cell system, a Co-IP-grade antibody or tag, or a written light–heavy contrast can also review the MtoZ Biolabs Protein-Protein Interaction Analysis Service for project-specific feasibility and analysis planning.
What a SILAC-Based Analysis Can Return
A completed run can return a peptide-identified protein table with paired intensities, a bait-recovery check, and a shortlist of outliers against the control channel. That is an analysis of proteins that survived lysis, wash, and elution with the bait. It is not a census of every protein that touched the bait in the cell.
Sonication or another harsh release step can raise total protein yield and still dissolve the complex that the Co-IP was meant to keep. In vivo labeling can reduce some later handling bias. It cannot repair an unmatched antibody, an empty-tag arm processed on a different day, or a search that treats every identification as an interactor. For what Co-IP-MS evidence means before a label is added, How to Identify Protein–Protein Interactions Using Mass Spectrometry provides a complementary evidence-type discussion.
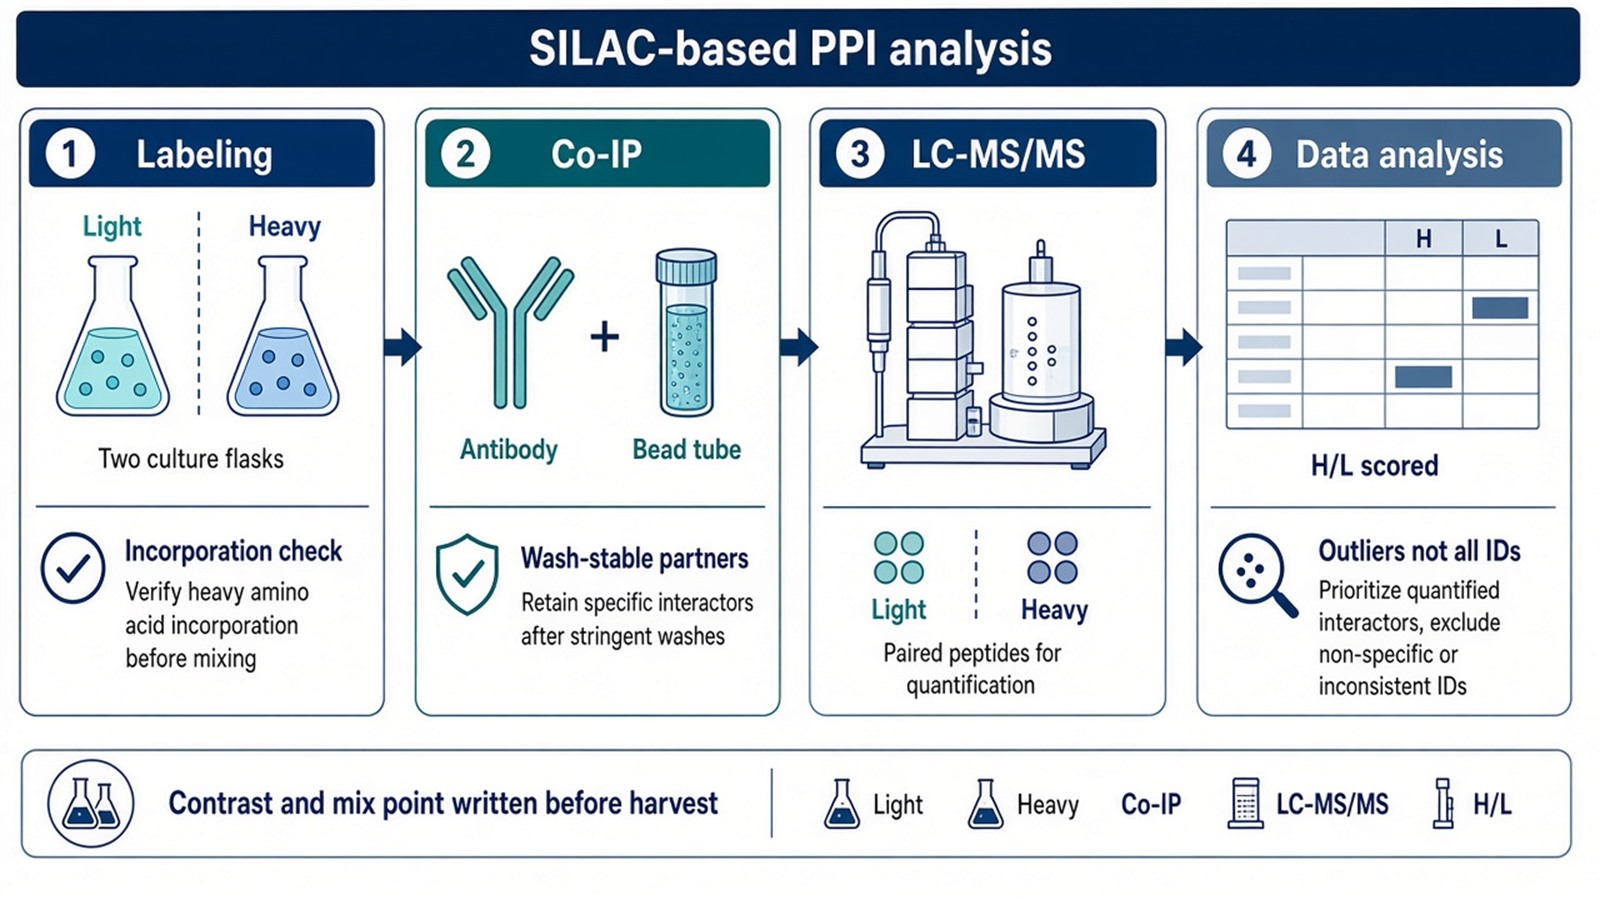
Figure 1. SILAC-based PPI analysis is labeling, capture, paired LC-MS/MS, then a scored table. The label does not convert every identification into an interactor.
Write the contrast and the mix point before the first harvest. Analysis cannot invent a control that was not captured.
The Four-Step SILAC-MS Workflow
The SILAC-based MS analysis workflow typically includes the following steps. Each step has a decision that later software cannot undo.
1. Sample preparation and labeling
Experimental and control cells are cultured in light- and heavy-labeled media and harvested after enough doublings for incorporation. A small digest should confirm that the heavy channel is no longer dominated by residual light peptide. Lysates are then prepared under a buffer that is intended to keep the complex, not to maximize total protein release. Complete disruption by sonication is a proteome-extraction habit. It is a poor default when the next step is Co-IP of a native assembly.
2. Co-immunoprecipitation
A specific antibody, or a tag handle, captures the target protein and proteins that remain bound after the stated wash. SILAC does not change what the resin retains. It only marks which culture the retained protein came from. Empty-tag, bead-only, or isotype arms belong in the opposite isotope state when the question is specificity. The same bait and the same antibody belong in both states when the question is a treatment contrast.
3. Mass spectrometry analysis
After Co-IP, eluates are usually mixed, digested with trypsin, and acquired by LC-MS/MS. Light and heavy versions of the same peptide are detected as a mass-shifted pair. The instrument records m/z and intensity. It does not assign biological meaning to the pair. Missed cleavage, co-isolation, and incomplete labeling still move the ratio.
4. Data analysis
Peptide pairs are rolled up to proteins and compared across the two states. The legitimate output is a ranked recovery table: which proteins move with the bait, or which proteins change recovery after treatment. SILAC does not convert that table into interaction strength. A large H/L can reflect better retention on the bead, higher prey abundance, or a bridged partner. A ratio near 1 is consistent with shared background. It is not proof that the protein is irrelevant in the cell.
Bait recovery should be inspected before prey ratios are interpreted. If the bait itself is unequal between channels, occupancy language is not available. Forward and reverse label-swaps test whether an outlier follows the biology rather than the isotope.
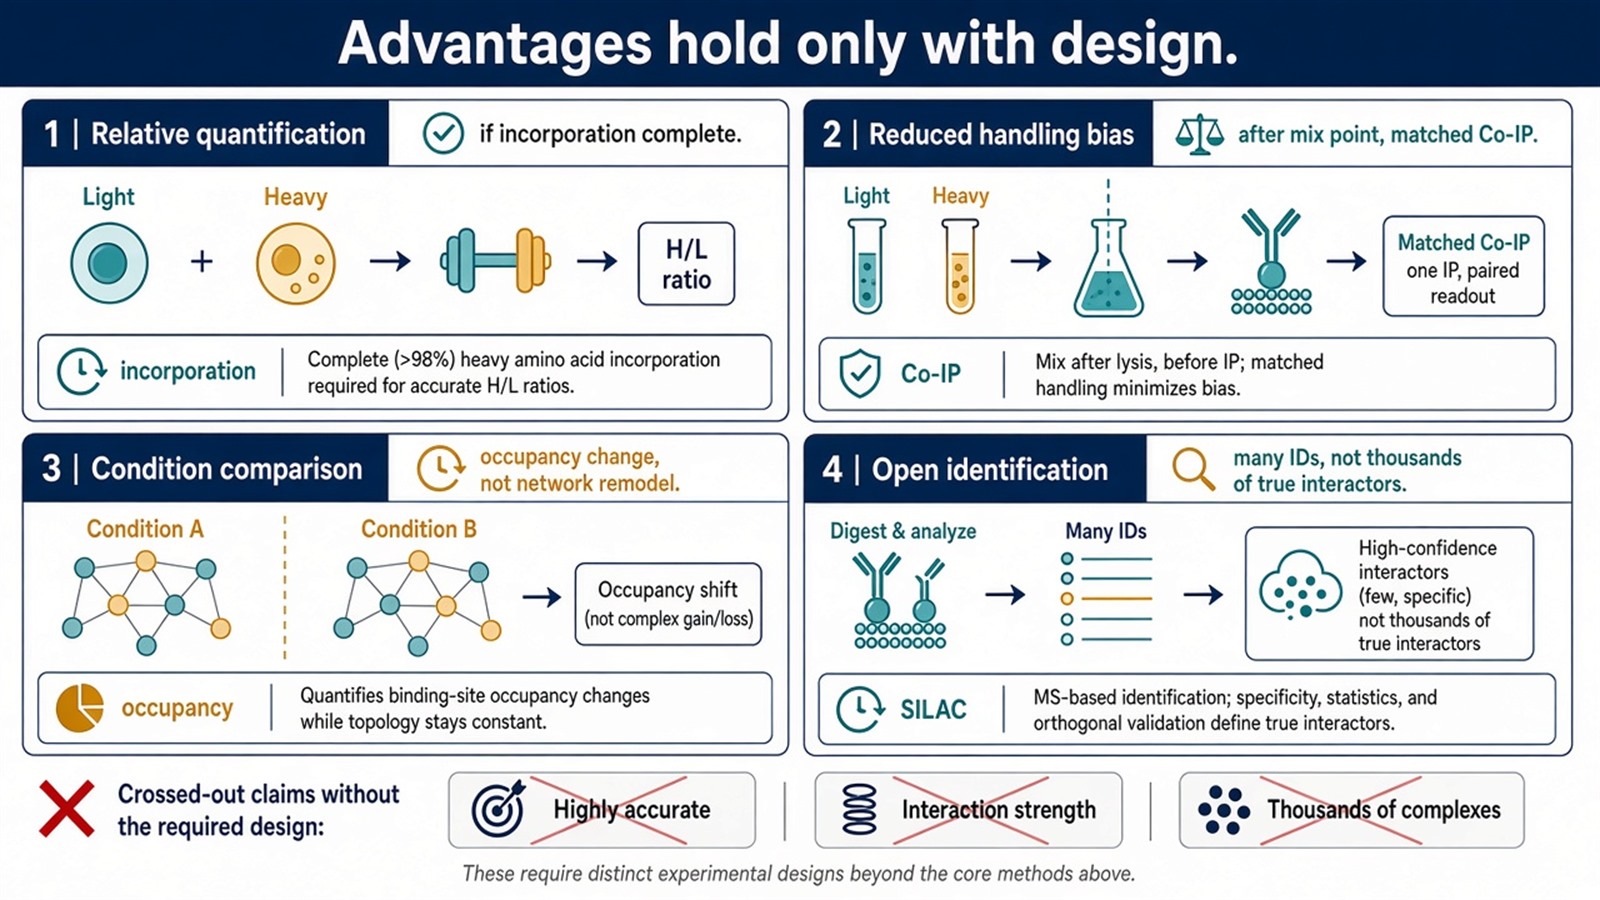
Figure 2. In-cell labeling, paired peptides, a treatment contrast, and an open identification list are usable only after incorporation, matched capture, and filtering. They are not automatic properties of the method name.
Advantages Restated as Design Conditions
The usual advantages of SILAC combined with MS are real only inside a written design.
1. Relative quantification
SILAC can give a chemically matched peptide pair and a usable relative ratio. That is not highly accurate measurement of interaction change under every condition. Incomplete labeling compresses outliers toward 1. Unequal bait recovery mislabels occupancy.
2. Reduced handling bias after mixing
Because the isotope is incorporated in living cells, mixed samples can share later digestion and acquisition. That reduces some sample-preparation error after the mix point. Error introduced by separate IPs, different bead lots, or unmatched lysis remains. An early mix can also allow subunit exchange and hide a true difference.
3. Condition-dependent comparison
A treatment contrast can show that recovery of a prey moved with stimulus, drug, or time. That is a change in the wash-stable neighborhood. It does not, by itself, reveal how a protein network was remodeled, and it does not assign a new edge as causal.
4. Open identification after one capture
LC-MS/MS can identify many proteins in a single eluate, which is the practical throughput of this path. Thousands of identifications are not thousands of interacting proteins. Most of the table is usually background, carry-over, or proteins that fail a control filter. High-throughput language here means an open, peptide-identified list, not an unbounded true-complex count.
How the Analysis Table Should Be Read
Start from the pair, not from a network drawing. Confirm incorporation, confirm bait recovery, then rank prey H/L against the intended contrast. Proteins that track the control channel stay in the background model. Proteins that track the bait, and that survive a label-swap or a second capture chemistry, are the candidates that can move to Western or reciprocal IP.
Do not export the unfiltered identification list into STRING or a pathway map and call that the SILAC analysis. The overlay can prioritize follow-up. It is not a substitute for the scored table. For control language that belongs with any quantitative AP-MS record, How to Improve Reliability and Reproducibility of PPI Experimental Results provides a more detailed design discussion.
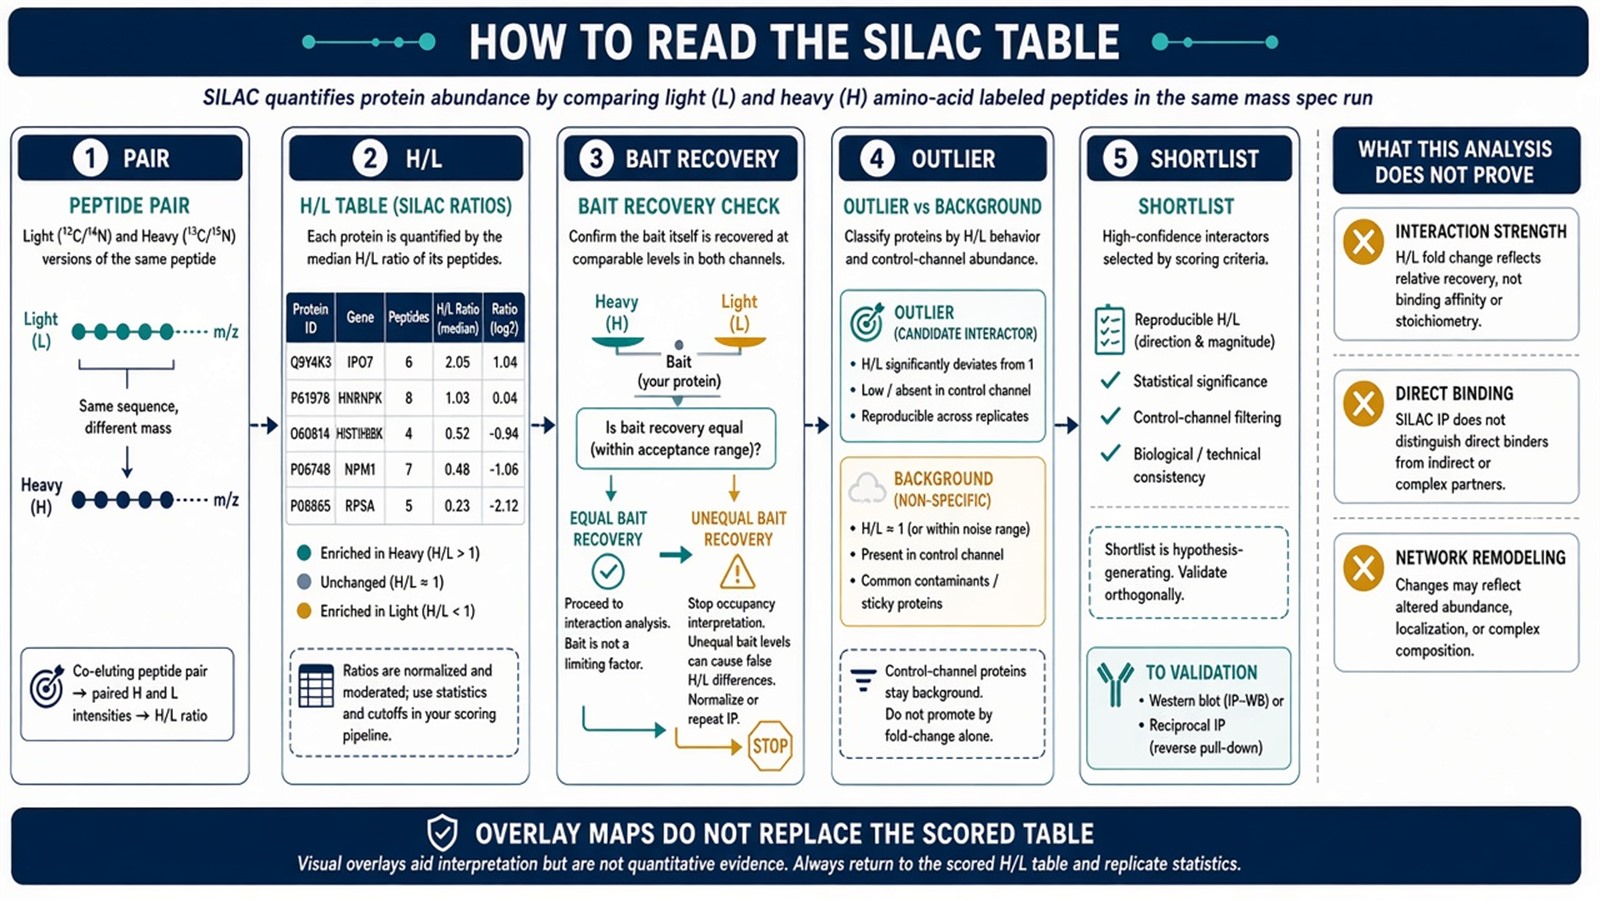
Figure 3. Analysis runs from paired peptides to an H/L table, a bait-recovery check, and an outlier shortlist. Strength, direct binding, and network remodeling are later claims.
Experimental Limits
SILAC-based analysis still requires proliferating cells that tolerate labeled medium. Tissue, fluid, and many primary cultures cannot be labeled through several doublings; those projects leave this workflow for a chemical label or a label-free comparison. Harsh lysis, weak antibodies, and overexpressed baits fail for the same reasons they fail in unlabeled Co-IP-MS. Diagnostic or target-nomination language sits outside ordinary research reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable SILAC-based analysis plan names the cell line, the light and heavy arms, the incorporation check, the Co-IP handle, the mix point, and the rule that turns pairs into a shortlist. MtoZ Biolabs scopes metabolic labeling, antibody or tag capture, Orbitrap-class LC-MS/MS, and paired-ratio reporting, with a written fallback when the cells will not label. The size of the identification list still depends on capture chemistry and filtering, not on calling the workflow high-throughput.
Once the four steps and the scoring rule are defined, the project can be scoped as a SILAC-based PPI analysis. Researchers can review the MtoZ Biolabs Protein-Protein Interaction Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What is SILAC-based PPI analysis using mass spectrometry?
It is metabolic light–heavy labeling, bait capture, LC-MS/MS of paired peptides, and a scored H/L table. It is not a general proof that two proteins bind.
2. Does in vivo labeling remove experimental error?
It can reduce some handling variance after samples are mixed. Separate IPs, unmatched lysis, and incomplete incorporation still move the result.
3. Can one SILAC-MS run detect thousands of interacting proteins?
One eluate can yield a large identification list. Interactors are the filtered outliers, not the raw count.
4. Does a changing H/L mean interaction strength changed?
No. It means recovery of that protein with the bait changed under the stated contrast. Strength is a later biophysical claim.
5. How does this page differ from a SILAC quantitative-methods page?
This page is the four-step analysis path and how to read its advantages. Ratio meaning and the choice among SILAC, TMT, and label-free comparison belong on the quantitative companion page.
Reference
- Ong S.E., Mann M. (2006). A practical recipe for stable isotope labeling by amino acids in cell culture (SILAC). Nat. Protoc., 1, 2650-2660. https://doi.org/10.1038/nprot.2006.427
- Blagoev B., Kratchmarova I., Ong S.E., et al. (2003). A proteomics strategy to elucidate functional protein-protein interactions applied to EGF signaling. Nat. Biotechnol., 21, 315-318. https://doi.org/10.1038/nbt790
- Vermeulen M., Hubner N.C., Mann M. (2008). High confidence determination of specific protein-protein interactions using quantitative mass spectrometry. Curr. Opin. Biotechnol., 19, 331-337. https://doi.org/10.1016/j.copbio.2008.06.001
- Hubner N.C., Bird A.W., Cox J., et al. (2010). Quantitative proteomics combined with BAC TransgeneOmics reveals in vivo protein interactions. J. Cell Biol., 189, 739-754. https://doi.org/10.1083/jcb.201001054
- Cox J., Mann M. (2008). MaxQuant enables high peptide identification rates, individualized p.p.b.-range mass accuracies and proteome-wide protein quantification. Nat. Biotechnol., 26, 1367-1372. https://doi.org/10.1038/nbt.1511
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
