



Protein interaction mass spectrometry reports names recovered after an enrichment or a covalent-capture step, then liquid chromatography–tandem mass spectrometry (LC-MS/MS). The sentence it can support is that a protein was recovered with this bait, this antibody pellet, or this covalent mark under these conditions. That sentence is narrower than binding occupancy, complex composition, interaction strength, or a functional-proteomics mechanism.
Affinity chromatography, co-precipitation, and yeast two-hybrid (Y2H) are the three strategies named on the original page. The first two become interaction-MS files only after a spectrometer reads the eluate. Y2H is a genetic reporter. Prey identity is usually recovered by sequencing, not by MS. Pairing Y2H with a later digest does not create a large-scale interaction-MS method, and it does not report mammalian complex composition or interaction strength. For the three MS routes that do share a spectrometer, How to Identify Protein-Protein Interactions Using Mass Spectrometry provides a complementary method-choice page.
Researchers who already have a bait, an immunoprecipitate, or a covalent-capture plan can also review the MtoZ Biolabs Protein-Protein Interaction Analysis Service for project-specific feasibility and analysis planning.
What the File Is Allowed to Say
Functional proteomics remains a field label. It is not an output of the search. A project can sit inside that field when the next assay is already named: a matched wild-type versus mutant recovery, a reciprocal capture, or a pairwise binding measurement. Regulatory mechanisms, disease molecular bases, and biopharmaceutical mechanisms of action are later overlays. They are not read from a protein list, and they are not granted by calling the spectrometer a cutting-edge tool.
Binding situation, in the original wording, has to be reduced to recovery. Identification of a name in one eluate is not occupancy. Enrichment against a matched empty-tag, bead-only, isotype, or ligase-only arm is a stronger, still provisional, candidate. Direct binding is a later assay. Strength, in the sense of a dissociation constant, is not a mass-spectrometry result. Relative ratios can rank a contrast. They do not report affinity.
Complex composition is not granted by naming proteins in an affinity eluate. One bait or one antibody pellet is a wash-stable neighborhood. Overlap across related baits can later mark a module. It is not a complete subunit list.
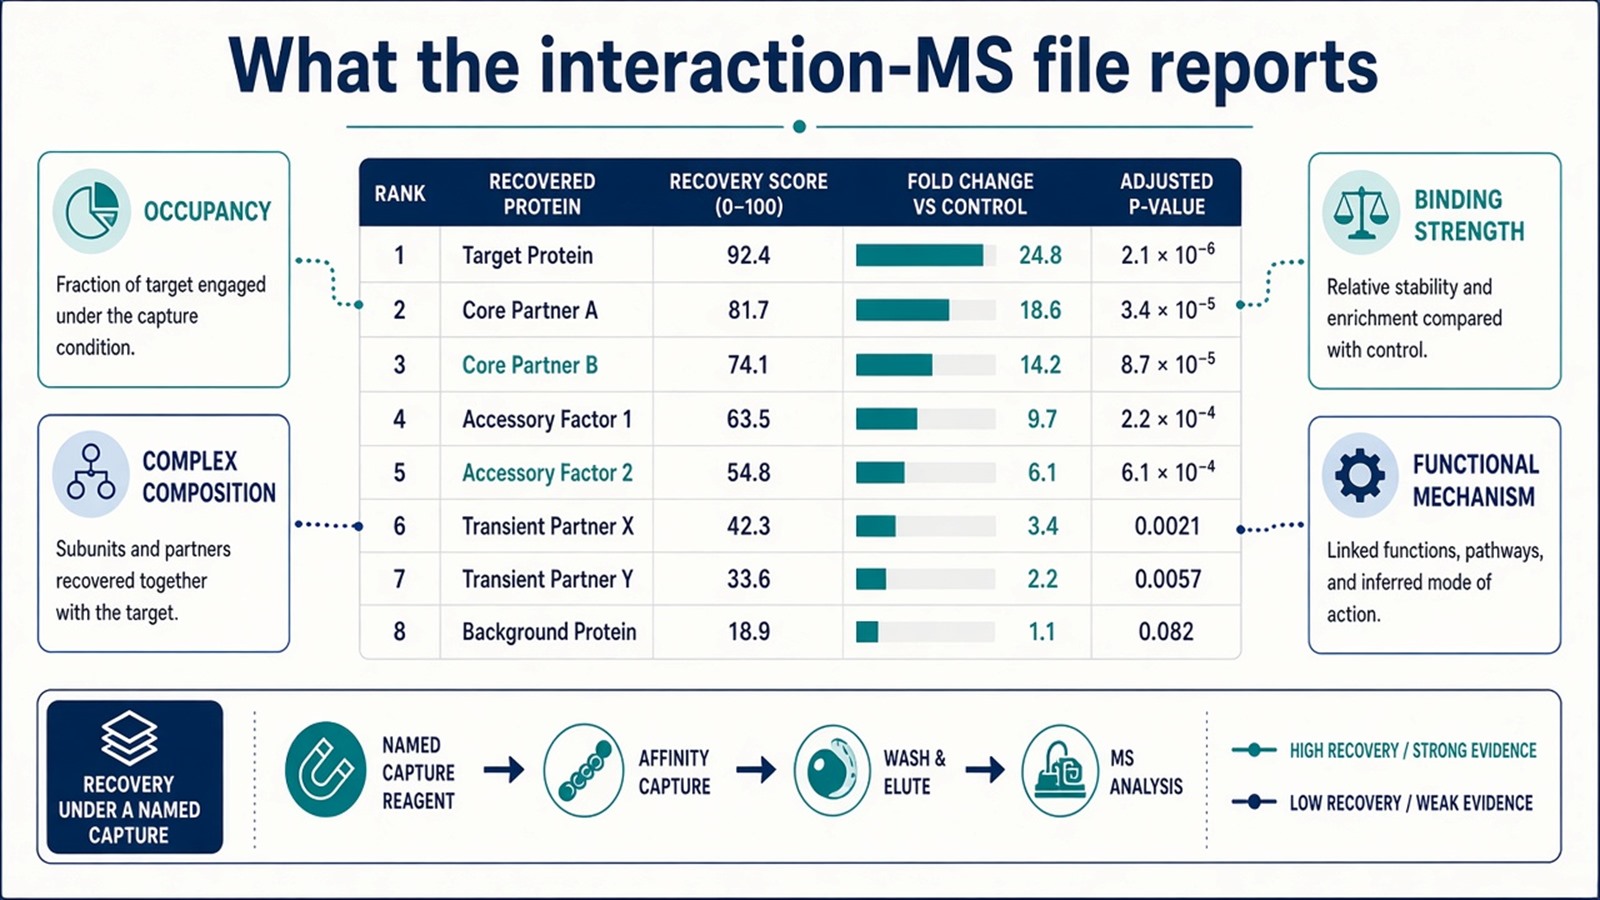
Figure 1. The file is a ranked recovery table. Binding occupancy, complex composition, interaction strength, and a functional mechanism are later overlays.
Write the next sentence as recovery under a named capture. Do not write it as strength, function, or drug action.
The Three Named Strategies, Recast
Affinity chromatography–mass spectrometry is tagged or ligand enrichment followed by LC-MS/MS. On this site that product is usually AP-MS or pull-down MS. The handle decides which protein entered the tube. The wash decides which associations remain. Identifying components in the eluate does not, by itself, explain interactions among those names, and it does not inventory a complex. For the interaction sentence that format can support, AP-MS Protein Interaction provides a complementary claim page.
Co-precipitation–mass spectrometry is Co-IP or IP followed by LC-MS/MS. The pellet reports proteins that survived lysis, antibody capture, and wash. It does not reveal whether a recovered name is a direct contact or an indirect hitchhiker. An isotype-matched antibody, matched by species, subclass, and micrograms, is the usual control. Bead-only is not a substitute when the question is antibody background. For that capture chain, How to Combine Co-IP with Mass Spectrometry for Protein Interaction Analysis? provides a complementary Co-IP-MS page.
Yeast two-hybrid–mass spectrometry, as written, mixes a genetic screen with a spectrometer. Y2H can nominate pairs that reconstruct in the yeast reporter. It is biased toward contacts that survive nuclear reconstitution in yeast, and it does not report a mammalian wash-stable pellet. It is not widely usable as a large-scale MS interaction method by adding a digest. If a third MS family is required, crosslinking MS or proximity-labeling MS is the usual addition, not Y2H.
Biopharmaceutical development remains an allowable study setting. A formulated antibody or a recombinant complex can be used as input when the capture and the control can still be written. Interaction-MS data from that input do not evaluate drug stability, optimize efficacy, or speak to safety. Those claims need dedicated analytical and pharmacological assays. A host-cell protein list in a process sample is a different experiment from an interaction-MS file.
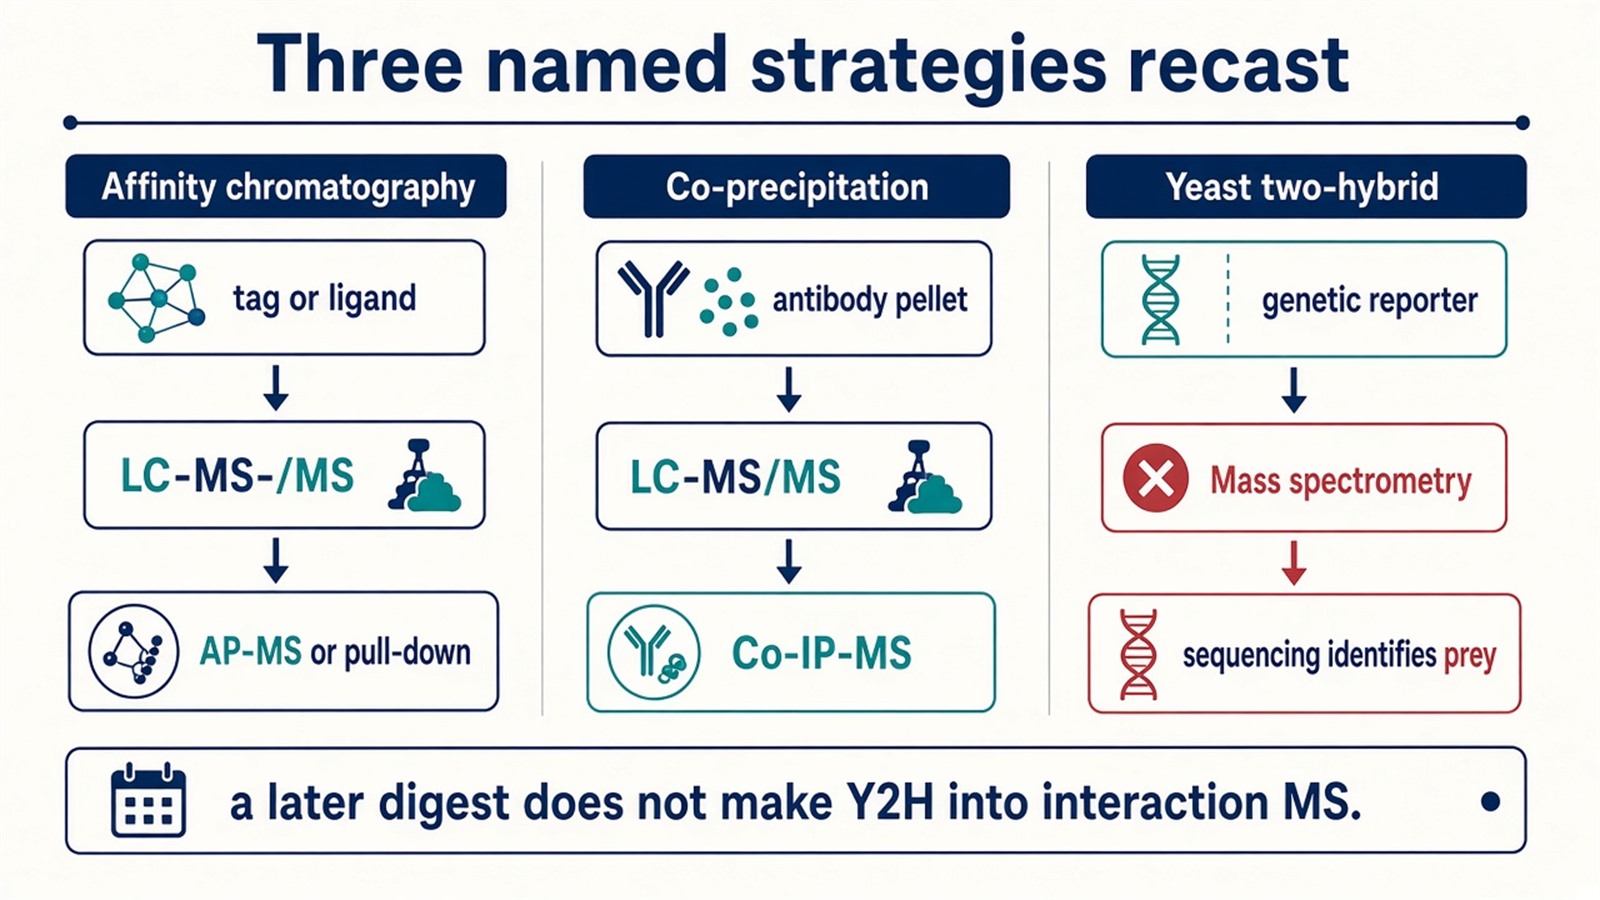
Figure 2. Affinity chromatography and co-precipitation feed LC-MS/MS. Y2H is a genetic reporter. A later digest does not convert it into interaction MS.
Experimental Limits
Protein interaction mass spectrometry reduces a project to one capture class, one control menu, and one claim level. Cutting-edge-tool, functional-proteomics-breakthrough, and disease-treatment language sits outside ordinary research reporting unless a separately validated assay is already named. A long identification list does not become a mechanism by being labeled functional proteomics. Further application of the same file will not, by accumulation, optimize a drug or treat a disease. Importance in biopharmaceutical research is a context sentence. It is not a measurement that the search returns, and it does not make yeast two-hybrid an MS strategy.
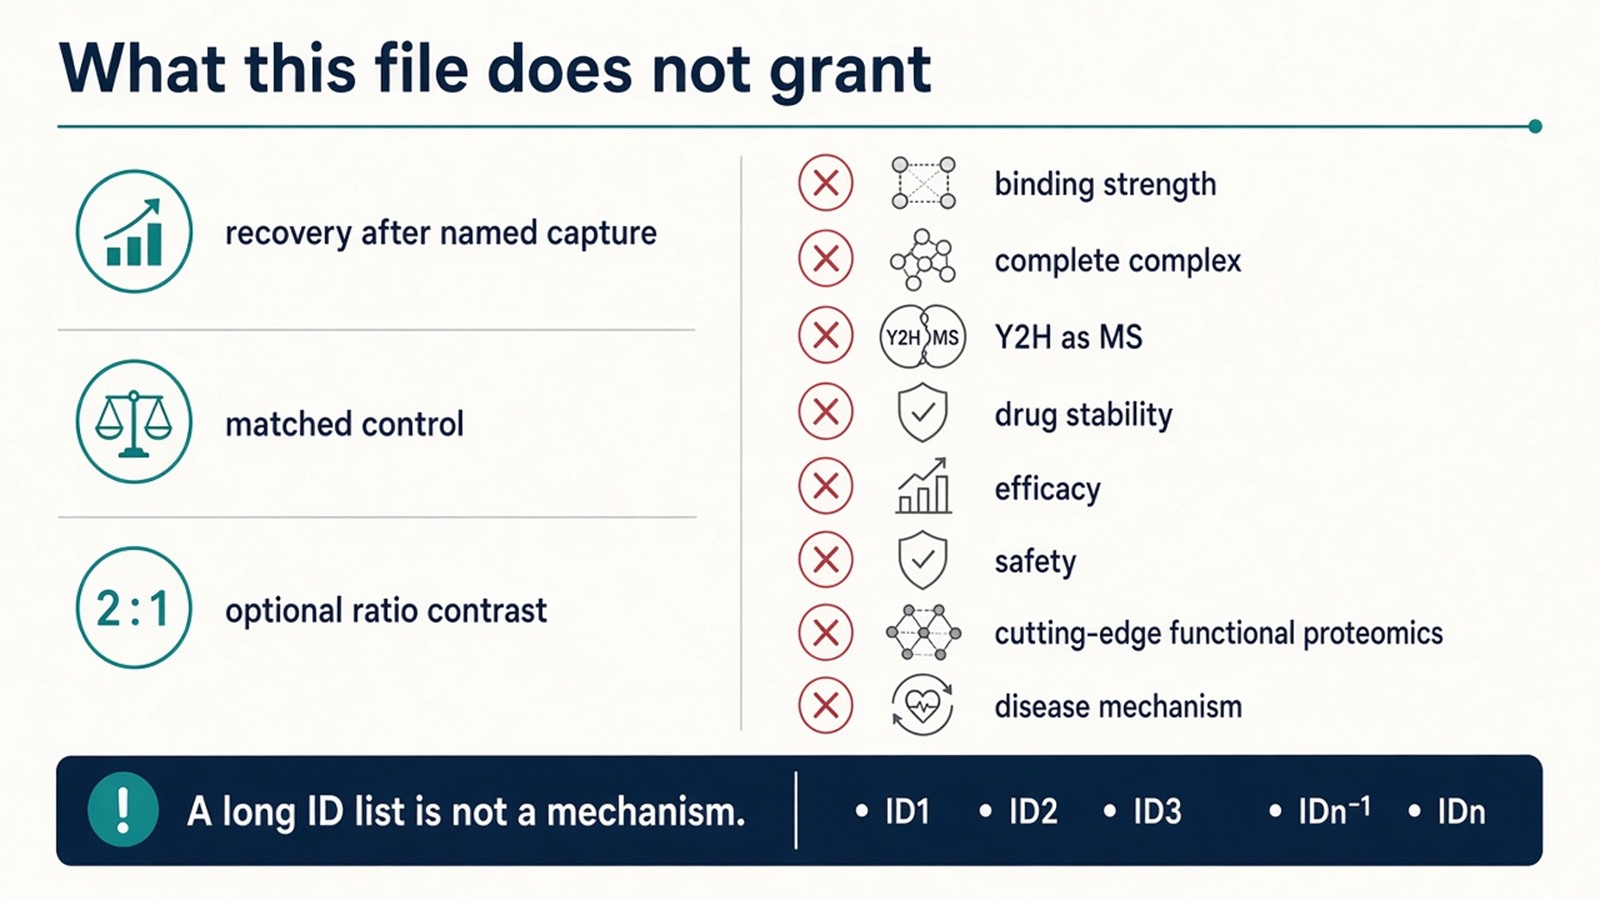
Figure 3. Keep: recovery after a named capture. Not granted: binding strength, complete complexes, Y2H as MS, biopharmaceutical stability, efficacy, or safety.
Project Planning and Service Scope
A usable request names the capture class, the control arm, and the sentence that will be allowed after the search. If the bait is tagged, the project is usually scoped as AP-MS. If an antibody pellet is already in hand, it is usually scoped as IP-MS or Co-IP-MS. If the needed sentence is a live-cell neighborhood or a residue pair, this overview should not force the sample through affinity chromatography. MtoZ Biolabs scopes those files on Orbitrap-class LC-MS/MS. Interpretability still follows those arms, not a cutting-edge heading. A biopharmaceutical sample does not change that split.
Once the next sentence is a tagged-bait co-enrichment table, this overview should hand off. Researchers who need that table can review the MtoZ Biolabs Affinity Purification-Mass Spectrometry Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. Does protein interaction mass spectrometry report binding strength?
No. Ratios can rank a contrast under the capture used. They do not report a dissociation constant.
2. Can the file define complex composition?
Not from one eluate. It ranks names that survived that capture. Overlap across baits is a later pattern.
3. Is yeast two-hybrid a mass-spectrometry interaction method?
No. Y2H is a genetic reporter. Prey is usually identified by sequencing.
4. Does Co-IP-MS distinguish direct from indirect contacts?
No. The pellet is wash-stable recovery. Direct versus indirect is a later assay.
5. Can interaction MS evaluate biopharmaceutical stability or safety?
No. Those are separate analytical and pharmacological claims. An interaction-MS table does not optimize efficacy.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Dunham W.H., Mullin M., Gingras A.C. (2012). Affinity-purification coupled to mass spectrometry: basic principles and strategies. Proteomics, 12, 1576-1590. https://doi.org/10.1002/pmic.201100523
- Fields S., Song O. (1989). A novel genetic system to detect protein-protein interactions. Nature, 340, 245-246. https://doi.org/10.1038/340245a0
- Roux K.J., Kim D.I., Raida M., Burke B. (2012). A promiscuous biotin ligase fusion protein identifies proximal and interacting proteins in mammalian cells. J. Cell Biol., 196, 801-810. https://doi.org/10.1083/jcb.201112098
- Leitner A., Walzthoeni T., Kahraman A., et al. (2010). Probing native protein structures by chemical cross-linking, mass spectrometry, and bioinformatics. Mol. Cell. Proteomics, 9, 1634-1649. https://doi.org/10.1074/mcp.R000001-MCP201
