



A targeted proteomics project reaches multiple reaction monitoring (MRM) when peptide ions can already be named and the remaining task is to record those ions with two sequential mass filters. On a triple quadrupole, quadrupole 1 (Q1) isolates a precursor m/z, quadrupole 2 (Q2) fragments that precursor by collision-induced dissociation (CID), and quadrupole 3 (Q3) isolates one product ion. The chromatographic trace of that precursor-to-product pair is the quantitative signal, usually reported as peak area. Selected reaction monitoring (SRM) names one such pair. MRM names the same experiment when several pairs are recorded in one run. The mechanism does not store a full fragment spectrum, does not recover unknown proteins, and does not report concentration unless a matched stable isotope-labeled internal standard (SIS) is present.
What an MRM or SRM Transition Is
A transition is a defined ion pair: one precursor m/z plus one product m/z, acquired at a stated collision energy. Lange et al. 2008 described two mass filters around a collision cell, with dwell time spent only on the chosen pair. IUPAC prefers the term SRM for that measurement. Proteomics papers still use MRM when many transitions share one liquid chromatography (LC) run. In practice, a laboratory asking how MRM works is asking how scheduled, multiplexed SRM is implemented on a triple quadrupole.
The peptides that enter those filters are surrogates of parent proteins after digestion, most often with trypsin. Proteotypic, unique sequences are preferred. Shared peptides, missed cleavage, and variable modifications change which protein the transition can represent. Antibody availability does not gate the assay. The assay is gated by whether the ion pair can be formed, isolated, and distinguished in the study matrix.
MRM is not a discovery acquisition mode. Proteins outside the scheduled list stay unmeasured. This page treats peptide MRM for targeted proteomics. The usual proteomics MRM route is electrospray ionization (ESI) after reversed-phase LC, so that each precursor elutes in a retention-time window that can be scheduled.
Researchers can refer to Advantages and Disadvantages of Multiple Reaction Monitoring for a more detailed trade-off framework.
How Q1, CID, and Q3 Create the Ion Current
The digest is separated by LC. At a given retention time, Q1 transmits ions inside a precursor isolation window, commonly near unit resolution on a triple quadrupole. Ions outside that window are rejected before fragmentation. Transmitted precursors enter Q2, a collision cell filled with a neutral gas. CID at a set collision energy breaks a fraction of those precursors into product ions. Q3 then transmits one product m/z window to the detector. The detector records ion current for that pair versus chromatographic time. Peak area of that trace is the quantitative value.
The two windows explain both selectivity and residual interference. Co-eluting ions whose precursor or product m/z falls outside the windows never reach the detector. Ions that share both windows still contribute to the same peak. Because a full MS2 spectrum is not stored, a contaminated product ion cannot be replaced from the same scan later. Collision energy, isolation width, and the identity of the product ion have to be chosen before the cohort is acquired.
An MRM signal is the ion current of one predefined precursor-to-product pair across the chromatographic peak.
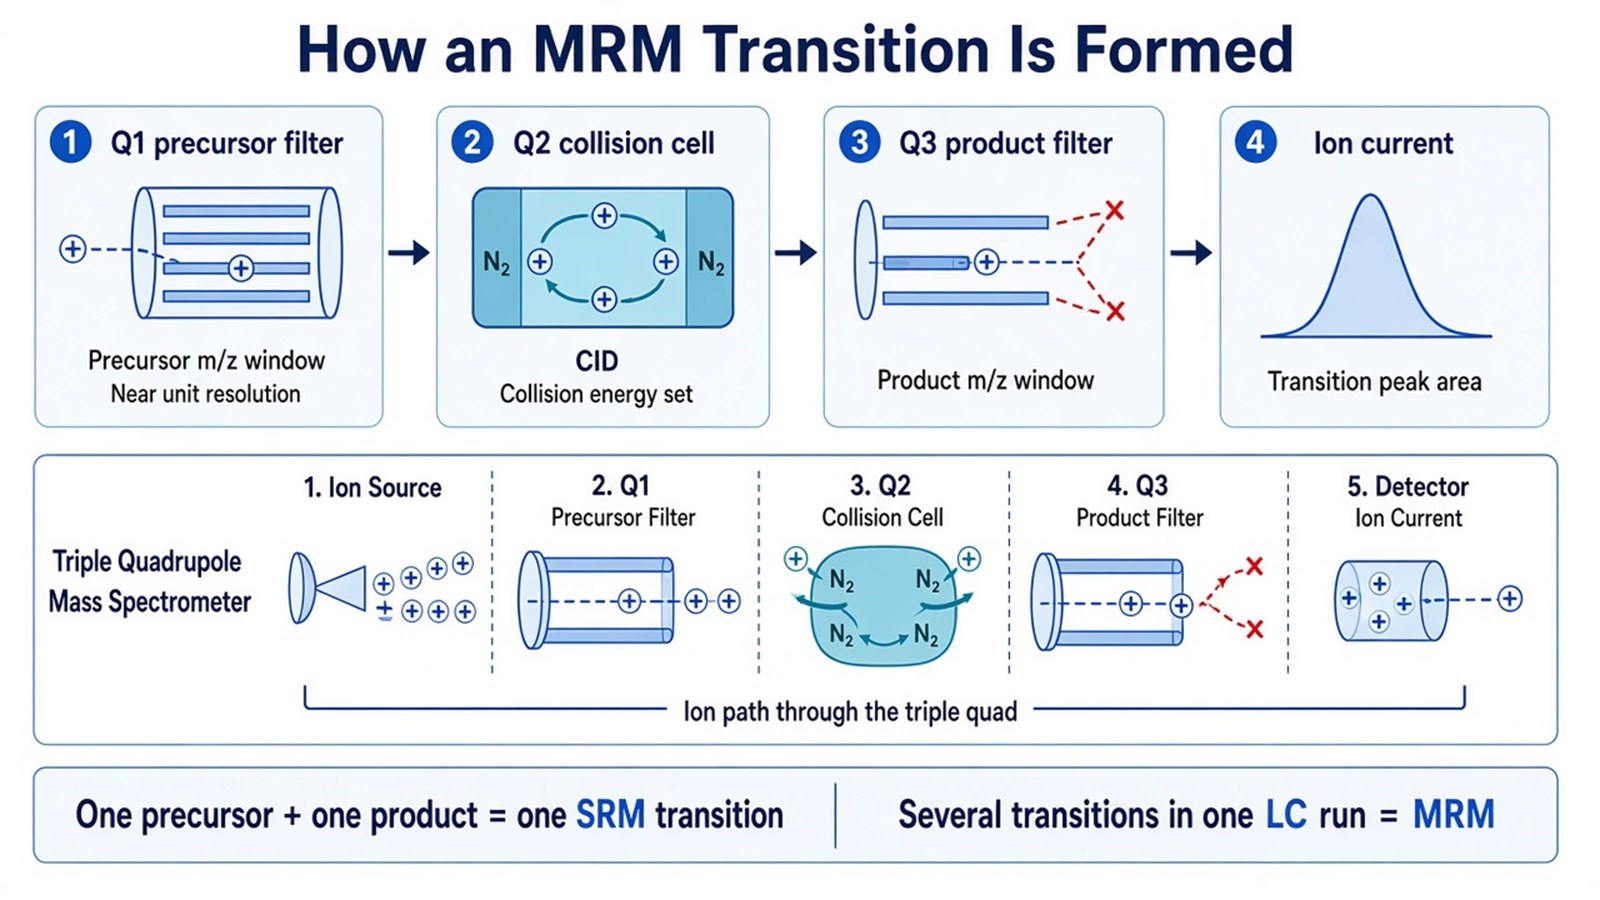
Figure 1. MRM transmits one precursor window, fragments that precursor by CID, transmits one product window, and quantifies from the ion-current peak area of that transition.
A quantitative transition is often paired with one or more qualifier transitions from the same precursor. Qualifier-to-quantifier intensity ratios, retention-time agreement, and, when an SIS is present, light-heavy co-elution are the practical tests that the peak is the intended peptide. Picotti and Aebersold 2012 treated those checks as part of the SRM measurement itself, not as optional decoration after a peak is integrated.
Parameters That Follow From Two Mass Filters
Dwell time is the interval Q1 and Q3 spend on one transition. Cycle time is the sum of dwells, plus overhead, for every transition that is active at that moment. Chromatographic peak width then sets how many points fall across the peak. Adding transitions shortens dwell or lengthens cycle. Either change can distort peak area if the peak is undersampled. Scheduled acquisition records a transition only near its expected retention time and recovers dwell for the rest of the gradient (Stahl-Zeng et al. 2007; Lange et al. 2008). Retention-time drift can make a scheduled window miss the peptide.
Collision energy controls how completely the precursor fragments into the chosen product. Too little energy leaves the precursor intact. Too much energy can destroy the product ion that Q3 is set to collect. Isolation width at Q1 and Q3 trades ion transmission against interference. Wider windows admit more signal and more co-isolated background. Those settings belong in the method record with the peak-area table.
Same-peptide peak areas can support a fold-change comparison when digestion, loading, and chromatography stay comparable. Amount or concentration requires a matched SIS. The heavy peptide occupies a second set of transitions, usually with the same product ions shifted by the isotope label. The SIS corrects for much of the ion suppression that affects the native peptide in the same scan. It does not correct digestion that failed before the standard was added.
In MtoZ Biolabs targeted proteomics offerings, MRM/SRM is typically run on a triple quadrupole. Parallel reaction monitoring (PRM) is typically run on a high-resolution quadrupole-Orbitrap or Q-TOF class instrument. The two routes share a closed precursor list. They differ in whether Q3 preselects one product ion or a high-resolution analyzer records the fragment spectrum.
Limits of the Mechanism and When to Use It
The mechanism cannot recover a different product ion after acquisition. It cannot measure proteins that were never placed on the list. Unit-resolution Q3 cannot separate product ions that share the same integer m/z window. Plasma and other high-dynamic-range matrices may still suppress the target even when the transition is correctly set. Prefractionation or enrichment can lower the usable limit and will add recovery variation that the later peak cannot diagnose.
PRM records all product ions of a scheduled precursor at high resolution, so some unit-resolution interferences can be dropped after the run (Peterson et al. 2012). That design costs fill time and multiplex capacity. A large, mature panel that must run many times on a triple quadrupole can still justify locking Q3 transitions once they are clean. Data-dependent acquisition (DDA) or data-independent acquisition (DIA) remains the route when the protein inventory is still open.
The useful test is whether named ion pairs can be scheduled, whether dwell time still samples each peak, and whether the claim matches the presence of an SIS.
Mechanism condition |
MRM/SRM often fits |
Reassess before locking the method |
|---|---|---|
Ion pair definition |
Precursor and product m/z can be named and optimized |
Only protein names exist, with no detectable peptide ions |
Spectrum storage |
Preselected transitions are acceptable; no full MS2 needed later |
Fragments still need to be chosen or swapped after acquisition |
Resolution |
Unit-resolution Q3 windows are clean in the study matrix |
Co-eluting ions share both Q1 and Q3 windows |
Cycle versus peak width |
Scheduled dwells yield enough points across each peak |
So many concurrent transitions that the peak is undersampled |
Quantitative claim |
Same-peptide relative comparison, or amount with SIS |
Copy number across different proteins, or amount without SIS |
Adjacent option |
Repeated triple-quadrupole panel after transitions are locked |
Open-proteome discovery, or a small panel that needs full MS2 |
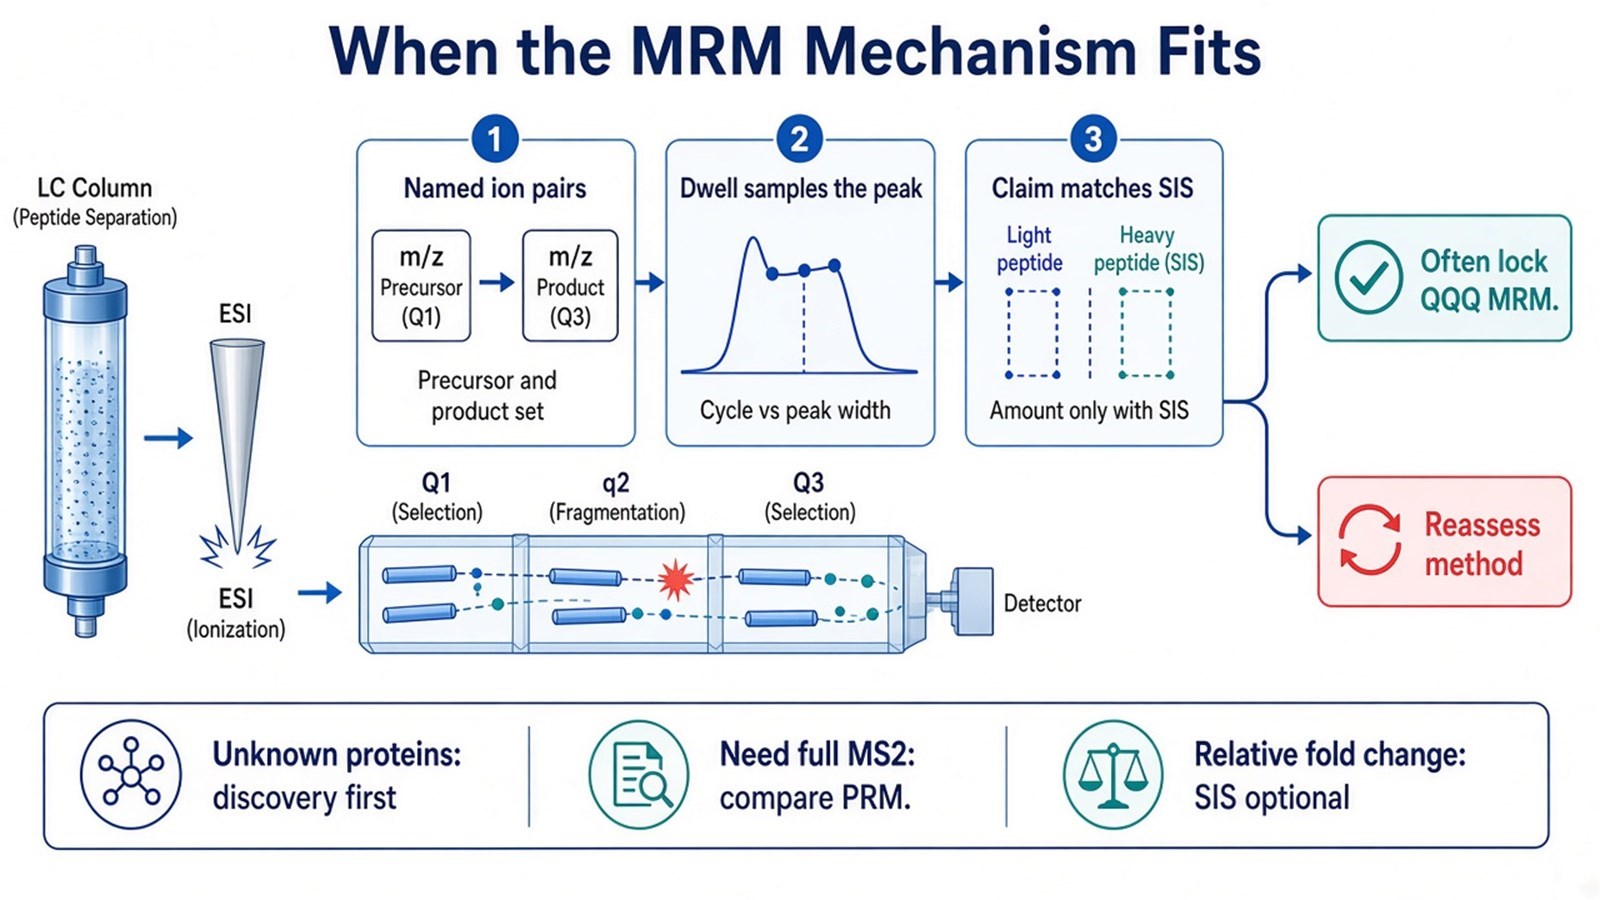
Figure 2. The MRM mechanism fits when ion pairs can be named, dwell time still samples each chromatographic peak, and the quantitative claim matches whether an SIS is present.
If names do not yet exist, keep discovery acquisition in place. If fragment confirmation in a complex digest is the scarce resource, researchers can refer to PRM LC-MS Targeted Proteomics before freezing Q3 ions.
Researchers who already have a closed transition list can also review the MtoZ Biolabs Multiple Reaction Monitoring Analysis Service for project-specific feasibility and analysis planning. When the remaining decision is whether MRM or PRM better matches target number and matrix complexity, researchers can also review the Targeted Proteomics Service for project-specific routing guidance.
Frequently Asked Questions
1. Why does MRM not store a fragment-ion spectrum?
Dwell time is spent transmitting one Q1 window and one Q3 window. Other product ions from the same precursor are not recorded in that scan. A contaminated transition cannot be replaced later from the same file. Choose quantifier and qualifier ions during method development.
2. Is MALDI the usual ionization for proteomics MRM?
Peptide MRM for this workflow is typically ESI after LC, so that retention time can be scheduled. MALDI can ionize peptides in other experiments. It does not by itself create the Q1-CID-Q3 filter sequence that defines MRM.
3. What is the difference between a quantifier and a qualifier transition?
The quantifier is the ion pair used for peak area. Qualifiers are additional products from the same precursor, used to check that the peak is the intended peptide. A qualifier ratio that disagrees with the reference weakens the quantitative value even if the quantifier peak is tall.
4. Does ion current equal protein concentration?
Ion current reports the monitored transition, including any interferent that passed both filters. Relative comparison of the same peptide can proceed without an SIS if digestion and chromatography remain comparable. Amount or concentration needs a matched heavy standard.
5. When should PRM be compared with this triple-quadrupole mechanism?
Compare PRM when the panel is still small or medium, fragments still need to be chosen, or unit-resolution Q3 is untrusted in the matrix. Keep MRM when a locked QQQ panel will be repeated at large scale after transitions are clean.
MtoZ Biolabs can provide multiple reaction monitoring and targeted proteomics analysis for predefined protein or peptide panels, including relative quantification and isotope-assisted absolute quantification when the study design requires matched internal standards.
If further evaluation of transition lists, dwell time, matrix interference, or PRM versus MRM routing is needed, project details can be submitted for a feasibility review.
