



After a discovery proteomics run, a laboratory often holds a short protein list that still needs confirmation in a new sample set. Multiple reaction monitoring (MRM) records predefined precursor-to-product ion transitions on a triple quadrupole mass spectrometer. The method can deliver high selectivity and a usable linear range for known peptides because two mass filters reject much of the matrix background. The same design also sets the cost: proteins outside the list stay unmeasured, each transition needs validation in the study matrix, and a co-eluting ion with the same transition can still distort the peak area. Advantages listed in instrument tutorials only hold after the target list, the matrix, and the quantification goal are specified.
Why MRM Trade-offs Appear After Discovery or Panel Design
A typical project reaches MRM after differential proteins have already been nominated, or after a pathway panel has already been written down. The question has shifted from which proteins can be observed to whether a closed peptide set can support a group comparison with enough selectivity. Generic claims such as high sensitivity or high throughput stop helping at this point, because both depend on how many transitions share the cycle time and whether the chosen product ions stay free of interference.
MRM, and selected reaction monitoring (SRM) when a single transition is discussed, is built for peptides that can be named before acquisition. Lange et al. 2008 described two sequential mass filters around a collision cell, with no full fragment-ion spectrum stored for each precursor. Researchers can refer to Mechanism of Multiple Reaction Monitoring for how Q1, CID, and Q3 produce the ion-current signal. Dwell time is spent on the chosen ion pair, so a contaminated product ion cannot be replaced from the same scan later. The decision for a principal investigator is whether that bargain matches the remaining samples, the target count, and the claim required for the paper.
How Q1 and Q3 Filtering Produce an MRM Transition
Proteins are digested, most often with trypsin, and one or more proteotypic peptides are treated as surrogates of the parent protein. In a triple quadrupole, quadrupole 1 (Q1) isolates a precursor m/z. Quadrupole 2 (Q2) fragments that precursor by collision-induced dissociation (CID). Quadrupole 3 (Q3) isolates one product ion. The chromatographic trace of that precursor-to-product pair is the quantitative signal, usually reported as peak area.
The two filters explain both the main advantage and the main limitation. Co-eluting ions whose precursor and product m/z fall outside the isolation windows are rejected, which is why MRM can remain selective in a complex digest. Ions that share the same transition within unit-resolution windows still reach the detector. Because a full MS2 spectrum is not stored, later inspection cannot recover a different product ion from the same scan. Collision energy, dwell time, and scheduling must be set before the cohort is acquired.
An MRM signal is the chromatographic peak area of a predefined transition.
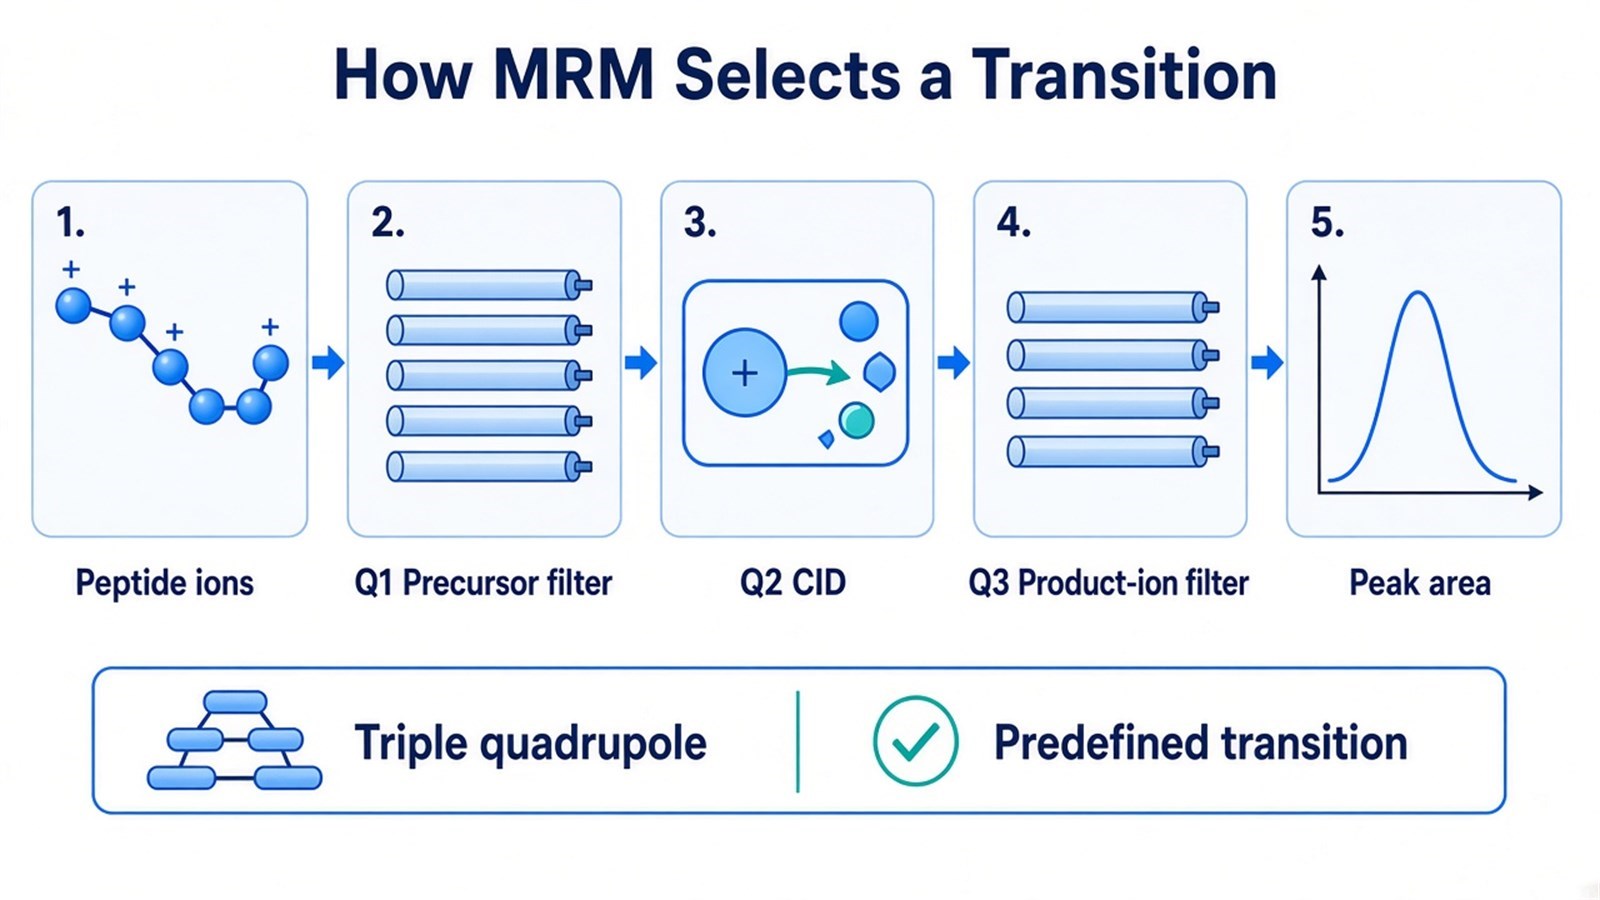
Figure 1. MRM records a predefined precursor-to-product transition on a triple quadrupole; the quantitative readout is the chromatographic peak area of that ion pair.
Advantages That Hold After Assay Conditions Are Set
The practical advantages of MRM are conditional. They appear when the peptide list is closed, transitions have been tested in the study matrix, and the same method will be repeated across the cohort.
Two-stage ion filtering in a complex digest
Q1 and Q3 isolation can suppress chemical background that would occupy a full-scan duty cycle. Lange et al. 2008 noted that non-scanning SRM can be about one to two orders of magnitude more sensitive than conventional full-scan acquisition for the monitored transitions, with a linear response that can extend across a wide dynamic range. That gain applies to the chosen ion pairs. An unvalidated transition is still not a protein concentration, and plasma peptides may still need enrichment or improved chromatography before they become usable.
Multiplex panels under a fixed cycle time
Several transitions can be monitored in one LC-MS/MS run, which is why MRM is used for protein panels. Useful panel size is limited by peak width, cycle time, and dwell time. Adding transitions shortens the time spent on each ion pair. Scheduled acquisition records a transition only near its expected retention time and can recover dwell time, as discussed by Stahl-Zeng et al. 2007 and Lange et al. 2008. Multiplexing helps when chromatography keeps peptides inside those windows. Retention-time drift can make scheduled transitions miss the peak.
Proteotypic peptides without an antibody
MRM measures a sequence-defined peptide. A new target can be added if a suitable peptide ionizes, fragments, and can be distinguished in the matrix. Antibody availability does not gate the assay. Specificity problems move to isobaric interference at the monitored transition, missed cleavage, and shared peptides between homologs. Sequence selection still has to exclude peptides with variable modifications unless the modification itself is the target.
Absolute amounts only with matched isotope standards
Relative peak areas of the same peptide can support a fold-change comparison when digestion, loading, and chromatography stay comparable. Absolute amount or concentration requires a matched stable isotope-labeled internal standard (SIS) at a known level, so that the light-to-heavy ratio can be converted to quantity. The SIS corrects for much of the ion suppression that affects the native peptide in the same scan. It does not correct incomplete digestion that occurred before the standard was added, and it does not create detectability for a peptide that remains below the assay limit.
Limitations That Change the Experimental Plan
The disadvantages of MRM show up as design constraints. They decide whether remaining sample should enter method development at all.
A closed target list cannot recover unknown proteins
Only scheduled precursors and product ions are recorded. A protein that was never nominated will not appear in the result table, even if it is abundant in the sample. MRM is a follow-up method for a list the study is willing to freeze. If the biological question still needs an open protein inventory, data-dependent acquisition (DDA) or data-independent acquisition (DIA) should remain the acquisition mode for that question.
Unit-resolution interference still reaches the peak area
Q3 on a triple quadrupole is a low-resolution mass filter. A co-eluting species that shares the precursor and product windows can inflate the peak area without any visual clue in a full spectrum, because no full spectrum is collected. Transition quality has to be demonstrated in the matrix: co-elution of light and heavy peaks when an SIS is used, consistent fragment ratios, Gaussian peak shape, and retention time agreement with a reference. Peterson et al. 2012 showed that parallel reaction monitoring (PRM) records all product ions at high resolution, which can separate some interferences that survive unit-resolution MRM. Interference risk is a reason to keep extra transitions during development and to drop contaminated ions before the study samples are queued.
Transition optimization, dwell time, and standard supply
Each peptide needs at least one quantitative transition and usually additional qualifier ions. Collision energy, precursor charge, and scheduling windows are optimized on a digest or on synthetic peptides, then confirmed in the real matrix. That work is the true time cost of MRM. If many targets are added late, dwell time falls or the method must be split into more injections. Absolute quantification adds a supply problem: heavy peptides must be available, their concentration must be known, and the spike-in level must sit in a linear range for each target. Relative comparison can proceed without an SIS. Amount or concentration claims cannot.
Matrix suppression and extra sample-preparation steps
Plasma, tissue, and other high-dynamic-range matrices can suppress peptide signal or bury the target under more abundant ions. Prefractionation or enrichment can lower the limit of quantification, and it also adds recovery variation that the later MRM peak cannot diagnose. Shi et al. 2016 treated enrichment as a sensitivity tool that trades throughput and additional variance. Sample-preparation load belongs in the MRM decision, especially when the panel must survive one common cleanup route.
How to Judge Whether MRM Is the Next Assay
The useful test is whether the study already has a closed peptide list, a repeated-sample design, and a plan to validate transitions in the same matrix that will be quantified.
Study condition |
MRM is often useful |
Reassess before locking the method |
|---|---|---|
Target definition |
Protein or peptide list is already closed |
Proteins of interest are still unknown |
Cohort design |
The same assay must be repeated across many samples |
The method would be built for a one-time look |
Interference control |
Transitions can be validated in the study matrix |
Co-eluting ions remain unresolved at unit resolution |
Quantification claim |
Same-peptide relative comparison across samples |
Amount or concentration without a matched SIS |
Panel size |
Scheduled transitions fit the chromatographic peak width |
So many ion pairs that dwell time collapses |
Adjacent option |
Mature triple-quadrupole panel |
Small panel in a complex digest that needs full MS2 confirmation |
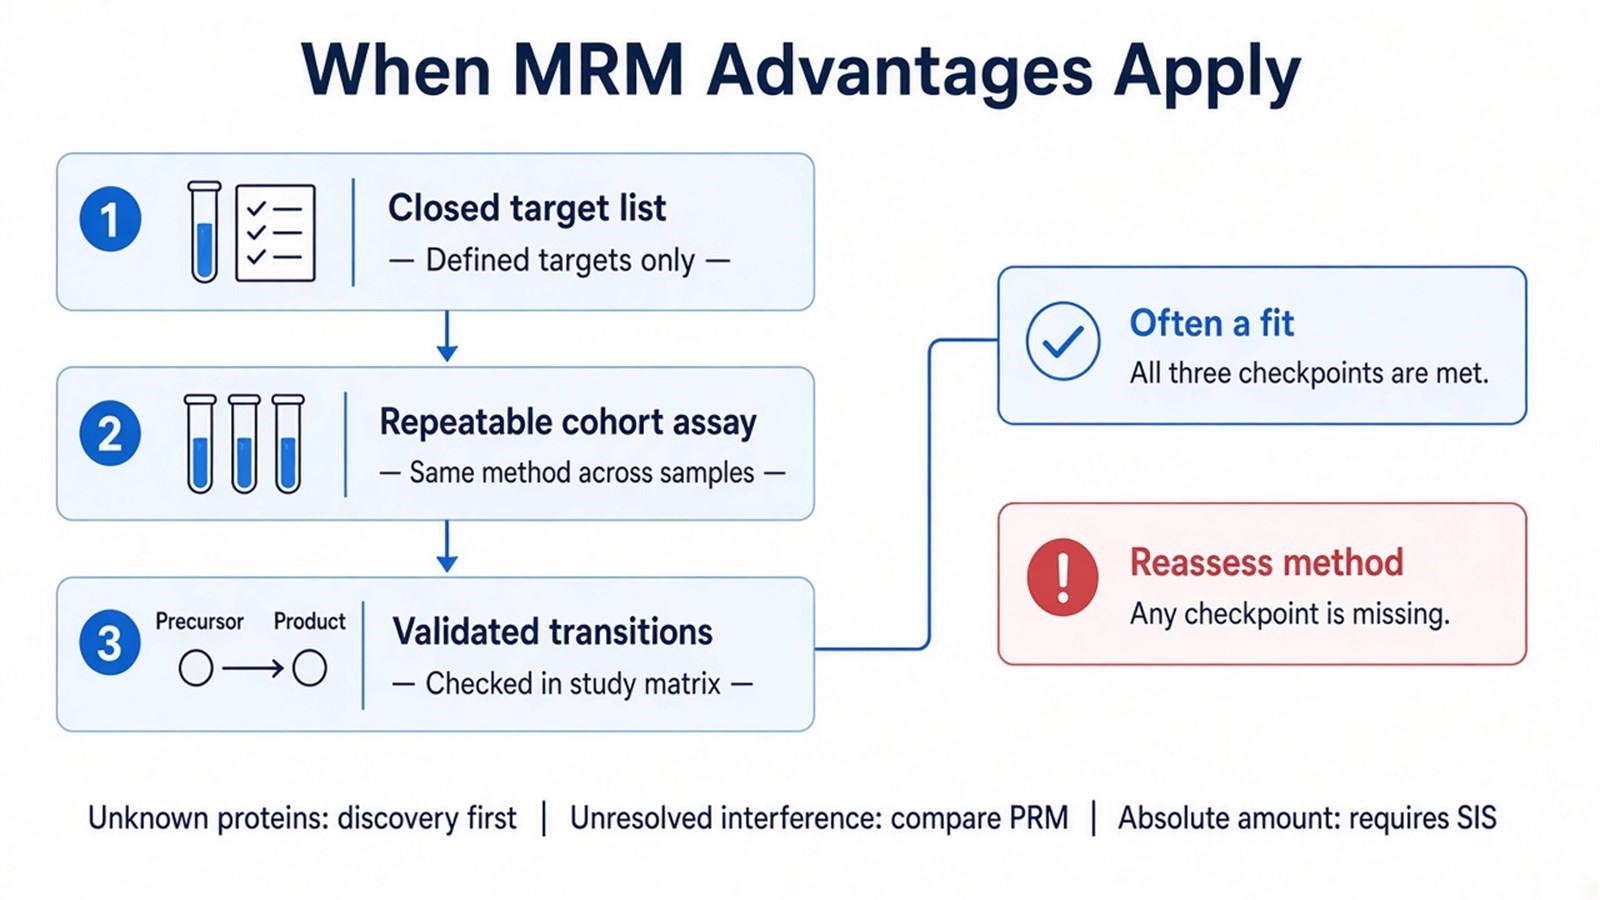
Figure 2. MRM advantages apply when the target list is closed, the assay will be repeated, and transitions have been validated in the study matrix.
When PRM or another targeted route should be compared
PRM isolates a precursor and records a high-resolution fragment-ion spectrum, so product ions do not have to be preselected with the same rigidity as MRM transitions. Peterson et al. 2012 argued that this design can reduce method-development effort and improve discrimination of co-isolated background. Shi et al. 2016 placed PRM as a high-resolution alternative for complex samples, while SRM on a triple quadrupole remains the conventional route for large, scheduled panels. In MtoZ Biolabs targeted proteomics offerings, PRM and MRM/SRM are separate acquisition choices.
A small candidate set in a difficult matrix often benefits from PRM during assay development, because fragment confirmation is available in the same run. Researchers can refer to Advantages and Disadvantages of MRM/PRM Techniques for a more detailed comparison of those acquisition routes. A large panel that will be run many times on a triple quadrupole can still justify the extra transition work of MRM after the ion pairs are clean. Discovery acquisition answers a different question.
What an MRM Result Can Support
A validated MRM peak area supports comparison of the same peptide across samples acquired with the same method. With an SIS, the light-to-heavy ratio can support an estimate of peptide amount, and protein amount can be inferred if digestion is complete enough for the chosen surrogate. Shared peptides, missed cleavage, variable modifications, and contaminated transitions weaken that inference. Peak area is not a copy-number ranking across different proteins, because peptides differ in digestion recovery and ionization.
Quality review should inspect peak shape, retention time, qualifier ratios, SIS recovery when a heavy peptide is present, and whether the calibrator or SIS amount sat in a linear range. A missing peak with a detectable SIS can support the statement that the native peptide was below the assay limit in that injection. A missing peak without a heavy counterpart can reflect a failed injection, a scheduling miss, or true absence. Batch effects still exist: digestion batches, cartridge lots, and column age can shift peak areas even when transitions are correct.
A validated transition supports same-peptide comparison across samples in one assay.
Researchers who already have a closed protein or peptide list can also review the MtoZ Biolabs Multiple Reaction Monitoring Analysis Service for project-specific feasibility and analysis planning. When the remaining decision is whether MRM or PRM better matches target number and matrix complexity, researchers can also review the Targeted Proteomics Service for project-specific routing guidance.
Frequently Asked Questions
1. Is MRM the same method as SRM?
SRM names one precursor-to-product transition. MRM names the same triple-quadrupole experiment when several transitions are recorded in one run. Lange et al. 2008 noted that IUPAC prefers SRM, while proteomics papers still use MRM for multiplexed assays. In practice, a laboratory asking about MRM advantages is asking about scheduled, multiplexed SRM on a triple quadrupole.
2. Does every MRM assay require isotope-labeled standards?
Relative comparison of the same peptide across samples can proceed without an SIS if digestion, loading, and chromatography remain comparable. Amount or concentration claims need a matched heavy peptide or protein standard. The standard should be added at a point that matches the claim: after digestion for peptide-level correction, or earlier if digestion recovery is part of what must be controlled.
3. Does a published MRM method transfer to a new sample matrix?
A published transition list is a starting point, not a finished assay in a new matrix. Retention time, interference, and peptide detectability can change with digestion conditions, sample type, and chromatography. The method still needs confirmation in the study matrix before cohort samples are queued.
4. Can MRM confirm candidates from a discovery proteomics experiment?
MRM can confirm nominated peptides if those peptides are detectable, unique enough, and distinguishable from interference in the confirmation matrix. The discovery fold-change does not transfer automatically. A new digest, a new chromatographic method, and a different quantitative signal mean the confirmation experiment has to be designed on its own, including which peptide represents the protein and what would count as agreement.
5. Why can an intense MRM peak still be a poor quantitative result?
Intensity reports ion current for the monitored transition, including any co-eluting interferent that passed both filters. A tall peak with a shoulder, a shifted retention time, a qualifier ratio that disagrees with the reference, or a light peak that does not overlay its SIS is weak evidence. Peak quality in the matrix matters more than raw height.
MtoZ Biolabs can provide multiple reaction monitoring and targeted proteomics analysis for predefined protein or peptide panels, including relative quantification and isotope-assisted absolute quantification when the study design requires matched internal standards.
If further evaluation of the target list, sample matrix, interference risk, or PRM versus MRM acquisition choice is needed, project details can be submitted for a feasibility review.
