



Optimizing immunoprecipitation Co-IP technology asks which proteins remain with an endogenous bait after antibody capture, Input and IgG contrast, and either targeted detection or LC-MS/MS identification. Optimization means titrating antibody, wash, and elution against those gates, not copying a harsher recipe until the blot looks clean. The work fits cells or tissues when an IP-competent antibody exists and the claim is co-recovery, not binary direct binding (Hein et al. 2015). Prefer this path when a first Co-IP already recovered the bait. Reassess when the antibody is Western-only, Input and IgG are unmatched, or extra washing is used as a substitute for IgG.
Researchers who already have a first Co-IP or IP-MS result and a written interaction claim can also review the MtoZ Biolabs IP-Mass Spectrometry Service for project-specific optimization review.
Optimize the Written Claim, Not the Cleanest Blot
Optimization is readable only after the sentence is written. Confirm a named pair, discover unknown partners, or compare the same bait across conditions. Those three claims change which variable is allowed to move.
A named-pair claim can accept a tighter wash once bait recovery is shown. An open-partner claim should keep milder native chemistry so associated proteins can remain, then rank names against IgG. A condition comparison must keep antibody, lysis, wash, and elution fixed so the same protein can be compared across arms. Copying a harsher mix because another bait looked clean is not optimization.
How those choices are locked before the first titration is covered in Designing an Efficient Co-immunoprecipitation (Co-IP) Protocol. Identification depth in a Co-IP-style eluate is typically hundreds to about a thousand proteins, depending on the sample. Depth is not a sign that optimization worked.
The path fits cultured cells or tissues when the antibody is supplied by the study. It is a poor first path when tagging is required because no IP-competent antibody exists, or when the claim is reconstituted binding of a purified domain. Drug-target screens and disease-mechanism maps are different experiment classes. They are not completed by calling a cleaner blot an optimized interactome.
Optimization of immunoprecipitation Co-IP reports a more readable co-recovery contrast. It does not certify that two proteins contact each other.
Antibody Specificity Is the First Variable to Change
Antibody specificity is the first optimization gate. If the capturing reagent does not recover the intended native bait, later lysis tweaks and a longer LC run will not create a Co-IP result. Application-specific validation remains necessary because Western performance does not establish IP performance (Uhlen et al. 2016). How to qualify that reagent is covered in How to Choose the Right Antibody for Co-IP Assays?.
Change one antibody variable at a time: clone, lot, amount, or bead pairing. Record those values next to the sample set. Extra antibody that fills the eluate with chain signal makes both blot and IP-MS noisier. Raising lysate input only to compensate for a Western-only clone is not optimization. Empty-tag controls from AP-MS do not replace antibody documentation on this path.
If no IP-competent antibody exists, stop optimizing Co-IP and change handle. Tagged AP-MS is a different analysis class.
Input and IgG Must Stay Matched While You Tune
Input and IgG are the denominators of every titration. They are not optional after one wash looks cleaner.
Input is taken before capture. It shows whether the bait existed in the extract and provides a baseline for enrichment. A missing bait band or protein group is an expression or extraction problem if Input is empty. It is a capture or elution problem if Input is positive. Without a fresh Input after a lysis change, the new recipe cannot be scored.
IgG or bead controls must use the same lysate batch, lysis, wash, elution, and, for IP-MS, the same digest and search. Preclearing is not a substitute for IgG. Beads without antibody detect plastic or resin carryover. They do not replace an isotype arm. Increasing wash count on the bait IP only, or eluting IgG unlike the bait IP, makes the titration unreadable.
Optimization check |
What a pass looks like |
What the change cannot say |
|---|---|---|
Antibody specificity |
Intended bait recovered after the new clone or amount |
Partner names from an off-target capture |
Input |
Bait present before capture when expected |
A missing IP read as a missing interaction |
IgG or bead |
Same wash, elution, and digest as the bait IP |
Every remaining name is bait-specific |
Enrichment efficiency |
Bait higher in bait IP than in Input and IgG |
A longer wash as a finished interactome |
MS compatibility |
Eluate digestible after the planned cleanup |
Residual SDS or salt left in the digest |
A practical optimization path is:
- Write the co-recovery sentence the blot or MS table must support.
- Confirm native bait recovery versus Input on the current recipe.
- Change one variable and apply that same recipe to matched IgG.
- Keep the change only if bait enrichment improved or background fell without losing the bait.
- Keep a shortlist for an orthogonal pair assay.
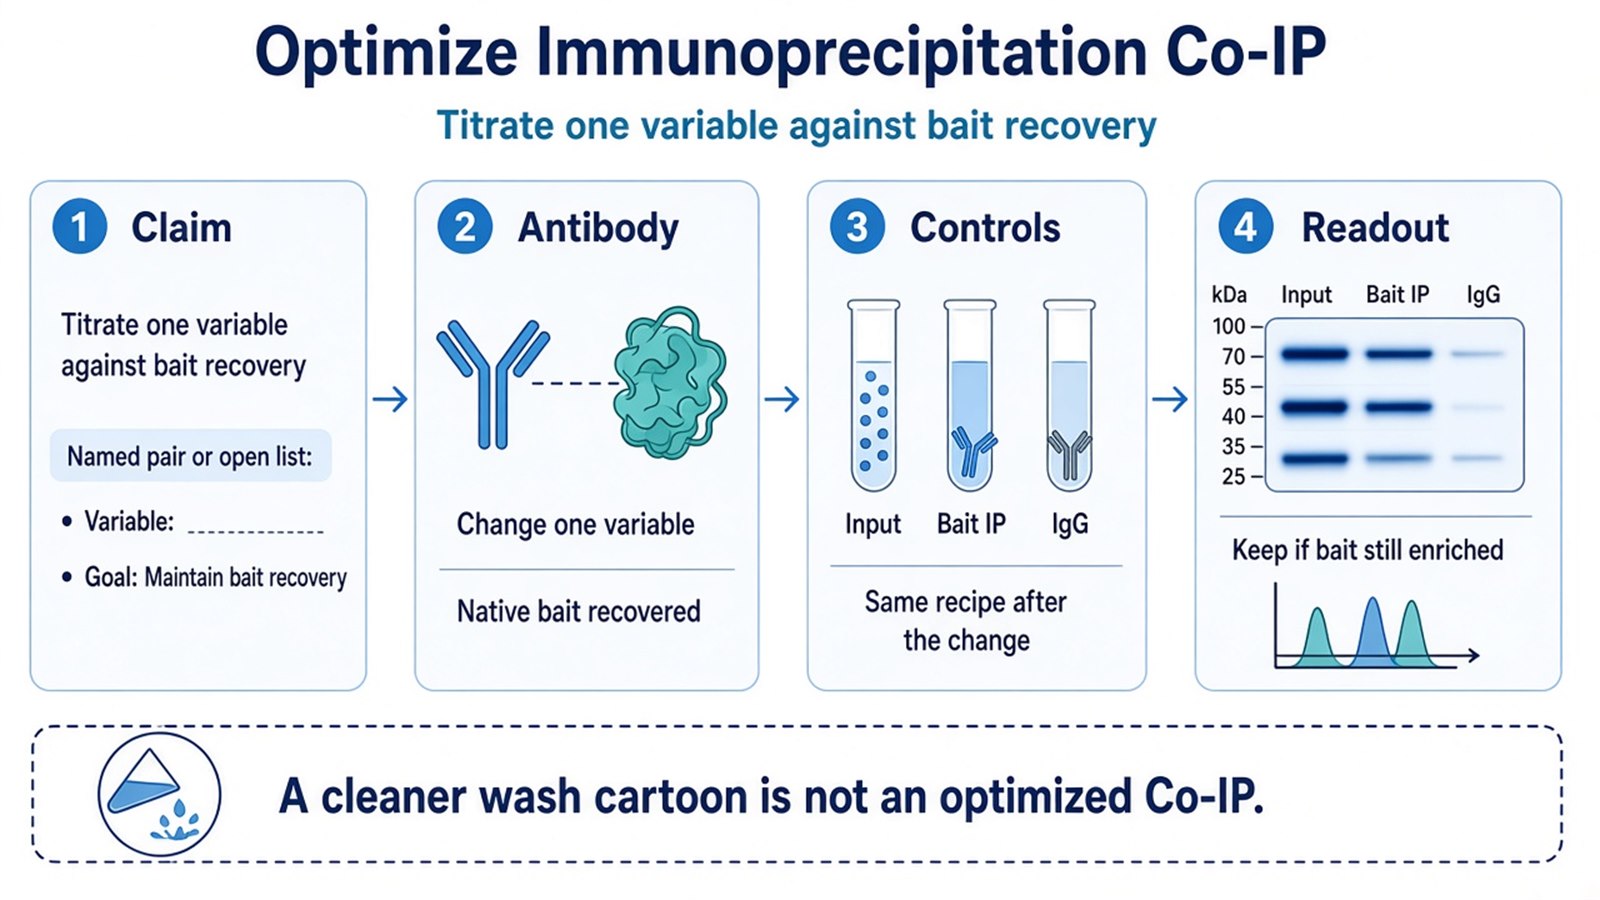
Figure 1. Optimizing Co-IP starts after bait recovery is shown. A cleaner wash cartoon is not an optimized result.
Enrichment Efficiency and Mass Spectrometry Credibility
Enrichment efficiency decides whether the last change worked before partners are interpreted. If the bait is weaker versus Input after a harsher wash, stop partner ranking and reverse that change. Relative intensity can compare the same protein across Input, IgG, and bait IP only when every arm used one recipe. Intensity does not rank unrelated proteins as tighter binders.
Acid elution, denaturing elution plus cleanup, or on-bead digestion can all finish an optimized IP-MS path when they leave a digestible, bait-positive sample. SDS-PAGE loading buffer does not go straight to LC-MS/MS. Residual SDS above a digestible level blocks trypsin. How matched arms stay comparable through digestion is covered in How to Prepare Immunoprecipitation Samples for Mass Spectrometry.
For IP-MS, identification of a protein is not a confidence call. Ranking requires bait recovery, Input-aware enrichment, and a declared contrast against IgG. Probabilistic scoring such as SAINT uses bait versus control counts to assign a probability that a prey is specifically co-purified (Choi et al. 2011). A contaminant catalog such as the CRAPome is a prior on control frequency (Mellacheruvu et al. 2013). Pathway software can annotate a shortlist. It cannot repair a missing IgG arm or convert a generic Co-IP cartoon into a validated interactome.
Optimized immunoprecipitation Co-IP supports candidate co-recovery after antibody, Input, IgG, and bait enrichment are shown. It does not certify a binary interactome.
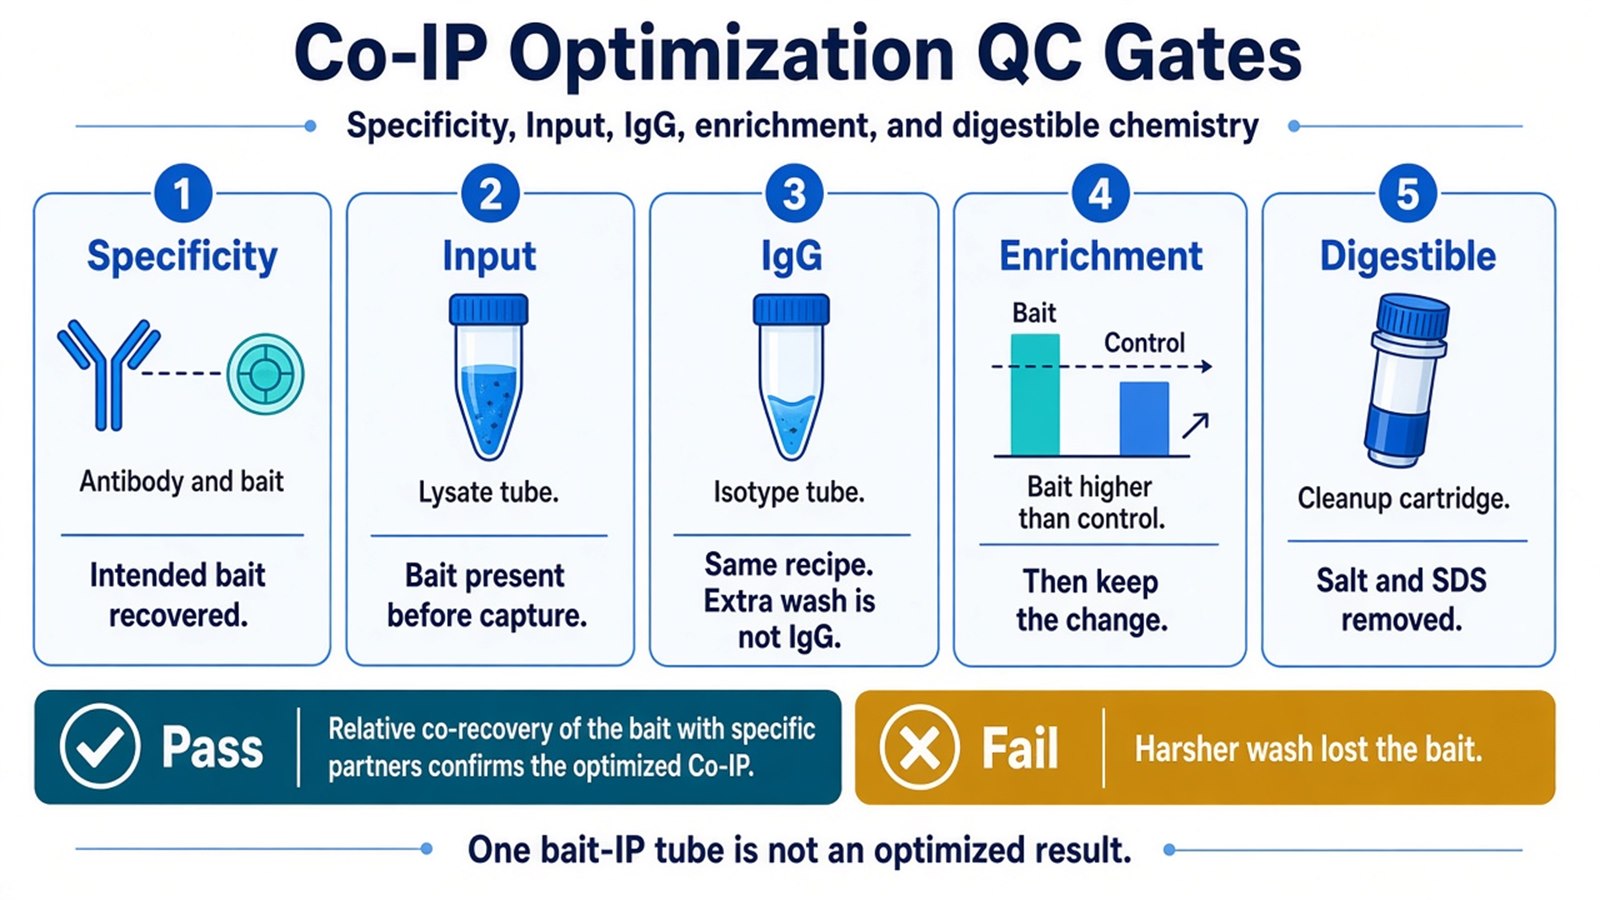
Figure 2. Optimization is a path outcome. Extra washing does not replace Input and IgG.
Common Optimization Biases and Result Boundaries
The common bias is writing Co-IP as highly specific because an antibody was used, then treating more washes as the optimization. Another bias is adding mass spectrometry to an unmatched IP and reading leftover names as a network. Over-washing can erase the association the claim needed. Under-washing, or a Western elution left in the tube, makes bait IP and IgG similar or undigestible.
Do not write a successful bait IP as proof that remaining proteins contact the bait. Do not write a negative table after a harsher wash as proof that no interaction exists in the cell. Absence can mean the wrong antibody, epitope masking, over-washing, or a bait that is not expressed in the tested state.
Prefer this optimization path when native bait recovery can be shown and every arm matches. Prefer tagged AP-MS when no IP-competent antibody exists. Prefer a defined-pair assay when the next sentence is direct contact.
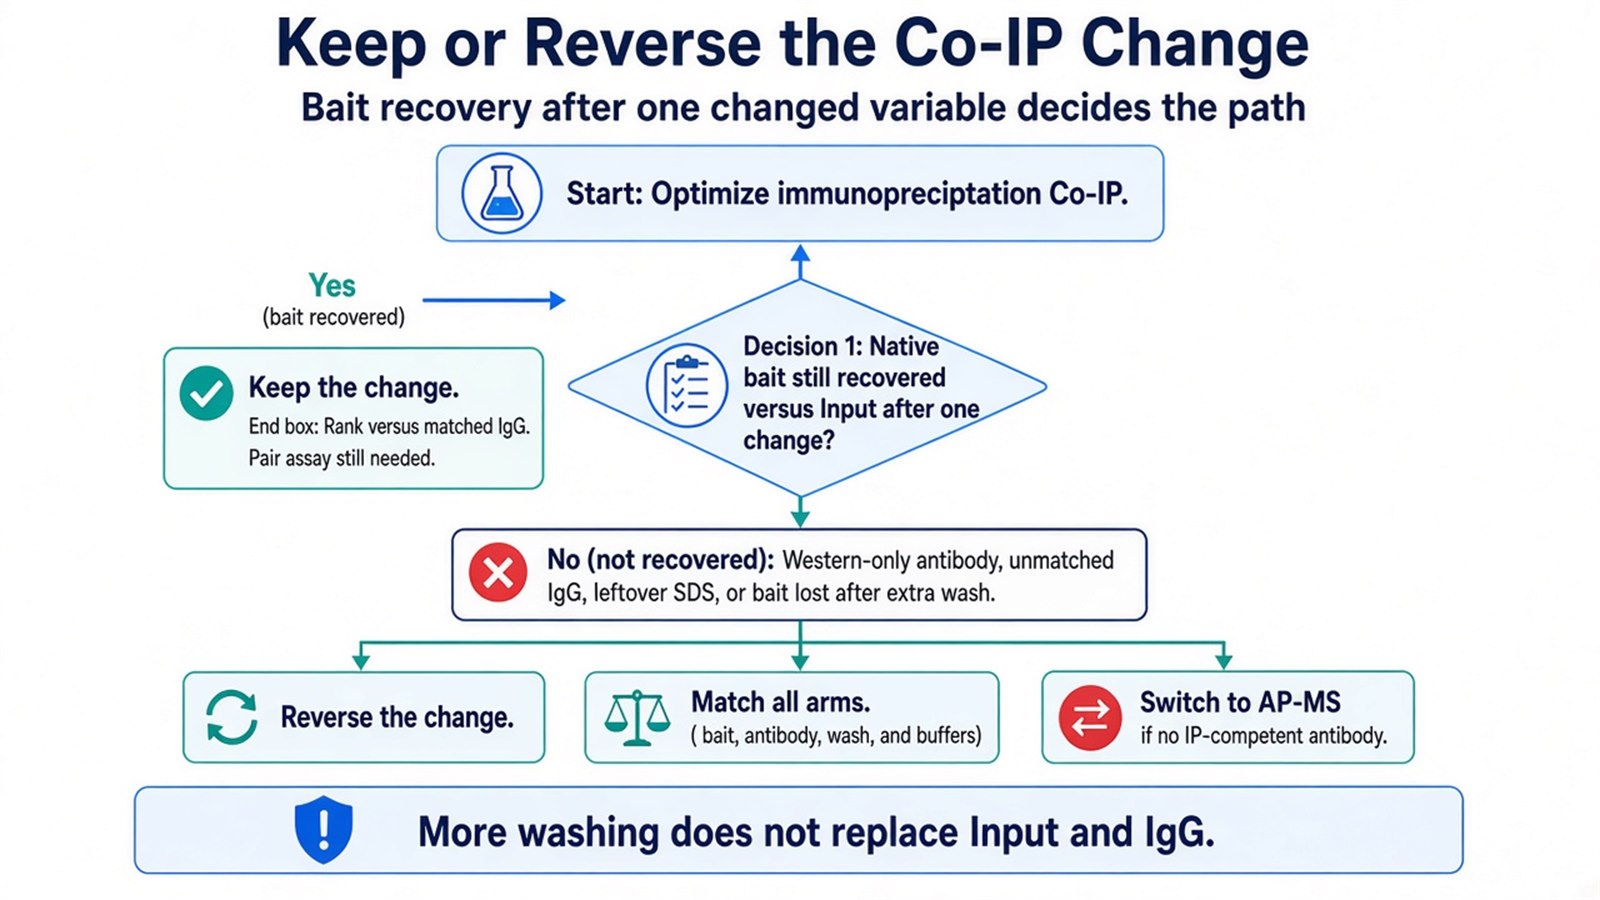
Figure 3. Keep an optimization only while native bait recovery is documented. A pathway map is not a completed Co-IP tune.
Once the claim, antibody status, matched Input and IgG, and the planned blot or digest are defined, the project can be scoped as an optimized Co-IP or IP-MS path. Researchers can review the MtoZ Biolabs IP-Mass Spectrometry Service for sample evaluation and path planning.
Frequently Asked Questions
1. Does more washing optimize Co-IP?
No. Extra washes on the bait IP only make enrichment unreadable. Keep IgG on the same recipe and stop if the bait falls.
2. Can I optimize by adding more antibody or lysate?
Not as a first move. Titrate amount only after the clone recovers the intended bait. Extra antibody often raises chain background.
3. Is a beads-only tube enough as the negative control?
No. Beads-only detects resin carryover. IgG must see the same wash, elution, and digest as the bait IP.
4. Does adding mass spectrometry confirm the interaction?
No. MS opens the eluate. Ranking still requires bait recovery and matched IgG.
5. When should optimization move to a pair assay?
Move when the next written sentence is direct contact or affinity. Keep the optimized shortlist as the input to that assay, not as the proof.
Reference
- M. Uhlen, A. Bandrowski, S. Carr, A. Edwards, J. Ellenberg, E. Lundberg, D. L. Rimm, H. Rodriguez, T. Hiltke, M. Snyder, T. Yamamoto (2016). A proposal for validation of antibodies. Nat. Methods, 13, 823-827. https://doi.org/10.1038/nmeth.3995
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- H. Choi, B. Larsen, Z.Y. Lin, A. Breitkreutz, D. Mellacheruvu, D. Fermin, Z.S. Qin, M. Tyers, A.C. Gingras, A.I. Nesvizhskii (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
