Advantages and Disadvantages of SILAC-Based Co-IP-MS in Protein Interaction Analysis
SILAC-based Co-IP-MS is the right quantitative method when the cells divide in culture, the two states name one contrast, and the next table must be a heavy-to-light (H/L) ratio on a Co-IP pellet. It is the wrong first method when the sample is tissue or a non-dividing primary culture, when more than two or three conditions must share one run, or when a nominated Western already answers the pair. The real advantages of combining stable isotope labeling by amino acids in cell culture (SILAC) with co-immunoprecipitation–mass spectrometry (Co-IP-MS) are chemically matched peptide pairs in one acquisition and, if lysates are mixed before one IP, a route that sends post-mix bead binders toward ratio 1. The real disadvantages sit in the same chemistry: days of labeling, an antibody-gated capture, a two- or three-state ceiling, arginine-to-proline conversion, and no culture SILAC on tissue. Precise quantification, non-invasive physiology, and high specificity are not properties of the combination. They are claims that have to be earned by incorporation, the mix point, and a matched Co-IP control.
A page that lists those slogans as inherent strengths, then lists antibody quality as the only operational cost, misleads the choice against TMT, label-free quantification (LFQ), or unlabeled Co-IP.
Researchers who already have a proliferating line, an IP-grade antibody or tag, and a written light–heavy contrast can also review the MtoZ Biolabs Co-IP-MS Protein Interaction Analysis Service for project-specific feasibility and analysis planning.
Advantages That Hold Only Under Stated Conditions
These points are useful when the condition in the same sentence is true. They are not built-in virtues.
Relative quantification in one acquisition, if the pair is complete
Light and heavy peptides share chromatography and ionization. After the mix point, later handling is common to both channels. That is an advantage over two unlabeled IPs run on different days. It is relative H/L, not absolute copies. Incomplete labeling compresses every ratio toward 1 and removes the advantage. For how that ratio is read as specificity versus occupancy, Quantitative Analysis of Protein-Protein Interactions Using SILAC and MS provides a complementary discussion.
Mix-before-IP contaminant filtering, if that mix point is used
When lysates are combined before one antibody capture, proteins that bind beads or Fc after mixing occupy both channels and approach H/L ≈ 1. Partners already on the bait in one state keep an unbalanced ratio. That filtering is a SILAC-Co-IP-MS advantage. It disappears if the two IPs are finished in separate tubes and mixed only at elution. For why the mix point is the principle decision, Principle of SILAC-Based Co-IP-MS in Protein Interaction Analysis provides a complementary principle discussion.
Metabolic label without a chemical tag, not a native-cell snapshot
Lysine and arginine enter protein during translation. There is no later TMT NHS chemistry on the peptide. That is “non-invasive” only relative to chemical labeling. Dialyzed serum, label-free medium, and five to eight doublings are still interventions. They do not return a true physiological interactome.
Cultured, dividing systems only
Mammalian lines, yeast, and some microbes that grow in defined medium can be labeled. That is the applicability, not “nearly all biological systems.” Drug-target and pathway language still needs a written bait and contrast.
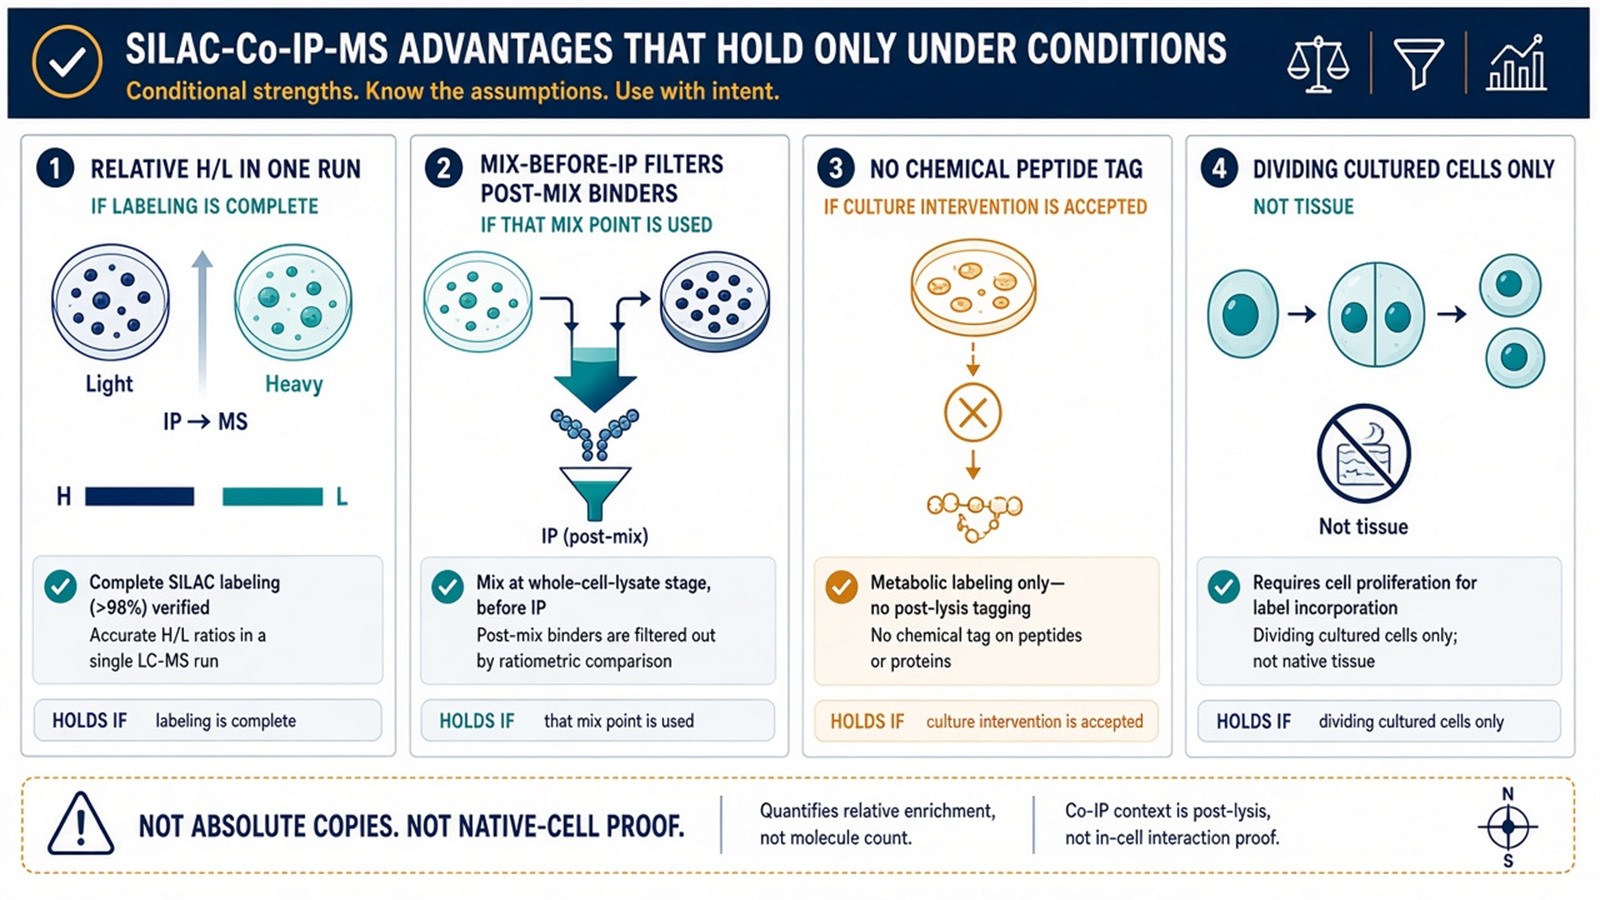
Figure 1. Matched peptide pairs and mix-before-IP filtering are advantages only under those conditions. They are not automatic precision or native-cell specificity.
Write the condition next to each advantage. An advantage without a condition is a slogan.
Disadvantages That Are Structural
These limits remain after a careful protocol. They are not beginner errors.
Days of labeling
Incorporation needs multiple doublings. Slow lines stretch the calendar. Medium must stay free of unlabeled Lys and Arg. That time cost is why a nominated pair is often cheaper as unlabeled Co-IP plus Western.
Antibody gate is unchanged
SILAC does not repair a blot-only antibody, an unmatched isotype, or a tag that collects handle binders. The Co-IP disadvantages remain: lysis is not the cell, the pellet is not a direct contact, and occupancy still hides scarce prey. For those unlabeled limits, Advantages and Disadvantages of Co-Immunoprecipitation in Protein Interaction Analysis provides a complementary choice discussion.
Two or three states, not a multiplex panel
Classic SILAC compares two channels; a medium pair adds a third. A dose series or a five-arm panel is TMT or a set of LFQ runs. Calling data analysis “too complex” is the weaker complaint. MaxQuant-class H/L tables are routine. The real analysis cost is incomplete label, arginine-to-proline satellites, and a mix point that was never written.
Tissues and most primary samples are out
Culture SILAC does not label a biopsy or a non-dividing isolate. Spike-in SILAC or another label is a different design. Input amount still follows bait occupancy; SILAC does not lower the protein needed for a detectable IP.
Heavy amino-acid cost
Lys8 and Arg10 add reagent cost that LFQ does not. The cost is justified when the mix-before-IP filter or the one-run pair is the reason for choosing SILAC. It is not justified as a default upgrade on every Co-IP-MS.
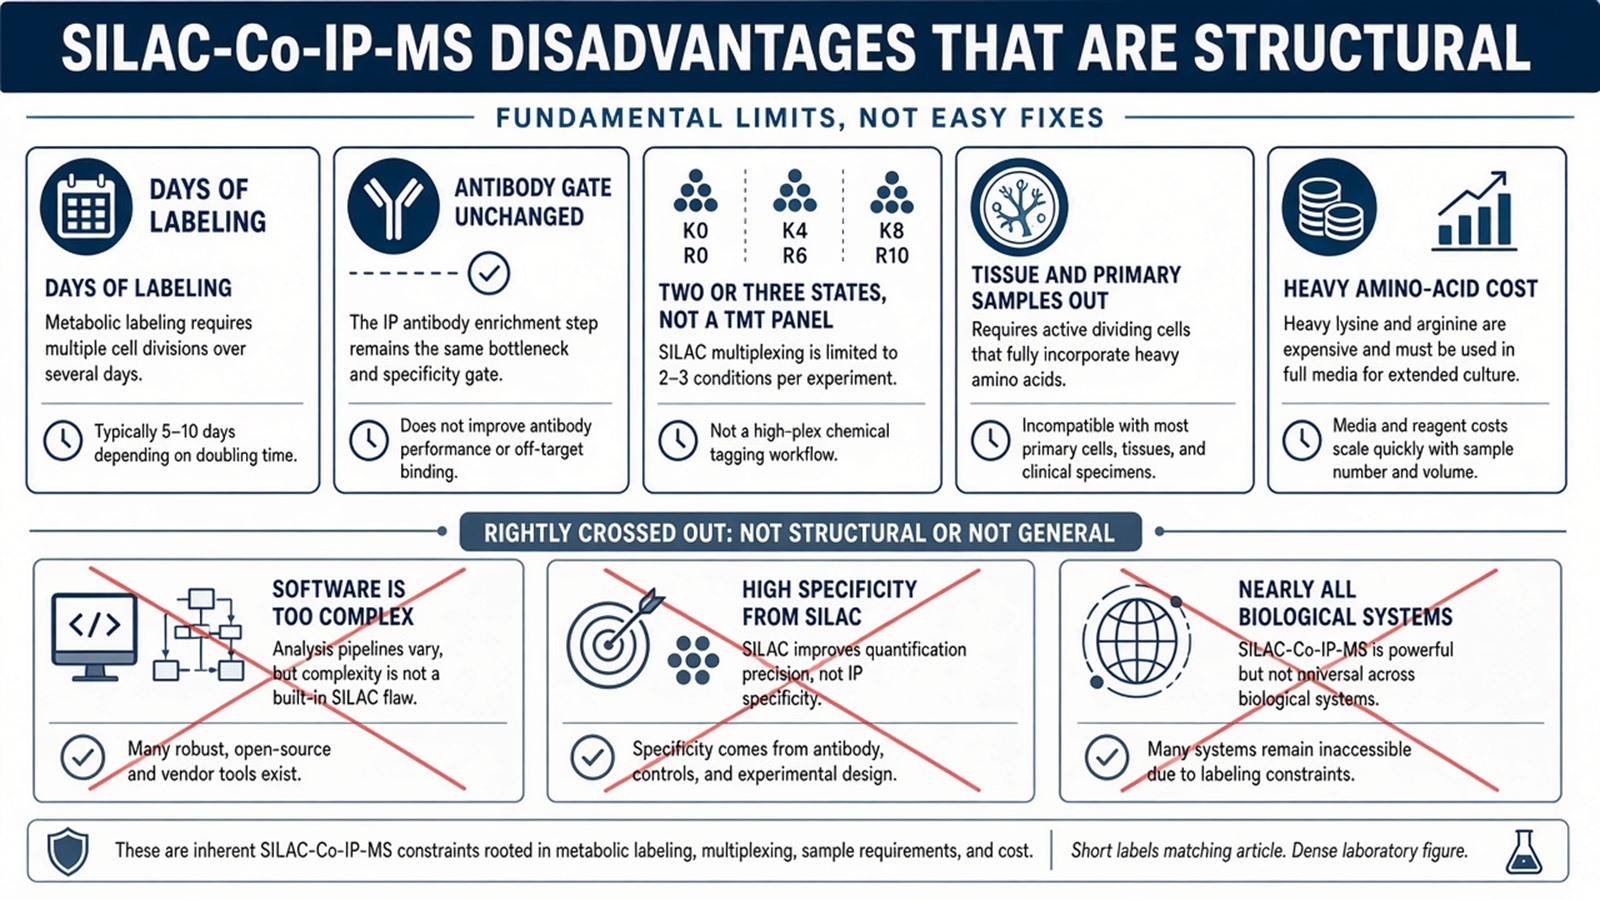
Figure 2. Time, antibody dependence, a two- or three-state ceiling, and sample class are structural. Software complexity is not the main limit.
When Another Method Is the Better First Choice
Unlabeled Co-IP plus Western is enough for a nominated pair with a good antibody.
LFQ Co-IP-MS fits tissues, primary cells, and designs that cannot wait for incorporation.
TMT or iTRAQ fits many conditions in one peptide-level multiplex when chemical labeling is acceptable.
In-cell crosslinking MS or proximity labeling is required when the claim is a residue or a neighborhood that does not survive lysis.
SILAC-Co-IP-MS remains useful after those methods as a ratio check in a labeled line. It should not be asked to replace them.
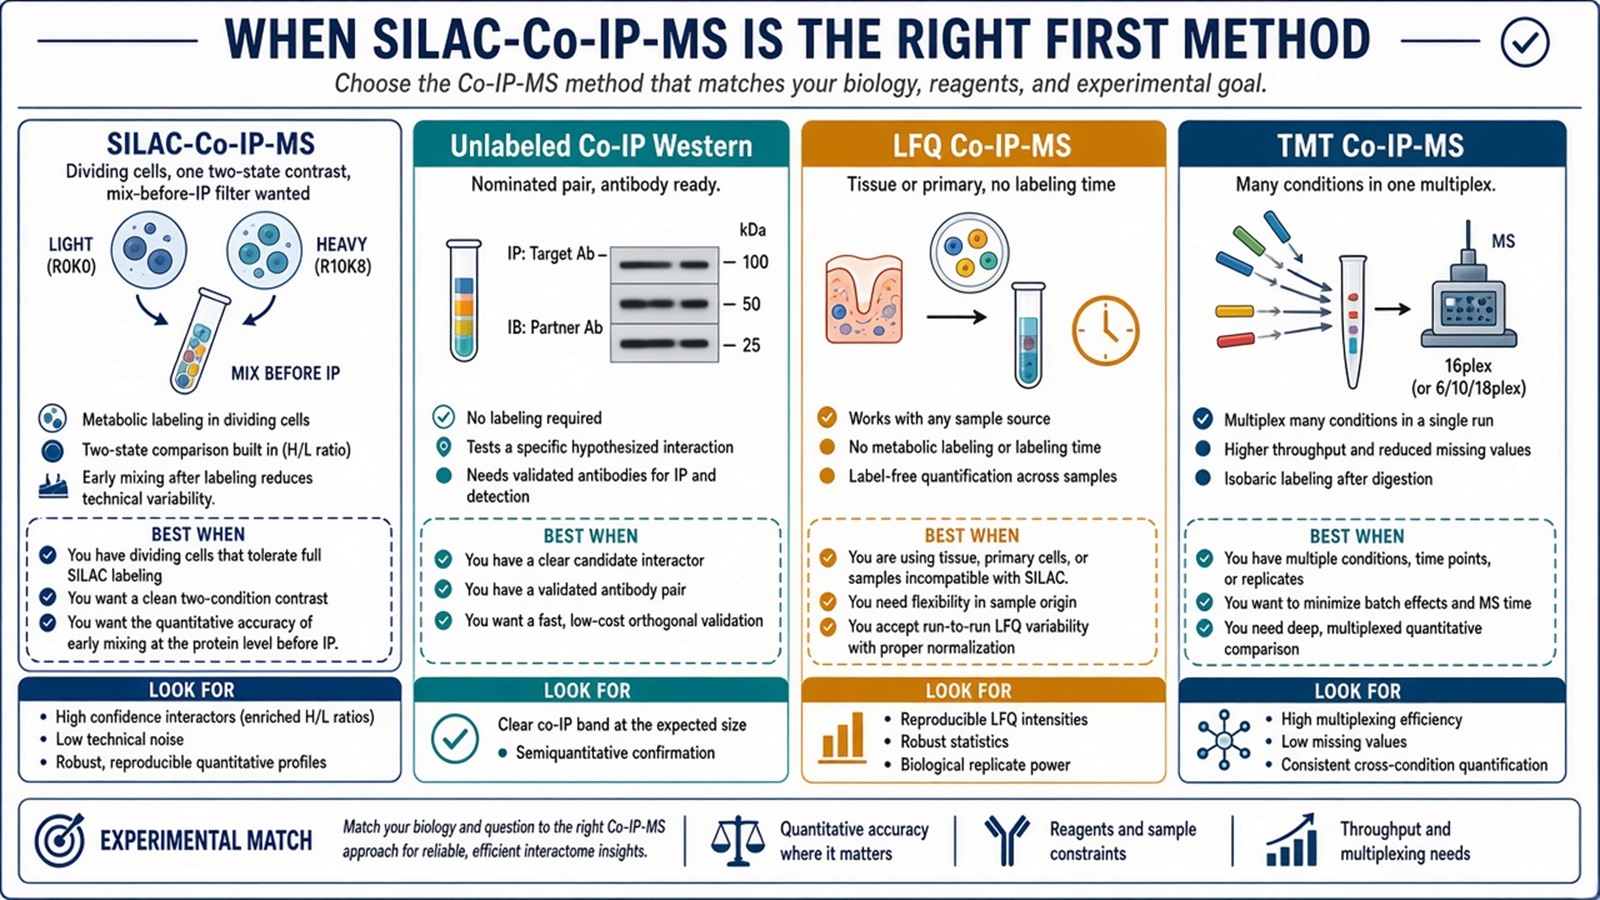
Figure 3. Use SILAC-Co-IP-MS when cells divide and the claim is one H/L contrast on a Co-IP pellet. Use another quantitative or unlabeled route when the sample or the panel does not fit.
Common Mis-statements
“Precise quantification” is a complete pair after a written mix point. It is not absolute abundance.
“Non-invasive labeling” means no NHS tag on the peptide. It does not mean the culture is unperturbed.
“High specificity and sensitivity” are Co-IP and MS properties. SILAC only pairs the channels.
“Applicable to nearly all biological systems” excludes tissue and most primary samples.
“Complex data analysis” overstates the table and understates incomplete labeling.
Experimental Limits
Advantages and disadvantages are the same mass pair read from two sides. Three biological replicates do not repair a 70% heavy channel. Diagnostic language sits outside ordinary research reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable plan names the claim first, then asks whether SILAC-Co-IP-MS advantages apply and whether its structural limits are acceptable. MtoZ Biolabs scopes labeled Co-IP-MS on Orbitrap-class LC-MS/MS when the cells divide, the contrast is written, and the mix point is chosen. Interpretability still follows those items, not the instrument name.
Once those limits are acceptable, the project can be bait-versus-control or treated-versus-control Co-IP-MS. Researchers who need a quantitative interactome plan after that decision can review the MtoZ Biolabs Protein-Protein Interaction Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What is the main advantage of SILAC-based Co-IP-MS?
A relative H/L on chemically matched peptides in one run, and mix-before-IP filtering of post-mix contaminants, when labeling is complete and the cells divide.
2. What is the main disadvantage?
The assay still needs an IP-grade antibody, days of incorporation, and a sample that can be labeled in culture. Tissue and large condition panels do not fit.
3. Does SILAC make Co-IP more specific?
No. It pairs two states. Specificity still follows the antibody, the wash, and the mix point.
4. Is SILAC better than TMT for every Co-IP-MS?
No. SILAC wins on a two-state cultured contrast and on mix-before-IP filtering. TMT wins when many arms must share one multiplex.
5. Can I skip SILAC and still quantify a Co-IP?
Yes. LFQ compares separate runs. A nominated Western compares two blots. Those routes trade the one-run pair for simpler sample class.
Reference
- Ong S.E., Blagoev B., Kratchmarova I., et al. (2002). Stable isotope labeling by amino acids in cell culture, SILAC, as a simple and accurate approach to expression proteomics. Mol. Cell. Proteomics, 1, 376-386. https://doi.org/10.1074/mcp.M200025-MCP200
- Mann M. (2006). Functional and quantitative proteomics using SILAC. Nat. Rev. Mol. Cell Biol., 7, 952-958. https://doi.org/10.1038/nrm2067
- Trinkle-Mulcahy L., Boulon S., Lam Y.W., et al. (2008). Identifying specific protein interaction partners using quantitative mass spectrometry and bead proteomes. J. Cell Biol., 183, 223-239. https://doi.org/10.1083/jcb.200805092
- Vermeulen M., Hubner N.C., Mann M. (2008). High confidence determination of specific protein-protein interactions using quantitative mass spectrometry. Curr. Opin. Biotechnol., 19, 331-337. https://doi.org/10.1016/j.copbio.2008.06.001
- Tackett A.J., DeGrasse J.A., Sekedat M.D., et al. (2005). I-DIRT, a general method for distinguishing between specific and nonspecific protein interactions. J. Proteome Res., 4, 1752-1756. https://doi.org/10.1021/pr050168c
Support Documents
Quantitative Proteomics: Label-Based and Label-Free Strategies
Compare label-based and label-free quantitative proteomics strategies and explore their applications to help identify a suitable approach for your research.