



AQUA (Absolute QUAntification) turns a predefined proteotypic peptide into a concentration readout by pairing it with a stable isotope-labeled synthetic standard of known amount. Relative discovery can show that a protein rises or falls between groups. An AQUA workflow is used when the project must report how much of that peptide is present in declared units such as fmol, ng/mL, or ppm.
The workflow is a closed-list assay sequence rather than an open proteome survey. It links peptide selection, labeled-standard design, sample preparation and spike timing, targeted LC-MS/MS acquisition, calibration, and concentration reporting. Skipping any of those links often produces a light-to-heavy ratio that looks quantitative but cannot support the intended absolute claim. Researchers planning an AQUA project can review the MtoZ Biolabs Absolute Quantitative Analysis (AQUA) Service for standard strategy and reporting-unit scoping before labeled peptides are ordered.
Overview of the AQUA Workflow
A practical AQUA workflow moves from target definition to a concentration report through a fixed set of stages. Each stage constrains what the next stage can correct.
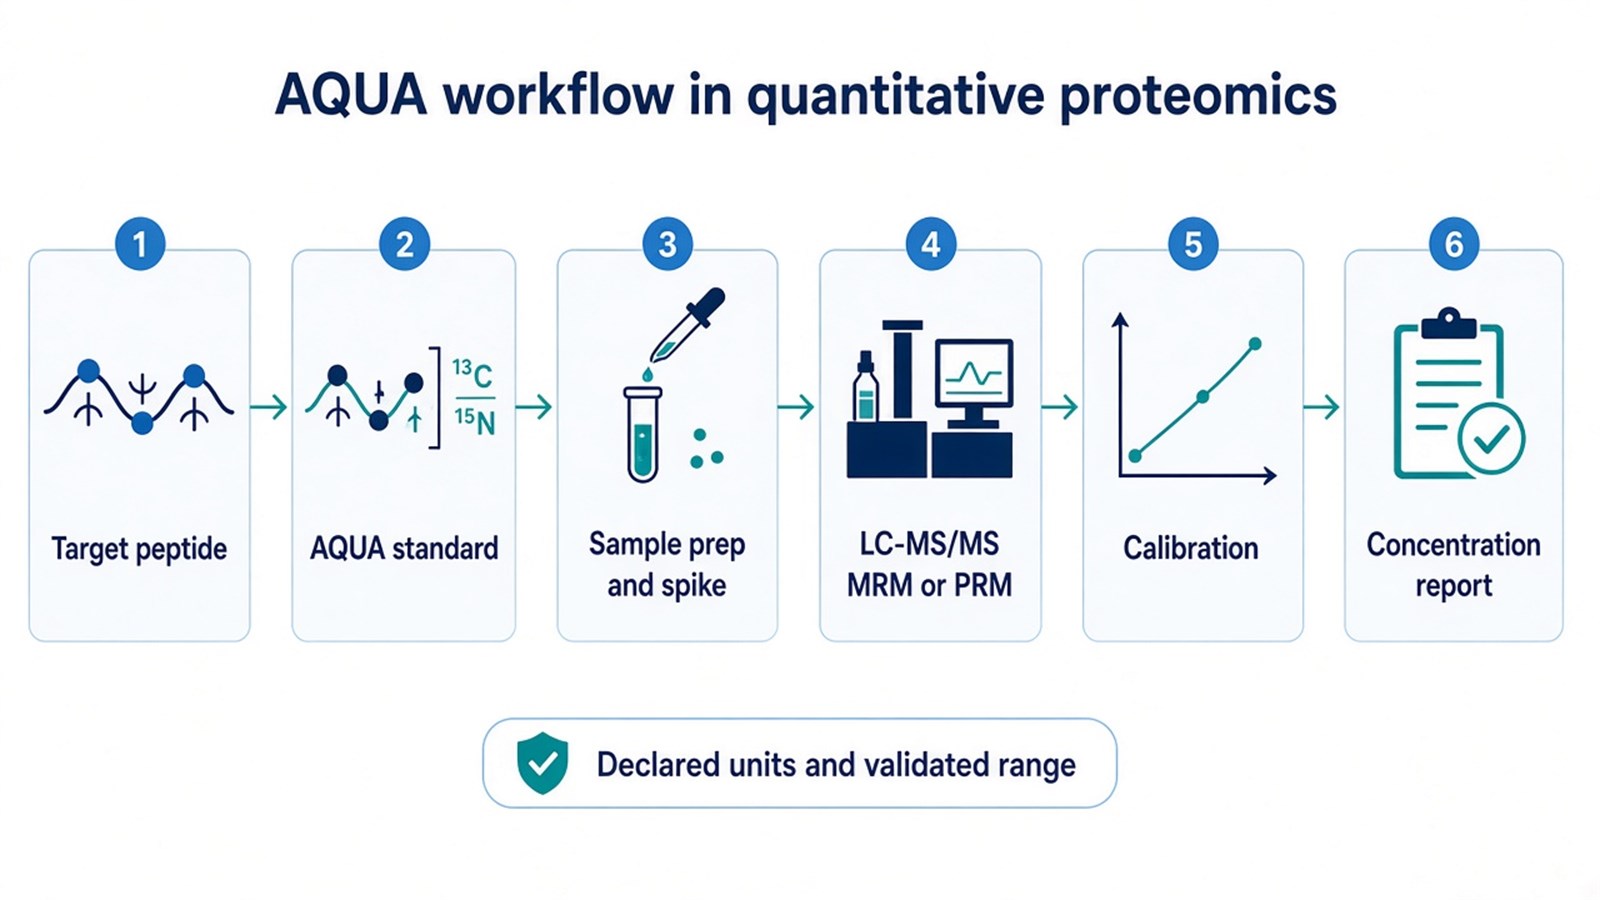
Figure 1. AQUA workflows convert a scheduled peptide and labeled standard into concentration only when calibration and reporting units are declared.
Typical stages include:
- Select proteotypic peptides for the closed target list.
- Synthesize or source AQUA standards matched to those sequences.
- Prepare samples and spike standards at a declared step.
- Acquire light and heavy signals by MRM/SRM or PRM.
- Apply a matrix-matched calibration model within a validated range.
- Report peptide concentration with QC flags for out-of-range samples.
The principle behind why these stages support an absolute claim is summarized in Principle of Absolute Quantitative Analysis.
Sample Preparation and Standard Addition
Sample preparation begins with protein extraction from the study matrix, followed by denaturation, reduction, and alkylation so disulfide bonds are opened and do not reform unpredictably. Proteins are then digested into peptides, commonly with trypsin or Trypsin/Lys-C, and cleaned for LC-MS/MS. Peptide recovery, digestion completeness, and matrix cleanup all influence the endogenous signal that will later be compared with the AQUA standard.
AQUA standards are synthetic peptides that match the target sequence but incorporate stable isotopes such as carbon-13 or nitrogen-15 at selected residues. They are designed to share retention time and ionization behavior with the endogenous peptide while remaining distinguishable by precursor or fragment m/z. The known amount of each standard is the reference that converts the light-to-heavy response into concentration.
Spike timing is part of the quantitative design, not a minor prep detail. A standard added after digestion can correct many post-digest handling and ionization effects, but it cannot reconstruct digestion yield lost earlier. A pre-digest spike can track more preparation steps for that peptide pair, yet still does not automatically equal intact-protein recovery unless the assay was designed for that claim. Protein-level absolute inference therefore requires additional assumptions beyond a post-digest peptide spike.
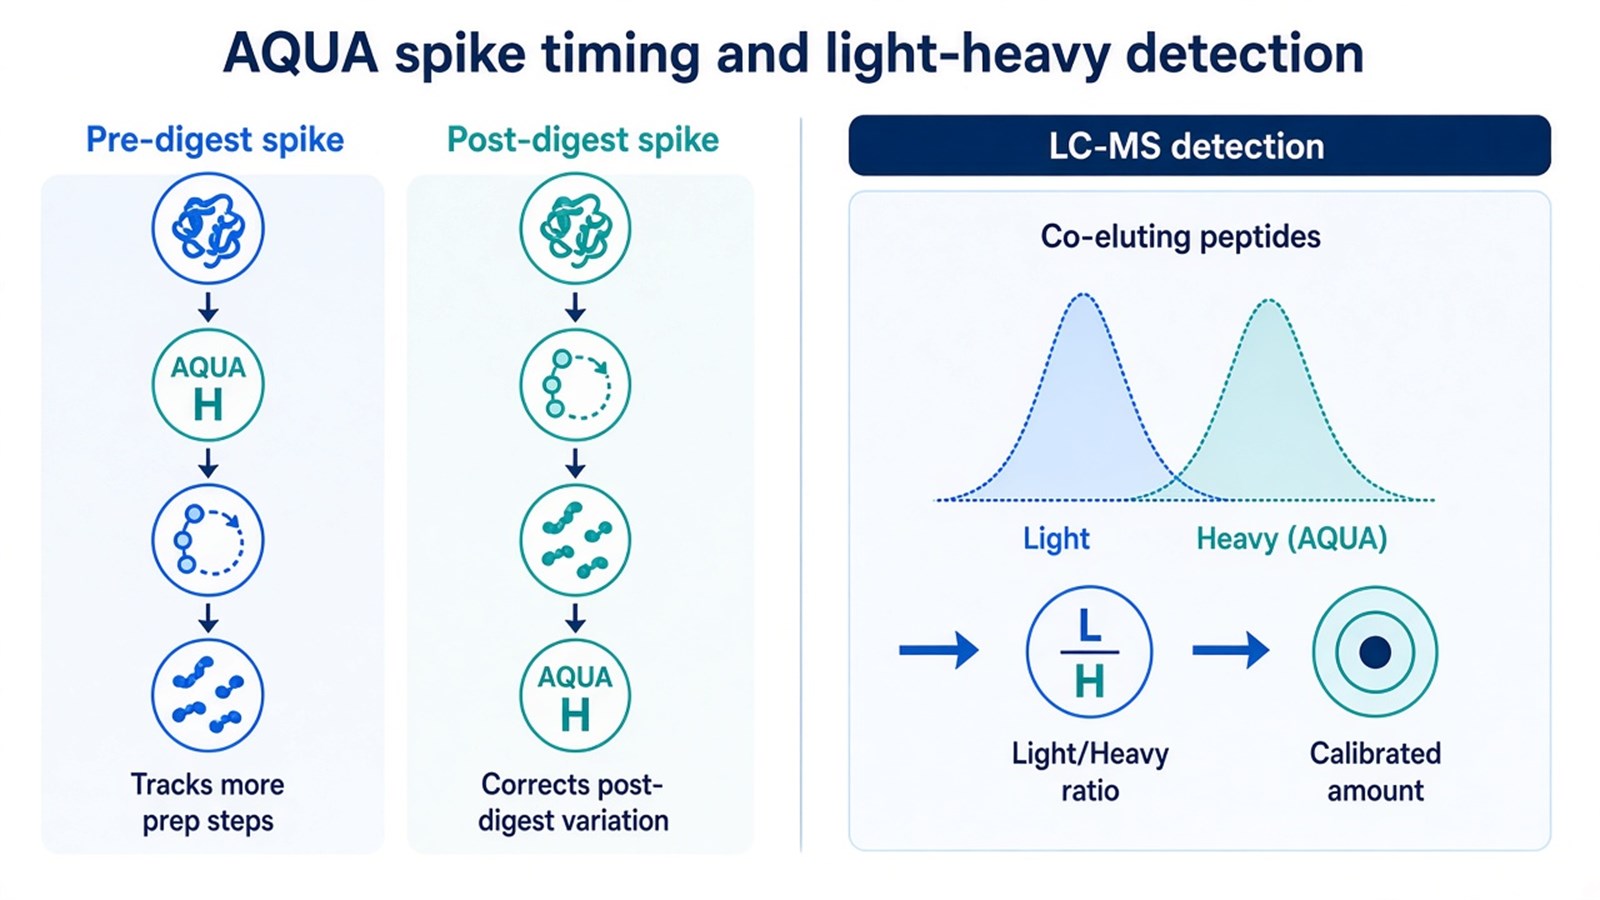
Figure 2. Pre-digest and post-digest AQUA spikes correct different parts of the workflow; both still require light-heavy detection and a declared calibration model.
Mass Spectrometry Analysis
Prepared samples are separated by liquid chromatography and measured by tandem mass spectrometry. In AQUA assays, acquisition is usually targeted rather than discovery-based. MRM/SRM on a triple-quadrupole platform monitors predefined precursor-to-product transitions for light and heavy peptides. PRM on a high-resolution platform isolates scheduled precursors and records product-ion spectra from which fragment chromatograms are extracted.
Because the AQUA peptide amount is known, the endogenous peptide amount is calculated from the paired response after a calibration model is applied. Raw peak-area comparison without a declared curve, matrix matching, and range check is not a complete absolute workflow. Complex matrices may require transition revision, retention-time scheduling, or a shift from MRM to PRM when interference limits selectivity.
Data Processing, Calibration, and Validation
After acquisition, targeted analysis software identifies light and heavy peptide signals, integrates peak areas, and relates the response to calibrators. Linear regression or another declared calibration model is used within the validated concentration interval. Background review, peak-shape checks, and replicate injections support reproducibility assessment, but they do not replace matrix-matched calibration.
Validation typically confirms that calibrators bracket expected sample concentrations, that QC recoveries fall inside agreed limits, and that samples outside the validated range are flagged rather than extrapolated as reportable amounts. Reporting units must match the preparation and normalization scheme used in the assay. A value labeled ng/mL is a different claim from fmol on column or ppm relative to total protein input.
Assay-design decisions that often decide whether calibration succeeds in the project matrix are covered in Design Absolute Protein Quantification Assay.
Reporting and Research Use
A complete AQUA report usually includes concentration tables for the closed peptide panel, calibration and QC summaries, method notes for standards and acquisition, and comments on surrogate-peptide assumptions. These outputs support biomarker candidate confirmation, host cell protein monitoring, drug-target stoichiometry questions, and biopharmaceutical specification comparison in research or CMC-supporting contexts.
Diagnostic cutoffs, commercial release decisions, and regulated clinical claims sit outside ordinary research AQUA reporting unless a separately validated assay framework is declared. Peptide-level concentration also does not automatically equal protein copy number when digestion is incomplete, modifications are unaccounted for, or shared peptides confuse protein inference.
Practical Limits Across the Workflow
Common failure points include ordering AQUA peptides before matrix peptide performance is reviewed, building buffer curves that do not transfer to plasma or lysate, mismatched spike timing for the intended claim, and treating relative PRM/MRM ratios as absolute values. Multiplex panels raise synthesis cost and review burden, so feasibility scoping before standard production usually reduces rework.
Conclusion
The workflow of AQUA in quantitative proteomics is a calibrated assay path: proteotypic peptide selection, labeled-standard design, declared spike timing, targeted LC-MS/MS, matrix-matched calibration, and concentration reporting inside a validated range. Each step exists to protect the absolute claim rather than to decorate a relative targeted method with a heavy peptide. When those links are declared and tested in the study matrix, AQUA can deliver amount units that relative fold-change tables cannot provide.
Researchers planning AQUA absolute quantification can contact MtoZ Biolabs to review targets, spike strategy, and matrix requirements, and can also consult the MtoZ Biolabs Absolute Quantitative Analysis (AQUA) Service for project-specific feasibility and standard planning.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
