



Many proteomics projects need an open discovery matrix first and fragment-level confirmation of a short peptide list later. DIA-PRM proteomics names that staged path. Data-independent acquisition (DIA) records scheduled-window fragment chromatograms for relative discovery across a broad inventory. Parallel reaction monitoring (PRM) then remeasures a closed precursor panel with high-resolution product-ion spectra. The mechanism is therefore sequential complementarity, not a single hybrid scan mode that is open and fully targeted at once.
Calling a project DIA-PRM does not convert DIA into a closed confirmation assay, and it does not convert PRM into proteome-wide surveying. Each stage keeps its own acquisition logic and quantitative claim. Researchers planning discovery followed by targeted confirmation can review the MtoZ Biolabs DIA-PRM Proteomics Service for staged feasibility and panel-scoping before the first cohort is queued.
What DIA-PRM Mechanism Means
DIA divides a predefined precursor mass range into consecutive isolation windows and fragments the ions in each window on a fixed schedule. Peptide quantities are recovered computationally from extracted fragment chromatograms and spectral-library or library-free matching. PRM isolates only scheduled precursors and stores full high-resolution MS2 for those targets, from which selected fragment chromatograms are integrated after acquisition.
The combined mechanism is a project handoff: DIA nominates candidates under an open relative-discovery claim; PRM confirms a shortlist under a closed-list claim. Principle-level framing of that sequence is available in Principle of DIA-PRM Proteomics.
DIA-PRM is a staged mechanism: open relative discovery, then closed-list confirmation.
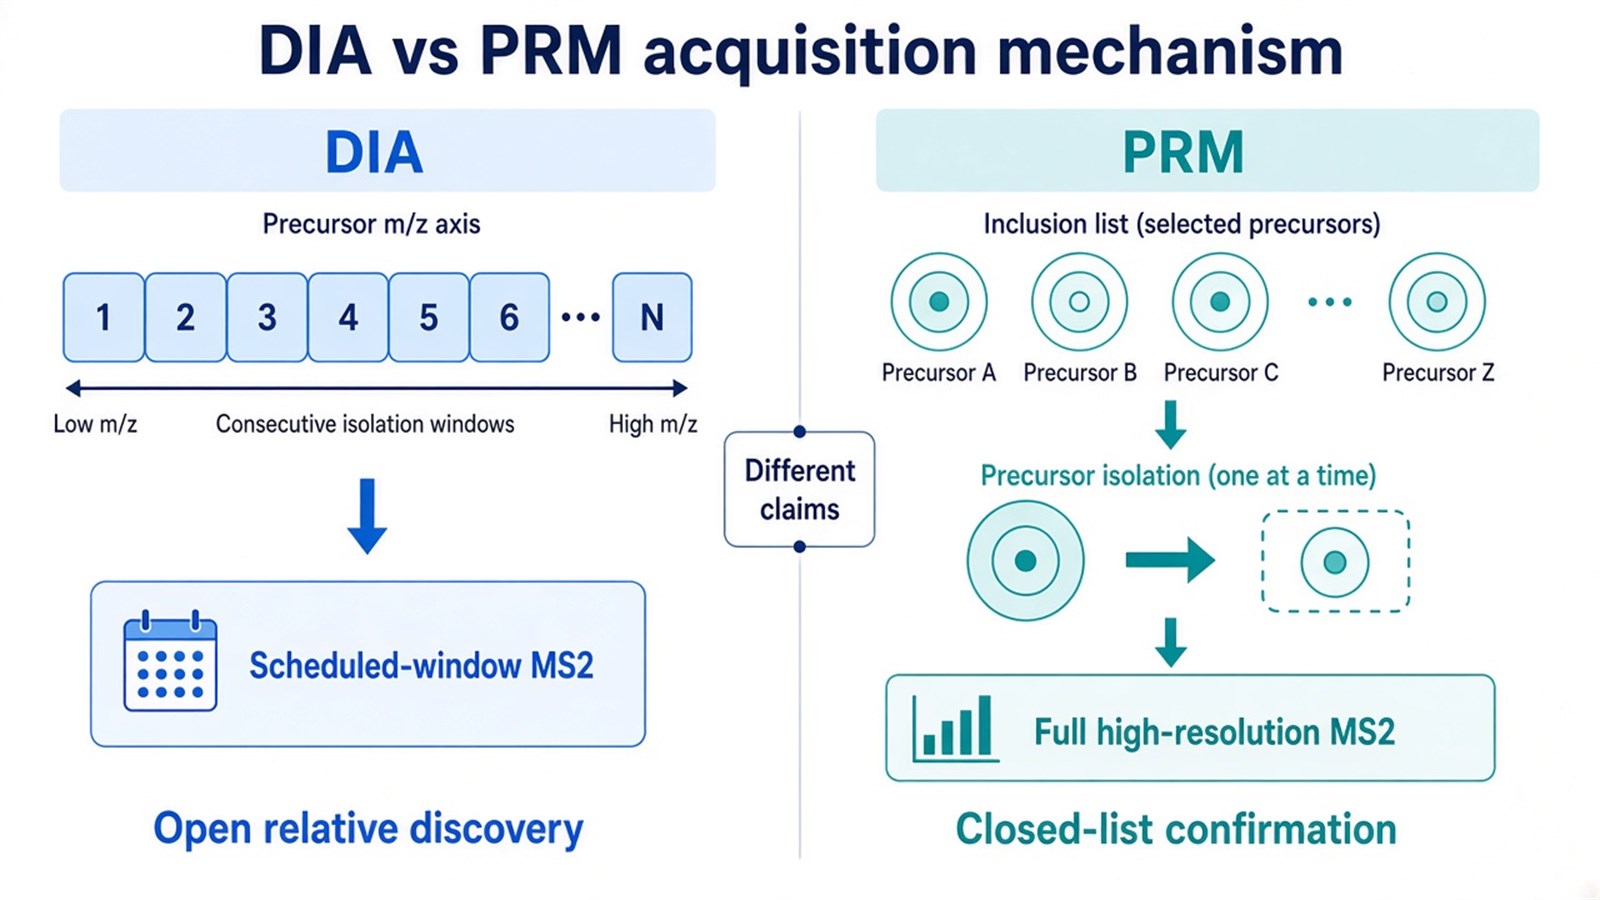
Figure 1. DIA and PRM use different acquisition mechanisms and support different quantitative claims.
Mechanism of the DIA Stage
1. Sample Preparation and Digestion
DIA analysis begins with protein extraction from the study matrix, followed by reduction, alkylation, and enzymatic digestion into peptides. Cleanup and input consistency matter because DIA recovers quantities from many co-fragmented ions; preparation bias at this stage propagates into the discovery matrix that later feeds the PRM shortlist.
2. Scheduled-Window Acquisition
Unlike data-dependent acquisition (DDA), which selects precursors stochastically by intensity ranking, DIA systematically covers the precursor range through consecutive windows. Within each window, precursor ions are fragmented together and the resulting product ions are recorded. This design improves measurement consistency across independent injections for comparative cohorts, but it also admits co-isolation interference inside a window.
3. Identification and Relative Quantification
DIA data volumes are large and require specialized processing. Software uses spectral libraries, predicted libraries, or library-free strategies to assign peptide evidence and extract fragment chromatograms for relative quantification. The default claim remains relative change of the same peptide or protein under matched preparation, not molar concentration.
Mechanism of the PRM Stage
1. Candidate Shortlist and Peptide Handoff
After DIA ranking, selected proteins are translated into proteotypic peptides for a closed inclusion list. Peptide choice, retention-time expectation, and matrix detectability determine whether confirmation can succeed. A protein significant in DIA may fail PRM if the wrong surrogate peptide is chosen or if preparation differs between stages.
2. Scheduled Precursor Isolation and Full MS2
PRM isolates each scheduled precursor and records a high-resolution product-ion spectrum. Quantification uses extracted fragment chromatograms rather than a pre-locked triple-quadrupole transition list. That mechanism supports interference review and fragment-level confirmation for the closed panel, within chromatographic peak width and MS2 cycle-time limits.
3. Optional Absolute Follow-Up
Ordinary PRM fragment areas remain relative unless stable isotope-labeled standards and a declared calibration model are introduced. Absolute concentration is an optional extension of the PRM stage, not an automatic property of DIA-PRM staging.
How the Two Mechanisms Are Integrated
A complete DIA-PRM project mechanism usually follows five linked steps.
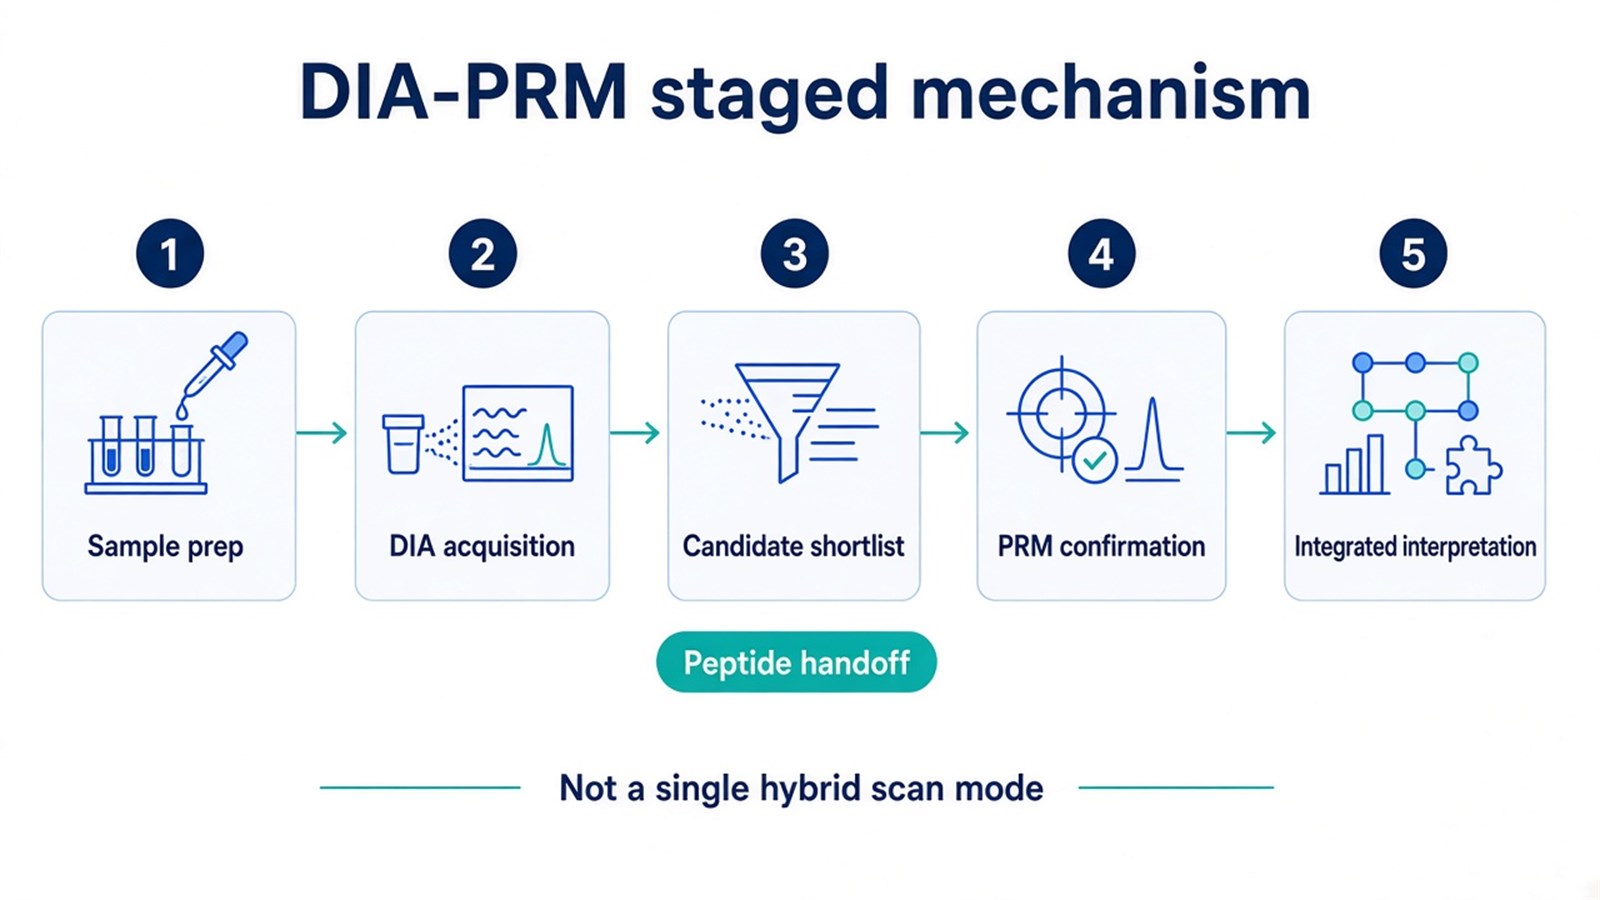
Figure 2. DIA-PRM integrates two experiments through peptide handoff; it is not a single hybrid scan mode.
- Prepare matched samples for discovery acquisition.
- Acquire DIA and build a relative discovery matrix.
- Shortlist candidates and design the PRM peptide panel.
- Confirm selected peptides by PRM under a closed inclusion list.
- Interpret discovery and confirmation claims separately, then integrate only where peptide identity and preparation are matched.
Data integration does not mean treating DIA intensities and PRM areas as interchangeable absolute values. It means checking whether the same biological conclusion survives both the open matrix and the closed remeasurement. Trade-offs of that staging decision are reviewed in Advantages and Disadvantages of DIA-PRM Proteomics.
Practical Limits of the Mechanism
Staging doubles planning load: discovery settings, contrast rules, and peptide handoff must be documented before confirmation begins. PRM multiplex is limited by chromatography and cycle time, so long candidate lists must be narrowed or split. Window interference in DIA and co-isolation inside a PRM precursor window are not erased by combining the methods. Diagnostic cutoffs remain outside ordinary research DIA or PRM reporting unless a separately validated assay framework is declared.
Keep DIA-only when open relative discovery is sufficient. Keep PRM-only when targets are already named and discovery breadth is not required.
Conclusion
The mechanism of DIA-PRM proteomics is sequential: DIA provides scheduled-window relative discovery across an open inventory, and PRM provides high-resolution fragment confirmation of a closed peptide shortlist. Reliable use depends on stating which claim belongs to which stage, matching preparation across handoff, and treating integration as confirmation of a shortlist rather than as proof that one acquisition file can be both fully open and fully targeted. Researchers evaluating that staged mechanism can contact MtoZ Biolabs to review discovery scope, panel size, and claim unit before study start, and can also consult the MtoZ Biolabs DIA-PRM Proteomics Service for staged feasibility and workflow planning.
